Related Experiment Video
Updated: May 3, 2026
Isolation, Characterization, And High Throughput Extracellular Flux Analysis of Mouse Primary Renal Tubular Epithelial Cells
Published on: June 20, 2018
X-linked, COL4A5 hypomorphic Alport mutations such as G624D and P628L may only exhibit thin basement membrane
A Pierides1, K Voskarides2, M Kkolou3
1Department of Nephrology, Hippocrateon Hospital, Nicosia, Cyprus ; Department of Biological Sciences and Molecular Medicine Research Center, University of Cyprus, Nicosia, Cyprus.

Abstract:
Alport syndrome (ATS) results from X-linked, COL4A5 mutations (85%) or from autosomal recessive homozygous or compound heterozygous COL4A3/A4 mutations (15%), associated with alternate thinning and thickening as well as splitting and lamellation of the glomerular basement membranes. In contrast, familial microhematuria with thin basement membranes is thought to result from heterozygous COL4A3/A4 mutations. This absolute separation may not always be true. Renal biopsies and molecular genetics were used to study microhematuric families in the Hellenic population we serve. The COL4A5 gene was studied by PCR and direct re-sequencing for new mutations, while PCR-RFLP was used to identify more carriers of known COL4A5 and COL4A3/A4 mutations. Molecular genetics in two undiagnosed microhematuric Cypriot families, revealed COL4A5 mutation P628L indicating X-linked ATS. Of nine males, seven developed end stage kidney disease (ESKD) between 31 and 56, while two are well at 51 and 57, exhibiting microhematuria and thin basement membrane nephropathy (TBMN). COL4A5 mutation G624D was also identified in six Greek families. Seventy five members had DNA tests and 37 proved positive. Four positive males developed ESKD at 61, 51, 50 and 39 years, while the remaining and all females showed only microhematuria. A literature search revealed eight papers with six similar hypomorphic COL4A5 mutations presenting as phenocopies of TBMN. In conclusion, X-linked COL4A5 ATS mutations produce a phenotypic spectrum with a) classical ATS with early onset ESKD, neurosensory deafness and ocular defects b) males with only ESKD and late deafness and c) males due to missense mutations, such as G624D and P628L that may only exhibit microhematuria, TBMN, mild chronic renal failure (CRF) or late onset ESKD. Consequently when investigating "benign familial hematuria" these and other similar X-linked COL4A5 mutations should also be searched for.
More Related Videos
 11:47
Using 2-Photon Microscopy to Quantify the Effects of Chronic Unilateral Ureteral Obstruction on Glomerular Processes
Published on: March 4, 2022
 08:46
Implementing Patch Clamp and Live Fluorescence Microscopy to Monitor Functional Properties of Freshly Isolated PKD Epithelium
Published on: September 1, 2015
Related Concept Videos
 01:05
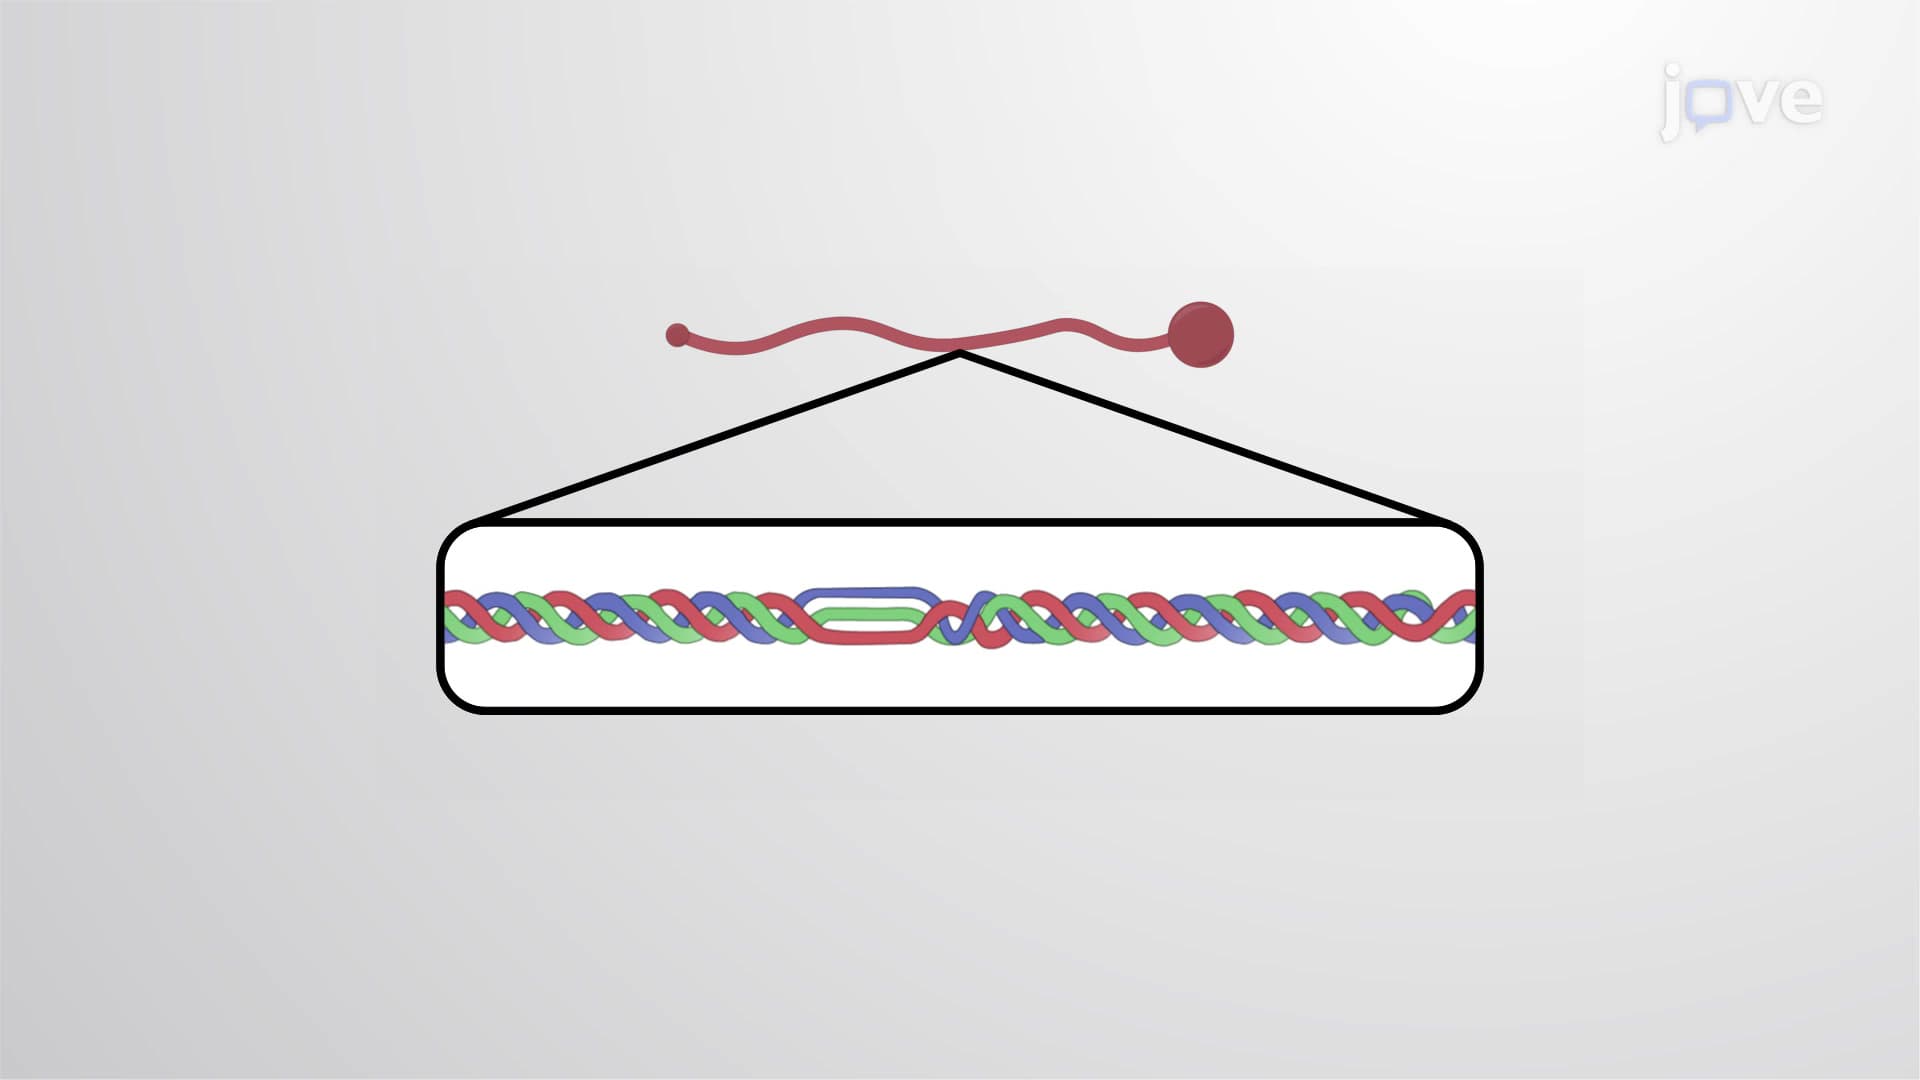
Type IV Collagen of Basal Lamina
A type IV collagen molecule has six alpha chains which can...
 01:20
Renal Corpuscle
Glomerulus: Structure and Function
The glomerulus is a tiny, intricate network of capillaries located at the beginning of the nephron. It's enveloped by the Bowman's capsule and receives its blood supply from an afferent arteriole, which divides into numerous...
 01:28
Diabetic Nephropathy
 02:41
Lethal Alleles
Lucien Cuénot discovered lethal alleles in 1905 while studying the inheritance of coat color in mice. The agouti gene is responsible for the color of the coat in mice. This gene codes for an agouti-signaling protein, which is responsible for melanin distribution in mammals. The wild-type allele gives rise to gray-brown coat color in mice, while the mutant allele gives rise to yellow coat color. In addition to coat color, the agouti gene is associated with the yellow...
 01:24
Nephrotic Syndrome I : Introduction
 01:10
Nephrons