Related Experiment Video
Updated: May 3, 2026
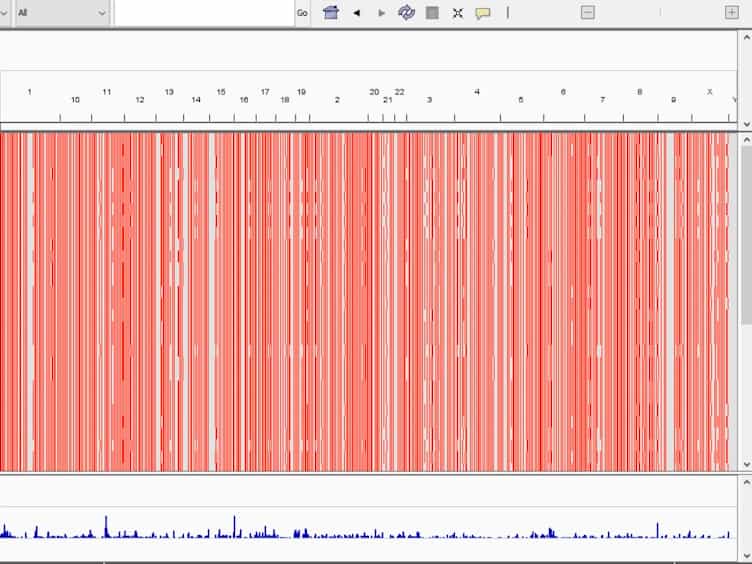
A Fast and Quantitative Method for Post-translational Modification and Variant Enabled Mapping of Peptides to Genomes
Published on: May 22, 2018
Improving read mapping using additional prefix grams
Jongik Kim, Chen Li, Xiaohui Xie1
1Department of Computer Science, University of California, Irvine, USA. xhx@ics.uci.edu.

Background:
Next-generation sequencing (NGS) enables rapid production of billions of bases at a relatively low cost. Mapping reads from next-generation sequencers to a given reference genome is an important first step in many sequencing applications. Popular read mappers, such as Bowtie and BWA, are optimized to return top one or a few candidate locations of each read. However, identifying all mapping locations of each read, instead of just one or a few, is also important in some sequencing applications such as ChIP-seq for discovering binding sites in repeat regions, and RNA-seq for transcript abundance estimation.
Results:
Here we present Hobbes2, a software package designed for fast and accurate alignment of NGS reads and specialized in identifying all mapping locations of each read. Hobbes2 efficiently identifies all mapping locations of reads using a novel technique that utilizes additional prefix q-grams to improve filtering. We extensively compare Hobbes2 with state-of-the-art read mappers, and show that Hobbes2 can be an order of magnitude faster than other read mappers while consuming less memory space and achieving similar accuracy.
Conclusions:
We propose Hobbes2 to improve the accuracy of read mapping, specialized in identifying all mapping locations of each read. Hobbes2 is implemented in C++, and the source code is freely available for download at http://hobbes.ics.uci.edu.
Related Concept Videos
 02:07
Improving Translational Accuracy
02:07Improving Translational Accuracy