Related Experiment Video
Updated: May 2, 2026

Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Prediction of hydration free energies for the SAMPL4 diverse set of compounds using molecular dynamics simulations
Oliver Beckstein1, Anaïs Fourrier, Bogdan I Iorga
1Department of Physics, Arizona State University, P.O. Box 871504, Tempe, AZ, 85287-1504, USA, oliver.beckstein@asu.edu.

Abstract:
All-atom molecular dynamics computer simulations were used to blindly predict the hydration free energies of a range of small molecules as part of the SAMPL4 challenge. Compounds were parametrized on the basis of the OPLS-AA force field using three different protocols for deriving partial charges: (1) using existing OPLS-AA atom types and charges with minor adjustments of partial charges on equivalent connecting atoms and derivation of new parameters for a number of distinct chemical groups (N-alkyl imidazole, nitrate) that were not present in the published force field; (2) calculation of quantum mechanical charges via geometry optimization, followed by electrostatic potential (ESP) fitting, using Jaguar at the LMP2/cc-pVTZ(-F) level; and (3) via geometry optimization and CHelpG charges (Gaussian09 at the HF/6-31G* level), followed by two-stage RESP fitting. The absolute hydration free energy was computed by an established protocol including alchemical free energy perturbation with thermodynamic integration. The use of standard OPLS-AA charges (protocol 1) with a number of newly parametrized charges and the use of histidine derived parameters for imidazole yielded an overall root mean square deviation of the prediction from the experimental data of 1.75 kcal/mol. The precision of our results appears to be mainly limited by relatively poor reproducibility of the Lennard-Jones contribution towards the solvation free energy, for which we observed large variability that could be traced to a strong dependence on the initial system conditions.
More Related Videos
 12:11
Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 02:42
Aqueous Solutions and Heats of Hydration
When ionic compounds dissolve in water, the ions in the solid separate and disperse uniformly throughout the solution because water molecules surround and solvate the ions, reducing the strong electrostatic forces between them. This process...
 02:27
Predicting Molecular Geometry
 02:45
Acid-Catalyzed Hydration of Alkenes
 01:20
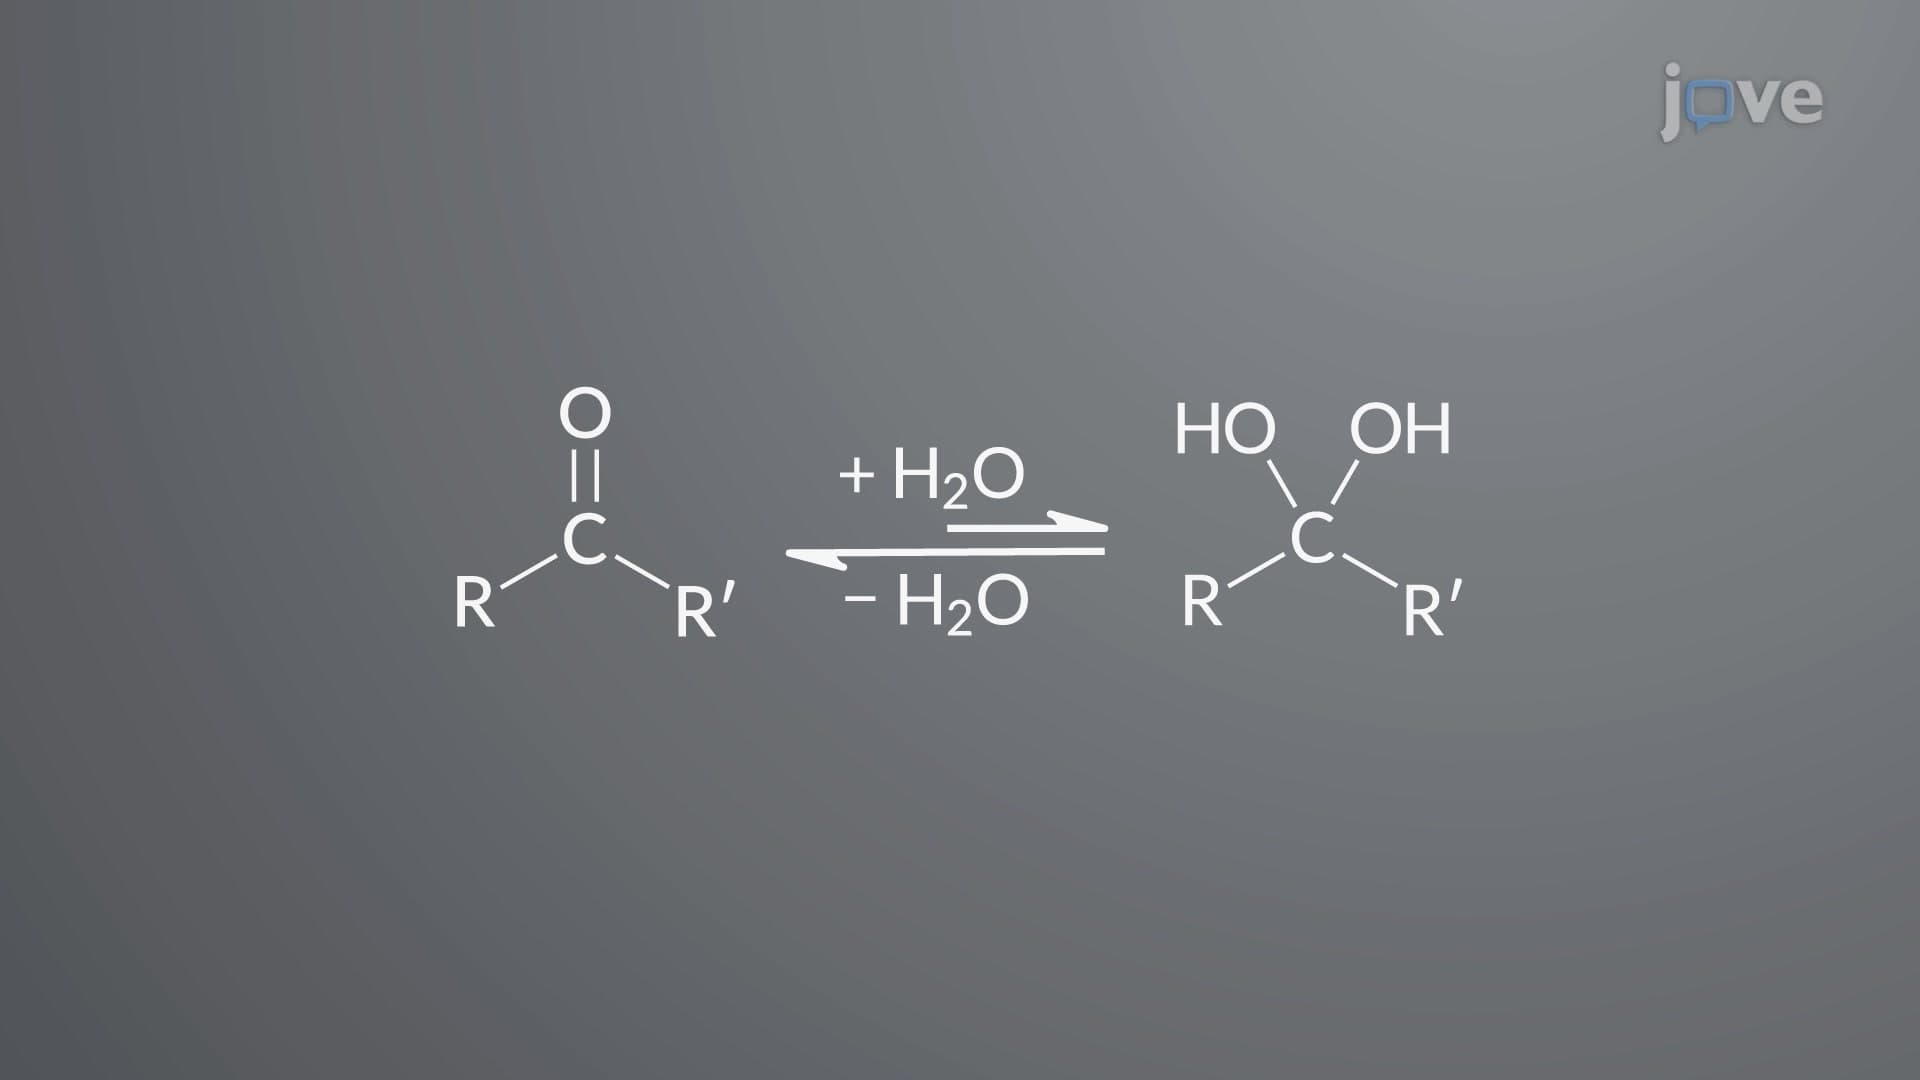
Aldehydes and Ketones with Water: Hydrate Formation
The formation of hydrates is a reversible reaction. Hydrate formation is influenced by steric and electronic factors accompanying the alkyl substituents on the carbonyl group: The rate of hydrate formation increases with a decrease in the number of alkyl groups attached to the carbonyl carbon. Hence,...
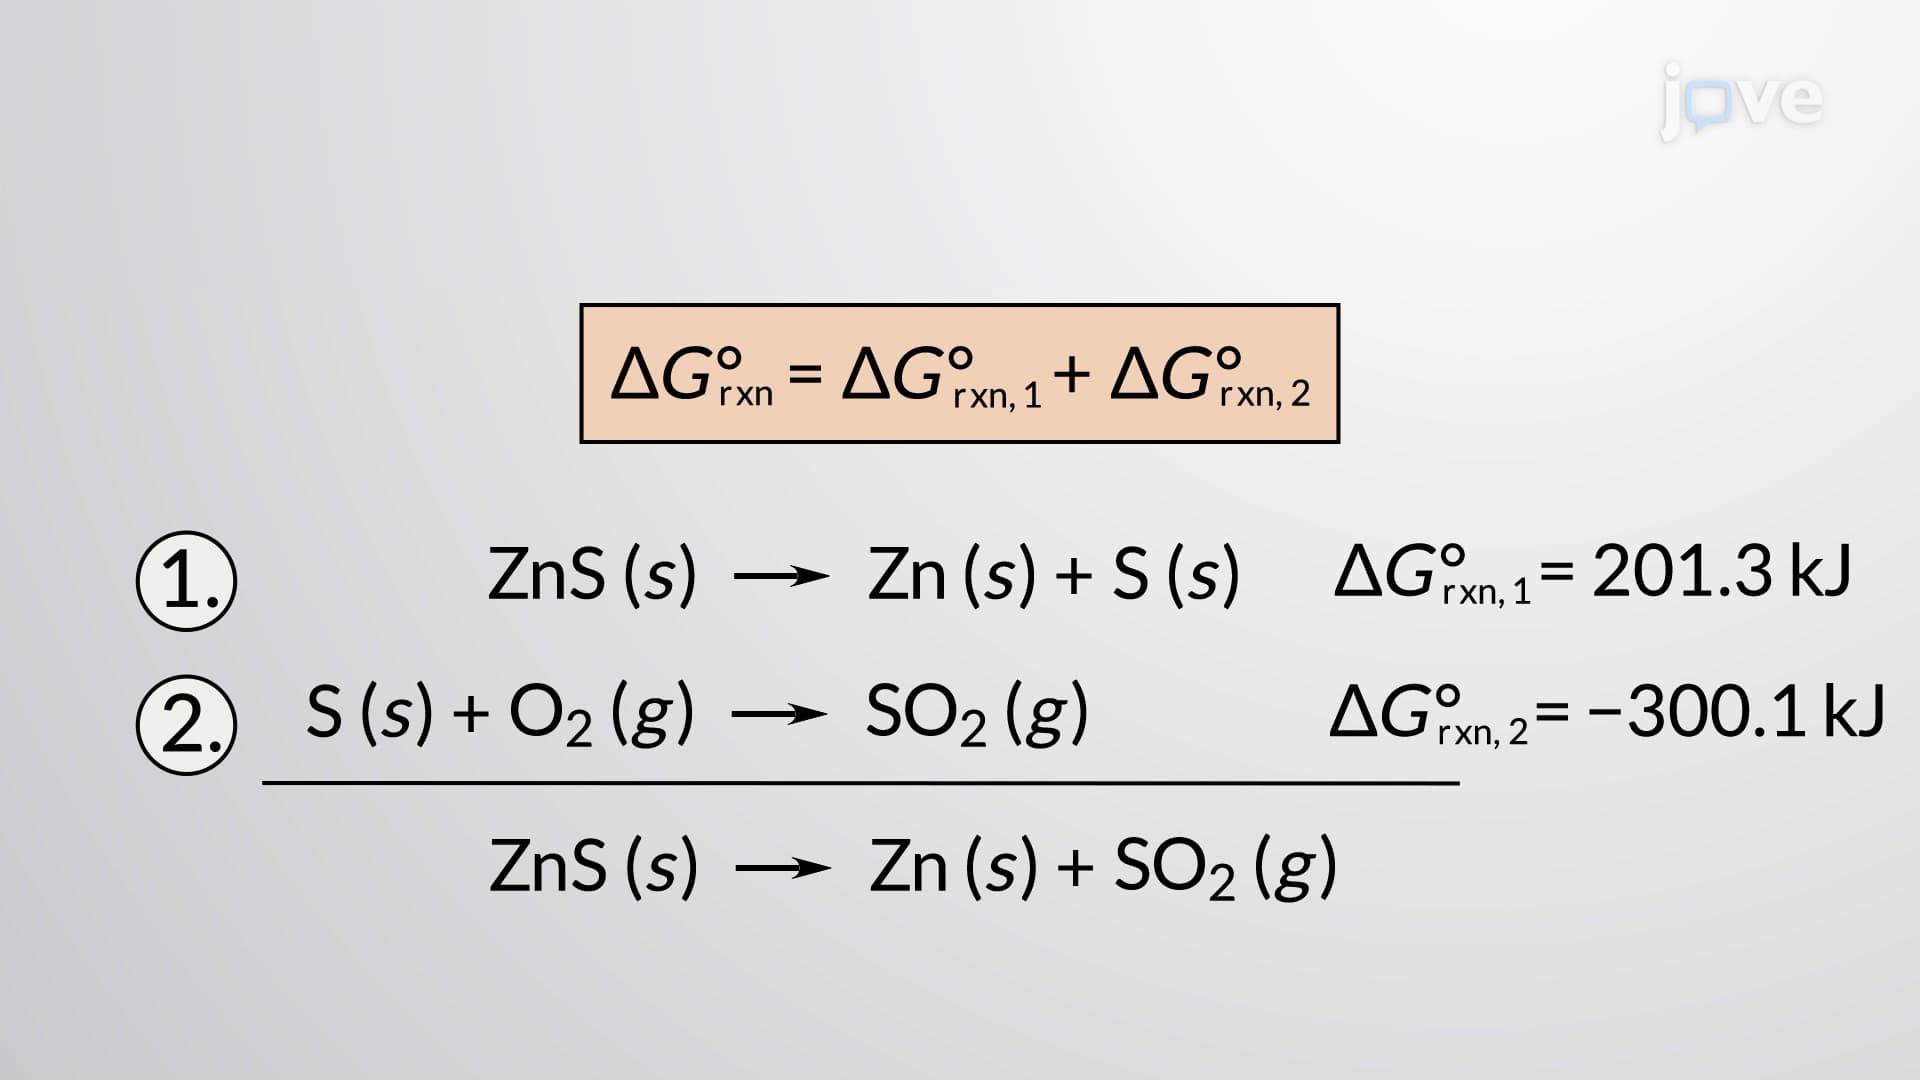 02:49
Calculating Standard Free Energy Changes
 02:34
Regioselectivity and Stereochemistry of Acid-Catalyzed Hydration