Related Experiment Video
Updated: May 2, 2026
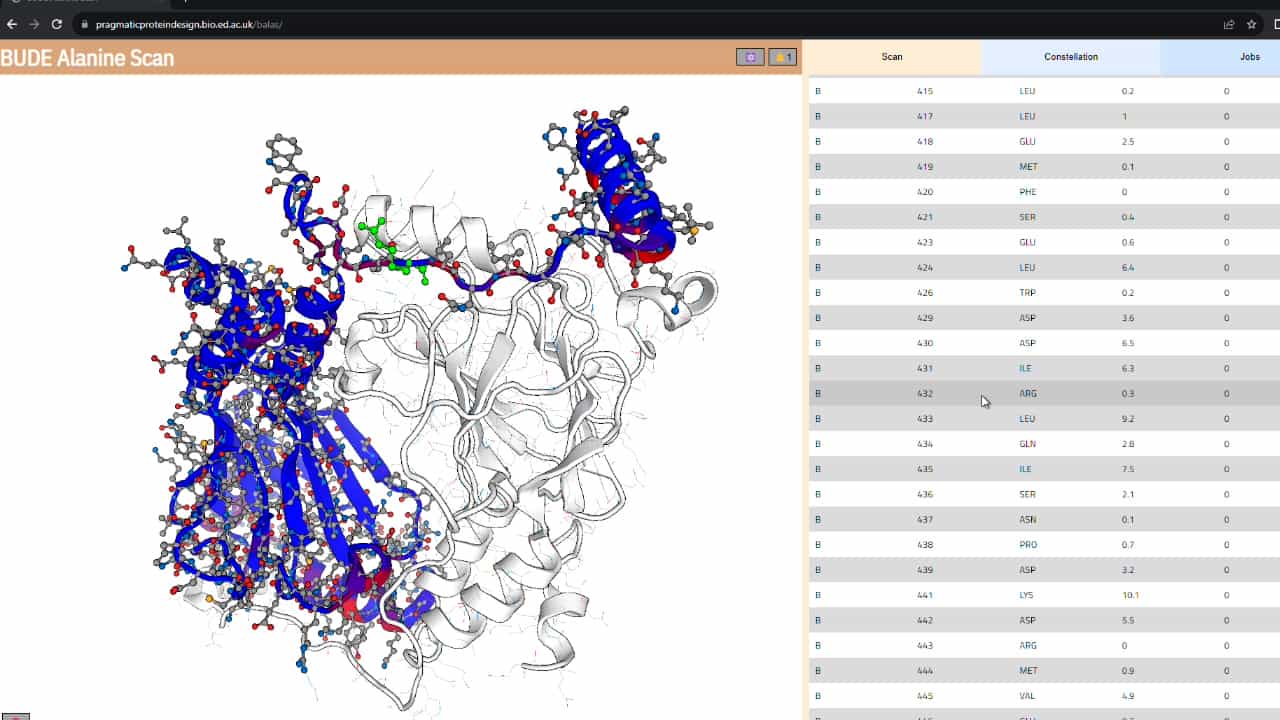
Author Spotlight: A Computational Approach to Decipher Amino Acid Preferences in Multispecific Protein-Protein Interactions
Published on: January 26, 2024
Prediction of protein interaction types based on sequence and network features

Background:
Protein interactions mediate a wide spectrum of functions in various cellular contexts. Functional versatility of protein complexes is due to a broad range of structural adaptations that determine their binding affinity, the number of interaction sites, and the lifetime. In terms of stability it has become customary to distinguish between obligate and non-obligate interactions dependent on whether or not the protomers can exist independently. In terms of spatio-temporal control protein interactions can be either simultaneously possible (SP) or mutually exclusive (ME). In the former case a network hub interacts with several proteins at the same time, offering each of them a separate interface, while in the latter case the hub interacts with its partners one at a time via the same binding site. So far different types of interactions were distinguished based on the properties of the corresponding binding interfaces derived from known three-dimensional structures of protein complexes.
Results:
Here we present PiType, an accurate 3D structure-independent computational method for classifying protein interactions into simultaneously possible (SP) and mutually exclusive (ME) as well as into obligate and non-obligate. Our classifier exploits features of the binding partners predicted from amino acid sequence, their functional similarity, and network topology. We find that the constituents of non-obligate complexes possess a higher degree of structural disorder, more short linear motifs, and lower functional similarity compared to obligate interaction partners while SP and ME interactions are characterized by significant differences in network topology. Each interaction type is associated with a distinct set of biological functions. Moreover, interactions within multi-protein complexes tend to be enriched in one type of interactions.
Conclusion:
PiType does not rely on atomic structures and is thus suitable for characterizing proteome-wide interaction datasets. It can also be used to identify sub-modules within protein complexes. PiType is available for download as a self-installing package from http://webclu.bio.wzw.tum.de/PiType/PiType.zip.
Related Concept Videos
 02:26
Protein Networks
These interactions can be represented through maps depicting protein-protein interaction networks, represented as nodes and edges. Nodes are circles that are representative of a protein,...
02:26Protein Networks
 02:04
Protein-protein Interfaces
02:04Protein-Protein Interfaces
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 02:47
Protein Families