Related Experiment Video
Updated: May 2, 2026

Ethylene Polymerizations Using Parallel Pressure Reactors and a Kinetic Analysis of Chain Transfer Polymerization
Published on: November 27, 2015
Vibrational quasi-degenerate perturbation theory with optimized coordinates: applications to ethylene and
1Theoretical Molecular Science Laboratory, RIKEN, 2-1 Hirosawa, Wako, Saitama 351-0198, Japan.

Abstract:
A perturbative extension to optimized coordinate vibrational self-consistent field (oc-VSCF) is proposed based on the quasi-degenerate perturbation theory (QDPT). A scheme to construct the degenerate space (P space) is developed, which incorporates degenerate configurations and alleviates the divergence of perturbative expansion due to localized coordinates in oc-VSCF (e.g., local O-H stretching modes of water). An efficient configuration selection scheme is also implemented, which screens out the Hamiltonian matrix element between the P space configuration (p) and the complementary Q space configuration (q) based on a difference in their quantum numbers (λpq = ∑s|ps - qs|). It is demonstrated that the second-order vibrational QDPT based on optimized coordinates (oc-VQDPT2) smoothly converges with respect to the order of the mode coupling, and outperforms the conventional one based on normal coordinates. Furthermore, an improved, fast algorithm is developed for optimizing the coordinates. First, the minimization of the VSCF energy is conducted in a restricted parameter space, in which only a portion of pairs of coordinates is selectively transformed. A rational index is devised for this purpose, which identifies the important coordinate pairs to mix from others that may remain unchanged based on the magnitude of harmonic coupling induced by the transformation. Second, a cubic force field (CFF) is employed in place of a quartic force field, which bypasses intensive procedures that arise due to the presence of the fourth-order force constants. It is found that oc-VSCF based on CFF together with the pair selection scheme yields the coordinates similar in character to the conventional ones such that the final vibrational energy is affected very little while gaining an order of magnitude acceleration. The proposed method is applied to ethylene and trans-1,3-butadiene. An accurate, multi-resolution potential, which combines the MP2 and coupled-cluster with singles, doubles, and perturbative triples level of electronic structure theory, is generated and employed in the oc-VQDPT2 calculation to obtain the fundamental tones as well as selected overtones/combination tones coupled to the fundamentals through the Fermi resonance. The calculated frequencies of ethylene and trans-1,3-butadiene are found to be in excellent agreement with the experimental values with a mean absolute error of 8 and 9 cm(-1), respectively.
More Related Videos

Related Concept Videos
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the...
 01:28
Stability of Conjugated Dienes
A comparison of the enthalpies of hydrogenation of dienes reveals that conjugated dienes release less heat on hydrogenation, rendering them more stable than their nonconjugated analogs.
 02:18
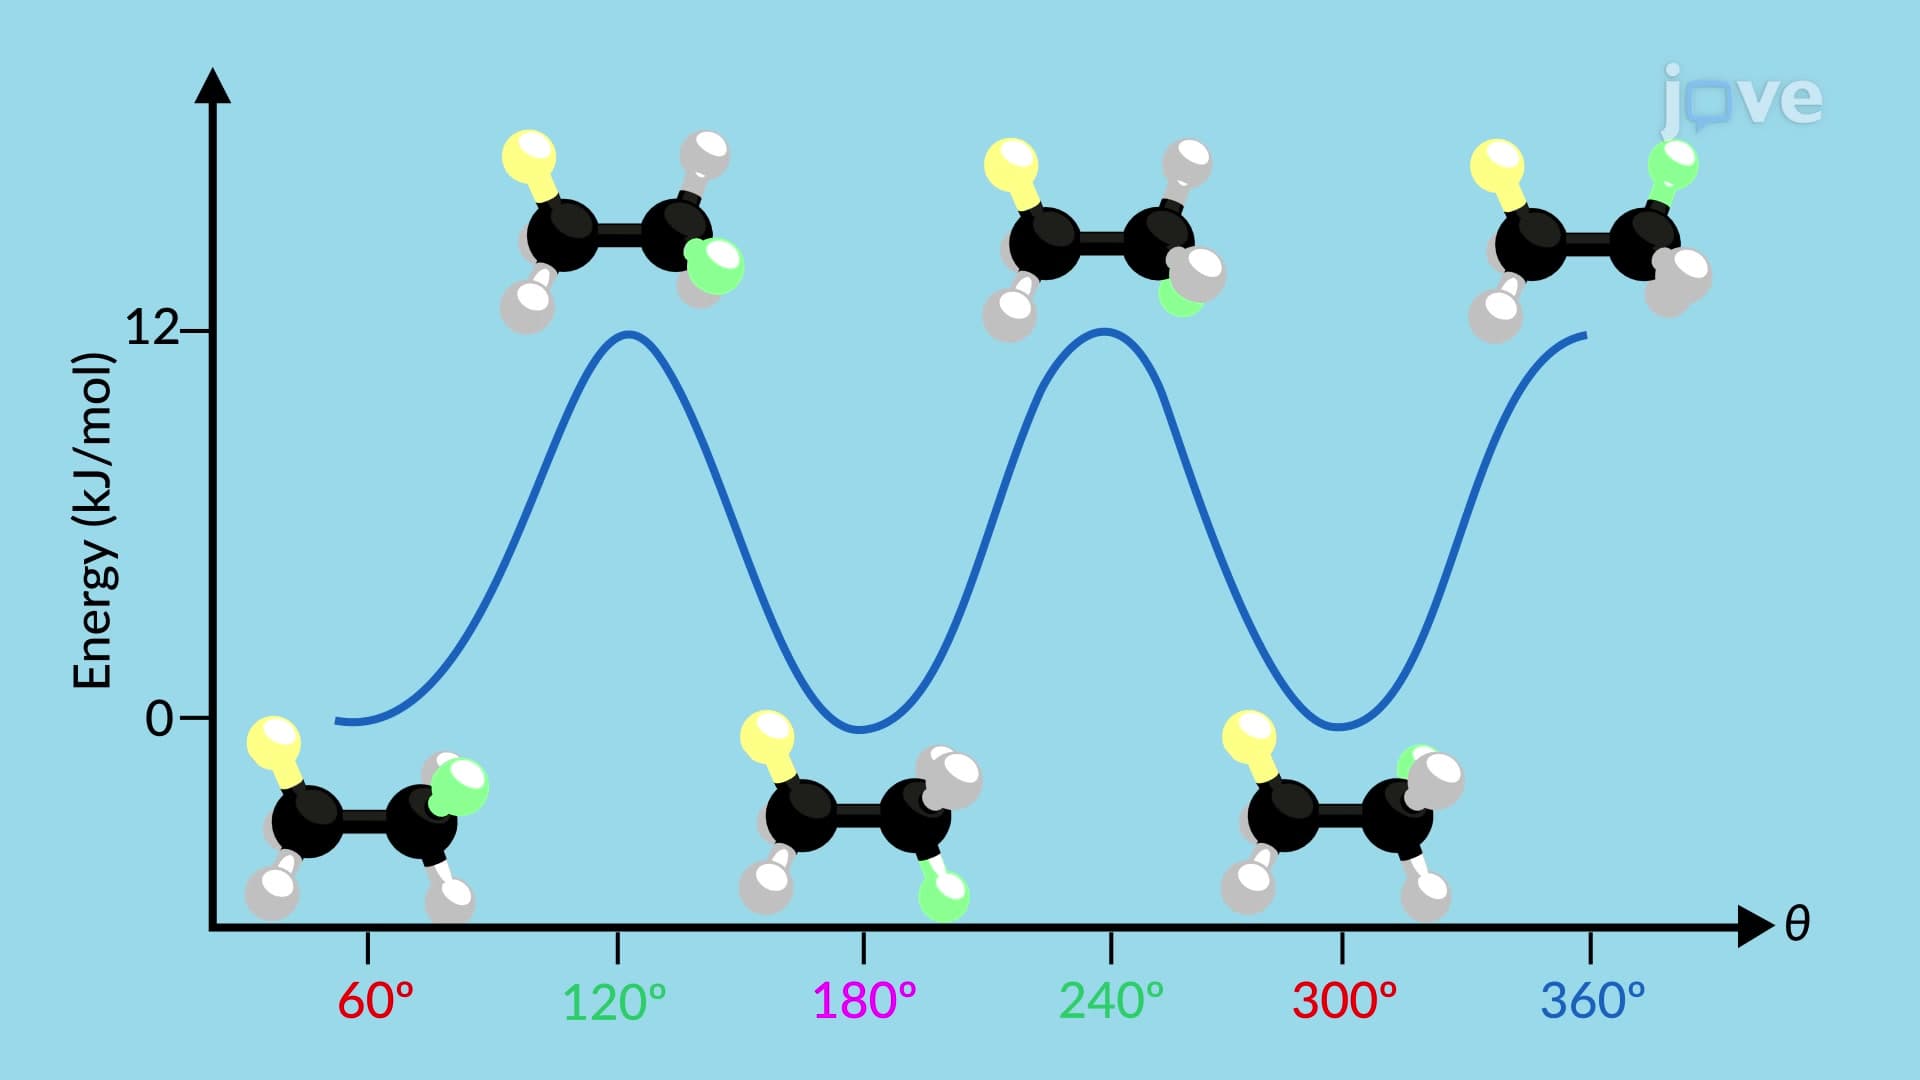
Conformations of Ethane and Propane
Staggered conformation is a low energy and more stable conformation with the C-H bonds on the front carbon placed at 60°dihedral angles relative to the C-H bonds on the back carbon, leading to a reduced torsional strain. In staggered...
 01:20
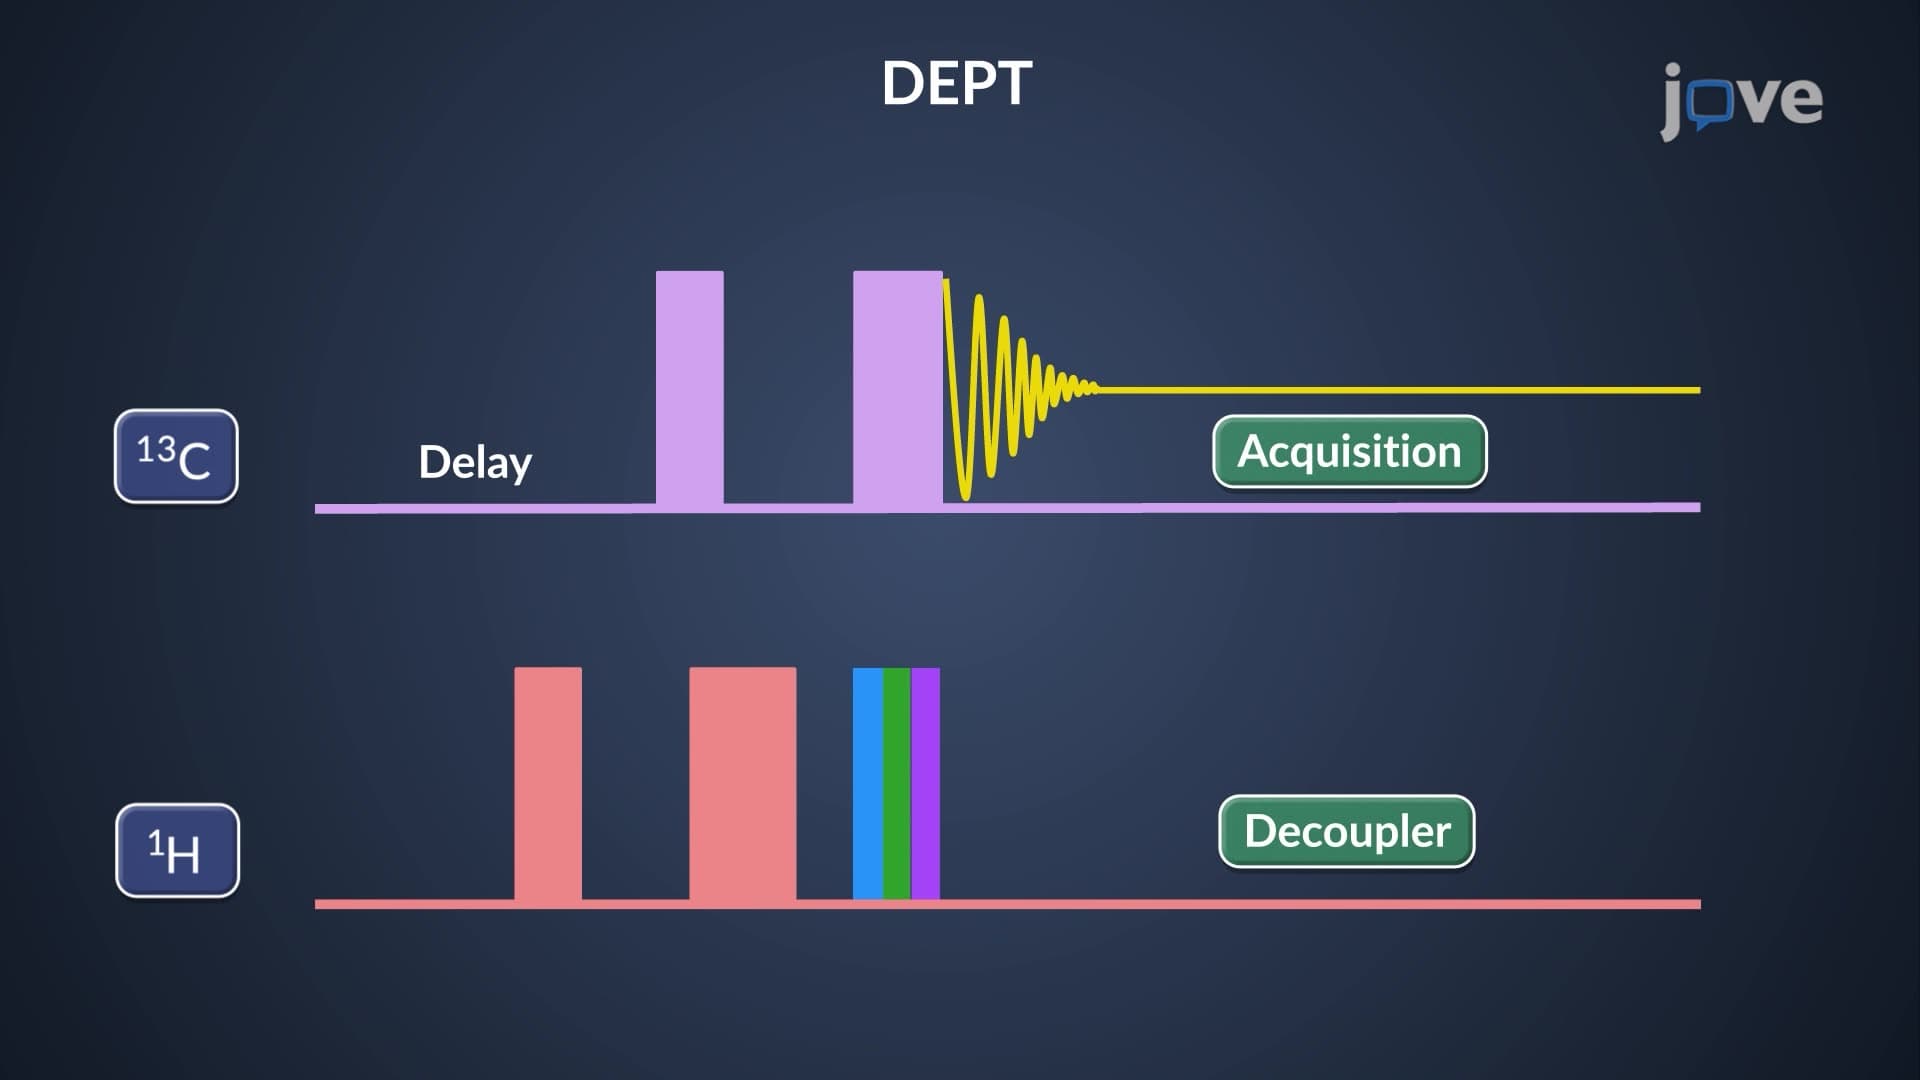
¹³C NMR: Distortionless Enhancement by Polarization Transfer (DEPT)
 01:17
Electrophilic 1,2- and 1,4-Addition of HX to 1,3-Butadiene
 01:16
Structure of Conjugated Dienes
Conjugated dienes are compounds characterized by the presence of alternating double and single bonds. In a conjugated system like 1,3-butadiene, the unhybridized 2p orbital on each carbon overlaps continuously, allowing the π electrons to be delocalized across the entire molecule. In contrast, this type of overlap does not occur in cumulated and isolated dienes, such as 2,3-pentadiene and 1,4-pentadiene, respectively. Instead, the π electrons remain localized between the...