Differential expression analysis on RNA-Seq count data based on penalized matrix decomposition
Related Experiment Video
Updated: May 2, 2026
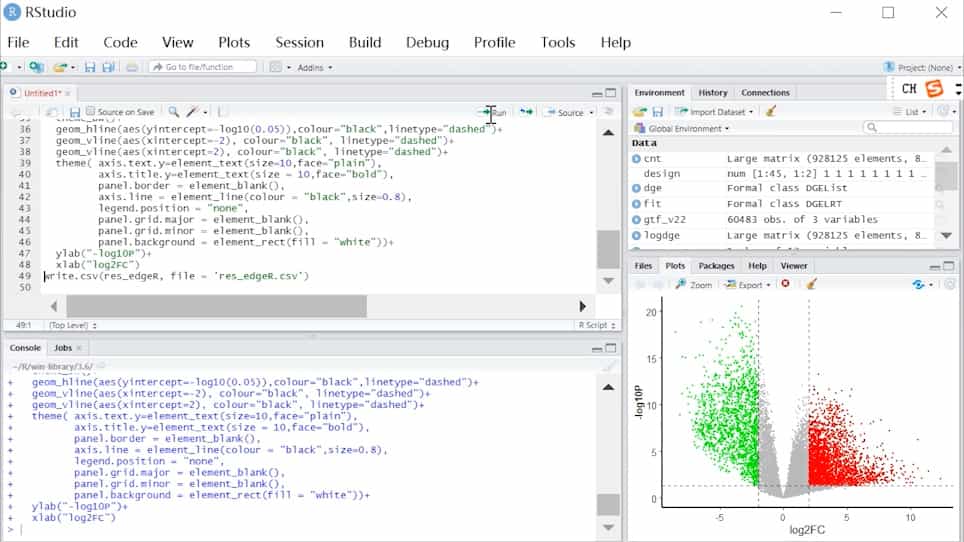
Three Differential Expression Analysis Methods for RNA Sequencing: limma, EdgeR, DESeq2
Published on: September 18, 2021

Abstract:
With the development of deep sequencing, vast amounts of RNA-Seq data have been generated. It is crucial how to extract and interpret the meaningful information contained in deep sequencing data. In this paper, based on penalized matrix decomposition (PMD), a novel method, named PMDSeq, was proposed to analyze RNA-seq count data. Firstly, to obtain the differential expression matrix, the matrix of RNA-seq count data was normalized. Secondly, the differential expression matrix was decomposed into three factor matrices. By imposing appropriate constraint on factor matrices, the PMDSeq method can highlight the differentially expressed genes. Thirdly, the proposed method can identify the differentially expressed genes based on the scaled eigensamples. Finally, we used gene ontology tools to check these differentially expressed genes. The experimental results on simulation and three real RNA-seq count data sets demonstrated the effectiveness of our method.
Related Concept Videos
 03:21
RNA-seq
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...
 02:24
Ribosome Profiling
Applications of ribosome profiling
Ribosome profiling has many applications, including in vivo monitoring of translation inside a particular organ or tissue type and quantifying new protein synthesis levels.
The technique...