Related Experiment Video
Updated: May 2, 2026

Methods for the Discovery of Novel Compounds Modulating a Gamma-Aminobutyric Acid Receptor Type A Neurotransmission
Published on: August 16, 2018
3D-QSAR/CoMFA-based structure-affinity/selectivity relationships of aminoalkylindoles in the cannabinoid CB1 and CB2
Jaime Mella-Raipán1, Santiago Hernández-Pino2, César Morales-Verdejo3
1Departamento de Química y Bioquímica, Facultad de Ciencias, Universidad de Valparaíso, Av. Gran Bretaña 1111, Playa Ancha, Valparaíso, Casilla 5030, Chile. jaime.mella@uv.cl.

Abstract:
A 3D-QSAR (CoMFA) study was performed in an extensive series of aminoalkylindoles derivatives with affinity for the cannabinoid receptors CB1 and CB2. The aim of the present work was to obtain structure-activity relationships of the aminoalkylindole family in order to explain the affinity and selectivity of the molecules for these receptors. Major differences in both, steric and electrostatic fields were found in the CB1 and CB2 CoMFA models. The steric field accounts for the principal contribution to biological activity. These results provide a foundation for the future development of new heterocyclic compounds with high affinity and selectivity for the cannabinoid receptors with applications in several pathological conditions such as pain treatment, cancer, obesity and immune disorders, among others.
More Related Videos
 09:09
Preparation and Delivery of Protein Microcrystals in Lipidic Cubic Phase for Serial Femtosecond Crystallography
Published on: September 20, 2016
 14:34
A Bilingual Computational Workflow for Identifying Potential PLK1 Inhibitors in American Sign Language and English
Published on: April 3, 2026
Related Concept Videos
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
 01:16
Adrenergic Agonists: Chemistry and Structure-Activity Relationship
Aromatic ring substitutions: Substituting the aromatic ring with –OH groups at positions 3 and 4 yields catecholamines (e.g., epinephrine), which have a high affinity for adrenoceptors. Hydrogen bonding between –OH groups and receptors enhances adrenergic activity.
Separation of...
 01:30
Quantitative Aspects of Drug-Receptor Interaction
 01:25
Drug-Receptor Bonds
In...
 02:37
Disubstituted Cyclohexanes: cis-trans Isomerism
In cyclohexane, the substituents can occupy different positions generating distinct isomers....
 02:29
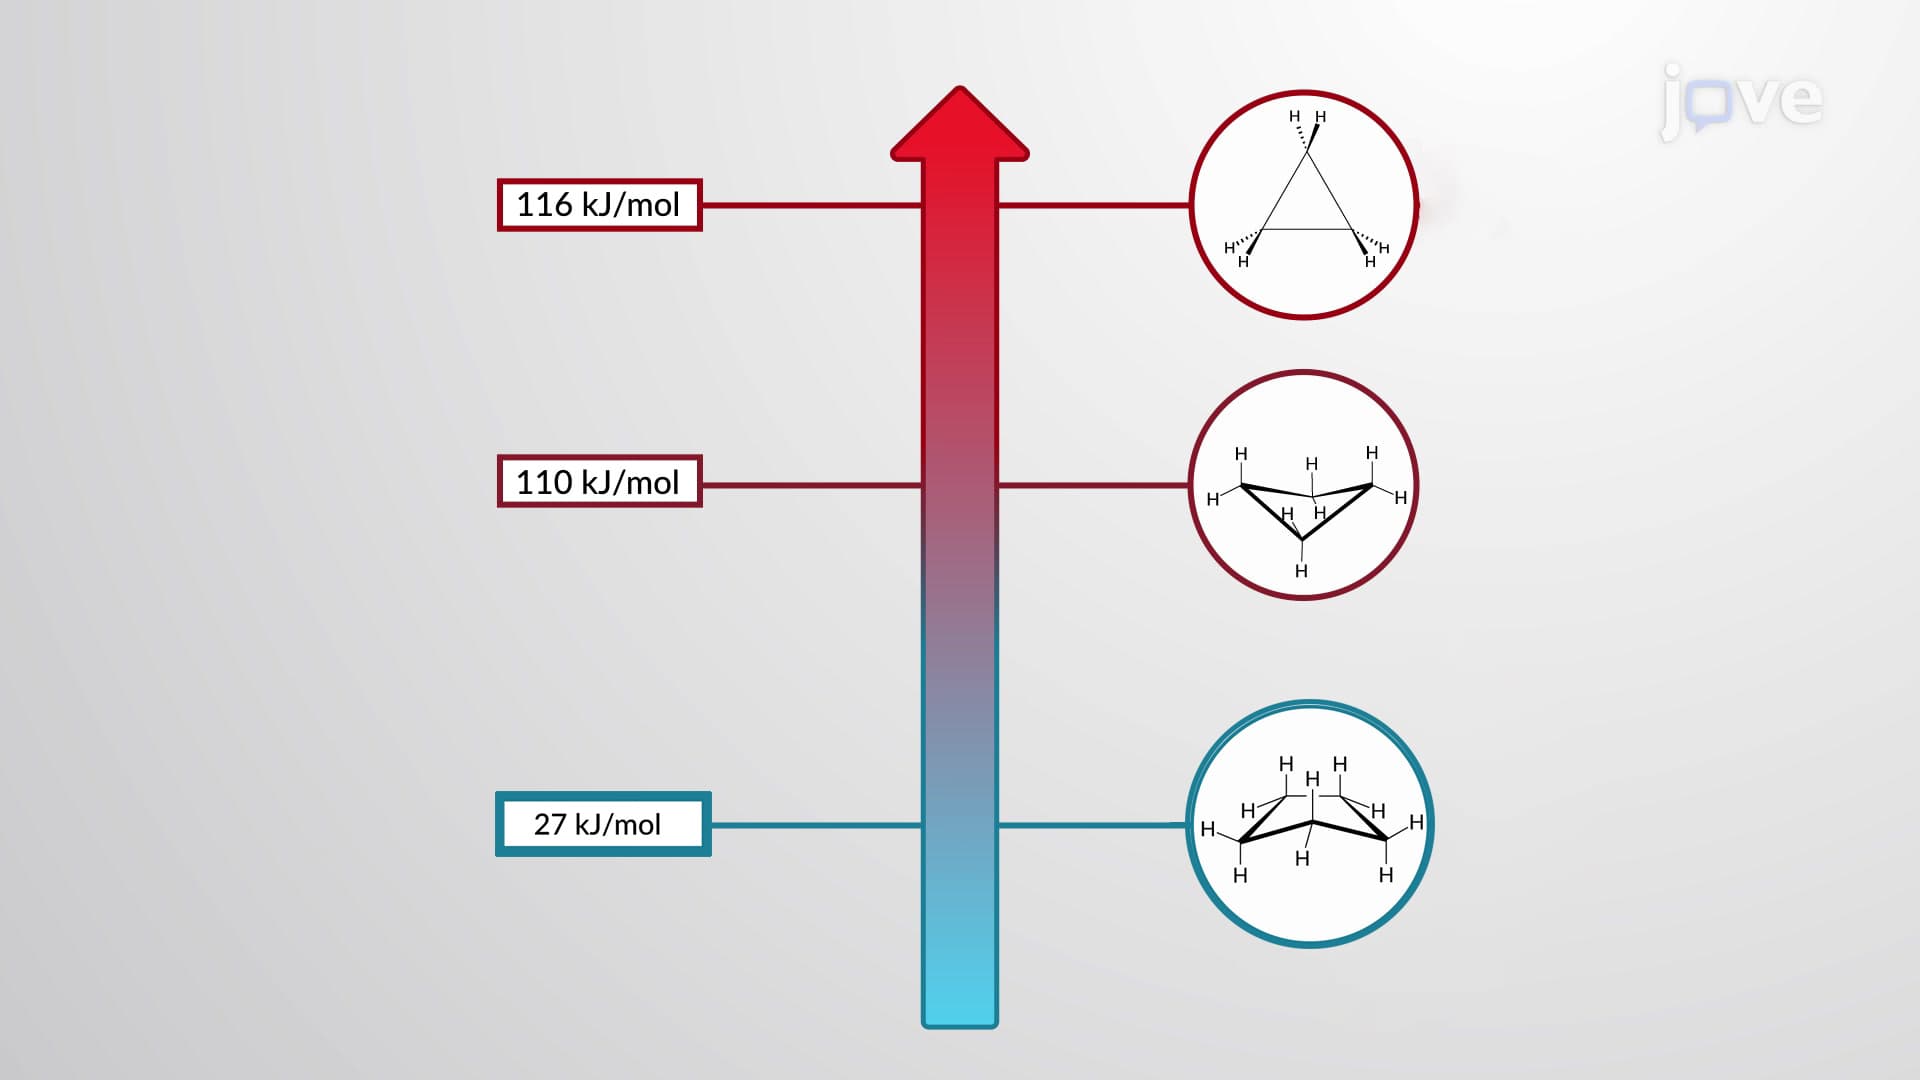
Conformations of Cycloalkanes