Related Experiment Video
Updated: May 2, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Electronic couplings for molecular charge transfer: benchmarking CDFT, FODFT, and FODFTB against high-level ab initio
Adam Kubas1, Felix Hoffmann1, Alexander Heck2
1Department of Physics and Astronomy, University College London, Gower Street, London WC1E 6BT, United Kingdom.

Abstract:
We introduce a database (HAB11) of electronic coupling matrix elements (H(ab)) for electron transfer in 11 π-conjugated organic homo-dimer cations. High-level ab inito calculations at the multireference configuration interaction MRCI+Q level of theory, n-electron valence state perturbation theory NEVPT2, and (spin-component scaled) approximate coupled cluster model (SCS)-CC2 are reported for this database to assess the performance of three DFT methods of decreasing computational cost, including constrained density functional theory (CDFT), fragment-orbital DFT (FODFT), and self-consistent charge density functional tight-binding (FODFTB). We find that the CDFT approach in combination with a modified PBE functional containing 50% Hartree-Fock exchange gives best results for absolute H(ab) values (mean relative unsigned error = 5.3%) and exponential distance decay constants β (4.3%). CDFT in combination with pure PBE overestimates couplings by 38.7% due to a too diffuse excess charge distribution, whereas the economic FODFT and highly cost-effective FODFTB methods underestimate couplings by 37.6% and 42.4%, respectively, due to neglect of interaction between donor and acceptor. The errors are systematic, however, and can be significantly reduced by applying a uniform scaling factor for each method. Applications to dimers outside the database, specifically rotated thiophene dimers and larger acenes up to pentacene, suggests that the same scaling procedure significantly improves the FODFT and FODFTB results for larger π-conjugated systems relevant to organic semiconductors and DNA.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Related Concept Videos
 02:28
Electronic Structure of Atoms
An atom comprises protons and neutrons, which are contained inside the dense, central core called the nucleus, with electrons present around the nucleus. Taking into account the wave–particle duality of electrons and the uncertainty in position around the nucleus, quantum mechanics provides a more accurate model for the atomic structure. It describes atomic orbitals as the regions around the nucleus where electrons of discrete energy exist, characterized by four quantum...
 01:22
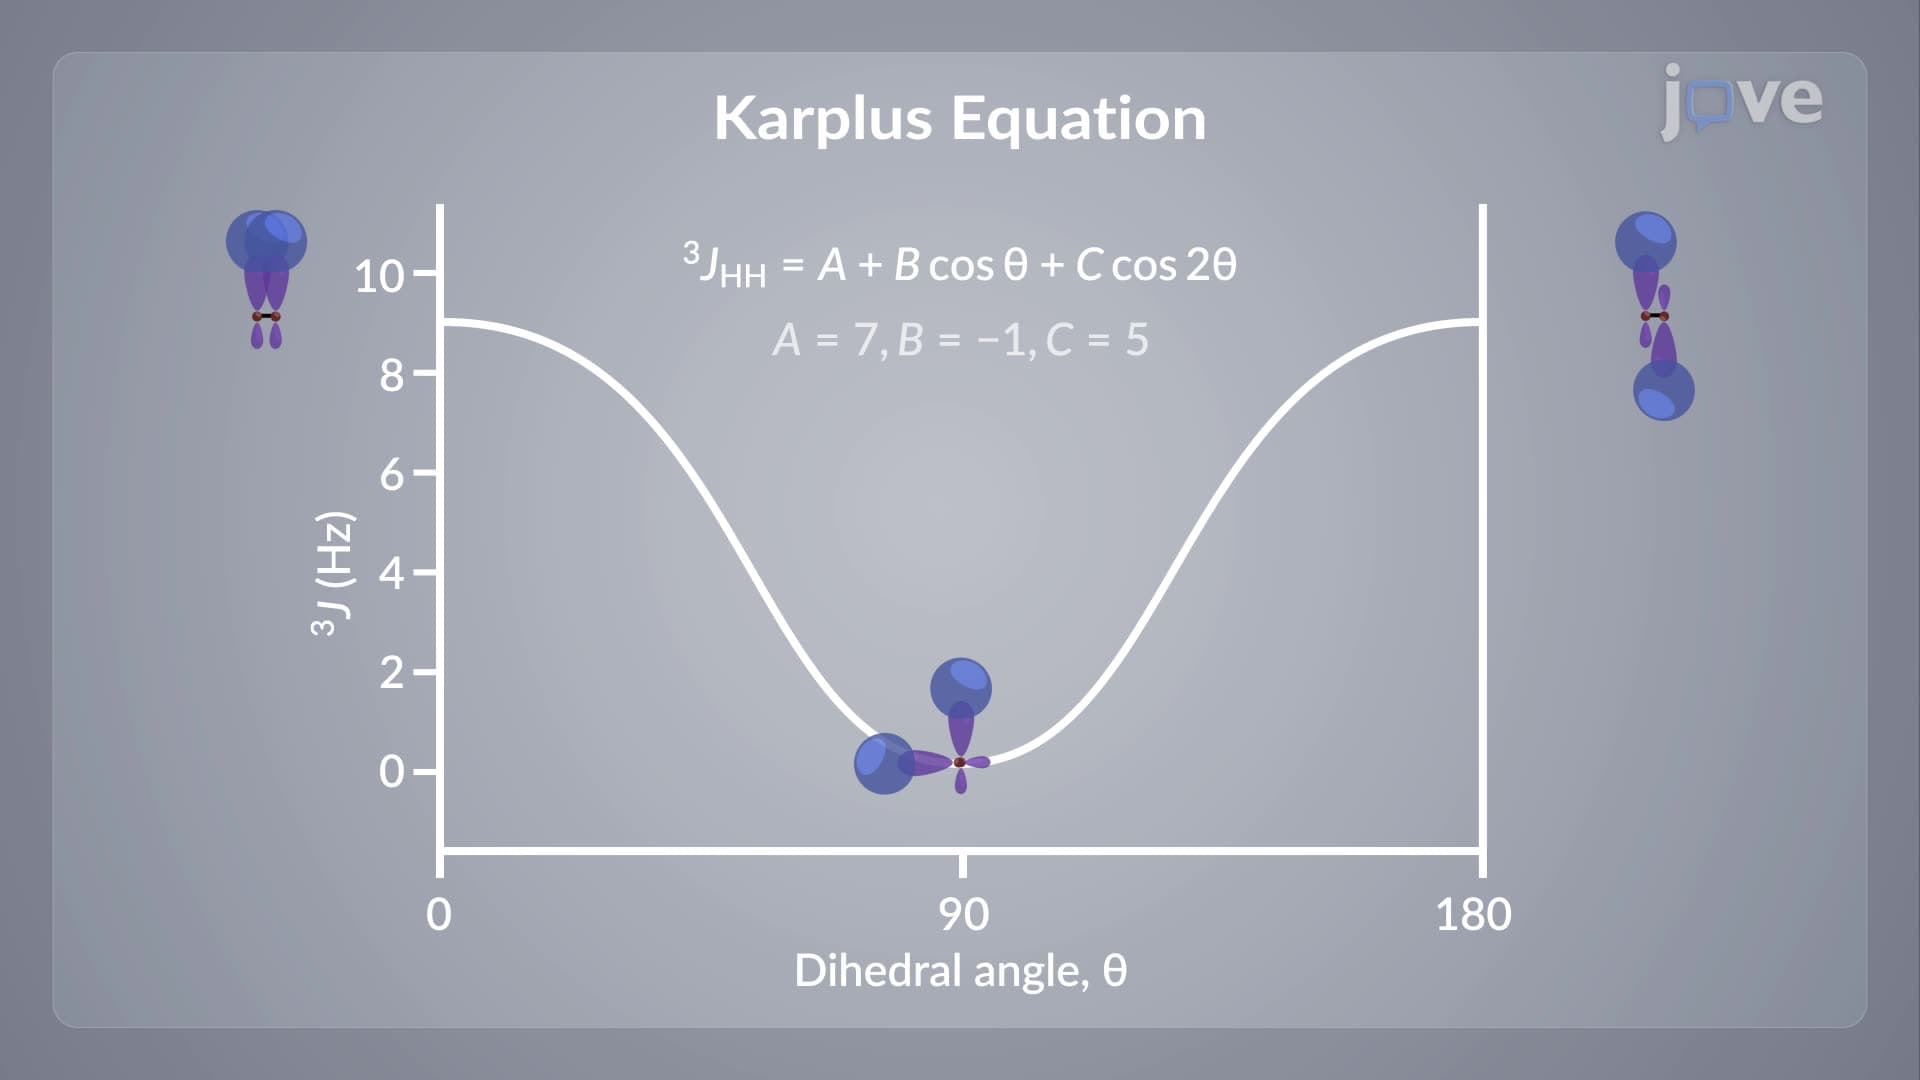
Spin–Spin Coupling: Three-Bond Coupling (Vicinal Coupling)
The extent of coupling depends on the C‑C bond length, the two H‑C‑C angles, any electron-withdrawing substituents, and the dihedral angle between the...
 01:27
π Electron Effects on Chemical Shift: Overview
 01:17
Chemical Shift: Internal References and Solvent Effects
The internal reference compound generally used in NMR spectroscopy is tetramethylsilane (TMS). TMS is preferred because it is chemically inert, soluble in NMR solvents, and easily removable. Also, the highly shielded methyl protons in TMS yield an intense...
 02:36
Molecular Geometry and Dipole Moments
 02:40
Chemical Bonds
Atoms participate in a chemical bond formation to acquire a completed valence-shell electron configuration similar to that of the noble gas nearest to it in atomic number. Ionic, covalent, and metallic bonds are some of the important types of chemical bonds. Bond energy and bond length determine the strength of a chemical bond.
Types of Chemical Bonds
An ionic bond is formed due to electrostatic attraction between cations and anions. Often, the ions are formed by the transfer of electrons...