Related Experiment Video
Updated: May 2, 2026
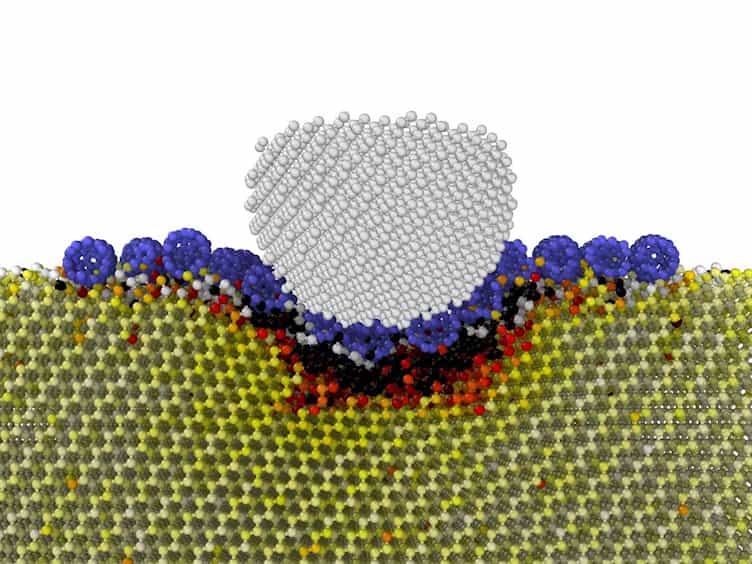

Probing C84-embedded Si Substrate Using Scanning Probe Microscopy and Molecular Dynamics
Published on: September 28, 2016
Molecular dynamics simulation of diffusion coefficients and structural properties of some alkylbenzenes in
Jinyang Wang1, Haimin Zhong1, Huajie Feng2
1KLGHEI of Environment and Energy Chemistry, School of Chemistry and Chemical Engineering, Sun Yat-sen University, Guangzhou 510275, People's Republic of China.

Abstract:
The binary infinite dilute diffusion coefficients, D₁₂(∞), of some alkylbenzenes (Ph-C(n), from Ph-H to Ph-C12) from 313 K to 333 K at 15 MPa in supercritical carbon dioxide (scCO2) have been studied by molecular dynamics (MD) simulation. The MD values agree well with the experimental ones, which indicate MD simulation technique is a powerful way to predict and obtain diffusion coefficients of solutes in supercritical fluids. Besides, the local structures of Ph-C(n)/CO2 fluids are further investigated by calculating radial distribution functions and coordination numbers. It qualitatively convinces that the first solvation shell of Ph-C(n) in scCO2 is significantly influenced by the structure of Ph-C(n) solute. Meanwhile, the mean end-to-end distance, the mean radius of gyration and dihedral angle distribution are calculated to gain an insight into the structural properties of Ph-C(n) in scCO2. The abnormal trends of radial distribution functions and coordination numbers can be reasonably explained in term of molecular flexibility. Moreover, the computed results of dihedral angle clarify that flexibility of long-chain Ph-C(n) is the result of internal rotation of C-C single bond (σ(c-c)) in alkyl chain. It is interesting that compared with n-alkane, because of the existence of benzene ring, the flexibility of alkyl chain in Ph-C(n) with same carbon atom number is significantly reduced, as a result, the carbon chain dependence of diffusion behaviors for long-chain n-alkane (n ≥ 5) and long-chain Ph-C(n) (n ≥ 4) in scCO2 are different.
More Related Videos
 06:37
Analyzing Melts and Fluids from Ab Initio Molecular Dynamics Simulations with the UMD Package
Published on: September 17, 2021
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Related Concept Videos
 03:48
Behavior of Gas Molecules: Molecular Diffusion, Mean Free Path, and Effusion
 01:18
Directing and Steric Effects in Disubstituted Benzene Derivatives
 02:33
Physical Properties of Alkanes
 01:27
Distribution of Molecular Speeds
 01:28
Reduction of Benzene to Cyclohexane: Catalytic Hydrogenation
 01:28
Debye–Huckel–Onsager Conductance Equation