Theoretical investigation on the crystal structures and electron transport properties of several nitrogen-rich
Caibin Zhao1, Hongguang Ge, Shiwei Yin
1Key Laboratory for Macromolecular Science of Shaanxi Province, School of Chemistry and Chemical Engineering, Shaanxi Normal University, Xi'an, Shaanxi, 710062, China.

Abstract:
Exploring and synthesizing new simple n-channel organic semiconductor materials with large electron mobility and high air stability have remained a major challenge and hot issue in the field of organic electronics. In the current work, the electron transport properties of four novel nitrogen-rich pentacene derivatives (PBD1, PBD2, PBD3, and PBD4) with two cyano groups as potential n-channel OFET materials have been investigated at the molecular and crystal levels by means of density functional theory (DFT) calculations coupled with the prediction of crystal structures and the incoherent charge-hopping model. Calculations reveal that the studied compounds, which possess low-lying frontier molecular energy levels, large ionization potentials and electron affinities, are very stable exposed to air. Based on predicted crystal structures, the average electron mobility at room temperature (T = 300 K) for PBD1, PBD2, PBD3, and PBD4 is predicted to be as high as 0.950, 0.558, 0.518, and 1.052 cm²·V⁻¹·s⁻¹, which indicate that these four compounds are more than likely to be promising candidates as n-type OFET materials under favorable device conditions. However, this claim needs experimental verification. In addition, the angular-dependent simulation for electron mobility shows that the electron transport is remarkably anisotropic in these molecular crystals and the maximum μ(e) appears along the crystal axis direction since molecules along this direction exhibit the close face-to-face stacking arrangement with short interplanar distances (~3.6-4.0 Å), which induces large electronic couplings.
More Related Videos
 06:16
Monitoring the Effects of Illumination on the Structure of Conjugated Polymer Gels Using Neutron Scattering
Published on: December 21, 2017
 08:43
Effect of Bending on the Electrical Characteristics of Flexible Organic Single Crystal-based Field-effect Transistors
Published on: November 7, 2016
Related Concept Videos
 01:18
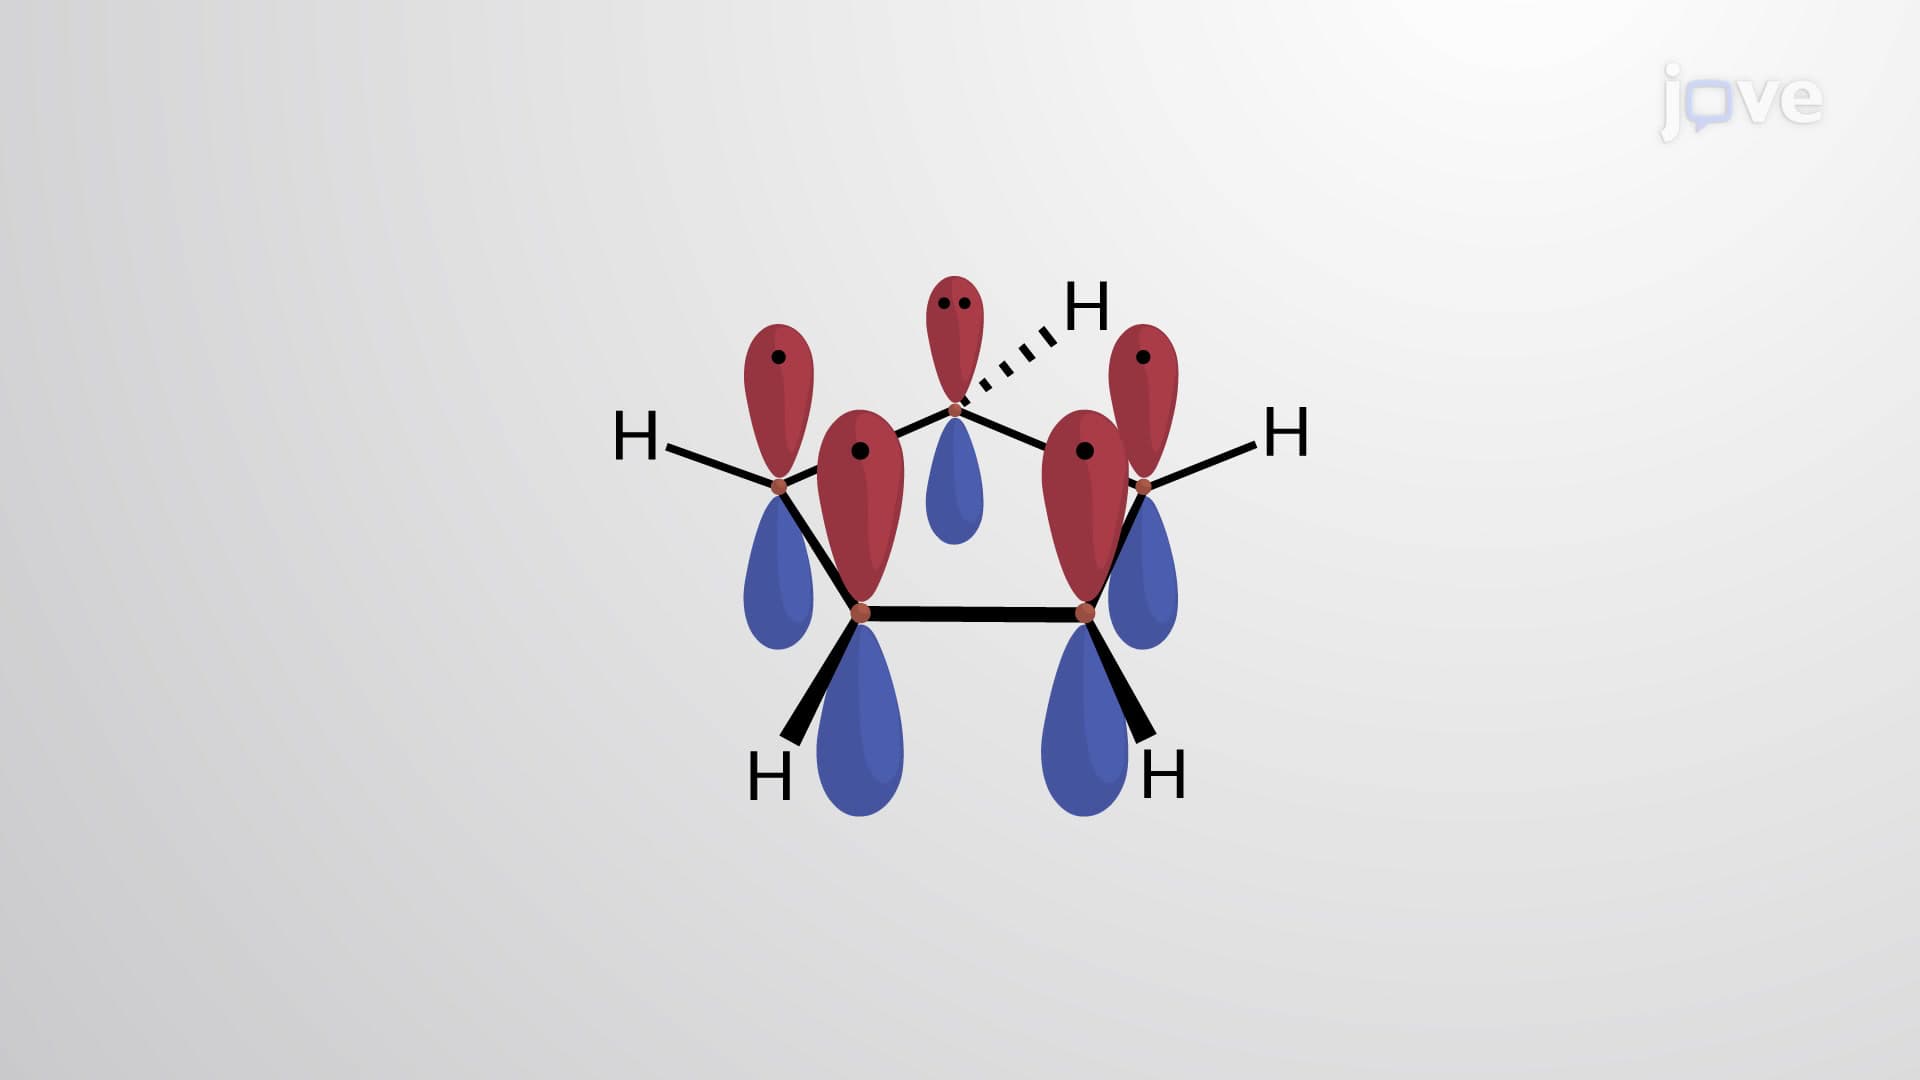
Aromatic Hydrocarbon Cations: Structural Overview
Removing one hydrogen from the intervening CH2 group...
 01:18
Aromatic Hydrocarbon Anions: Structural Overview
Due to the absence of continuous...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than...
 01:16
Structure of Conjugated Dienes
Conjugated dienes are compounds characterized by the presence of alternating double and single bonds. In a conjugated system like 1,3-butadiene, the unhybridized 2p orbital on each carbon overlaps continuously, allowing the π electrons to be delocalized across the entire molecule. In contrast, this type of overlap does not occur in cumulated and isolated dienes, such as 2,3-pentadiene and 1,4-pentadiene, respectively. Instead, the π electrons remain localized between the...
 01:26
Thermal and Photochemical Electrocyclic Reactions: Overview
 01:28
Stability of Conjugated Dienes
A comparison of the enthalpies of hydrogenation of dienes reveals that conjugated dienes release less heat on hydrogenation, rendering them more stable than their nonconjugated analogs.