Related Experiment Video
Updated: Apr 30, 2026

Author Spotlight: Analysis of Fluorescent-Stained Lipid Droplets with 3D Reconstruction for Hepatic Steatosis Assessment
Published on: June 2, 2023
Lysosomal acid lipase deficiency: wolman disease and cholesteryl ester storage disease
Anna Tylki-Szymańska1, Agnieszka Jurecka2
1Department of Pediatrics, Nutrition and Metabolic Diseases, The Children's Memorial Health Institute, Warsaw, Poland.

Abstract:
Cholesteryl ester storage disease (CESD, OMIM #278000) and Wolman disease (OMIM #278000) are autosomal recessive lysosomal storage disorders caused by a deficient activity of lysosomal acid lipase (cholesteryl ester hydrolase, LAL). Human lysosomal acid lipase is essential for the metabolism of cholesteryl esters and triglycerides. In Wolman disease, LAL activity is usually absent, whereas CESD usually presents some residual LAL activity. In infants, poor weight gain, massive hepatosplenomegaly, calcified adrenal glands (present about 2/3 of the time), vomiting, diarrhea and failure to thrive are indicative of Wolman disease. The clinical picture is more variable in CESD. Hepatomegaly and/or elevation of liver transaminases are almost always present. Hepatic steatosis often leads to fibrosis and cirrhosis. Other signs often include splenomegaly, high total cholesterol and LDL-cholesterol, elevated triglycerides, and low HDL-cholesterol. The diagnosis of LAL deficiency requires clinical experience and specialized laboratory tests. The diagnosis is based on finding deficient activity of acid lipase and/or molecular tests. Pilot screening projects using dried blood spot testing in 1) children with atypical fatty liver disease in the absence of overweight, 2) patients with dyslipidaemia and presence of hepatomegaly and/or elevated transaminases, 3) newborns/neonates with hepatomegaly and abdominal distension/failure to thrive/elevated transaminases are currently underway. Early diagnosis is particularly important for the enzyme replacement therapy. Human trials with recombinant LAL are currently ongoing, raising the prospect for specific correction of LAL deficiency in this progressive and often debilitating disorder.
More Related Videos
 10:16
In Vitro Enzyme Measurement to Test Pharmacological Chaperone Responsiveness in Fabry and Pompe Disease
Published on: December 20, 2017
 07:03
Author Spotlight: Establishing MASLD Cell Models for Investigating Disease Mechanisms and the Lipid-Lowering Effects of Koumiss
Published on: July 19, 2024
Related Concept Videos
 01:22
Lysosomal Hydrolases
 01:24
Overview of Lipid Metabolism
Lipolysis: The Breakdown of Lipids:
Lipolysis is the process of breaking down lipids, particularly triglycerides, into glycerol and fatty acids. This process typically occurs in the adipose tissue and is triggered by various hormones, including glucagon and...
 01:25
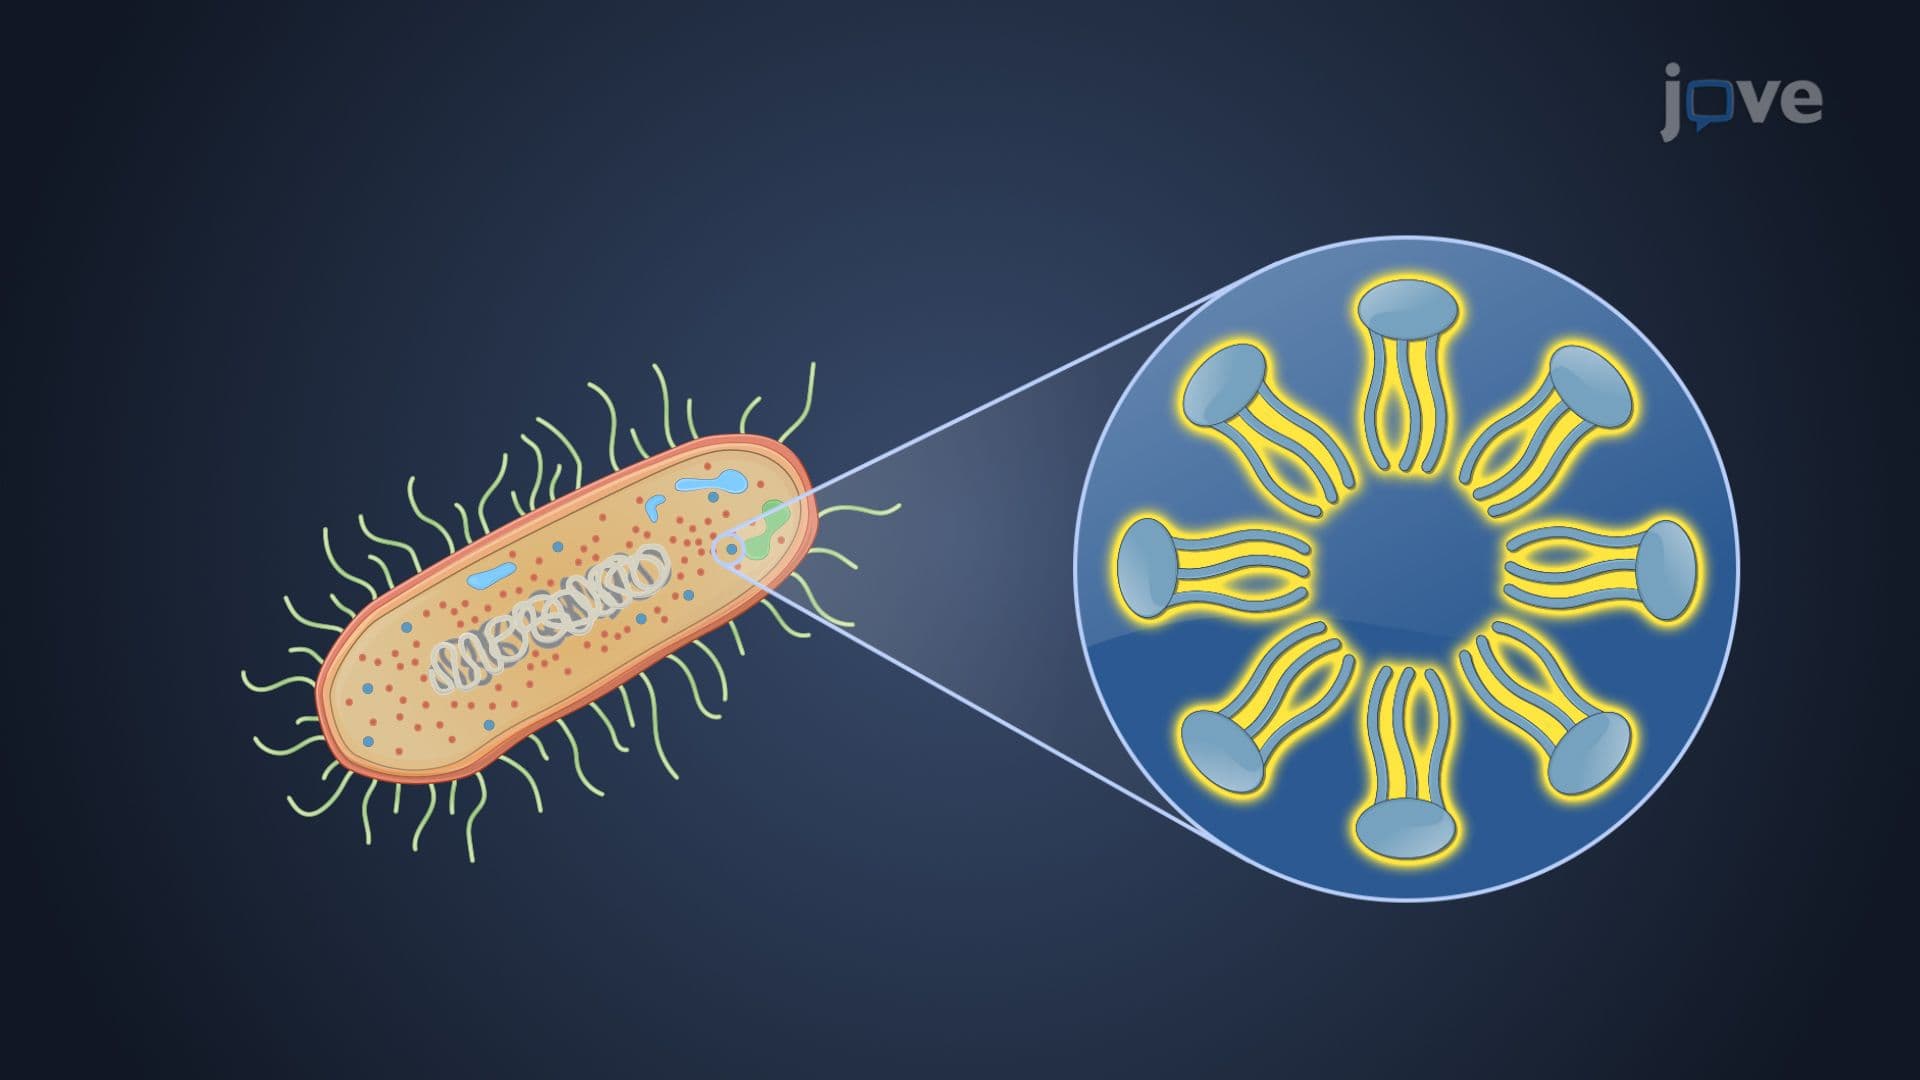
Lipid Catabolism
 01:06
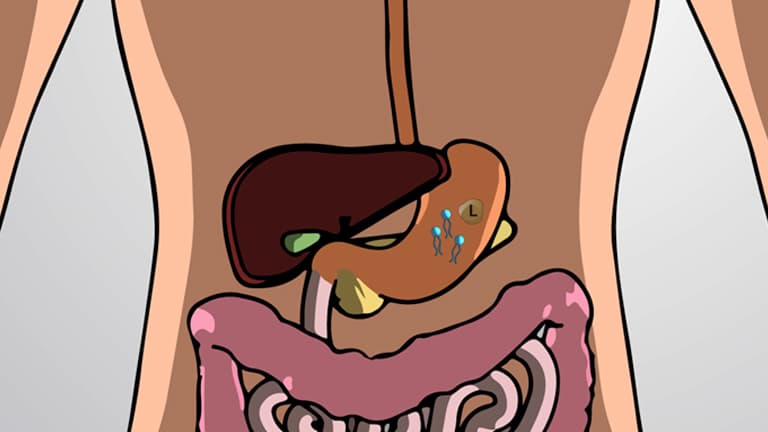
Lipid Digestion
 01:24
Lipid Absorption
These breakdown products bind with bile salts and lecithin to form micelles, which quickly pass between microvilli to come in close contact with the apical...
 01:20
Inborn Errors of Metabolism