
Genetics of mitochondrial respiratory chain deficiencies
1Inserm U1163, hôpital Necker-Enfants-Malades, institut Imagine, université Paris Descartes-Sorbonne Paris Cité, 149, rue de Sèvres, 75015 Paris, France.

Insights
Respiratory chain (RC) deficiency, crucial for ATP synthesis, can manifest diverse symptoms due to dual genetic origins. Nuclear gene mutations are increasingly identified as causes, expanding our understanding beyond mitochondrial DNA defects.
Area of Science:
- Biochemistry
- Genetics
- Cellular Respiration
Background:
- Oxidative phosphorylation (ATP synthesis via the respiratory chain) is vital for energy supply and functions from birth.
- Its dual genetic origin (nuclear and mitochondrial DNA) theoretically allows symptoms in any organ, at any age, with any inheritance pattern.
- Historically, mitochondrial DNA mutations were solely implicated in RC disorders, a view now evolving.
Purpose of the Study:
- To highlight the expanding role of nuclear gene mutations in respiratory chain disorders.
- To underscore the complexity of RC disorders due to their genetic origins.
- To correct the misconception that only mitochondrial DNA mutations cause these conditions.
Main Methods:
- Review of genetic databases and literature on respiratory chain disorders.
- Analysis of reported cases linking nuclear gene mutations to RC dysfunction.
- Comparative analysis of nuclear and mitochondrial DNA contributions to RC assembly and function.
Main Results:
- A growing number of disease-causing mutations are identified in nuclear genes affecting RC components.
- Nuclear genes encode not only subunits but also proteins crucial for holoenzyme biogenesis and mtDNA metabolism.
- This expands the known genetic basis of respiratory chain deficiencies.
Conclusions:
- Respiratory chain disorders result from mutations in both nuclear and mitochondrial DNA.
- The understanding of RC disorder etiology is broadening to include a significant contribution from nuclear genes.
- Accurate diagnosis requires consideration of both genetic origins.
Abstract:
Oxidative phosphorylation, i.e. ATP synthesis by the oxygen-consuming respiratory chain (RC), supplies most organs and tissues with a readily usable energy source, and is already fully functioning before birth. This means that, in theory, RC deficiency can give rise to any symptom in any organ or tissue at any age and with any mode of inheritance, due to the twofold genetic origin of RC components (nuclear DNA and mitochondrial DNA). It has long been erroneously believed that RC disorders originate from mutations of mtDNA as, for some time, only mutations or deletions of mtDNA could be identified. However, the number of disease-causing mutations in nuclear genes is now steadily growing. These genes not only encode the various subunits of each complex, but also the ancillary proteins involved in the different stages of holoenzyme biogenesis, including transcription, translation, chaperoning, addition of prosthetic groups and assembly of proteins, as well as the various enzymes involved in mtDNA metabolism.
More Related Videos
Related Concept Videos
 02:59
Animal Mitochondrial Genetics
 01:30
The Electron Transport Chain
Inhibitors of the electron transport chain
Rotenone, a widely used pesticide, prevents electron transfer from Fe-S cluster to ubiquinone or Q...
 01:48
ATP Synthase: Mechanism
 01:46
Electron Transport Chain: Complex I and II
ROS generation is regulated and maintained at moderate levels necessary...
 01:41
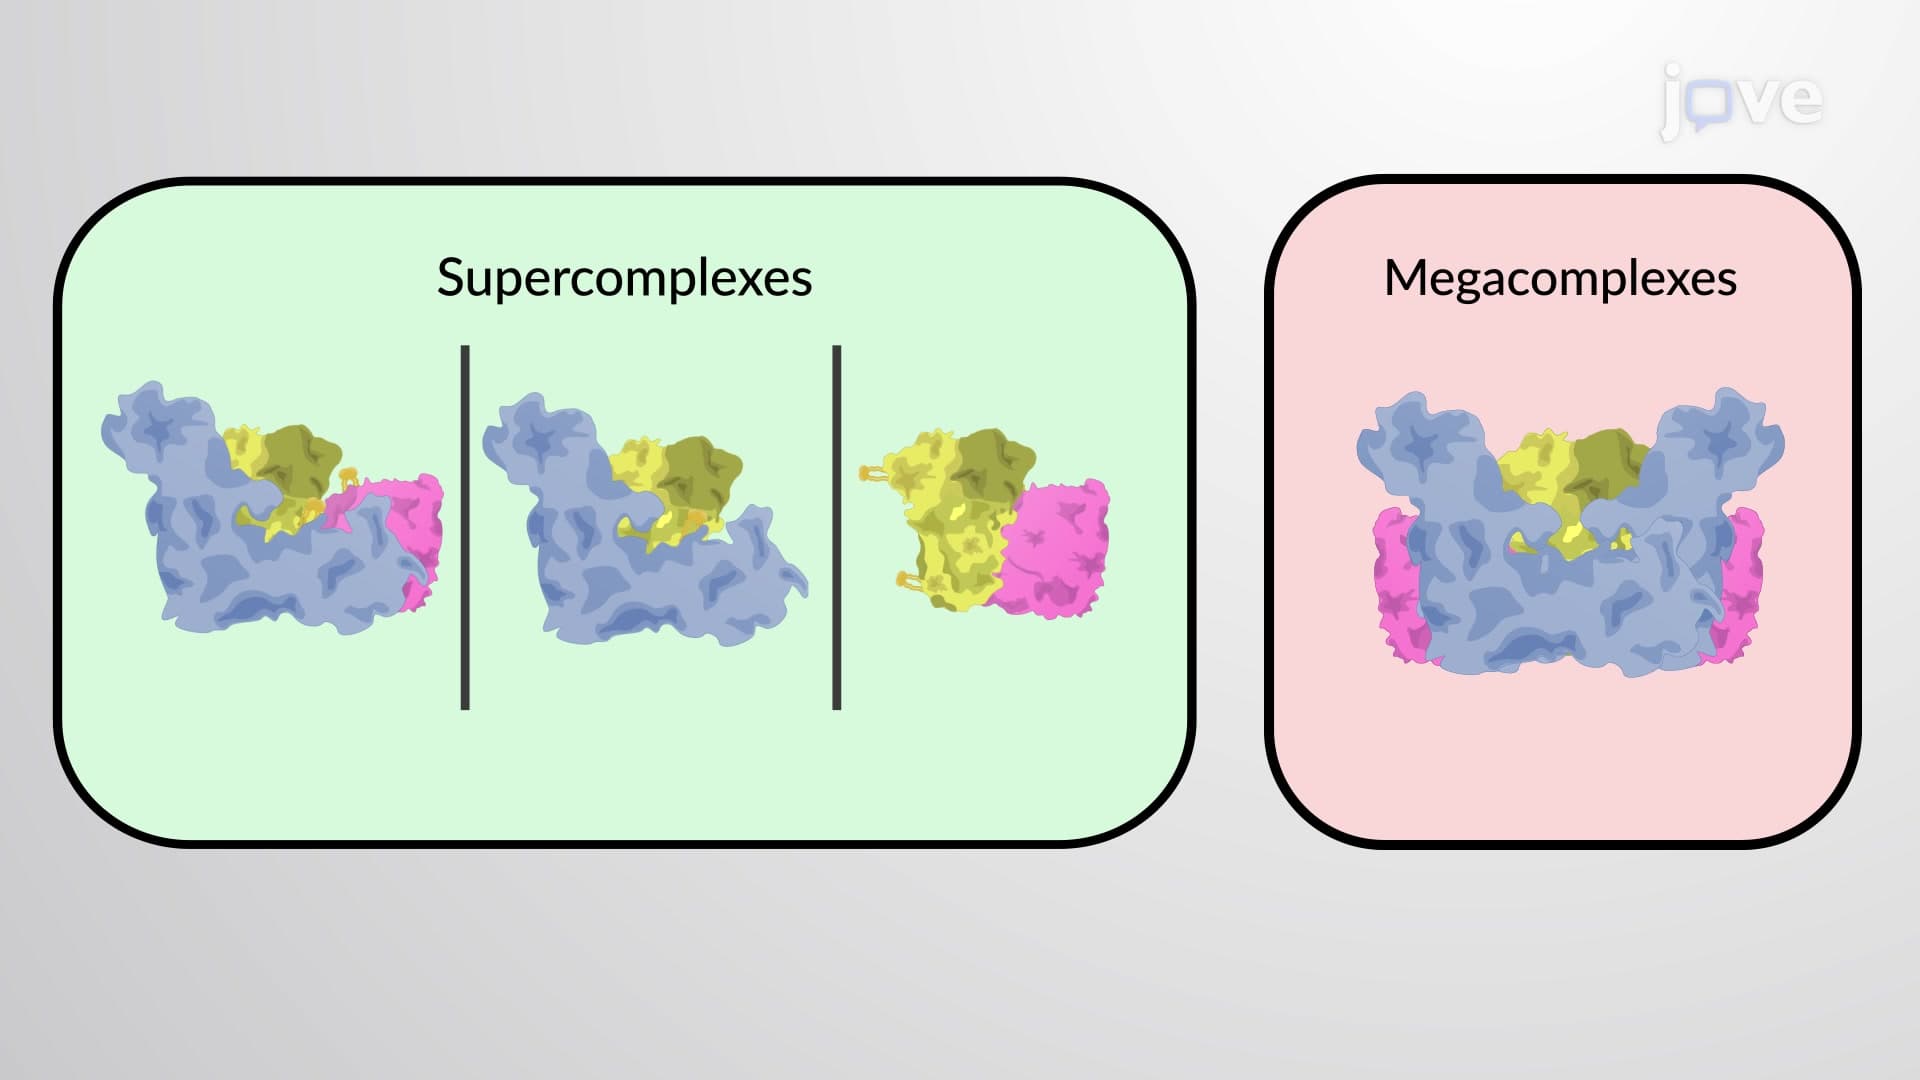
The Supercomplexes in the Crista Membrane
 01:37
Mitochondria