Related Experiment Video
Updated: Apr 30, 2026

Toxicity Screens in Human Retinal Organoids for Pharmaceutical Discovery
Published on: March 4, 2021
Involvement of organic cation transporter 1 and CYP3A4 in retrorsine-induced toxicity
Meijuan Tu1, Liping Li1, Hongmei Lei1
1Laboratory of Pharmaceutical Analysis and Drug Metabolism, Zhejiang Province Key Laboratory of Anti-Cancer Drug Research, College of Pharmaceutical Sciences, Zhejiang University, Hangzhou, China.

Abstract:
Retrorsine (RTS) is a hepatotoxic pyrrolizidine alkaloid present in plants of the Senecio genus. The present study is aimed at clarifying the role of organic cation transporters (OCTs) in the liver disposition of RTS, and the coupling of OCT1 and cytochrome P450 (CYP) 3A4 in the hepatotoxicity of RTS. MDCK or LLC-PK1 cells stably expressing liver uptake or efflux transporters were used to investigate the interaction of RTS with these transporters. Primary cultured rat hepatocytes (PCRH) and double-transfected MDCK-hOCT1-CYP3A4 cells were used to determine the contribution of OCT1 and CYP3A4 to the toxicity of RTS. The results showed that RTS inhibited the OCT1-mediated 1-methyl-4-phenylpyridinium (MPP(+)) uptake in MDCK-hOCT1 cells with the IC50 of 2.25±0.30μM. The uptake of RTS in MDCK-hOCT1 cells and PCRH was significantly inhibited by OCT1 inhibitors, while hOCT3, human multidrug and toxin extrusion (hMATE) transporter 1, multidrug resistance 1 (MDR1), and breast cancer resistance protein (BCRP) showed weak or no obvious interaction with RTS. The toxic effect of RTS on the PCRH was attenuated by OCT1 inhibitors, quinidine and (+)-tetrahydropalmatine ((+)-THP). Compared to mock cells, MDCK-CYP3A4 cells showed a decrease in viability after being treated with RTS. Furthermore, RTS showed a more severe toxicity in the OCT1/CYP3A4 double-transfected cells compared to all other cells. Our data suggests that OCT1 mediates the liver-specific uptake of RTS, and plays an important role in RTS-induced hepatotoxicity together with CYP3A4. Consequently, the OCT1 inhibitors could be applied to protect the liver from the toxicity of RTS.
Insights
Retrorsine (RTS) is a liver toxin. Organic cation transporter 1 (OCT1) and cytochrome P450 3A4 (CYP3A4) mediate RTS uptake and toxicity. OCT1 inhibitors may protect the liver from RTS damage.
Area of Science:
- Pharmacology and Toxicology
- Drug Metabolism and Transport
Background:
- Retrorsine (RTS) is a hepatotoxic pyrrolizidine alkaloid found in Senecio plants.
- The liver's role in RTS disposition and the mechanisms of its toxicity are not fully understood.
Purpose of the Study:
- To elucidate the role of organic cation transporters (OCTs) in the liver uptake of RTS.
- To investigate the interplay between OCT1 and cytochrome P450 3A4 (CYP3A4) in RTS-induced hepatotoxicity.
Main Methods:
- Utilized cell lines (MDCK, LLC-PK1) expressing various transporters to study RTS interactions.
- Employed primary cultured rat hepatocytes (PCRH) and double-transfected MDCK-hOCT1-CYP3A4 cells to assess toxicity.
- Investigated RTS inhibition of OCT1-mediated substrate uptake and the effects of OCT1 inhibitors on RTS toxicity.
Main Results:
- RTS inhibited OCT1-mediated uptake of 1-methyl-4-phenylpyridinium (MPP+) with an IC50 of 2.25±0.30μM.
- OCT1 inhibitors significantly reduced RTS uptake in cells and attenuated RTS-induced toxicity in primary hepatocytes.
- RTS exhibited increased toxicity in cells co-expressing OCT1 and CYP3A4, suggesting a synergistic effect.
Conclusions:
- OCT1 is crucial for the liver-specific uptake of RTS.
- OCT1 and CYP3A4 collaboratively contribute to RTS-induced hepatotoxicity.
- OCT1 inhibitors show potential as a therapeutic strategy to mitigate liver damage from RTS.
More Related Videos
 05:03
Author Spotlight: Investigating Physiological Functions of Vitamin A Transporters Using HPLC-Based Vitamin A Profiling
Published on: December 27, 2024
 10:44
Mass Spectrometry and Luminogenic-based Approaches to Characterize Phase I Metabolic Competency of In Vitro Cell Cultures
Published on: March 28, 2017
Related Concept Videos
 01:16
Drug toxicity: Idiosyncratic Reactions
 01:30
Drug toxicity: Drug–Drug Interaction
 01:24
Drug Toxicity: Risk factors
 01:25
Pharmacokinetics: Drug–Drug Interactions
 01:16
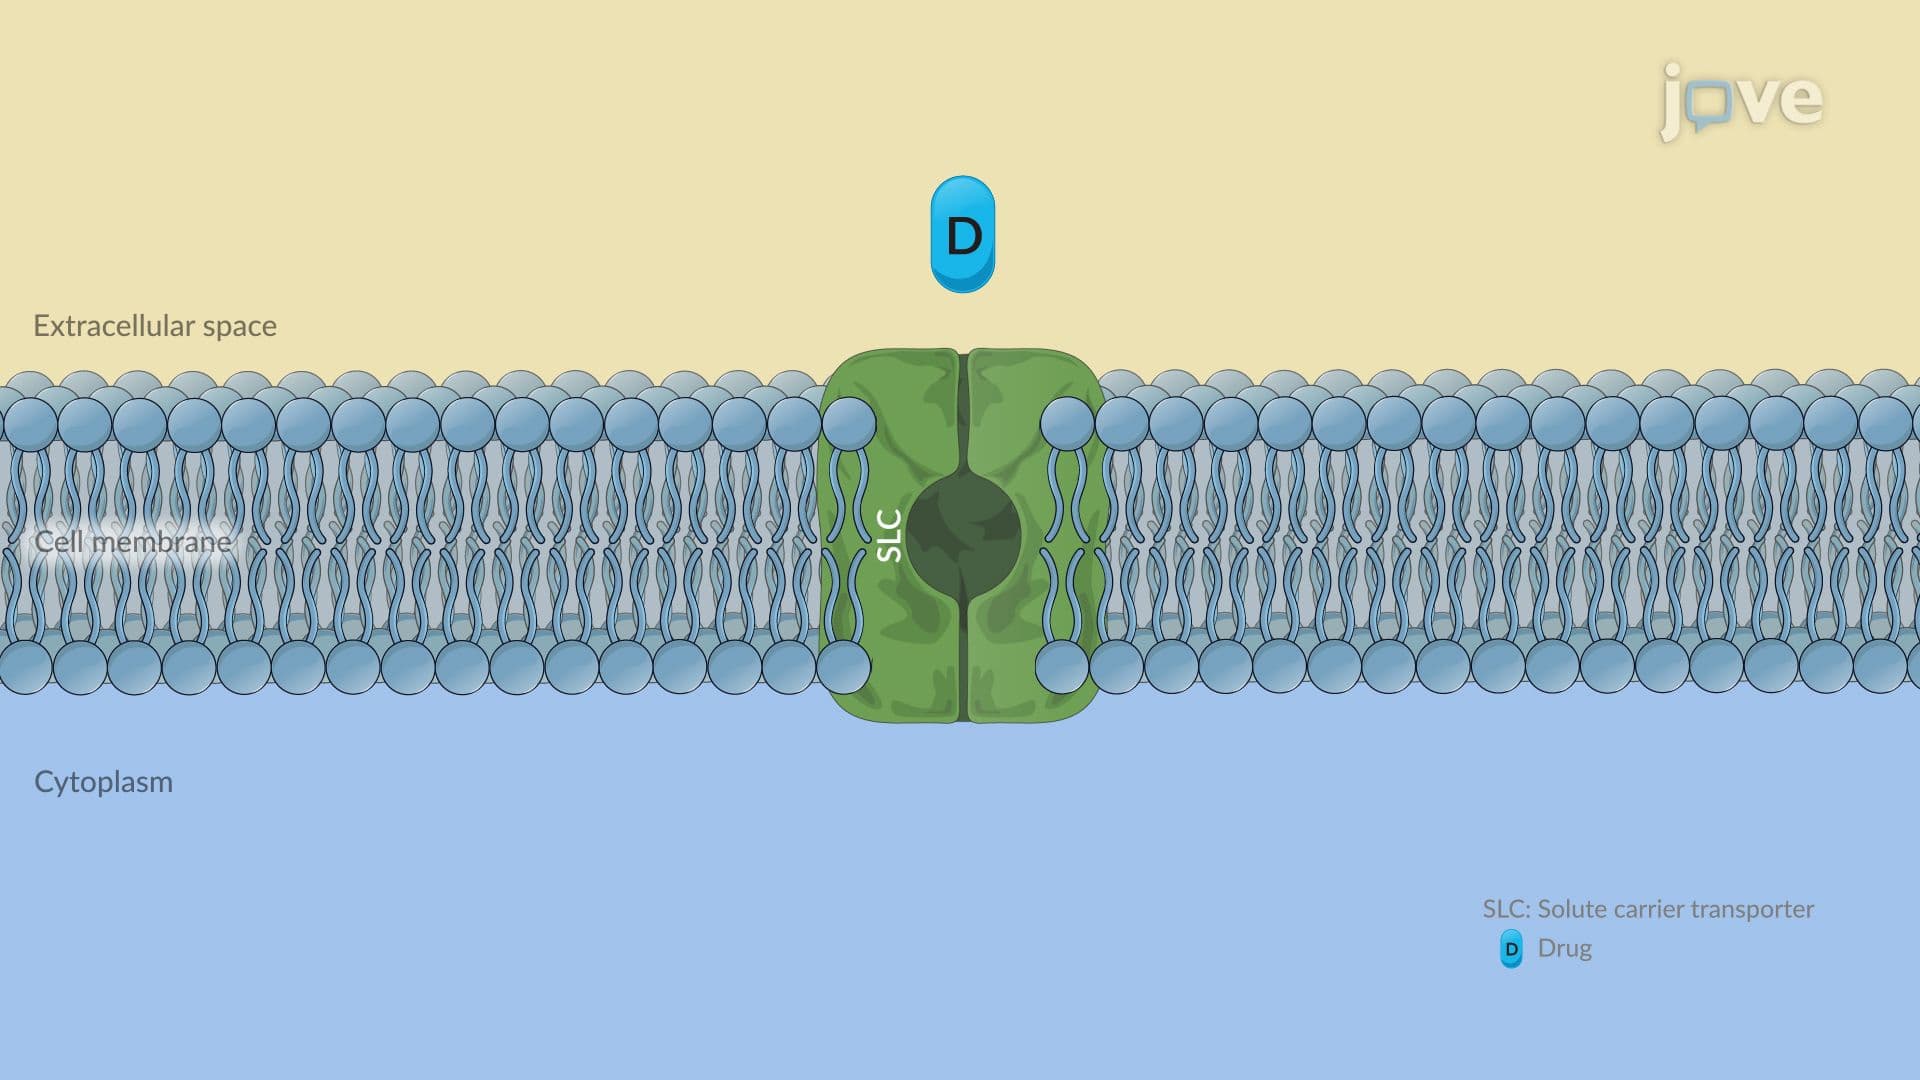
Pharmacogenetics of Drug Transporters: P-Glycoprotein and Solute Carrier Transporters
 01:24
Drug Toxicity: Dose-Dependent Reactions