Related Experiment Video
Updated: Apr 30, 2026


Isotopic Effect in Double Proton Transfer Process of Porphycene Investigated by Enhanced QM/MM Method
Published on: July 19, 2019
Quantum Mechanical Calculation of Noncovalent Interactions: A Large-Scale Evaluation of PMx, DFT, and SAPT Approaches
Amanda Li1, Hari S Muddana2, Michael K Gilson2
1Skaggs School of Pharmacy and Pharmaceutical Sciences, University of California San Diego , 9500 Gilman Drive, La Jolla, California 92093-0736, United States ; Department of Bioengineering, University of California San Diego , 9500 Gilman Drive, La Jolla, California 92093-0419, United States.

Abstract:
Quantum mechanical (QM) calculations of noncovalent interactions are uniquely useful as tools to test and improve molecular mechanics force fields and to model the forces involved in biomolecular binding and folding. Because the more computationally tractable QM methods necessarily include approximations, which risk degrading accuracy, it is essential to evaluate such methods by comparison with high-level reference calculations. Here, we use the extensive Benchmark Energy and Geometry Database (BEGDB) of CCSD(T)/CBS reference results to evaluate the accuracy and speed of widely used QM methods for over 1200 chemically varied gas-phase dimers. In particular, we study the semiempirical PM6 and PM7 methods; density functional theory (DFT) approaches B3LYP, B97-D, M062X, and ωB97X-D; and symmetry-adapted perturbation theory (SAPT) approach. For the PM6 and DFT methods, we also examine the effects of post hoc corrections for hydrogen bonding (PM6-DH+, PM6-DH2), halogen atoms (PM6-DH2X), and dispersion (DFT-D3 with zero and Becke-Johnson damping). Several orders of the SAPT expansion are also compared, ranging from SAPT0 up to SAPT2+3, where computationally feasible. We find that all DFT methods with dispersion corrections, as well as SAPT at orders above SAPT2, consistently provide dimer interaction energies within 1.0 kcal/mol RMSE across all systems. We also show that a linear scaling of the perturbative energy terms provided by the fast SAPT0 method yields similar high accuracy, at particularly low computational cost. The energies of all the dimer systems from the various QM approaches are included in the Supporting Information, as are the full SAPT2+(3) energy decomposition for a subset of over 1000 systems. The latter can be used to guide the parametrization of molecular mechanics force fields on a term-by-term basis.
More Related Videos
 12:11
Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 02:36
Molecular Geometry and Dipole Moments
 01:24
Van der Waals Interactions
 01:10
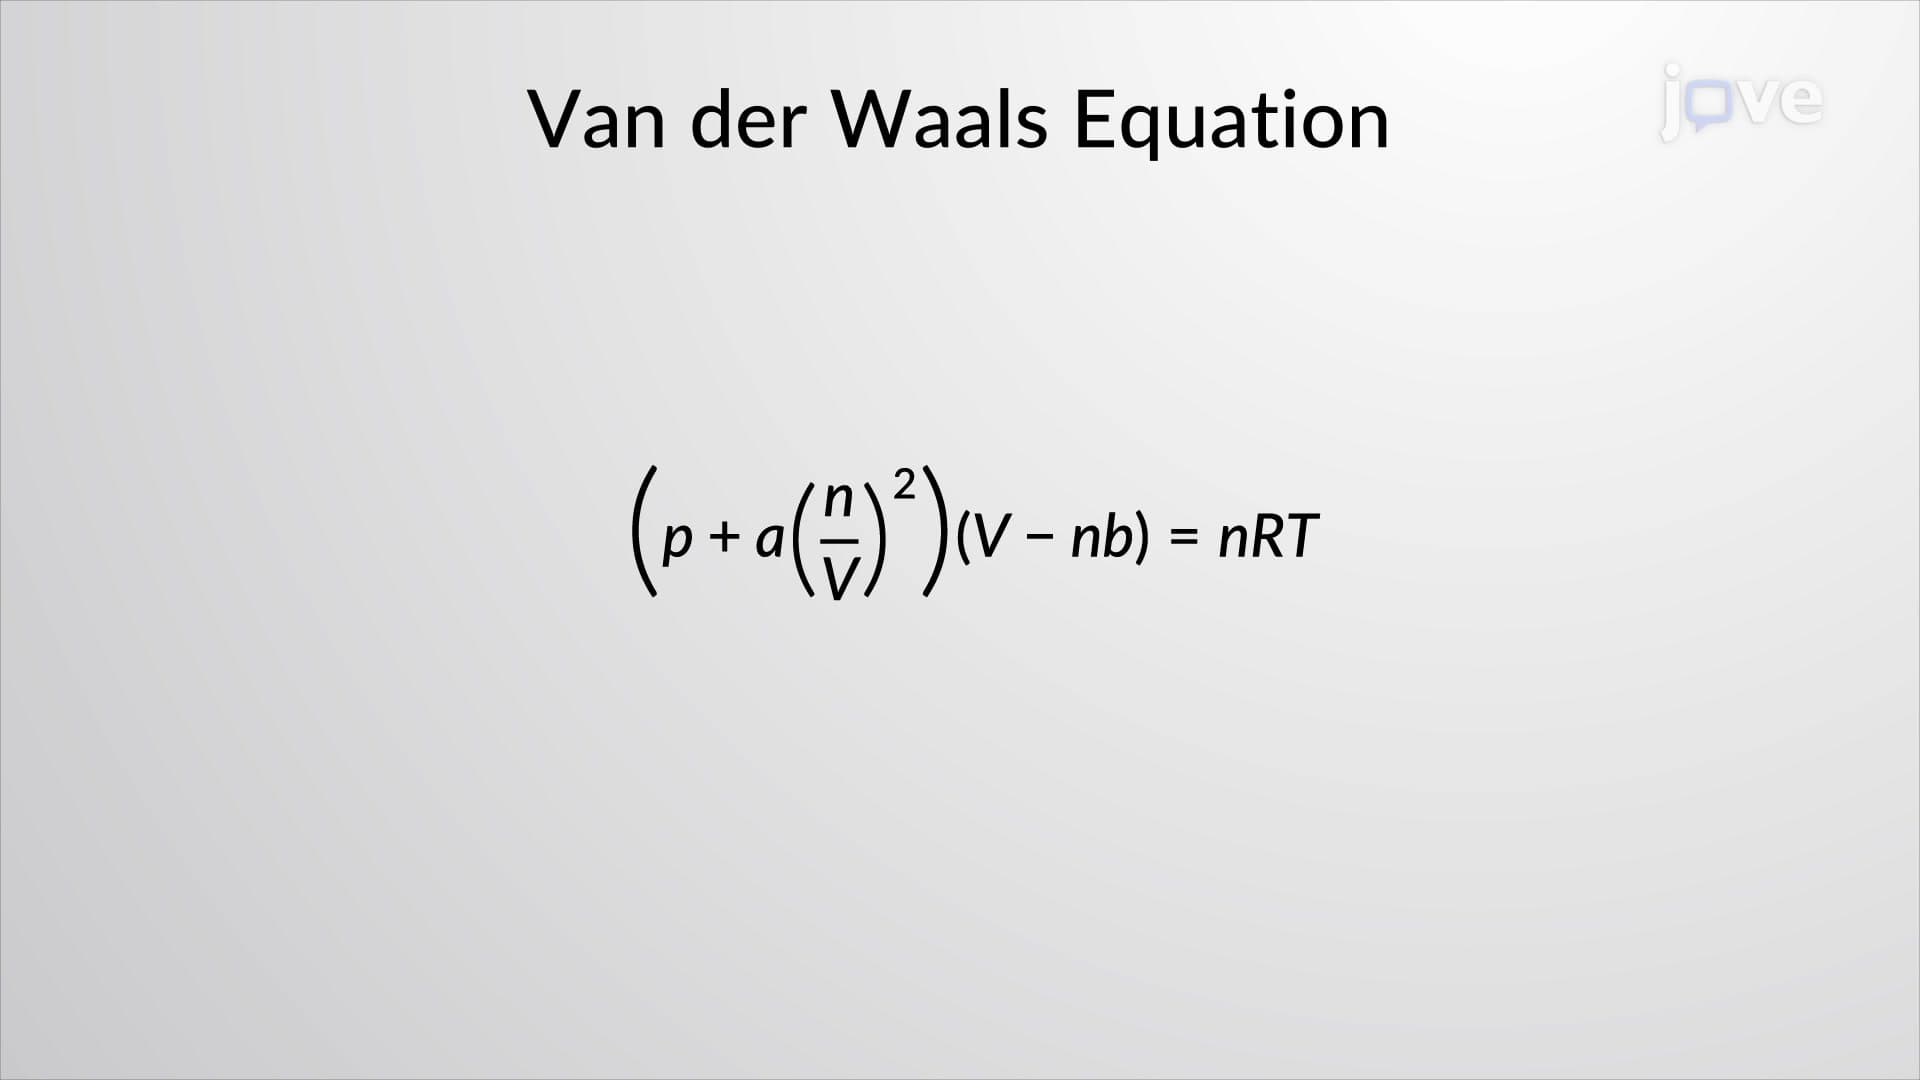
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the...
 02:40
MO Theory and Covalent Bonding
 02:38
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
 01:26
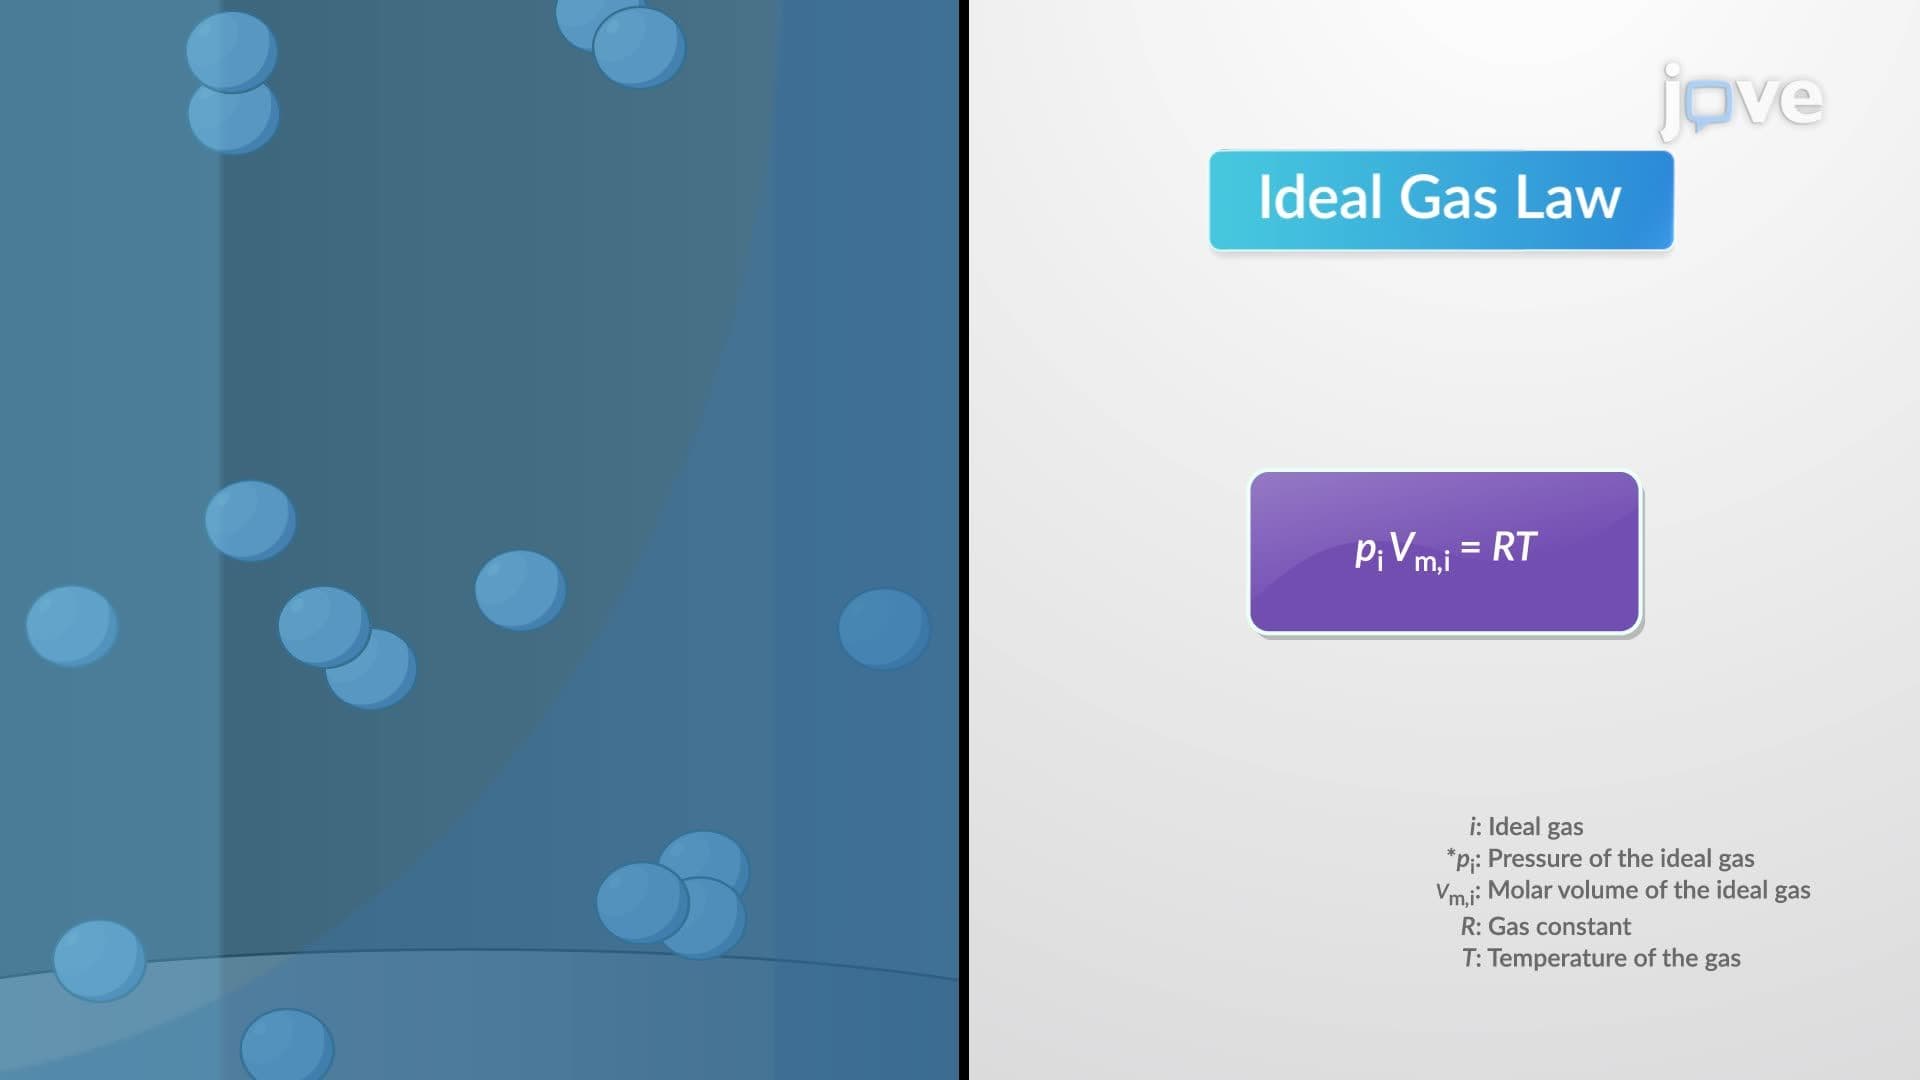
The Van der Waals Equation