

Introducing constricted variational density functional theory in its relaxed self-consistent formulation
Mykhaylo Krykunov1, Mike Seth1, Tom Ziegler1
1Department of Chemistry, University of Calgary, University Drive 2500, Calgary, Alberta T2N 1N4, Canada.

Abstract:
We have applied the relaxed and self-consistent extension of constricted variational density functional theory (RSCF-CV-DFT) for the calculation of the lowest charge transfer transitions in the molecular complex X-TCNE between X = benzene and TCNE = tetracyanoethylene. Use was made of functionals with a fixed fraction (α) of Hartree-Fock exchange ranging from α = 0 to α = 0.5 as well as functionals with a long range correction (LC) that introduces Hartree-Fock exchange for longer inter-electronic distances. A detailed comparison and analysis is given for each functional between the performance of RSCF-CV-DFT and adiabatic time-dependent density functional theory (TDDFT) within the Tamm-Dancoff approximation. It is shown that in this particular case, all functionals afford the same reasonable agreement with experiment for RSCF-CV-DFT whereas only the LC-functionals afford a fair agreement with experiment using TDDFT. We have in addition calculated the CT transition energy for X-TCNE with X = toluene, o-xylene, and naphthalene employing the same functionals as for X = benzene. It is shown that the calculated charge transfer excitation energies are in as good agreement with experiment as those obtained from highly optimized LC-functionals using adiabatic TDDFT. We finally discuss the relation between the optimization of length separation parameters and orbital relaxation in the RSCF-CV-DFT scheme.
More Related Videos
 08:44
Measurements of Long-range Electronic Correlations During Femtosecond Diffraction Experiments Performed on Nanocrystals of Buckminsterfullerene
Published on: August 22, 2017
 12:11
Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Related Concept Videos
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:38
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
 02:42
Valence Bond Theory
 02:45
Valence Bond Theory
 01:28
Debye–Huckel–Onsager Conductance Equation
 01:10
Van der Waals Equation
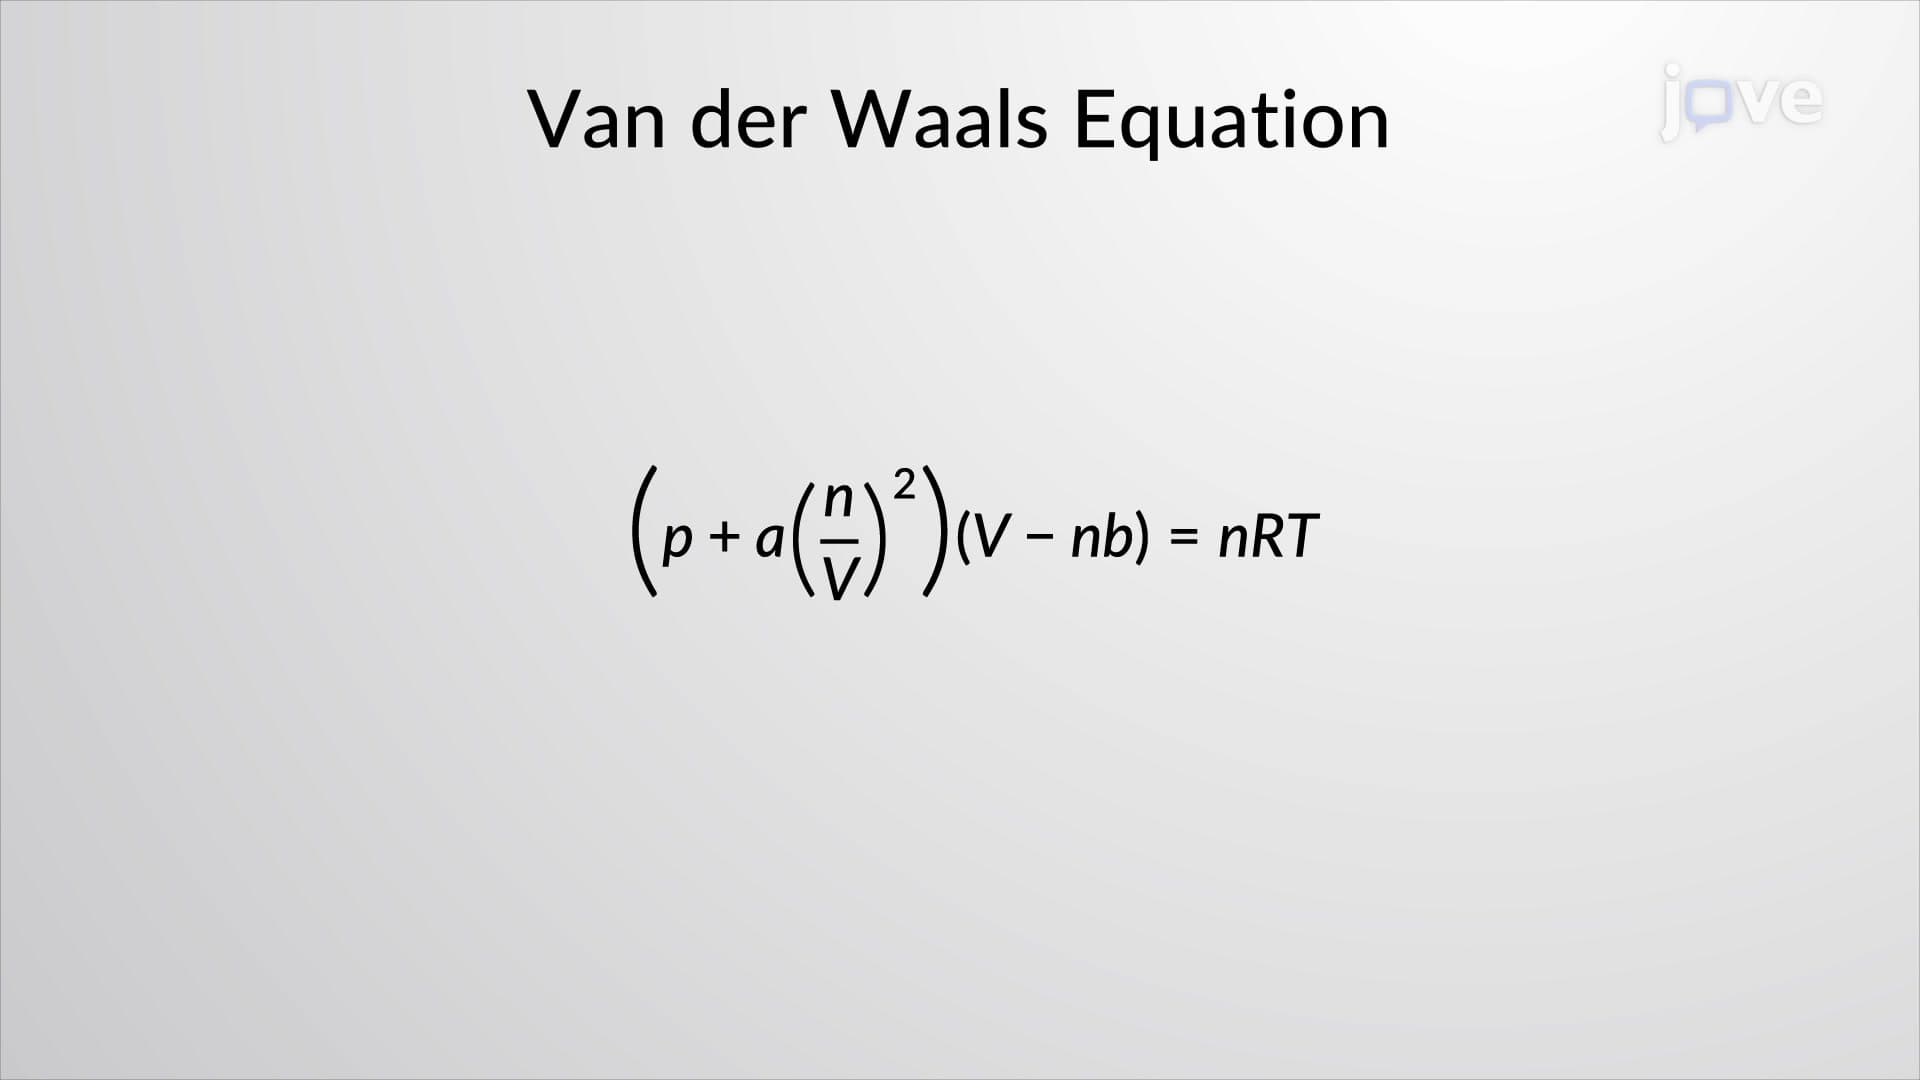
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the...