Related Experiment Video
Updated: Apr 29, 2026


Application of I TASSER, trRosetta, UCSF Chimera, HADDOCK server, and HEX loria for De Novo and In Silico Design of Proteins
Published on: July 8, 2025
An In silico Based Comparison of Drug Interactions in Wild and Mutant Human β-tubulin through Docking Studies
Selvaakumar Chellasamy1, Sudheer M M Mohammed2
1Department of Biotechnology and Bioinformatics, Padmashree Dr.D.Y. Patil University, Navi Mumbai, India.

Background:
Tubulin protein being the fundamental unit of microtubules is actively involved in cell division thus making them a potential anti-cancer drug target. In spite of many reported drugs against tubulin, few of them have started developing resistance in human β-tubulin due to amino acid substitutions.
Methods:
In this study we generated three mutants (F270V, A364T and Q292E) using Modeller9v10 which were targeted with compounds from higher and lower plants along with marine isolates using iGEMDOCK2.0 to identify their residual interactions.
Results:
The mutant F270V does not bring in any increase in the binding affinity in comparison with the taxol-wild type due to their conservative substitutions. However, it increases the volume of the active site. A364T mutant brings a better binding among few of the marine and higher plants isolates due to the substitution of the non-reactive methyl group with the polar residue. But this leads to reduced active site volume. Finally the mutant Q292E from epothilone binding site brings a remarkable change in drug binding in the mutants in comparison with the wild type due to the substitution of uncharged residue with the charged one. But as such there was no change in the volume of the active site observed in them.
Conclusion:
Lower plants extracts were reported to exhibit better interactions with the taxol and epothilone binding sites. Whereas marine and higher plants isolates shows significant interactions only in the wild type instead of the mutants. In addition to this, the residual substitutions were also found to alter the conformations of the active sites in mutants.
Insights
Drug resistance in cancer therapy is a growing concern. This study explores how specific mutations in human beta-tubulin affect drug binding, revealing potential strategies to overcome resistance using plant and marine compounds.
Area of Science:
- Biochemistry
- Pharmacology
- Structural Biology
Background:
- Tubulin, a key component of microtubules, is a validated anti-cancer drug target.
- Drug resistance in human beta-tubulin arises from amino acid substitutions, necessitating novel therapeutic approaches.
Purpose of the Study:
- To investigate the impact of specific amino acid substitutions (F270V, A364T, Q292E) in human beta-tubulin on drug binding.
- To identify potential drug candidates from plant and marine sources that can overcome tubulin-based drug resistance.
Main Methods:
- Generated three human beta-tubulin mutants (F270V, A364T, Q292E) using Modeller9v10.
- Screened compounds from higher plants, lower plants, and marine isolates for interactions with tubulin mutants using iGEMDOCK2.0.
Main Results:
- The F270V mutation showed conservative substitutions, increasing active site volume but not binding affinity compared to the taxol-wild type.
- The A364T mutation improved binding with some marine and higher plant compounds due to polar residue substitution, but reduced active site volume.
- The Q292E mutation, located in the epothilone binding site, significantly altered drug binding compared to the wild type due to substitution of an uncharged with a charged residue, without changing active site volume.
Conclusions:
- Lower plant extracts demonstrated favorable interactions with both taxol and epothilone binding sites.
- Marine and higher plant isolates showed significant interactions with wild-type tubulin but not with the generated mutants.
- Amino acid substitutions in tubulin mutants were observed to alter active site conformations, influencing drug interactions.
More Related Videos
 08:49
Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
 14:34
A Bilingual Computational Workflow for Identifying Potential PLK1 Inhibitors in American Sign Language and English
Published on: April 3, 2026
Related Concept Videos
 02:04
Protein-protein Interfaces
 02:18
The Equilibrium Binding Constant and Binding Strength
 02:40
Ligand Binding Sites
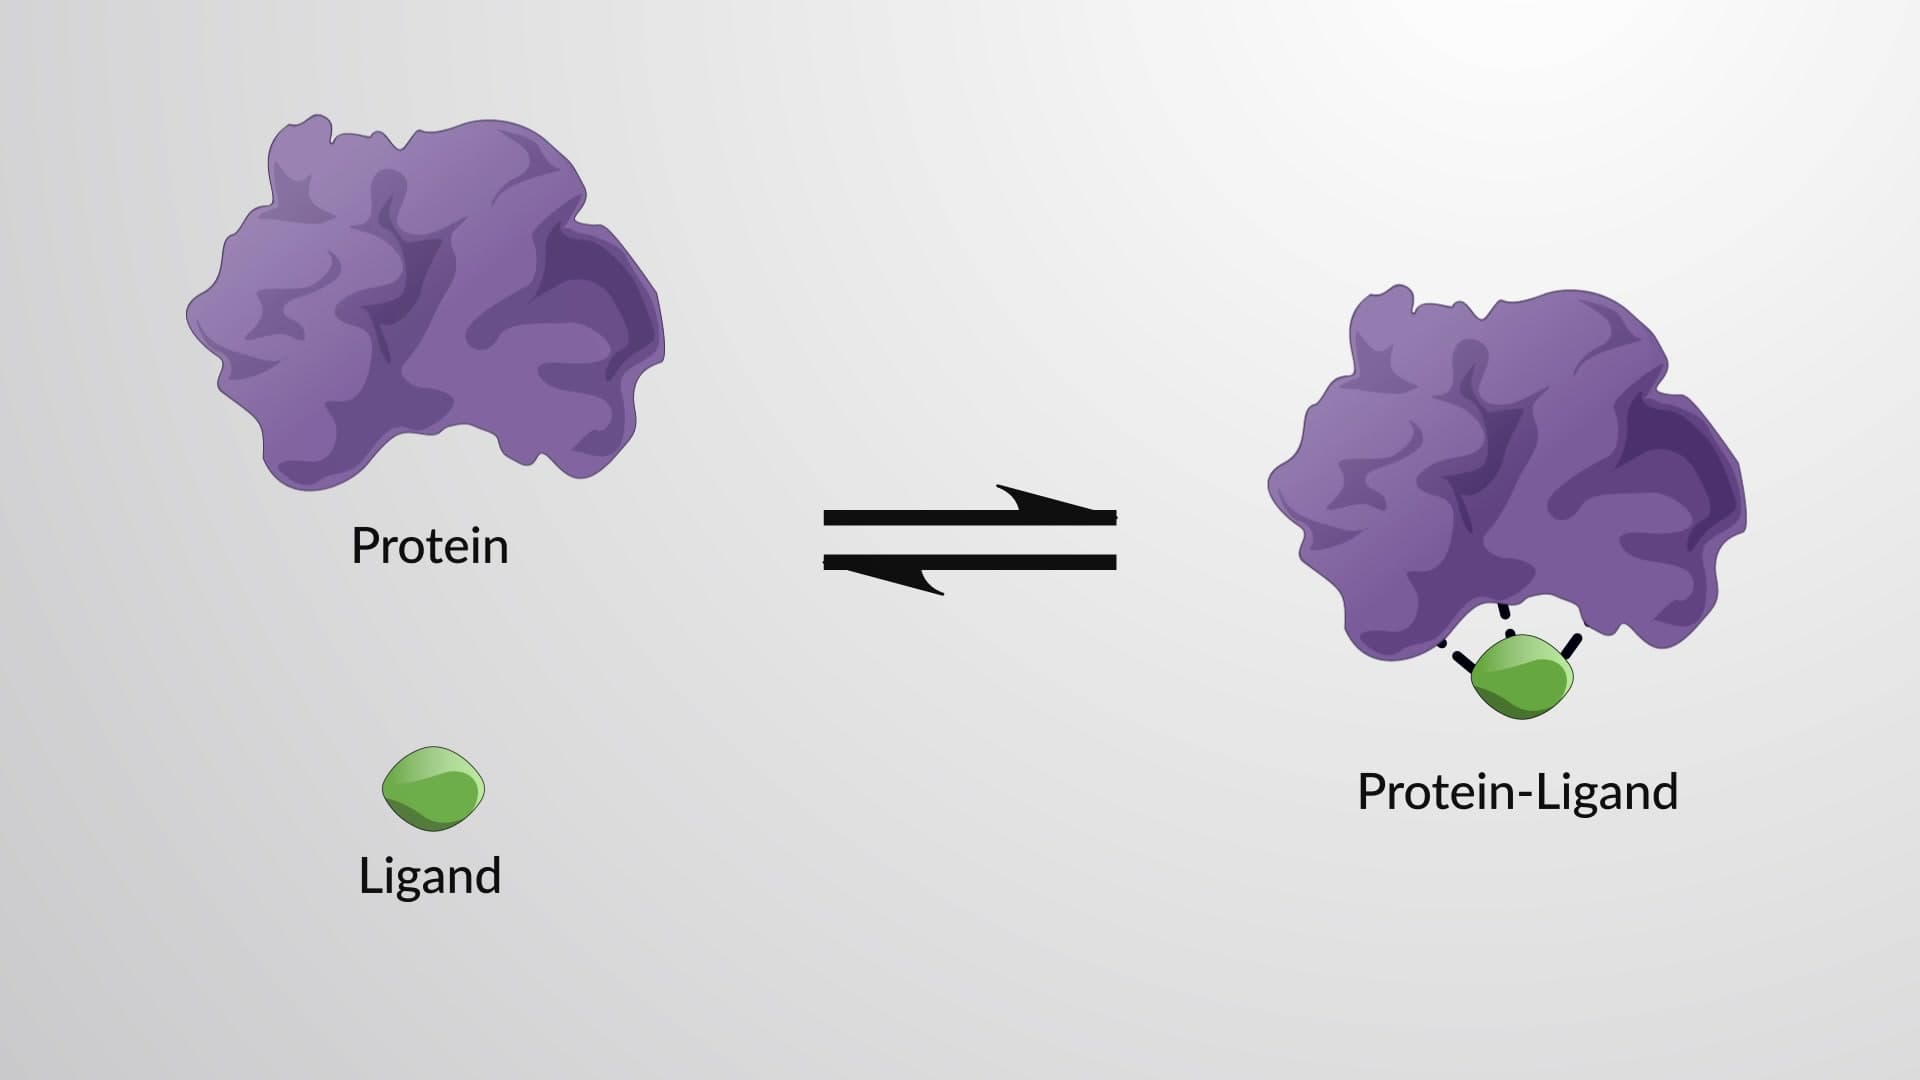
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...