Related Experiment Video
Updated: Apr 29, 2026

Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
Fractional charge behavior and band gap predictions with the XYG3 type of doubly hybrid density functionals
Neil Qiang Su1, Weitao Yang, Paula Mori-Sánchez
1Collaborative Innovation Center of Chemistry for Energy Materials, Shanghai Key Laboratory of Molecular Catalysis and Innovative Materials, MOE Laboratory for Computational Physical Science, Department of Chemistry, Fudan University , Shanghai 200433, China.

Abstract:
In this work, we examine the fractional charge behaviors of doubly hybrid (DH) functionals. By plotting the ground-state energies E and energy derivatives for atoms and molecules with fractional electron numbers N, we directly quantify the delocalization errors of some representative DH functionals such as B2PLYP, XYG3, and XYGJ-OS. Numerical assessments on ionization potentials (IPs), electron affinities (EAs), and fundamental gaps, from either integer number calculations or energy derivative calculations, are provided. It is shown that the XYG3 type of DH functionals gives good agreement between their energy derivatives and the experimental IPs, EAs, and gaps, as expected from their nearly straight line fractional charge behaviors.
More Related Videos
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than...
 02:35
Band Theory
The energy difference between these bands is known as the band gap.
Conductor, Semiconductor,...
 01:30
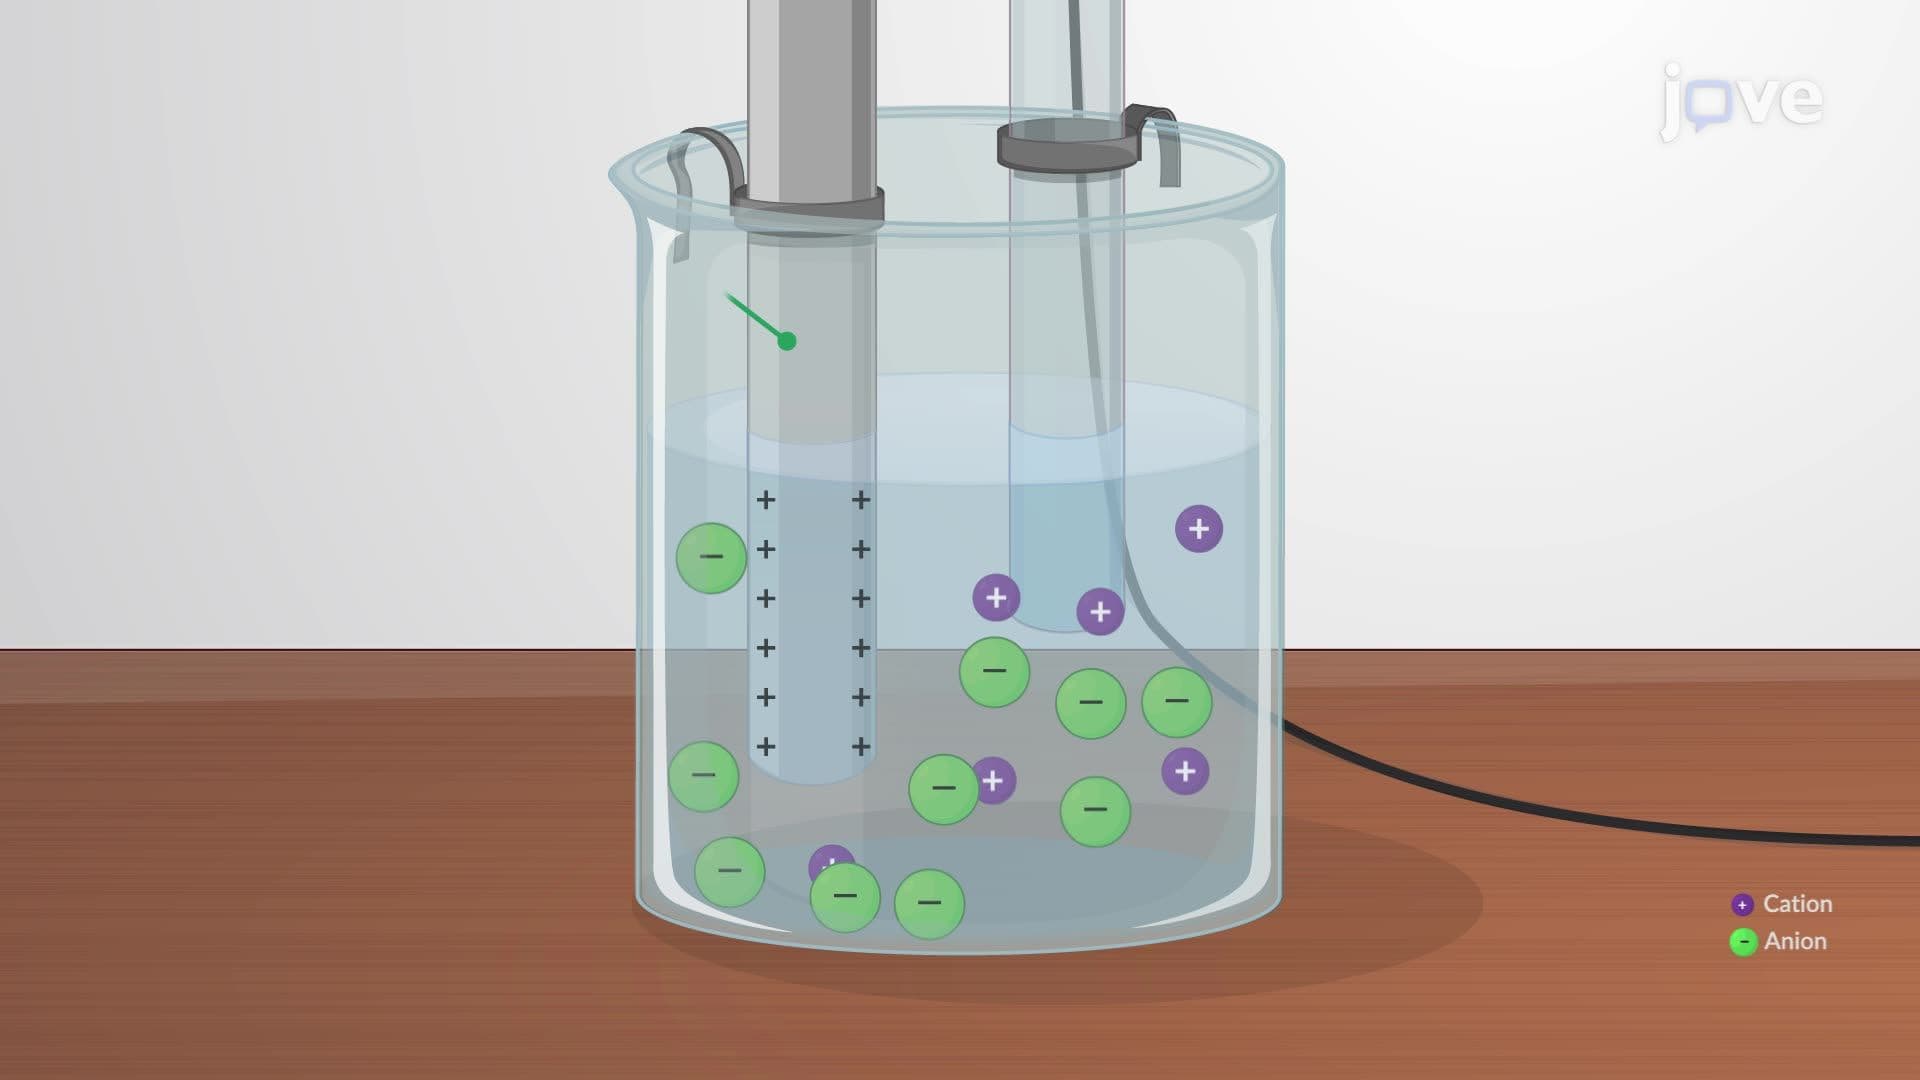
The Electrical Double Layer
 02:42
Valence Bond Theory
 02:45
Valence Bond Theory
 01:12
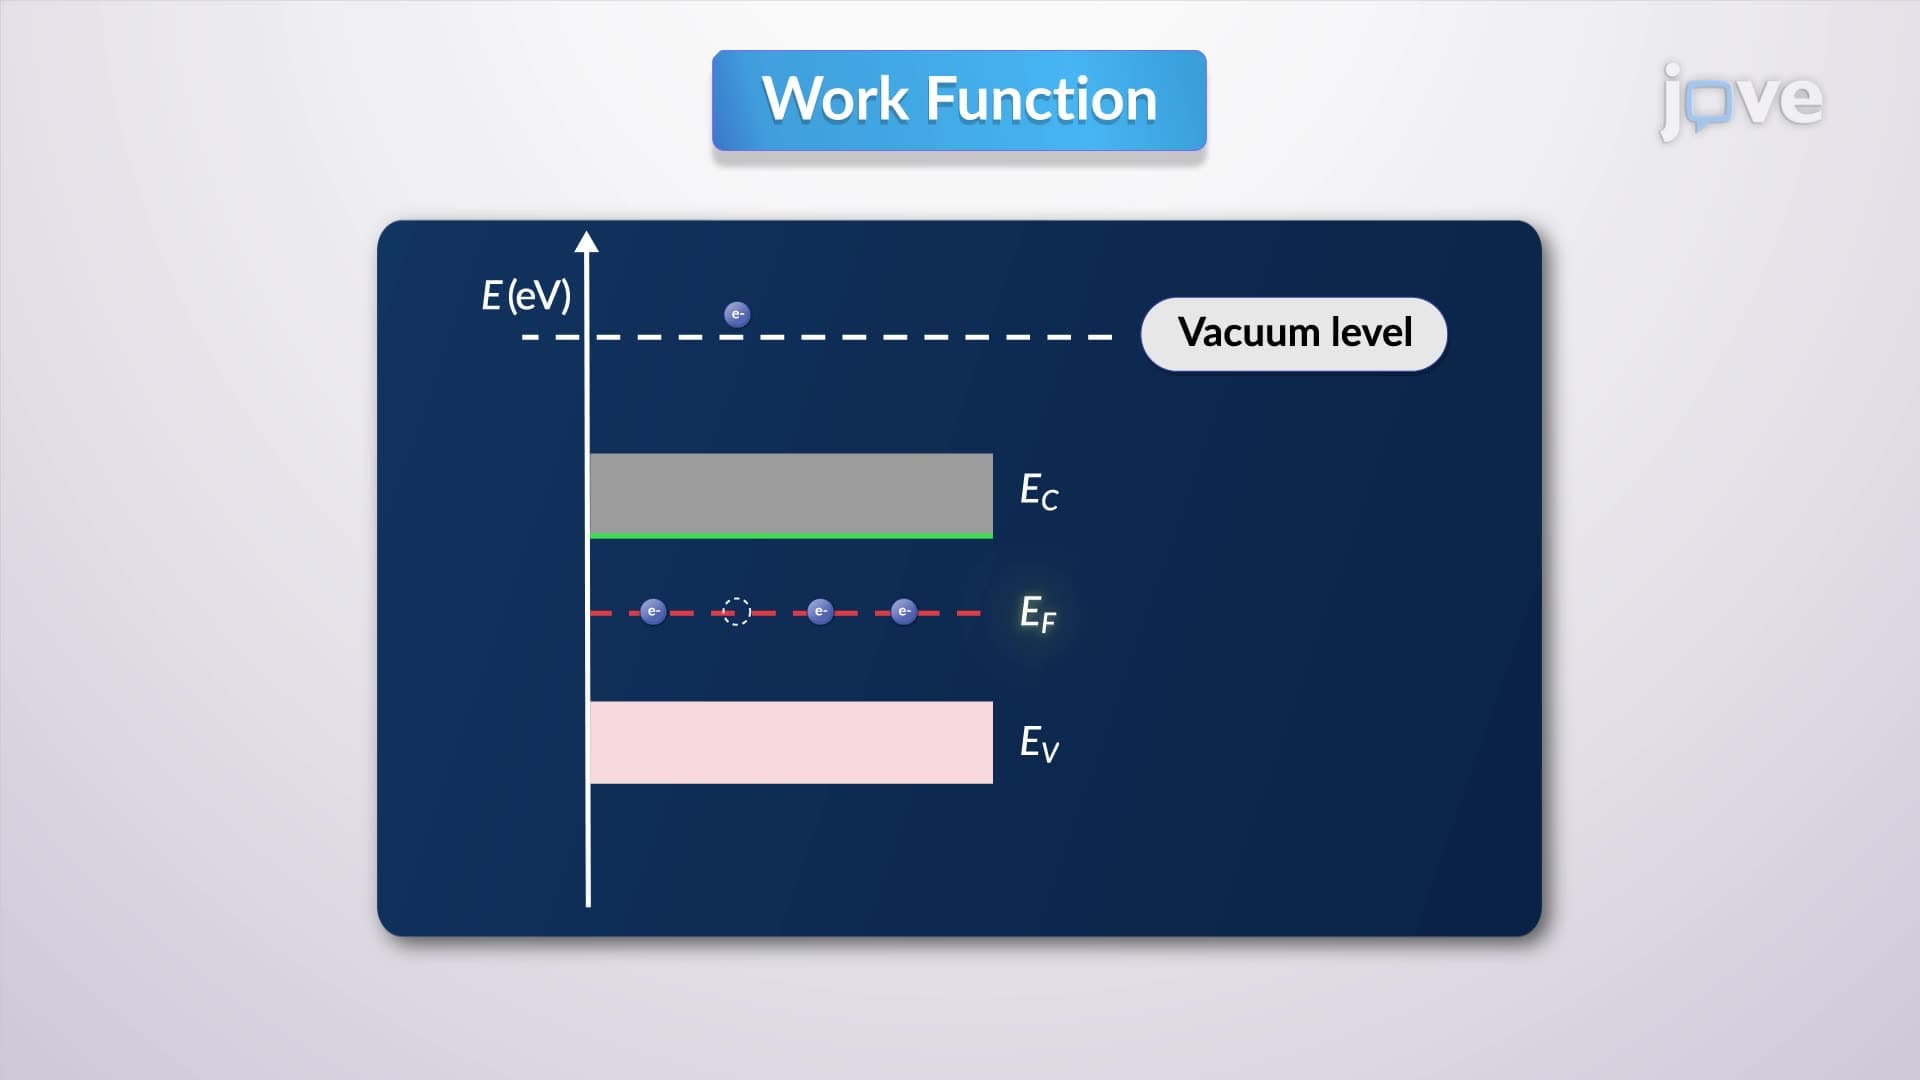
Fermi Level Dynamics
Electron affinity in semiconductors refers to the energy gap between the minimum of its conduction band and the vacuum level and it is a critical parameter in determining how easily a semiconductor can accept additional electrons.
The work...