Related Experiment Video
Updated: Apr 28, 2026

Detection of Glycosaminoglycans by Polyacrylamide Gel Electrophoresis and Silver Staining
Published on: February 25, 2021
Molecular characteristics of patients with glycosaminoglycan storage disorders in Russia
Dimitry A Chistiakov1, Kirill V Savost'anov2, Lyudmila M Kuzenkova3
1Department of Medical Nanobiotechnology, Pirogov Russian State Medical University, 117997 Moscow, Russia; Department of Molecular Genetic Diagnostics, Division of Laboratory Medicine, Institute of Pediatrics, Research Center for Children's Health, 119991 Moscow, Russia.

Background:
The mucopolysaccharidoses (MPSs) are rare genetic disorders caused by mutations in lysosomal enzymes involved in the degradation of glycosaminoglycans (GAGs). In this study, we analyzed a total of 48 patients including MPSI (n=6), MPSII (n=18), MPSIIIA (n=11), MPSIVA (n=3), and MPSVI (n=10).
Methods:
In MPS patients, urinary GAGs were colorimetrically assayed. Enzyme activity was quantified by colorimetric and fluorimetric assays. To find mutations, all IDUA, IDS, SGSH, GALNS, and ARSB exons and intronic flanks were sequenced. New mutations were functionally assessed by reconstructing mutant alleles with site-directed mutagenesis followed with expression of wild-type and mutant genetic variants in CHO cells, measuring enzymatic activity, and Western blot analysis of protein expression of normal and mutated enzymes in cell lysates.
Results:
A total of five novel mutations were found including p.Asn348Lys (IDUA) in MPSI, p.Tyr240Cys (GALNS) in MPSIVA, and three ARSB mutations (p.Gln110*, p.Asn262Lysfs*14, and pArg315*) in MPSVI patients. In case of mutations p.Asn348Lys, p.Asn262Lysfs*14, and p.Gln110*, no mutant protein was detected while activity of the mutant protein was <1% of that of the normal enzyme. For p.Tyr240Cys, a trace of mutant protein was observed with a remnant activity of 3.6% of the wild-type GALNS activity. For pArg315*, a truncated 30-kDa protein that had 7.9% of activity of the normal ARSB was detected.
Conclusions:
These data further enrich our knowledge of the genetic background of MPSs.
Insights
This study identified five novel mutations in mucopolysaccharidoses (MPS) patients, enhancing understanding of these rare genetic disorders. The findings detail specific genetic variants and their impact on enzyme function.
Area of Science:
- Biochemistry
- Genetics
- Molecular Biology
Background:
- Mucopolysaccharidoses (MPS) are rare genetic disorders.
- Caused by mutations in lysosomal enzymes crucial for glycosaminoglycans (GAGs) degradation.
- Study analyzed 48 patients across MPS I, II, IIIA, IVA, and VI types.
Purpose of the Study:
- To identify and characterize novel mutations in MPS patients.
- To functionally assess the impact of identified mutations on enzyme activity.
- To expand the knowledge of the genetic basis of various MPS types.
Main Methods:
- Colorimetric and fluorimetric assays for urinary GAGs and enzyme activity.
- Sequencing of exons and intronic flanks for IDUA, IDS, SGSH, GALNS, and ARSB genes.
- Functional assessment using site-directed mutagenesis, cell expression (CHO cells), enzymatic assays, and Western blot analysis.
Main Results:
- Identified five novel mutations: p.Asn348Lys (IDUA), p.Tyr240Cys (GALNS), and three ARSB mutations (p.Gln110*, p.Asn262Lysfs*14, pArg315*).
- Mutations p.Asn348Lys, p.Asn262Lysfs*14, and p.Gln110* resulted in undetectable mutant protein and <1% enzyme activity.
- p.Tyr240Cys showed trace protein with 3.6% activity; pArg315* yielded a truncated protein with 7.9% activity.
Conclusions:
- The identified mutations contribute to the genetic diversity of MPS disorders.
- Functional analyses confirm the pathogenic nature of the novel variants.
- Findings enrich the understanding of genotype-phenotype correlations in MPS.
Related Concept Videos
 01:22
Lysosomal Hydrolases
 01:05
Proteoglycans
 01:23
Glycosaminoglycans
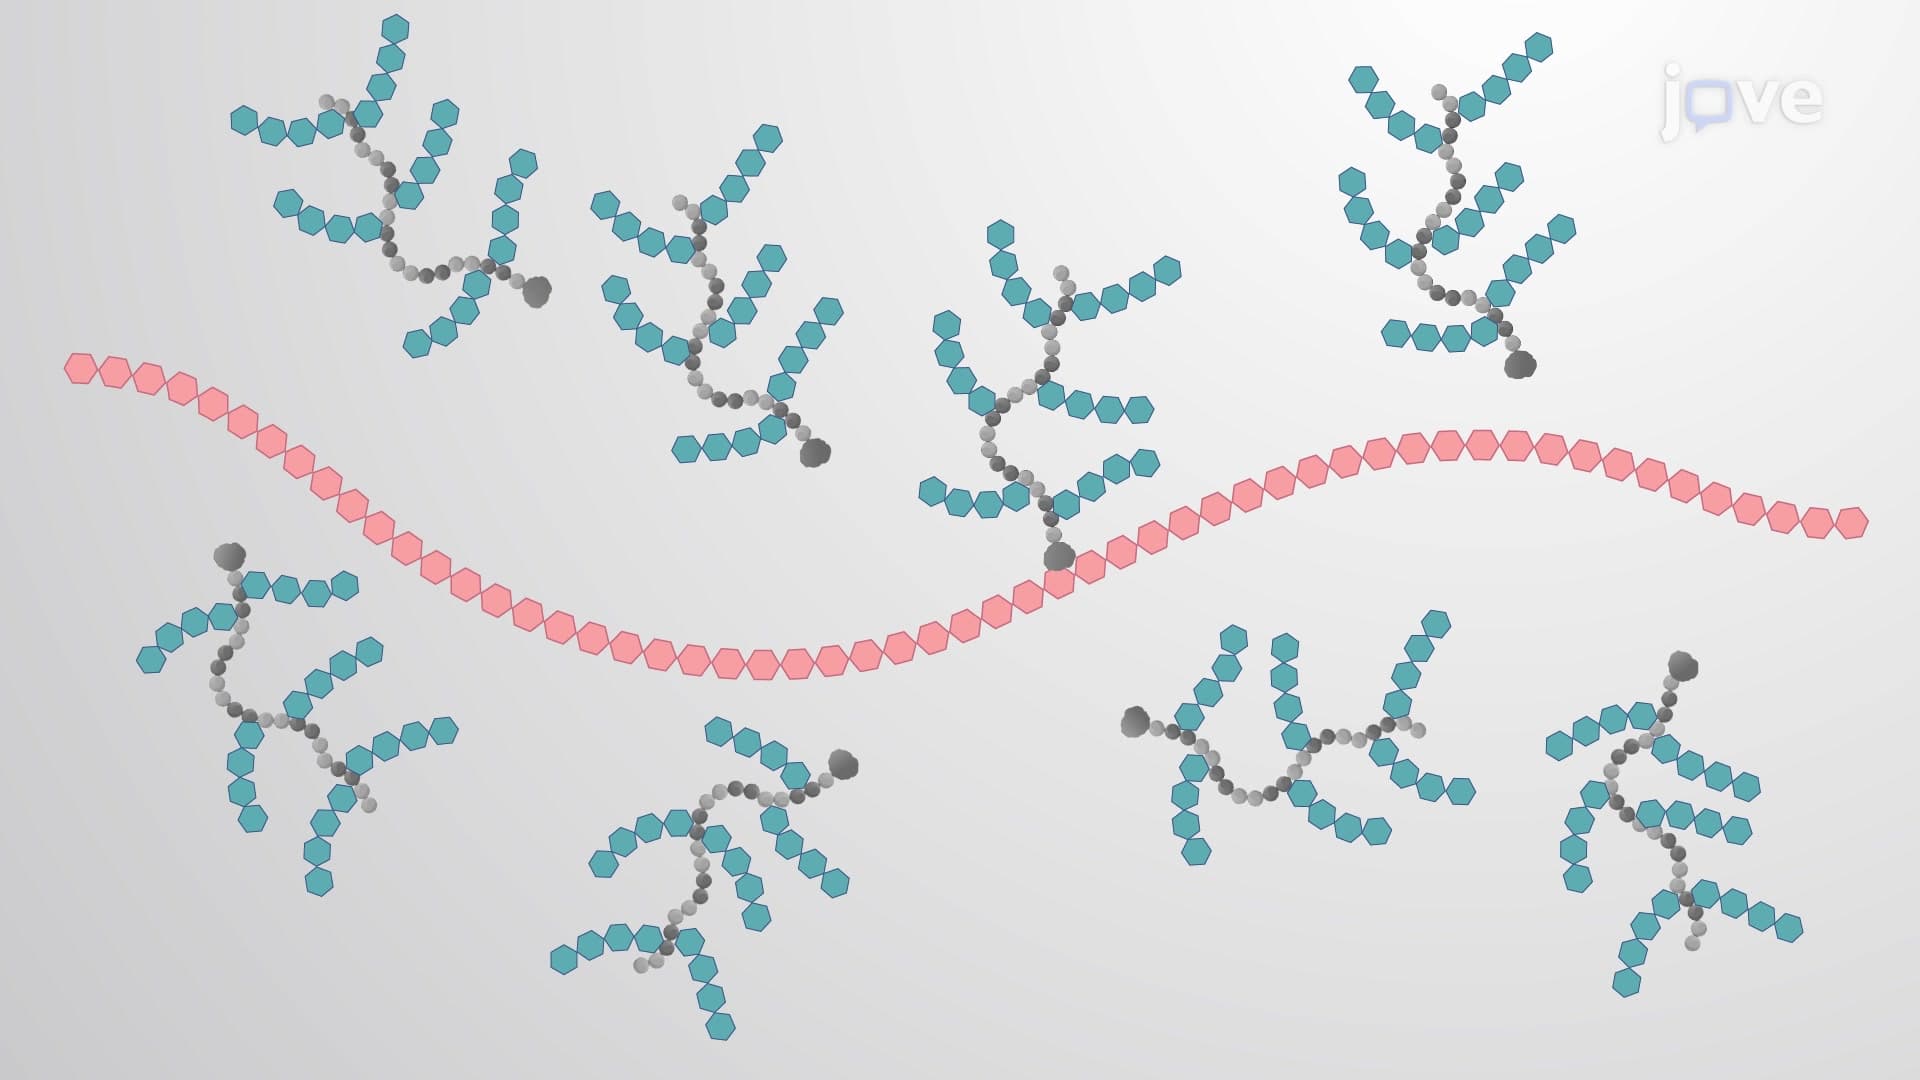
GAGS are found in the extracellular matrix of vertebrates, invertebrates, and bacteria. Due to their polar nature they attract water, and serve as excellent lubricants or shock absorbers in an animal body.
Hyaluronic...
 01:24
Oligosaccharide Assembly
Multiple sugar molecules that may or may...
 01:27
Glucose Transporters
Facilitated diffusion-glucose transporters (GLUTs) are encoded by the solute-linked carrier (SLC) family 2, subfamily A gene family, or SLC2A. The 14 GLUT protein members are distributed into three classes:
 01:25
Protein Glycosylation
Glycosylation occurs in...