Related Experiment Video
Updated: Apr 28, 2026


Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
Molecular basis of agonist docking in a human GPR103 homology model by site-directed mutagenesis and
1Inserm U982, Laboratory of Neuronal and Neuroendocrine Cell Differentiation and Communication, Neurotrophic Factors and Neuronal Differentiation Team, Institute for Research and Innovation in Biomedicine (IRIB); Cell Imaging Platform of Normandy (PRIMACEN), IRIB; Normandie Univ, France.

Background And Purpose:
The neuropeptide 26RFa and its cognate receptor GPR103 are involved in the control of food intake and bone mineralization. Here, we have tested, experimentally, the predicted ligand-receptor interactions by site-directed mutagenesis of GPR103 and designed point-substituted 26RFa analogues.
Experimental Approach:
Using the X-ray structure of the β2 -adrenoceptor, a 3-D molecular model of GPR103 has been built. The bioactive C-terminal octapeptide 26RFa(19-26) , KGGFSFRF-NH2 , was docked in this GPR103 model and the ligand-receptor complex was submitted to energy minimization.
Key Results:
In the most stable complex, the Phe-Arg-Phe-NH2 part was oriented inside the receptor cavity, whereas the N-terminal Lys residue remained outside. A strong intermolecular interaction was predicted between the Arg(25) residue of 26RFa and the Gln(125) residue located in the third transmembrane helix of GPR103. To confirm this interaction experimentally, we tested the ability of 26RFa and Arg-modified 26RFa analogues to activate the wild-type and the Q125A mutant receptors transiently expressed in CHO cells. 26RFa (10(-6) M) enhanced [Ca(2+) ]i in wild-type GPR103-transfected cells, but failed to increase [Ca(2+) ]i in Q125A mutant receptor-expressing cells. Moreover, asymmetric dimethylation of the side chain of arginine led to a 26RFa analogue, [ADMA(25) ]26RFa(20-26) , that was unable to activate the wild-type GPR103, but antagonized 26RFa-evoked [Ca(2+) ]i increase.
Conclusion And Implications:
Altogether, these data provide strong evidence for a functional interaction between the Arg(25) residue of 26RFa and the Gln(125) residue of GPR103 upon ligand-receptor activation, which can be exploited for the rational design of potent GPR103 agonists and antagonists.

Related Concept Videos
 01:29
The Two-State Receptor Model
The binding affinity of a drug determines its interaction with...
 01:15
G Protein-coupled Receptors
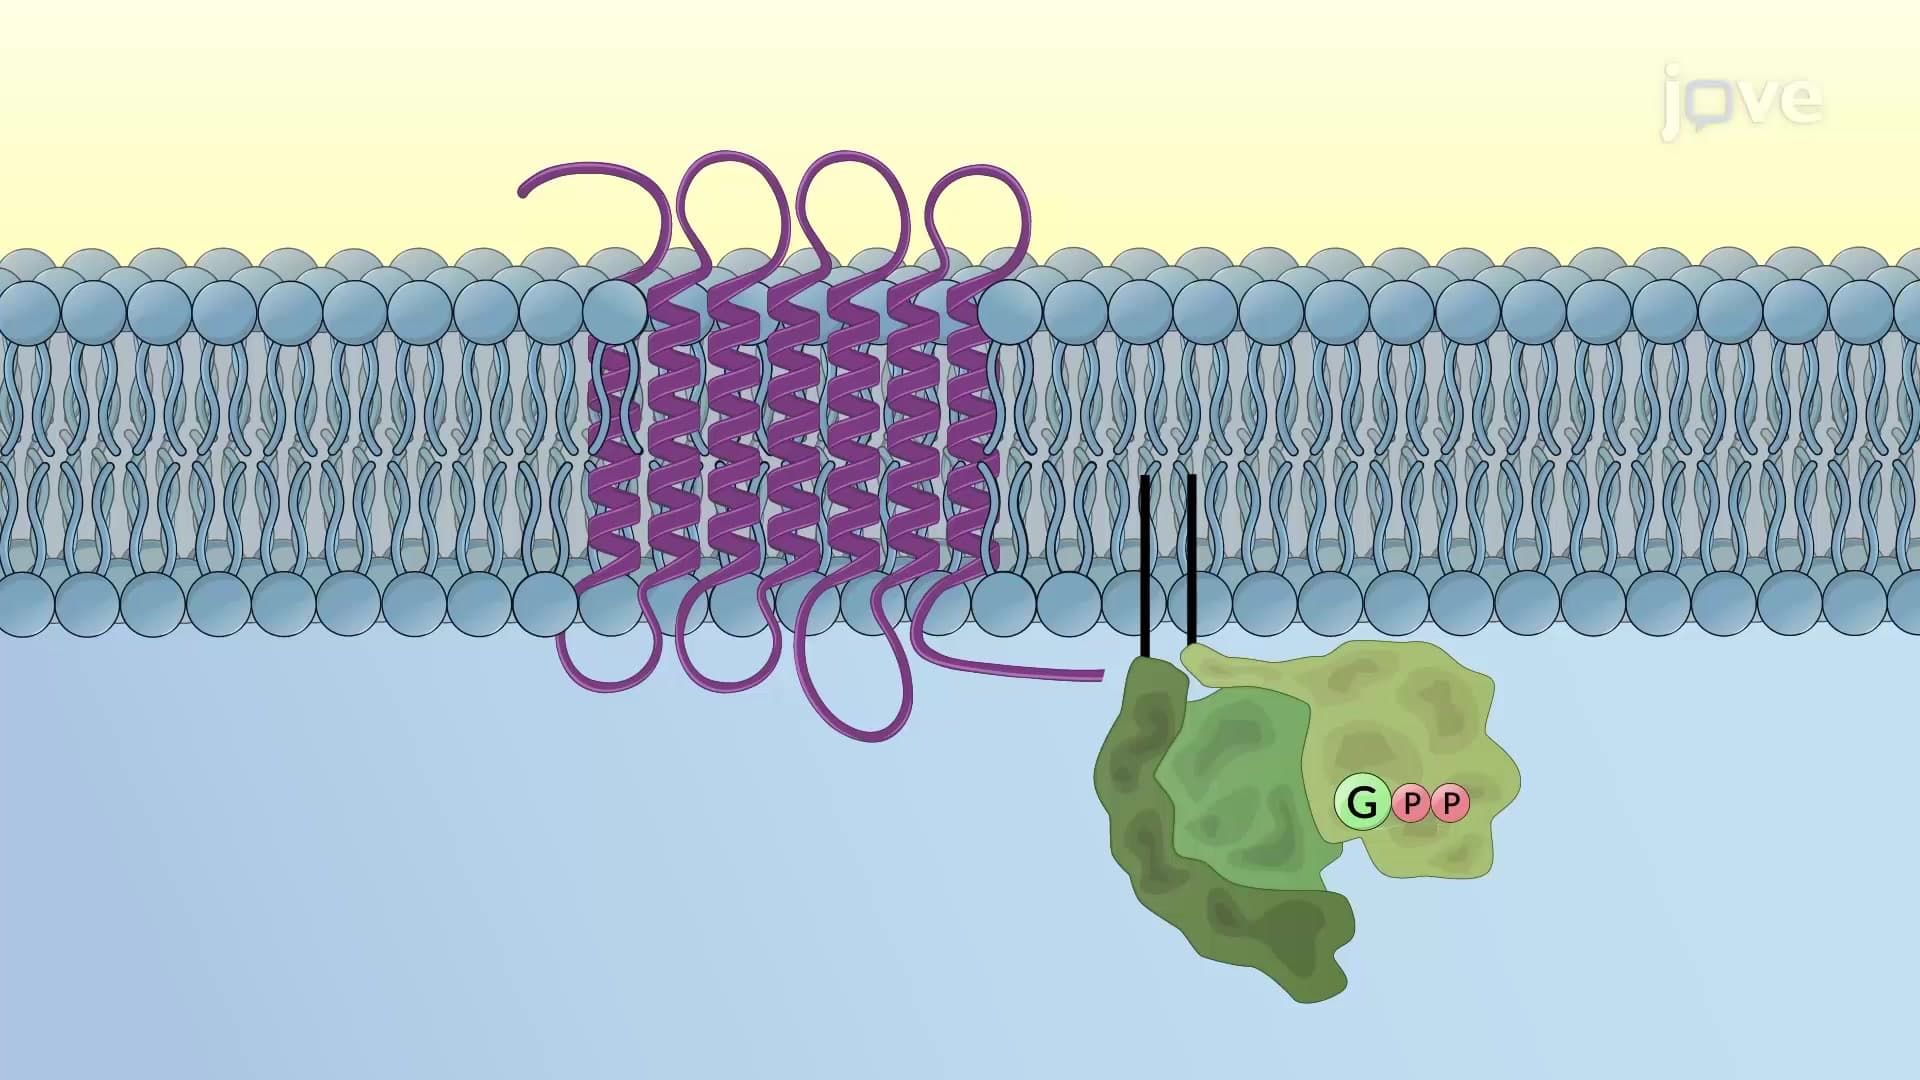
GPCRs are also called heptahelical, 7TM, or serpentine receptors, and consist of seven (H1-H7) transmembrane alpha-helices that span the bilayer to form a cylindrical core. The transmembrane helices are connected by three extracellular loops and three...
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
 00:49
Ligand Binding and Linkage
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 01:25
Drug-Receptor Interaction: Agonist
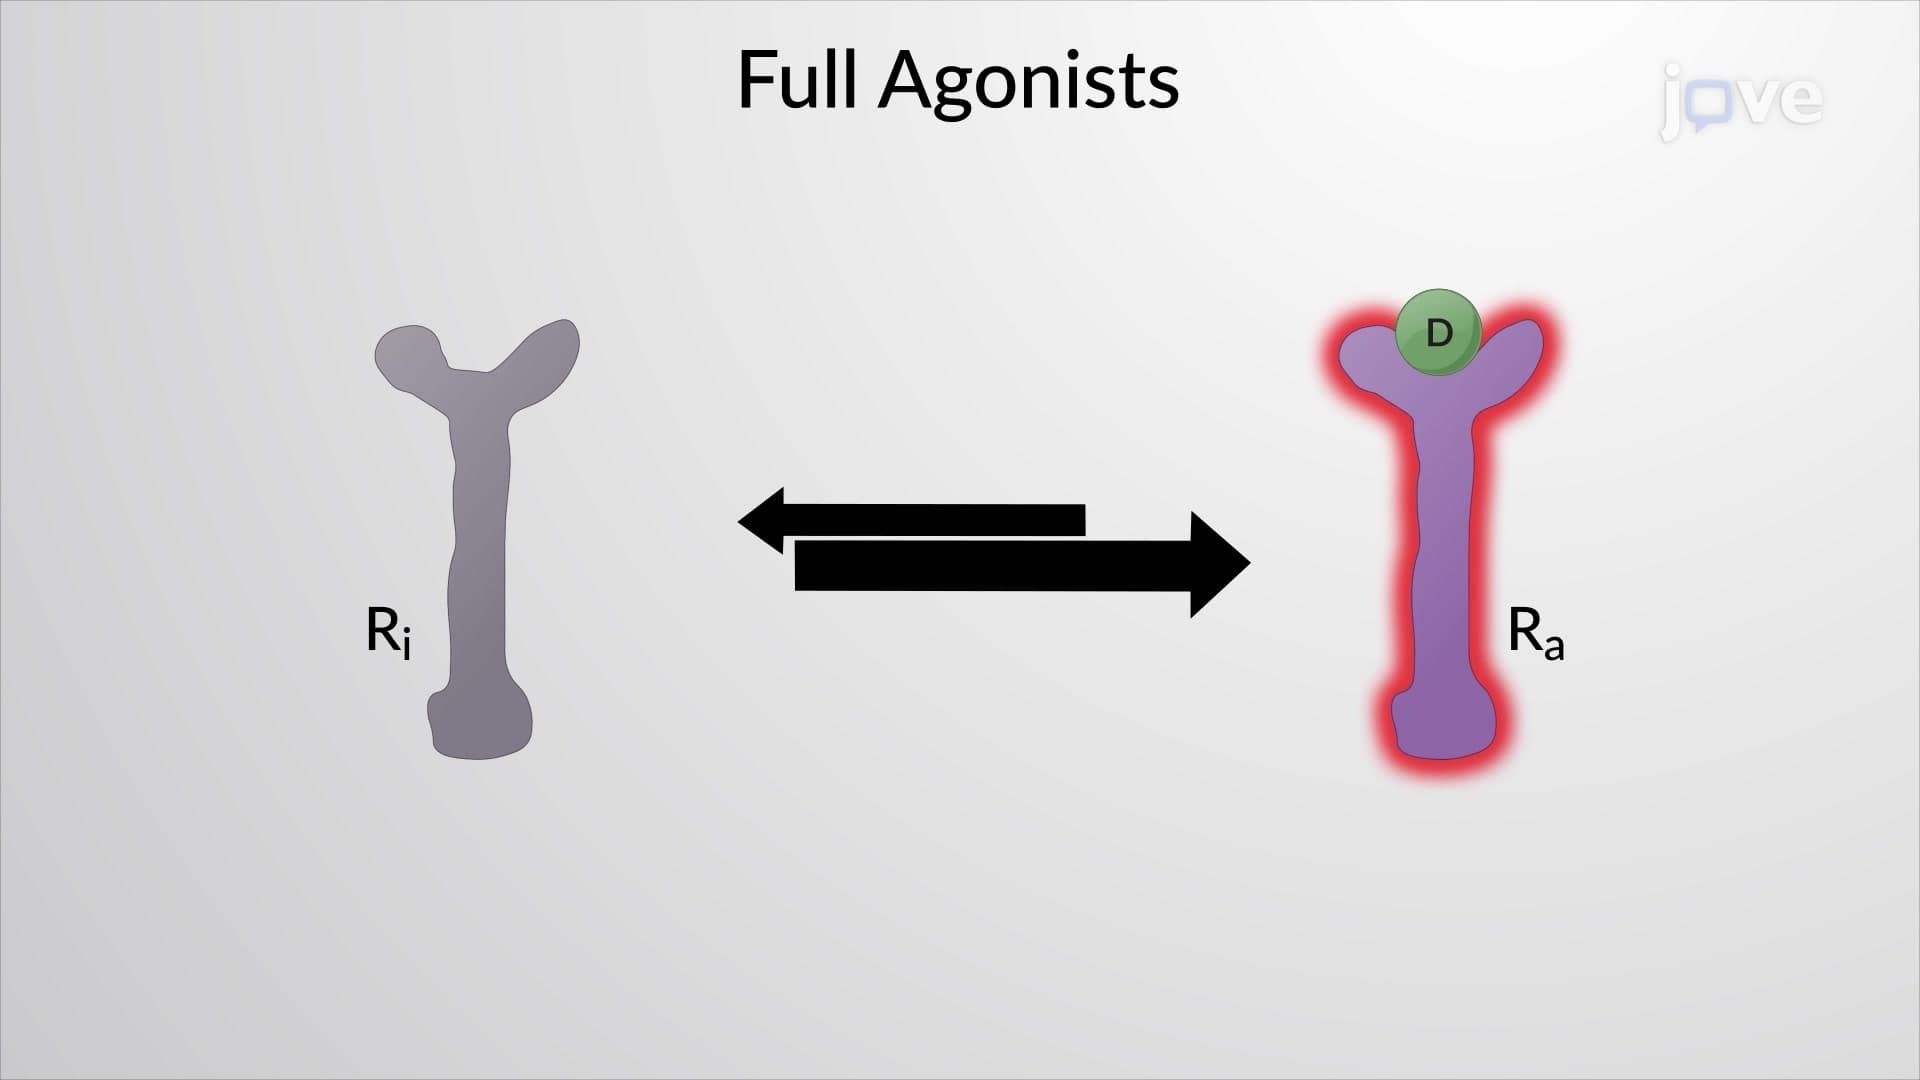
Agonists can bind to receptors in different ways. Some agonists bind directly to the receptor's active site, mimicking the endogenous...