Related Experiment Video
Updated: Apr 27, 2026


In Vivo Modeling of the Morbid Human Genome using Danio rerio
Published on: August 24, 2013
Integrating EMR-linked and in vivo functional genetic data to identify new genotype-phenotype associations
Jonathan D Mosley1, Sara L Van Driest2, Peter E Weeke1
1Department of Medicine, Vanderbilt University, Nashville, Tennessee, United States of America.

Abstract:
The coupling of electronic medical records (EMR) with genetic data has created the potential for implementing reverse genetic approaches in humans, whereby the function of a gene is inferred from the shared pattern of morbidity among homozygotes of a genetic variant. We explored the feasibility of this approach to identify phenotypes associated with low frequency variants using Vanderbilt's EMR-based BioVU resource. We analyzed 1,658 low frequency non-synonymous SNPs (nsSNPs) with a minor allele frequency (MAF)<10% collected on 8,546 subjects. For each nsSNP, we identified diagnoses shared by at least 2 minor allele homozygotes and with an association p<0.05. The diagnoses were reviewed by a clinician to ascertain whether they may share a common mechanistic basis. While a number of biologically compelling clinical patterns of association were observed, the frequency of these associations was identical to that observed using genotype-permuted data sets, indicating that the associations were likely due to chance. To refine our analysis associations, we then restricted the analysis to 711 nsSNPs in genes with phenotypes in the On-line Mendelian Inheritance in Man (OMIM) or knock-out mouse phenotype databases. An initial comparison of the EMR diagnoses to the known in vivo functions of the gene identified 25 candidate nsSNPs, 19 of which had significant genotype-phenotype associations when tested using matched controls. Twleve of the 19 nsSNPs associations were confirmed by a detailed record review. Four of 12 nsSNP-phenotype associations were successfully replicated in an independent data set: thrombosis (F5,rs6031), seizures/convulsions (GPR98,rs13157270), macular degeneration (CNGB3,rs3735972), and GI bleeding (HGFAC,rs16844401). These analyses demonstrate the feasibility and challenges of using reverse genetics approaches to identify novel gene-phenotype associations in human subjects using low frequency variants. As increasing amounts of rare variant data are generated from modern genotyping and sequence platforms, model organism data may be an important tool to enable discovery.
More Related Videos
 11:35
Screening for Functional Non-coding Genetic Variants Using Electrophoretic Mobility Shift Assay EMSA and DNA-affinity Precipitation Assay DAPA
Published on: August 21, 2016
 09:37
Navigating MARRVEL, a Web-Based Tool that Integrates Human Genomics and Model Organism Genetics Information
Published on: August 15, 2019
Related Concept Videos
 01:29
Pharmacogenomics: Identification of New Drug Targets
 01:11
Genome-wide Association Studies-GWAS
GWAS does not require the identification of the target gene involved in...
 01:29
Pharmacogenetic Phenotypes: Alterations in Pharmacokinetics, Drug Targets and Biologic Milieu
 02:46
Genetic Screens
Forward genetic screens
Forward or “classical” genetic screens involve creating random mutations in an organism’s DNA using radiation, mutagens, or insertion of additional bases, which...
 01:29
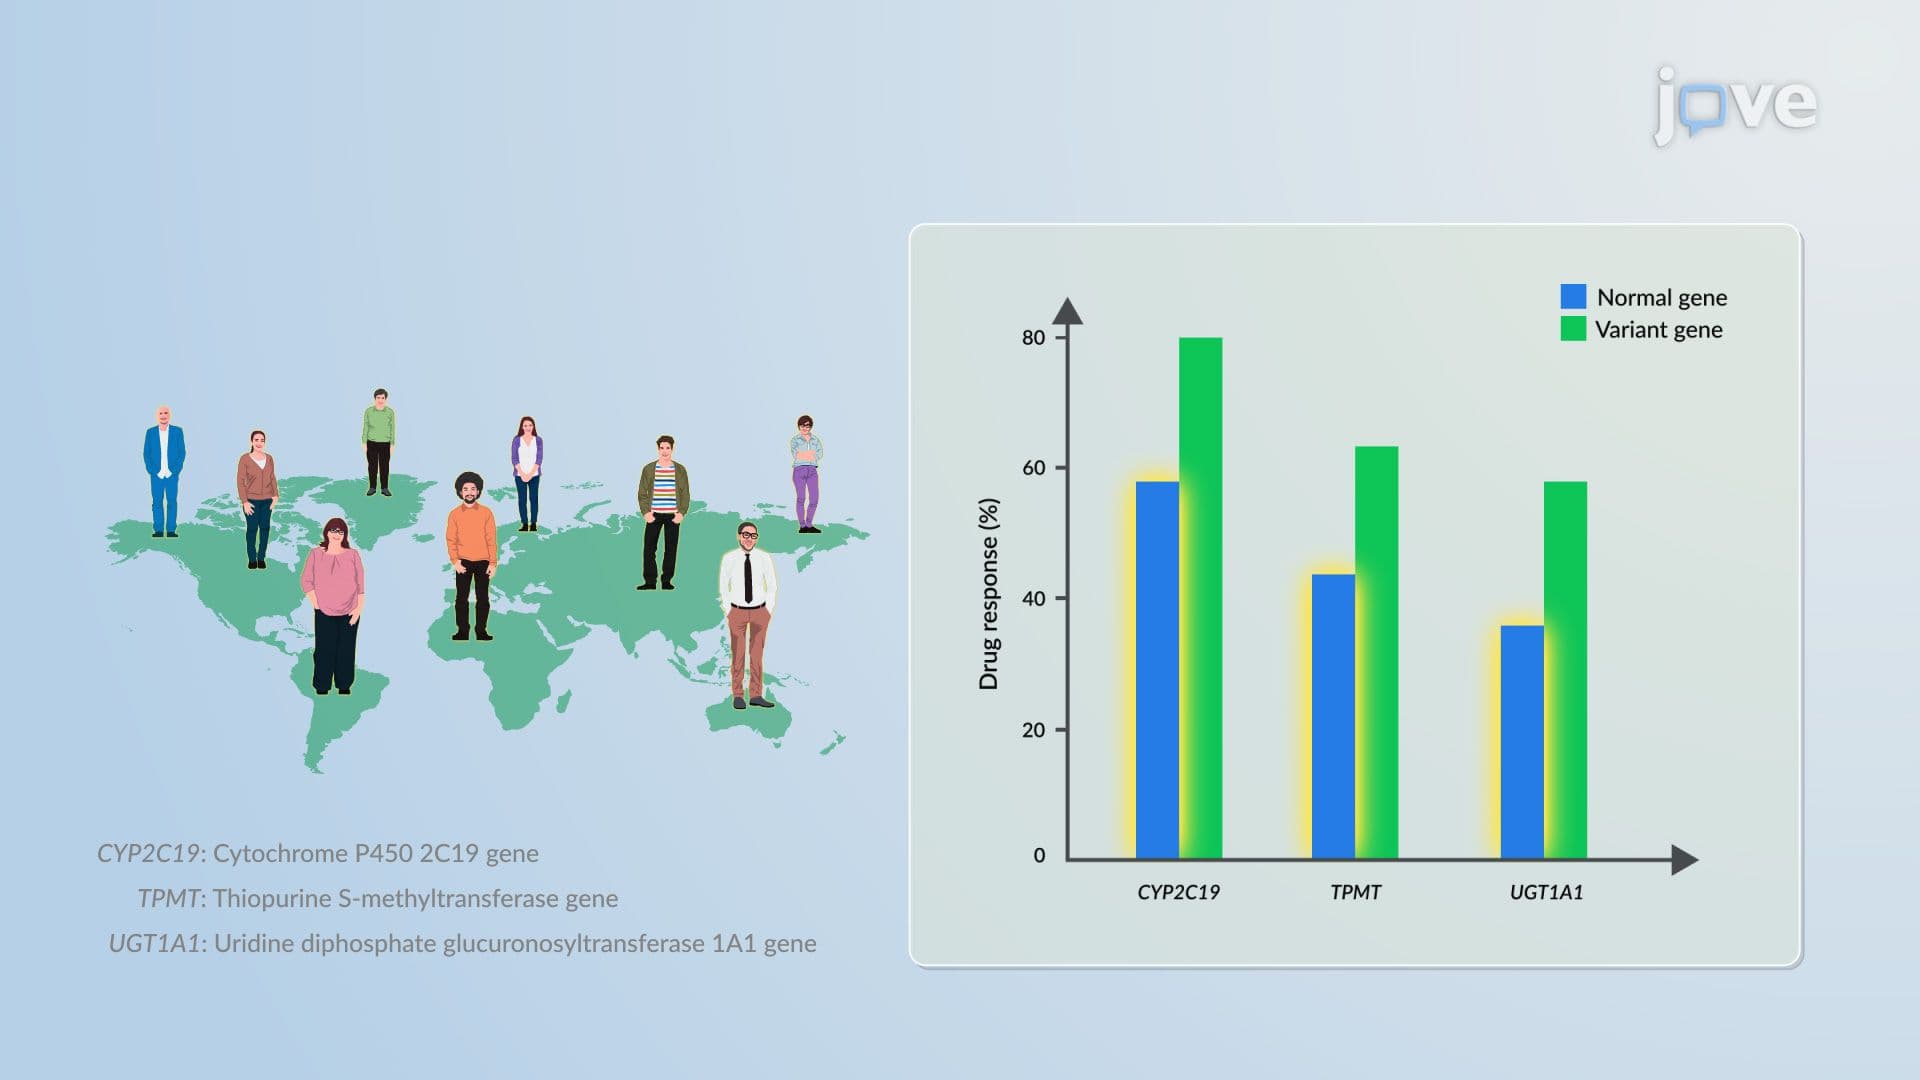
Pharmacogenetics and Pharmacogenomics: Overview
 01:27
Principles of Pharmacogenetics: Types of Genetic Variants