Related Experiment Video
Updated: Apr 27, 2026

Quantitative Atomic-Site Analysis of Functional Dopants/Point Defects in Crystalline Materials by Electron-Channeling-Enhanced Microanalysis
Published on: May 10, 2021
Spectroscopic distinctions between two types of Ce(3+) ions in X2-Y2SiO5: a theoretical investigation
Jun Wen1, Chang-Kui Duan, Lixin Ning
1School of Physics and Electronic Engineering, Anqing Normal University , Anqing 246011, People's Republic of China.

Abstract:
The Ce(3+) ions occupying the two crystallographically distinct Y(3+) sites both with C1 point group symmetry in the X2-Y2SiO5 (X2-YSO) crystal are discriminated by their spectroscopic properties calculated with ab initio approaches and phenomenological model analyses. Density functional theory (DFT) calculations with the supercell approach are performed to obtain the local structures of Ce(3+), based on which the wave function-based embedded cluster calculations at the CASSCF/CASPT2 level are carried out to derive the 4f → 5d transition energies. From the ab initio calculated energy levels and wave functions, the crystal-field parameters (CFPs) and the anisotropic g-factor tensors of Ce(3+) are extracted. The theoretical results agree well with available experimental data. The structural and spectroscopic properties for the two types of Ce(3+) ions in X2-YSO are thus distinguished in terms of the calculated local atomic structures, 4f → 5d transition energies, and spectral parameters.
More Related Videos
 07:24
Hyperspectral Imaging as a Tool to Study Optical Anisotropy in Lanthanide-Based Molecular Single Crystals
Published on: April 14, 2020
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Related Concept Videos
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than...
 03:26
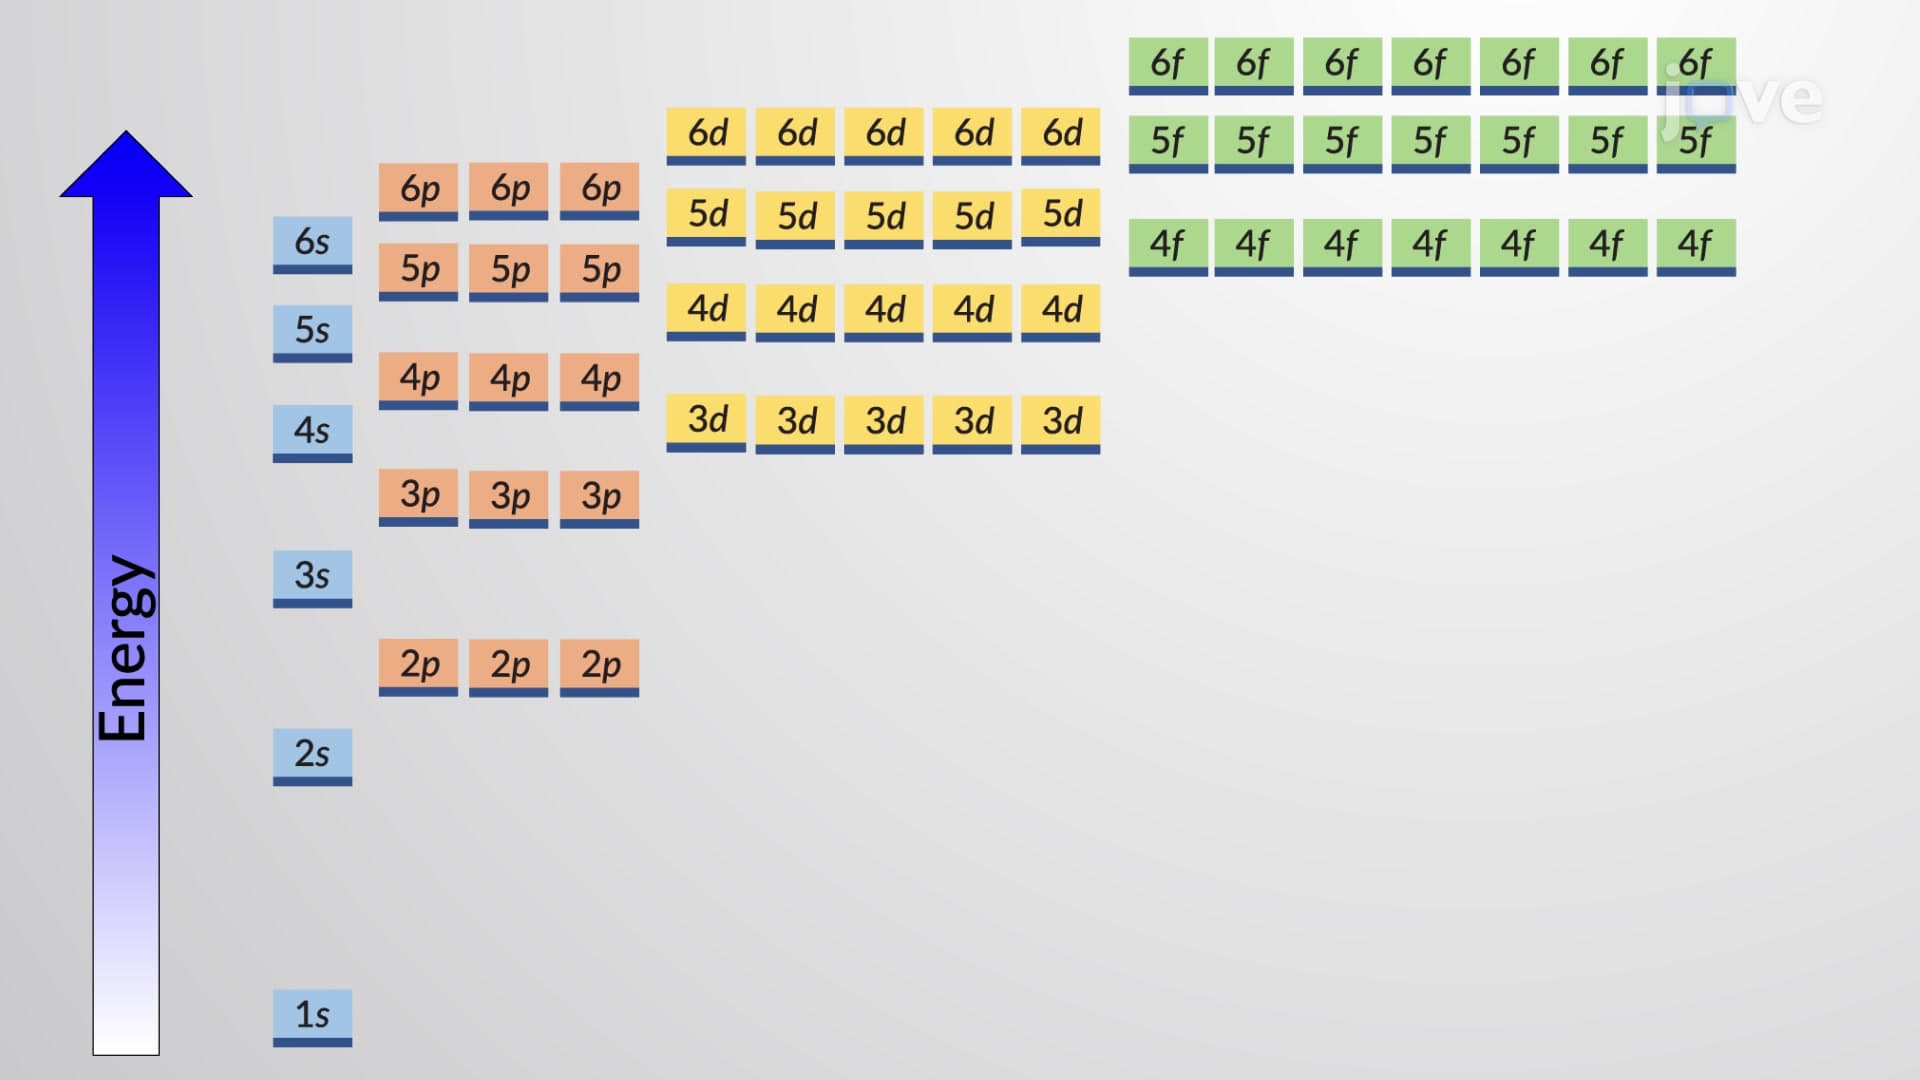
Electron Configuration of Multielectron Atoms
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:42
Valence Bond Theory
 02:54
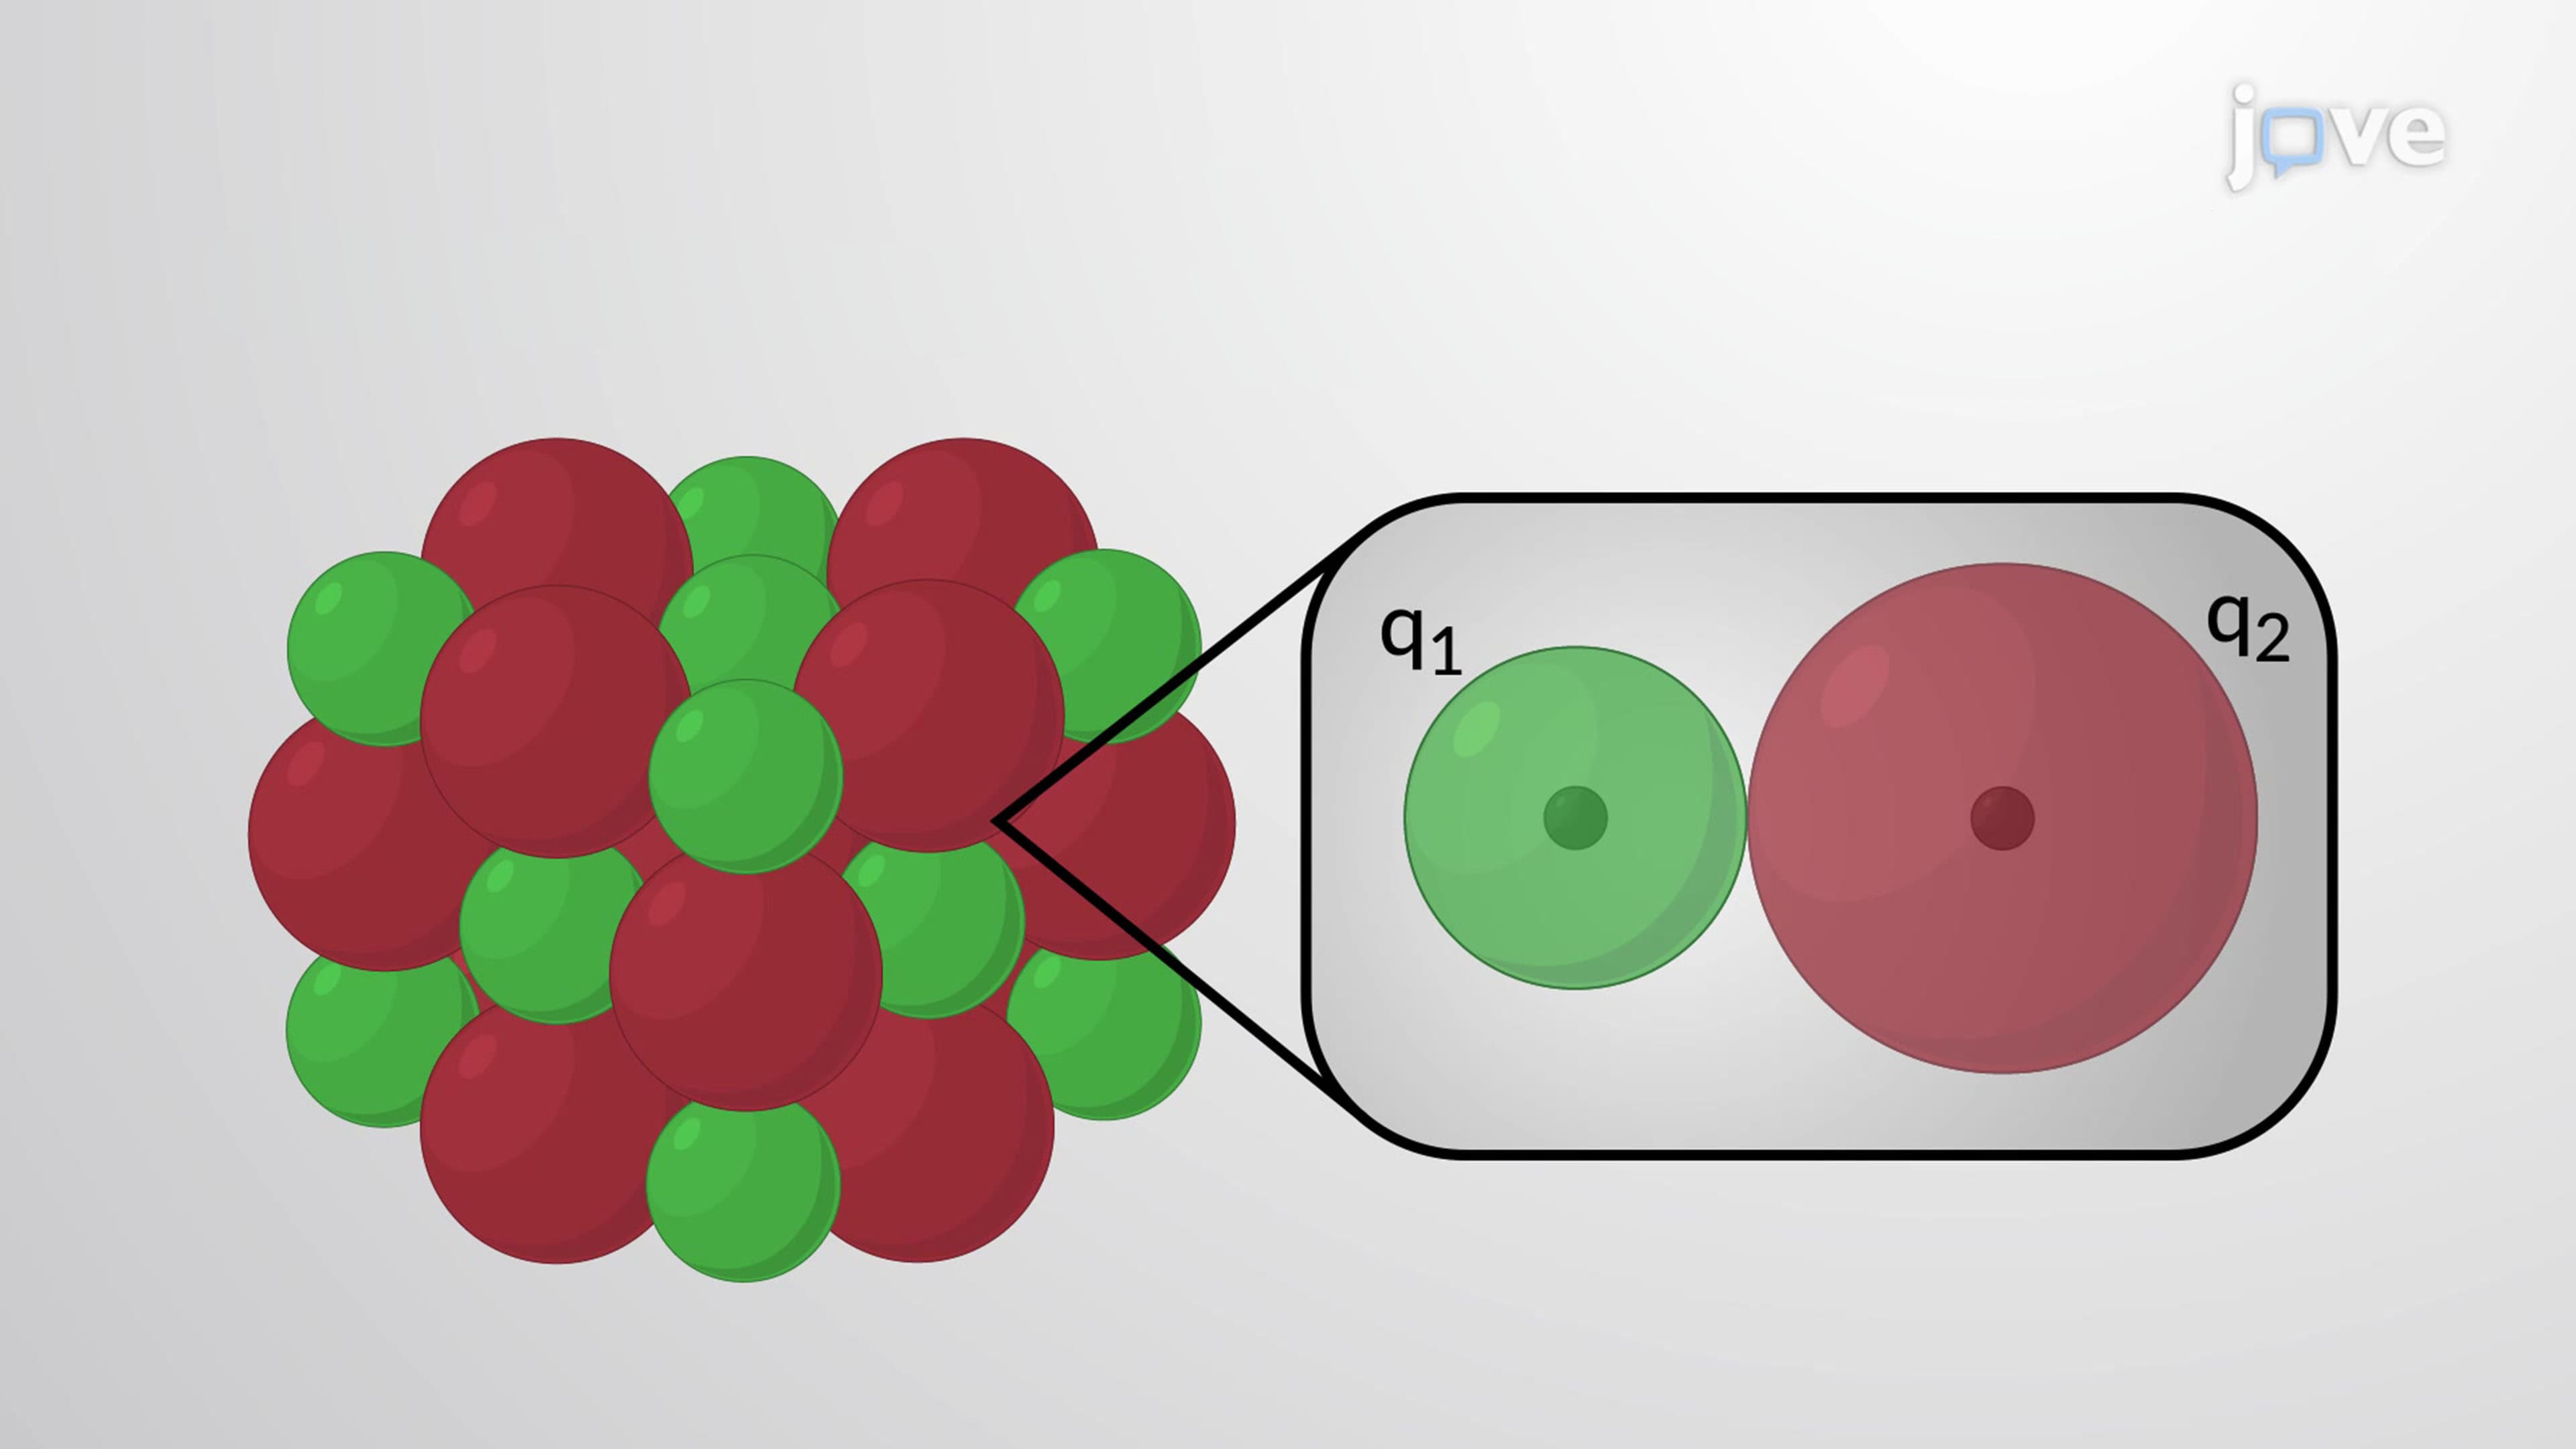
Trends in Lattice Energy: Ion Size and Charge
 03:02
Colors and Magnetism
When atoms or molecules absorb light at the proper frequency, their electrons are excited to higher-energy orbitals. For many main group atoms and molecules, the absorbed photons are in the ultraviolet range of the electromagnetic spectrum, which cannot be detected by the human eye. For coordination compounds, the energy difference between the d orbitals often allows photons in the visible range to be absorbed and emitted, which is seen as colors by the human...