Related Experiment Video
Updated: Apr 27, 2026

Comprehensive Characterization of Extended Defects in Semiconductor Materials by a Scanning Electron Microscope
Published on: May 28, 2016
Efficient and accurate treatment of weak pairs in local CCSD(T) calculations. II. Beyond the ring approximation
Martin Schütz1, Oliver Masur1, Denis Usvyat1
1Institut für Physikalische und Theoretische Chemie, Universität Regensburg, Regensburg D-93040, Germany.

Abstract:
In order to arrive at linear scaling of the computational cost with molecular size, local coupled cluster methods discriminate pairs of local molecular orbitals according to the spatial separation R of the latter. Only strong pairs are treated at the full coupled cluster level, whereas for weak pairs a lower level of theory (usually Møller-Plesset perturbation theory of second order, MP2) is used. Yet an MP2 treatment of weak pairs is inadequate in certain situations (for example, for describing π-stacking), which calls for an improved but still inexpensive method for dealing with the weak pairs. In a previous contribution, we proposed as a substituent for MP2 the LrCCD3 method, which is based on ring coupled cluster doubles (ring-CCD) and includes all third-order diagrams with energy contributions decaying not quicker than R(-6). In the present work, we explore a still more accurate method, which is based on the same principles. It turned out to be essential to abandon the restriction to ring-CCD, i.e., to include further CCD diagrams beyond the ring approximation. The occurring intermediates turn out to be formally very similar to LMP2 density matrices, such that an efficient evaluation of these non-ring CCD diagrams is possible. Furthermore, a computationally cheap a posteriori estimate for the fourth-order singles contribution to the weak pair energy, which also exhibits a decay behavior of R(-6), is introduced. The resulting method, denoted as LCCD[S]-R(-6), indeed provides a substantial improvement in accuracy over the previous LrCCD3 method at a relatively modest additional computational cost.

Related Concept Videos
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than...
 01:22
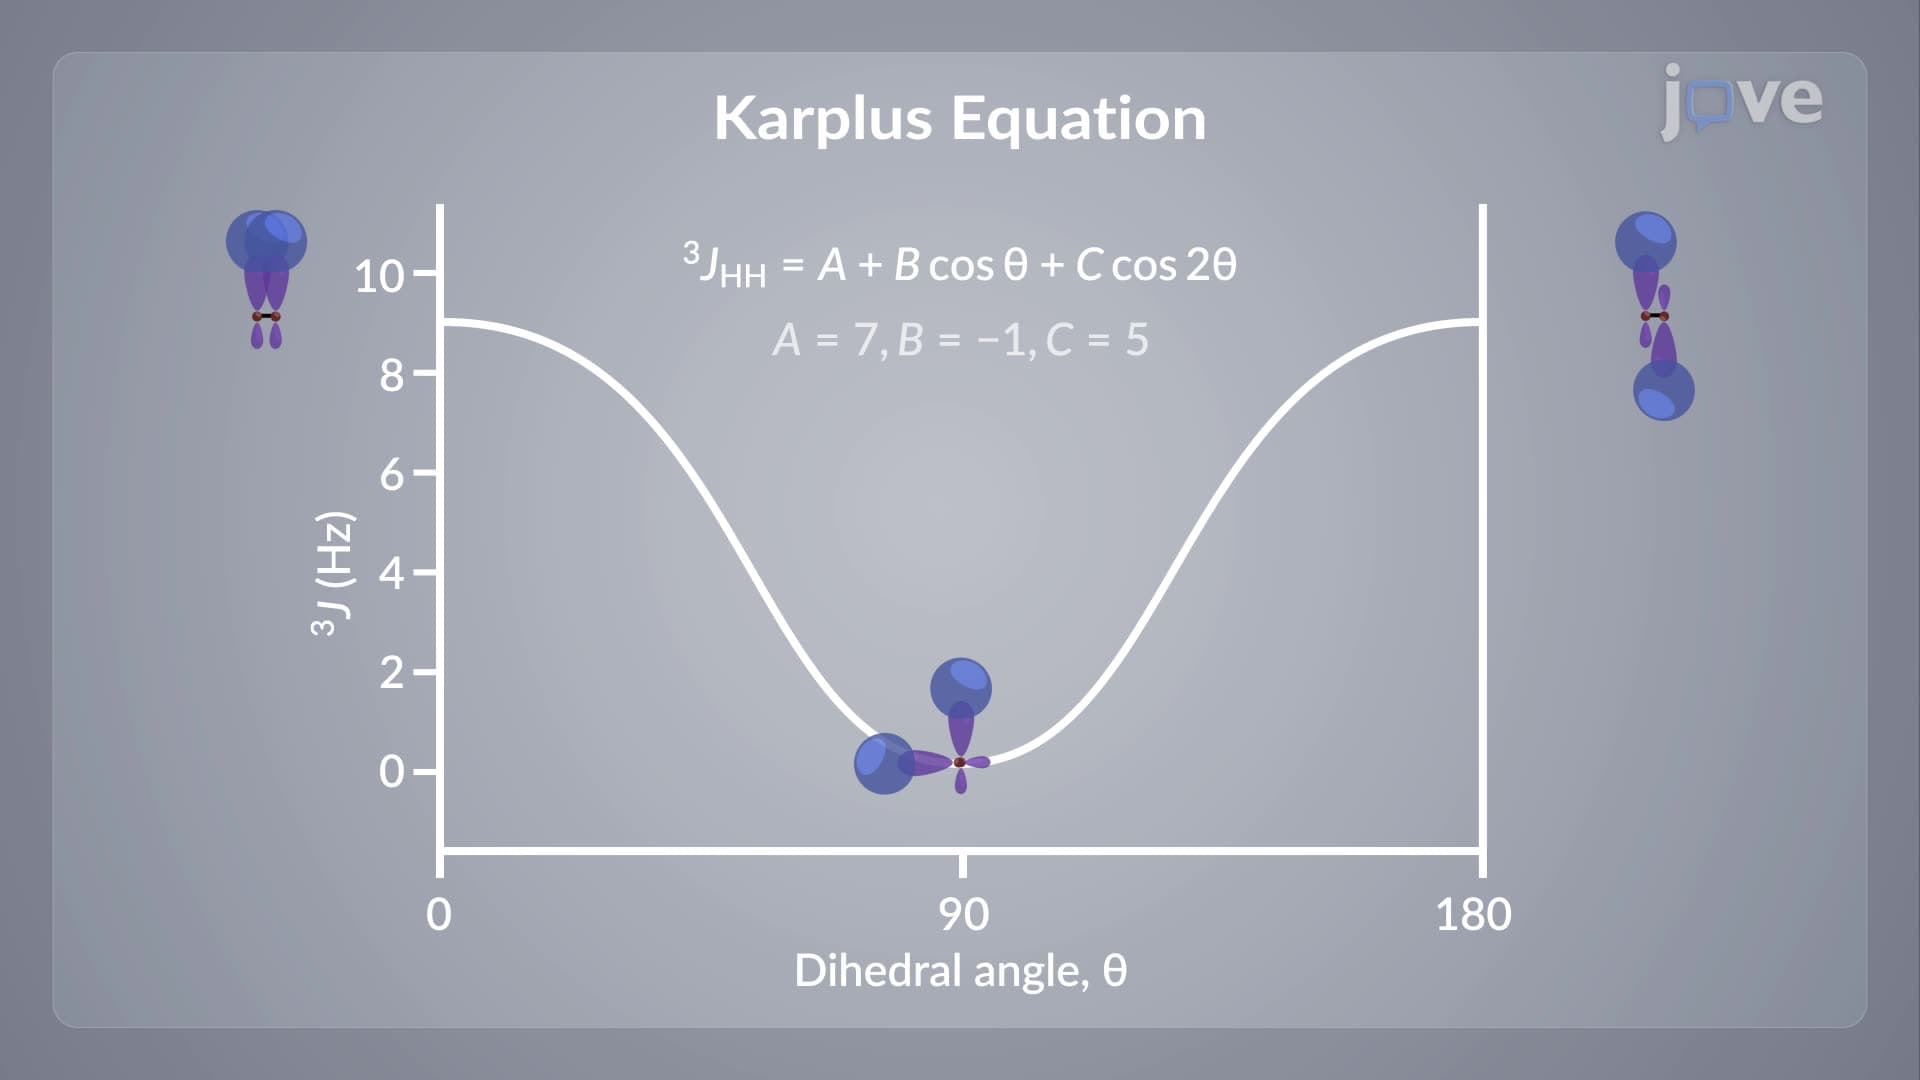
Spin–Spin Coupling: Three-Bond Coupling (Vicinal Coupling)
The extent of coupling depends on the C‑C bond length, the two H‑C‑C angles, any electron-withdrawing substituents, and the dihedral angle between the...
 03:21
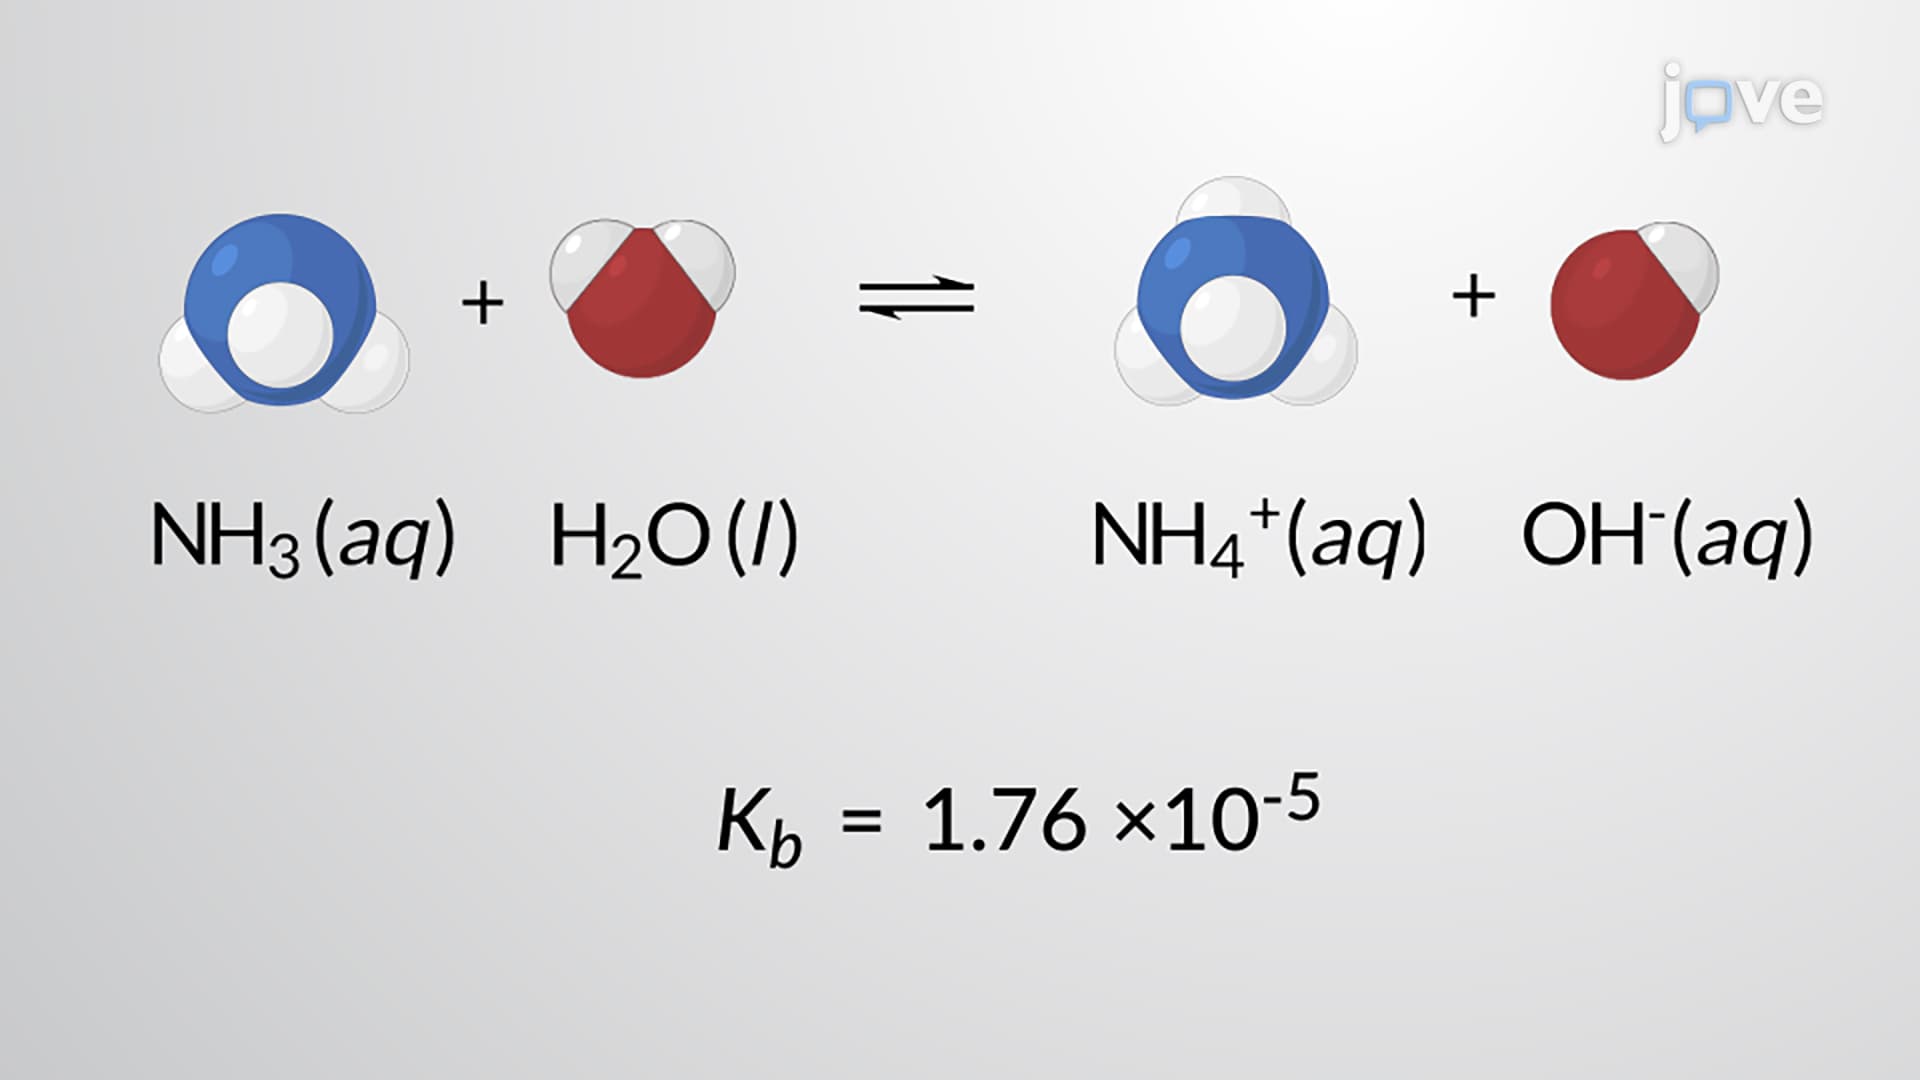
Weak Base Solutions
 01:20
Spin–Spin Coupling: Two-Bond Coupling (Geminal Coupling)
The central atom need not be NMR-active because its electrons are affected by the electron polarization of the spin-active atoms. However, spin information is transmitted less effectively than in one-bond coupling, and 2J values are usually weaker than 1J values. The energy of...
 01:27
¹H NMR: Long-Range Coupling
In alkenes, spin information is communicated via σ–π overlap, as seen in allylic (four-bond) and homoallylic (five-bond) couplings. These coupling interactions are stronger when the σ bond is parallel to the alkene...
 01:34
Woodward–Hoffmann Selection Rules and Microscopic Reversibility