Related Experiment Video
Updated: Oct 21, 2026
Modeling Encephalopathy of Prematurity Using Prenatal Hypoxia-ischemia with Intra-amniotic Lipopolysaccharide in Rats
Published on: November 20, 2015
Neonatal pyruvate dehydrogenase deficiency with lipoate responsive lactic acidaemia and hyperammonaemia
D J Byrd1, H P Krohn, L Winkler
1Kinderklinik der Medizinischen Hochschule, Hannover, Federal Republic of Germany.

Abstract:
A 2-day-old girl developed a severe lactic acidosis with a normal lactate/pyruvate ratio and hyperammonaemia. Plasma arginine and citrulline levels were below the limit of detection. In muscle total pyruvate dehydrogenase complex (PDHC) and pyruvate decarboxylase (E1) activities were reduced to a fraction of lower control values. The acute neonatal period was bridged with peritoneal dialysis, dichloroacetate therapy, supplements of arginine and branched chain amino acids, a complete vitamin B complex and lipoic acid. Lactate homeostasis responded to pharmacological supplements of lipoic acid. At age 1 year the child was hypotonic, showed severe developmental retardation, optic atrophy and cranial dysmorphism. She died aged 1 year 8 months with signs of respiratory paralysis but with normal lactate levels under assisted breathing. Pathological findings at autopsy were suggestive of Leigh syndrome, interstitial pneumonia and extensive fatty infiltration of hepatocytes. Regression analysis of data from 187 plasma amino acid determinations from the patient over a period of 1 year 8 months revealed a persistent-imbalance involving alanine, glutamic acid, glutamine, proline, citrulline and branched chain amino acids. Aspects of acute and long-term therapy in this patient and some implications of the imbalances in plasma amino acids are discussed.
More Related Videos
Related Concept Videos
 01:15
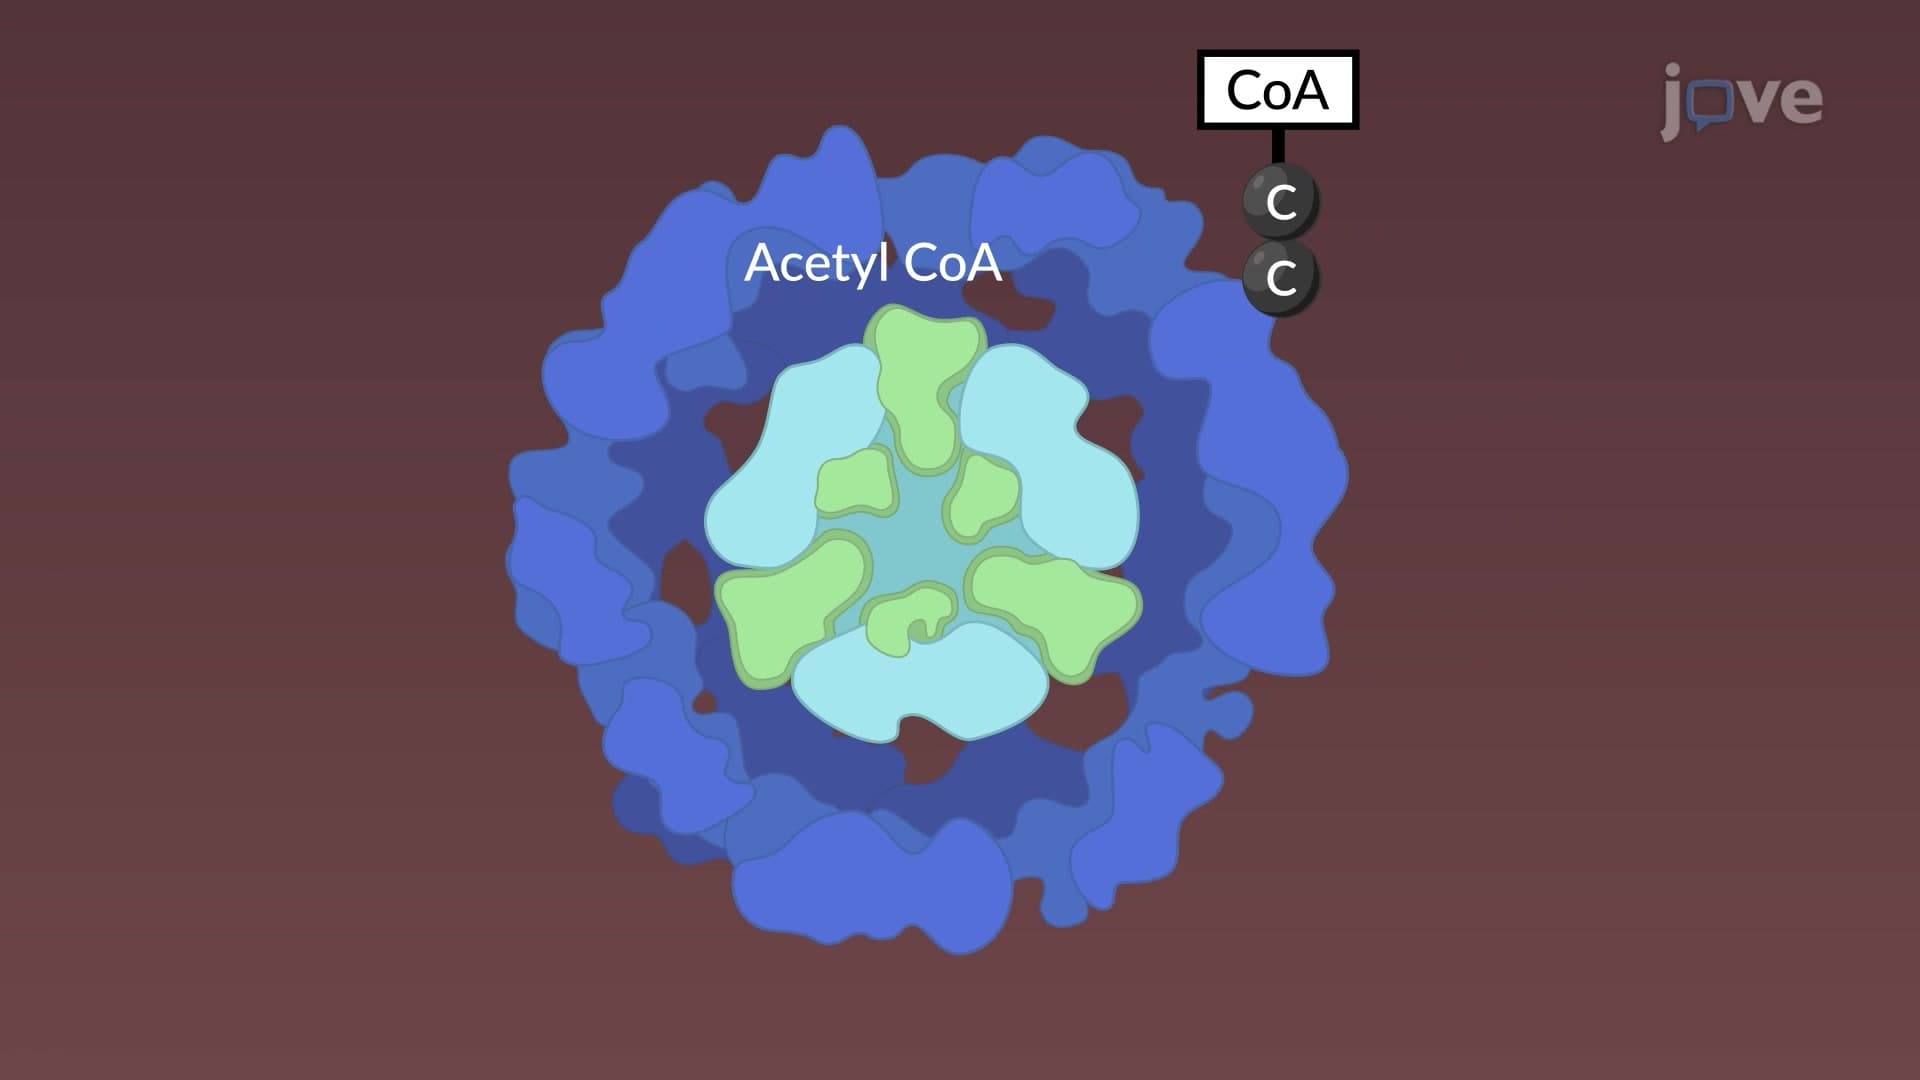
Pyruvate Oxidation
First, the enzyme pyruvate dehydrogenase removes the carboxyl group from pyruvate and releases it as carbon dioxide. The stripped molecule is then oxidized and releases electrons, which are then picked up by NAD+...
 01:23
Urea Cycle
 01:22
Lysosomal Hydrolases
 01:20
Fates of Pyruvate
In aerobic organisms, pyruvate is metabolized via the citric acid cycle to produce reduced coenzymes NADH and FADH2. These coenzymes are then oxidized in the electron transport chain to produce ATP and, in the process, regenerate the NAD+ and FAD. As seen in some cell types and organisms, fermentation...
 01:20
Inborn Errors of Metabolism
 01:29
Hepatic Encephalopathy