Related Experiment Video
Updated: Apr 27, 2026

Methods for the Discovery of Novel Compounds Modulating a Gamma-Aminobutyric Acid Receptor Type A Neurotransmission
Published on: August 16, 2018
Analysis of γ-aminobutyric acid (GABA) type A receptor subtypes using isosteric and allosteric ligands
1Department of Molecular and Medical Pharmacology, David Geffen School of Medicine at UCLA, Room CHS 23-120, 650 Young Drive South, Los Angeles, CA, 90095-1735, USA, rolsen@mednet.ucla.edu.

Abstract:
The GABAA receptors (GABAARs) play an important role in inhibitory transmission in the brain. The GABAARs could be identified using a medicinal chemistry approach to characterize with a series of chemical structural analogues, some identified in nature, some synthesized, to control the structural conformational rigidity/flexibility so as to define the 'receptor-specific' GABA agonist ligand structure. In addition to the isosteric site ligands, these ligand-gated chloride ion channel proteins exhibited modulation by several chemotypes of allosteric ligands, that help define structure and function. The channel blocker picrotoxin identified a noncompetitive channel blocker site in GABAARs. This ligand site is located in the transmembrane channel pore, whereas the GABA agonist site is in the extracellular domain at subunit interfaces, a site useful for low energy coupled conformational changes of the functional channel domain. Also in the trans-membrane domain are allosteric modulatory ligand sites, mostly positive, for diverse chemotypes with general anesthetic efficacy, namely, the volatile and intravenous agents: barbiturates, etomidate, propofol, long-chain alcohols, and neurosteroids. The last are apparent endogenous positive allosteric modulators of GABAARs. These binding sites depend on the GABAAR heteropentameric subunit composition, i.e., subtypes. Two classes of pharmacologically very important allosteric modulatory ligand binding site reside in the extracellular domain at modified agonist sites at other subunit interfaces: the benzodiazepine site, and the low-dose ethanol site. The benzodiazepine site is specific for certain subunit combination subtypes, mainly synaptically localized. In contrast, the low-dose (high affinity) ethanol site(s) is found at a modified benzodiazepine site on different, extrasynaptic, subtypes.
Related Concept Videos
 01:30
Ligand-Gated Ion Channel Receptor: Gating Mechanism
 01:31
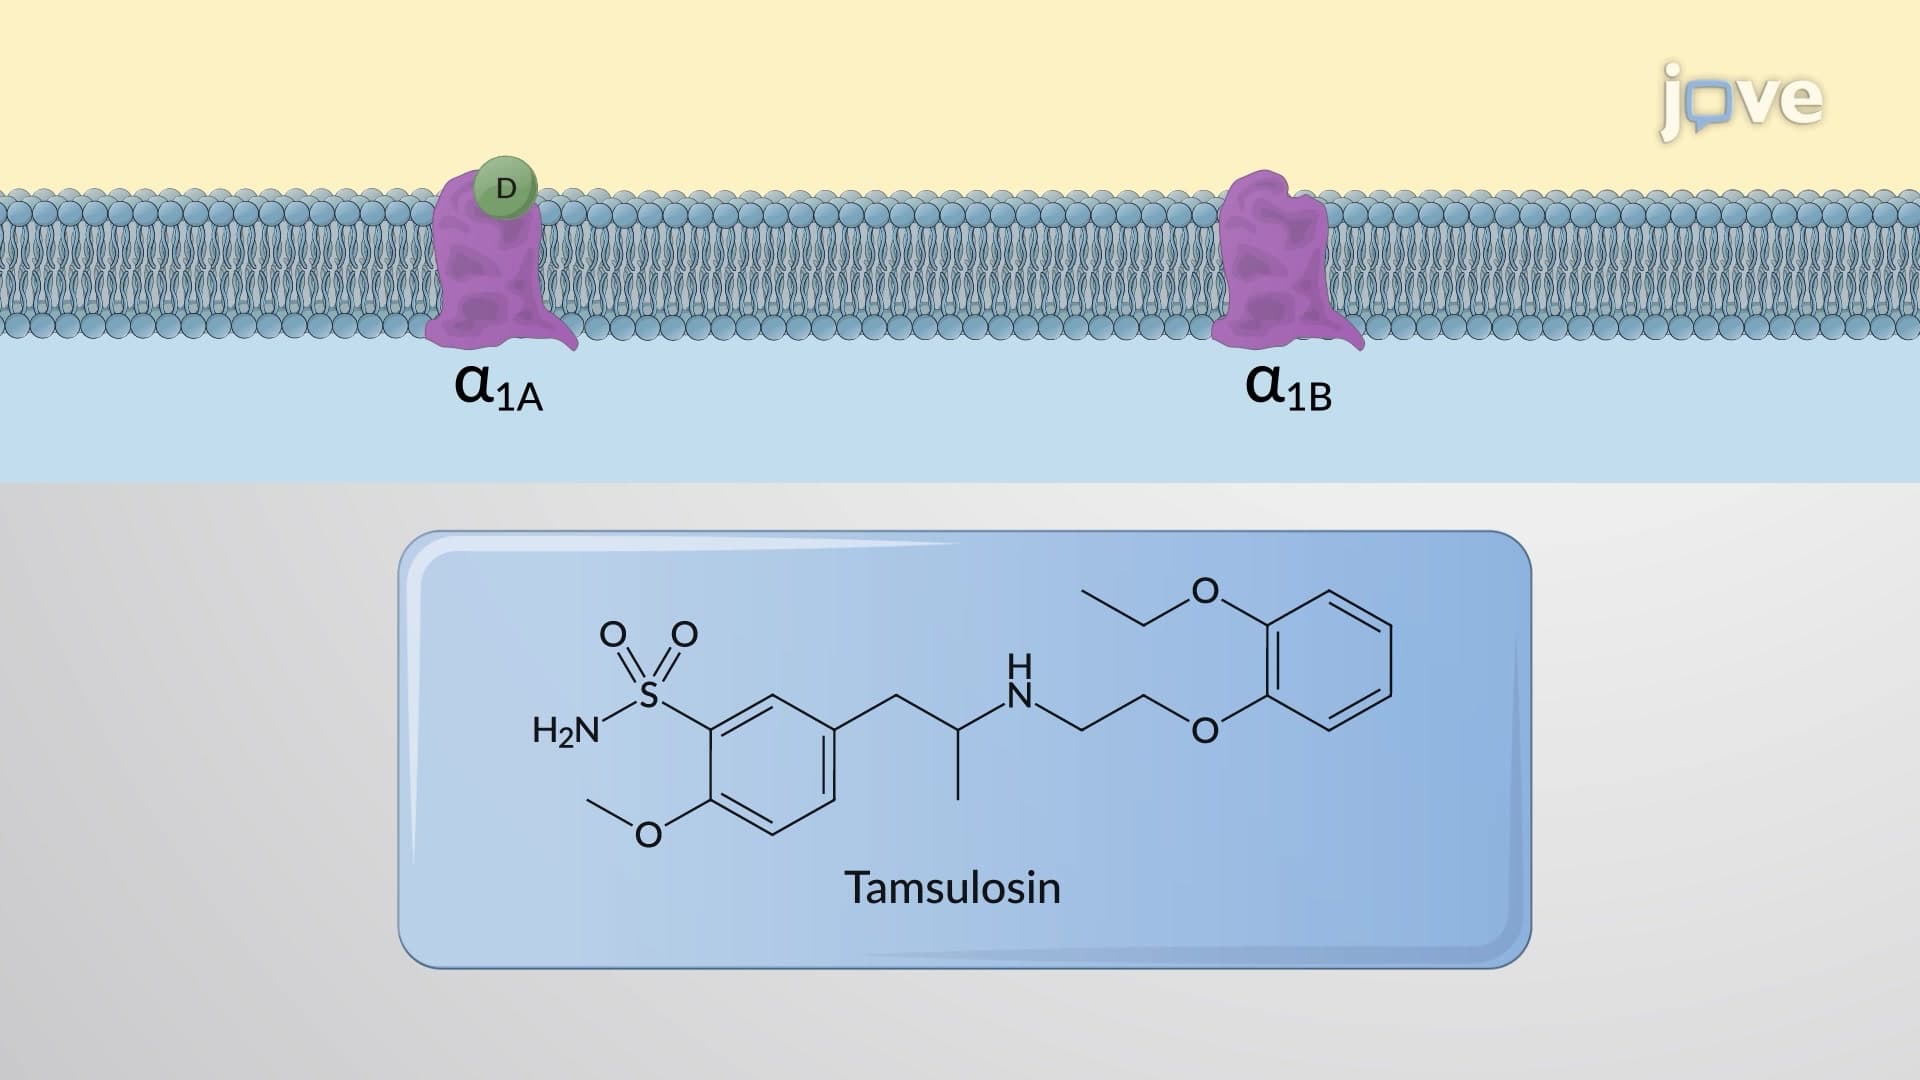
Adrenergic Receptors: ɑ Subtype
Adrenaline ≥ Noradrenaline >> Isoprenaline
α-adrenoceptors are further divided into α1 and α2-adrenoceptors.
α1-Adrenoceptors: These receptors are located postsynaptically on the effector organs and cause constriction of smooth muscle mediated by activation of phospholipase...
 01:29
The Two-State Receptor Model
The binding affinity of a drug determines its interaction with...
 01:21
G-protein Coupled Receptors
 01:19
Ligand-gated Ion Channels
Three Subfamilies of Ligand-gated Ion Channels
Ligand-gated ion channels fall into three subfamilies. The 'Cys-loop' includes the nicotinic acetylcholine receptors, γ-aminobutyric acid (GABA), glycine, and 5-hydroxytryptamine receptors. The second one is the 'Pore-loop' channels that...
 01:30
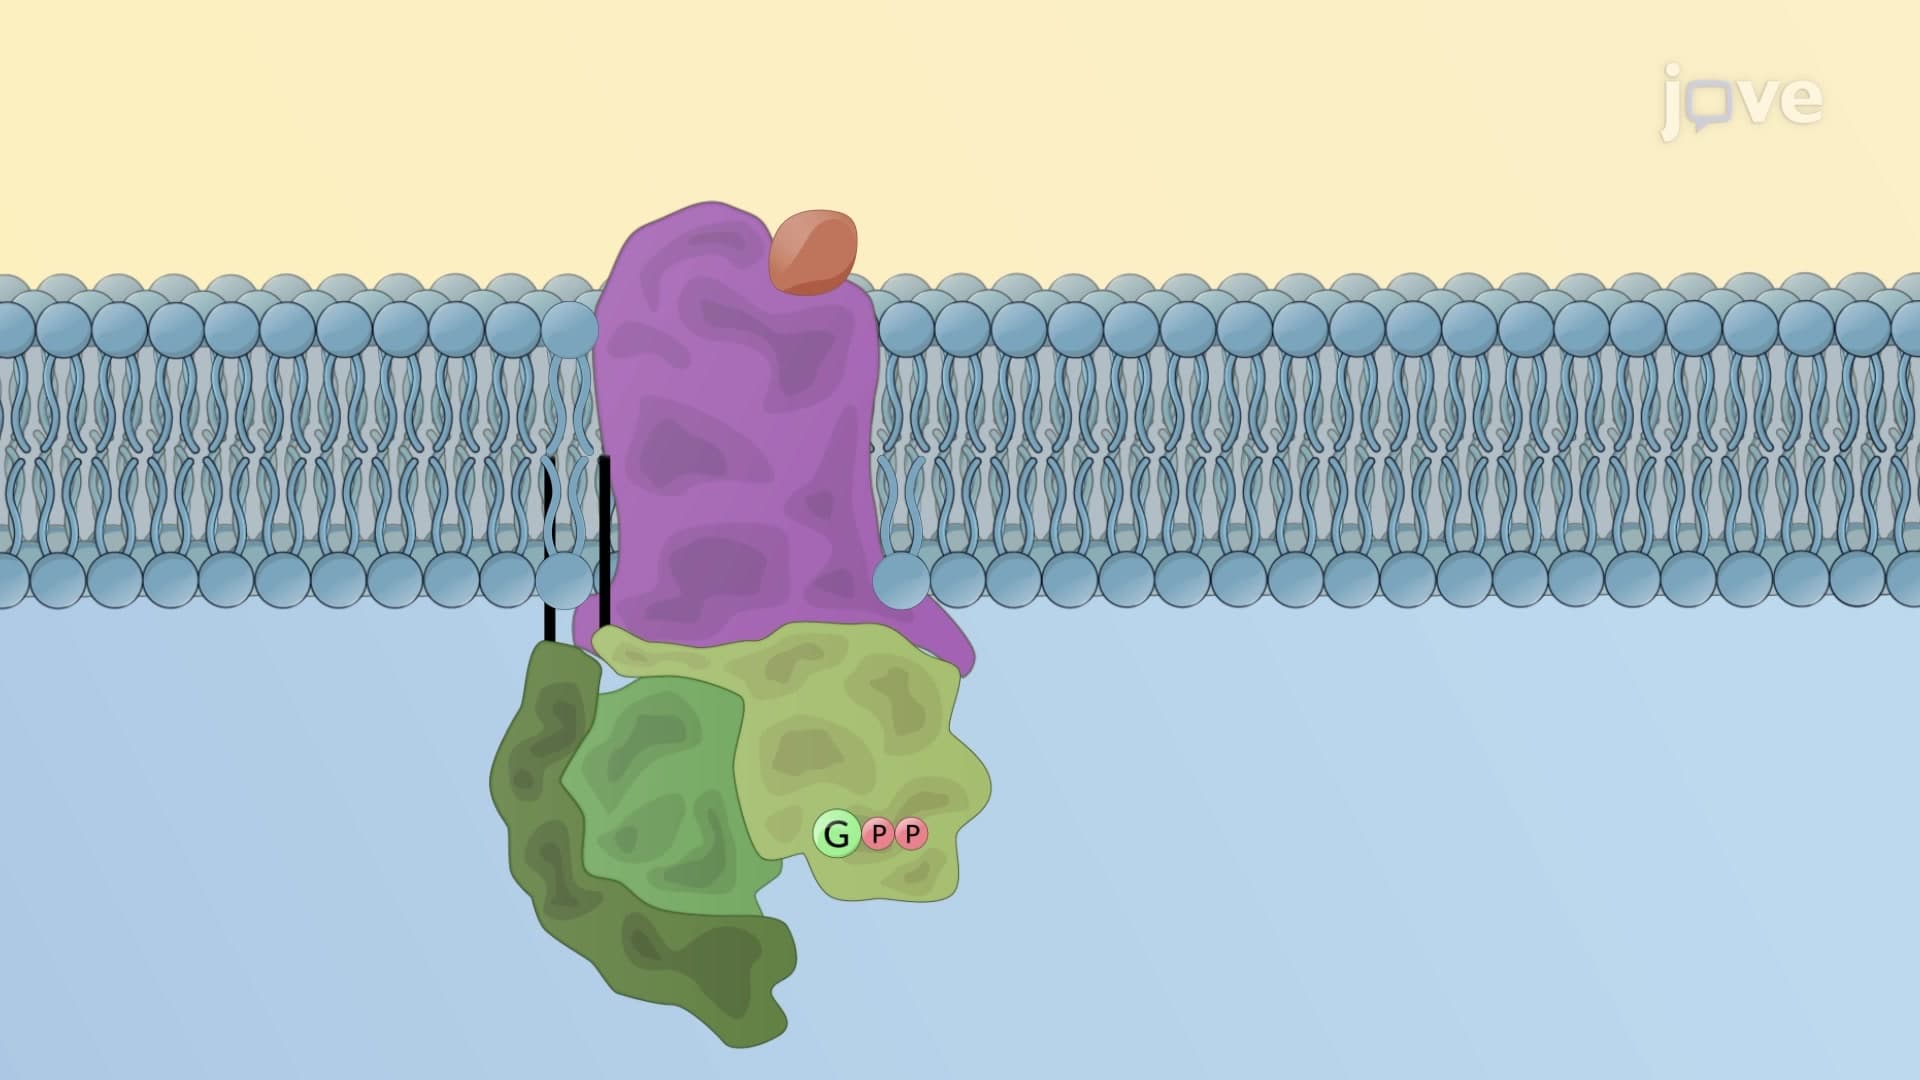
Transducer Mechanism: G Protein–Coupled Receptors
GPCRs are also called heptahelical,...