Related Experiment Video
Updated: Apr 27, 2026
Protein Film Infrared Electrochemistry Demonstrated for Study of H2 Oxidation by a [NiFe] Hydrogenase
Published on: December 4, 2017
New redox states observed in [FeFe] hydrogenases reveal redox coupling within the H-cluster
Agnieszka Adamska-Venkatesh1, Danuta Krawietz, Judith Siebel
1Max-Planck-Institut für Chemische Energiekonversion, Stiftstrasse 34-36, 45470 Mülheim an der Ruhr, Germany.

Abstract:
Active [FeFe] hydrogenases can be obtained by expressing the unmaturated enzyme in Escherichia coli followed by incubation with a synthetic precursor of the binuclear [2Fe] subcluster, namely: [NEt4]2[Fe2(adt)(CO)4(CN)2] (adt = [S-CH2-NH-CH2-S](2-)). The binuclear subsite Fe2(adt)(CO)3(CN)2 is attached through a bridging cysteine side chain to a [4Fe-4S] subcluster already present in the unmaturated enzyme thus yielding the intact native "H-cluster". We present FTIR electrochemical studies of the [FeFe] hydrogenase from Chlamydomonas reinhardtii, CrHydA1, maturated with the precursor of the native cofactor [Fe2(adt)(CO)4(CN)2](2-) as well as a non-natural variant [Fe2(pdt)(CO)4(CN)2](2-) in which the bridging amine functionality is replaced by CH2. The obtained active enzyme CrHydA1(adt) shows the same redox states in the respective potential range as observed for the native system (E(ox/red) = -400 mV, E(red/sred) = -470 mV). For the Hox → Hred transition the reducing equivalent is stored on the binuclear part, ([4Fe-4S](2+)Fe(II)Fe(I) → [4Fe-4S](2+)Fe(I)Fe(I)), while the Hred → Hsred transition is characterized by a reduction of the [4Fe-4S] part of the H-cluster ([4Fe-4S](2+)Fe(I)Fe(I) → [4Fe-4S](+)Fe(I)Fe(I)). A similar transition is reported here for the CO inhibited state of the H-cluster: ([4Fe-4S](2+)Fe(I)Fe(II)CO → [4Fe-4S](+)Fe(I)Fe(II)CO). An FTIR electrochemical study of the inactive variant with the pdt ligand, CrHydA1(pdt), identified two redox states H(pdt)-ox and H(pdt)-"red". Both EPR and FTIR spectra of H(pdt)-ox are virtually identical to those of the H(adt)-ox and the native Hox state. The H(pdt)-"red" state is also characterized by a reduced [4Fe-4S] subcluster. In contrast to CrHydA1(adt), the H(pdt)-ox state of CrHydA1(pdt) is stable up to rather high potentials (+200 mV). This study demonstrates the distinct redox coupling between the two parts of the H-cluster and confirms that the [4Fe-4S]H subsite is also redox active and as such an integral part of the H-cluster taking part in the catalytic cycle.
More Related Videos
 10:24
Defining Hsp33's Redox-regulated Chaperone Activity and Mapping Conformational Changes on Hsp33 Using Hydrogen-deuterium Exchange Mass Spectrometry
Published on: June 7, 2018
 06:01
EPR Monitored Redox Titration of the Cofactors of Saccharomyces cerevisiae Nar1
Published on: November 26, 2014
Related Concept Videos
 01:23
Redox Equilibria: Overview
 01:24
Redox Reactions
 01:27
Redox Reactions
 01:30
Ladder Diagrams: Redox Equilibria
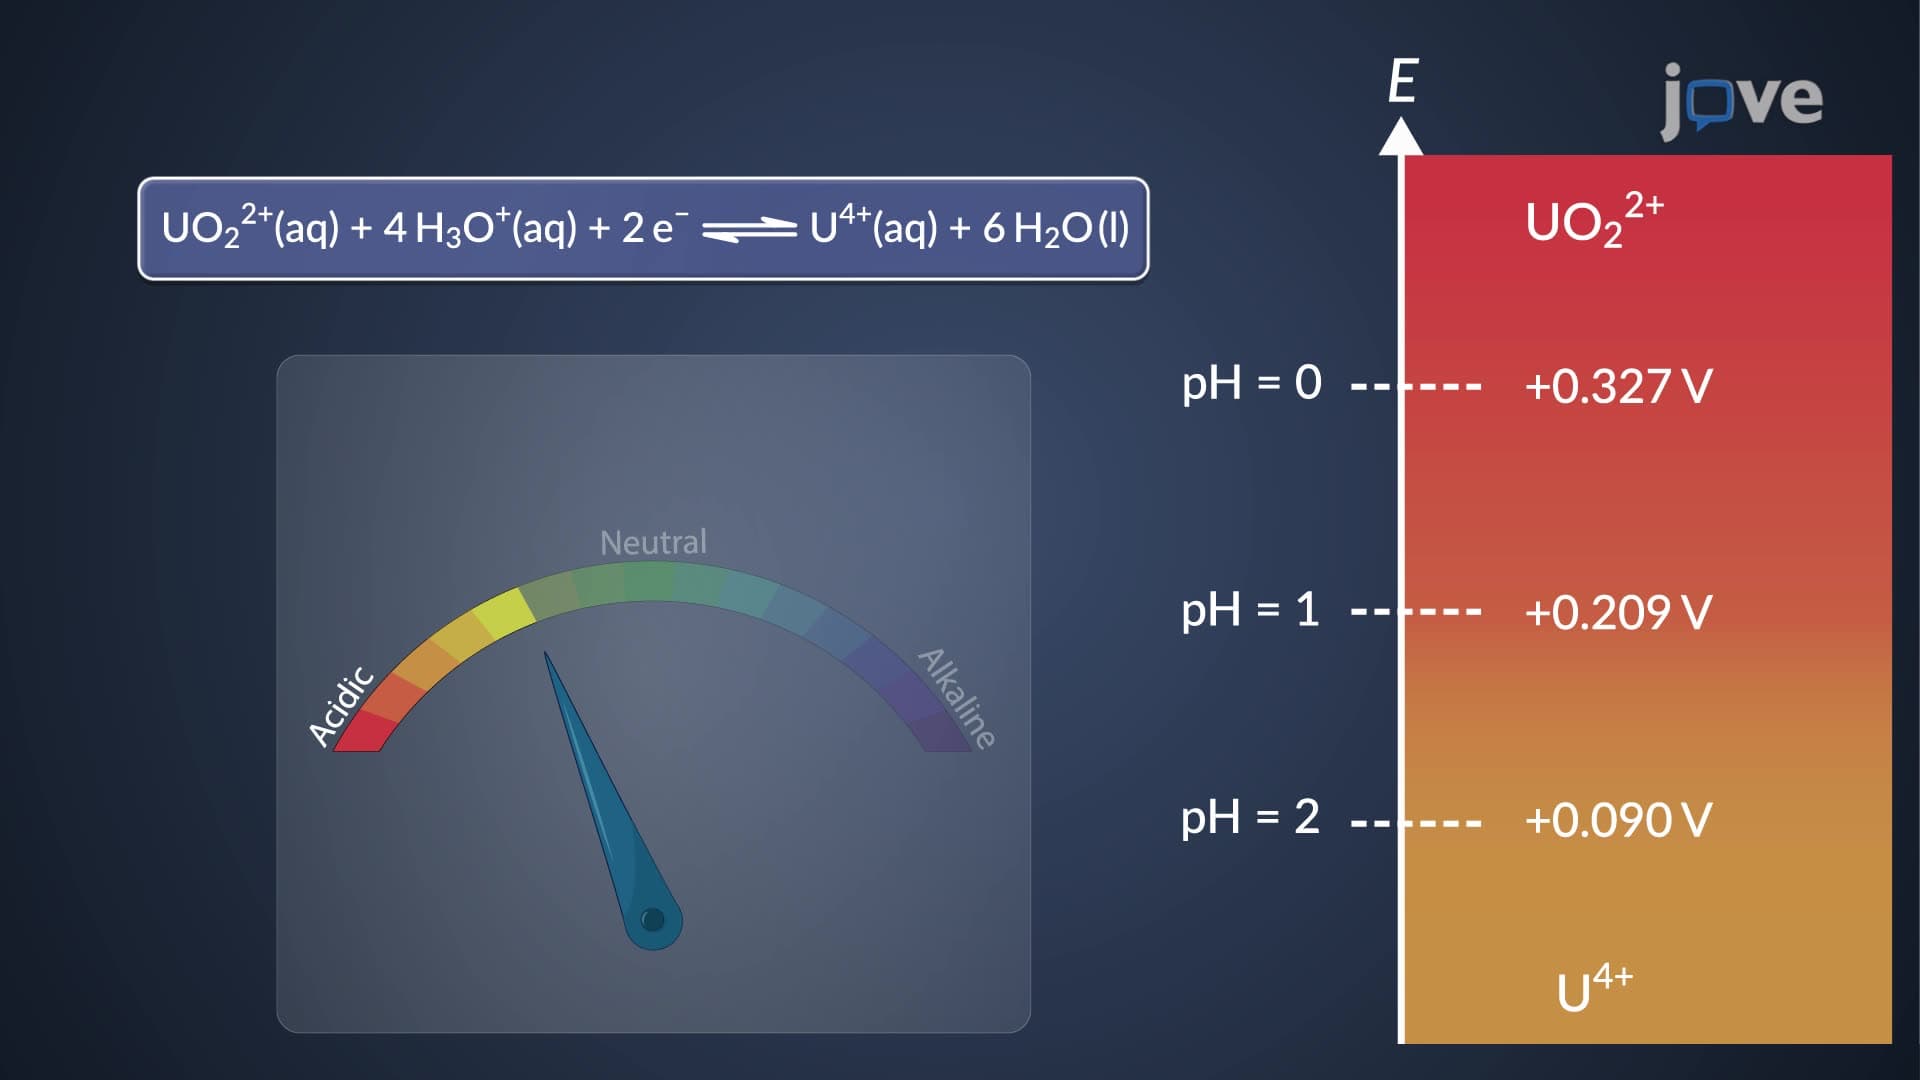
Consider the Fe3+/Fe2+ half-reaction, which has a standard-state potential of +0.771 V. At potentials more positive than +0.771 V, Fe3+ predominates, whereas Fe2+...
 01:41
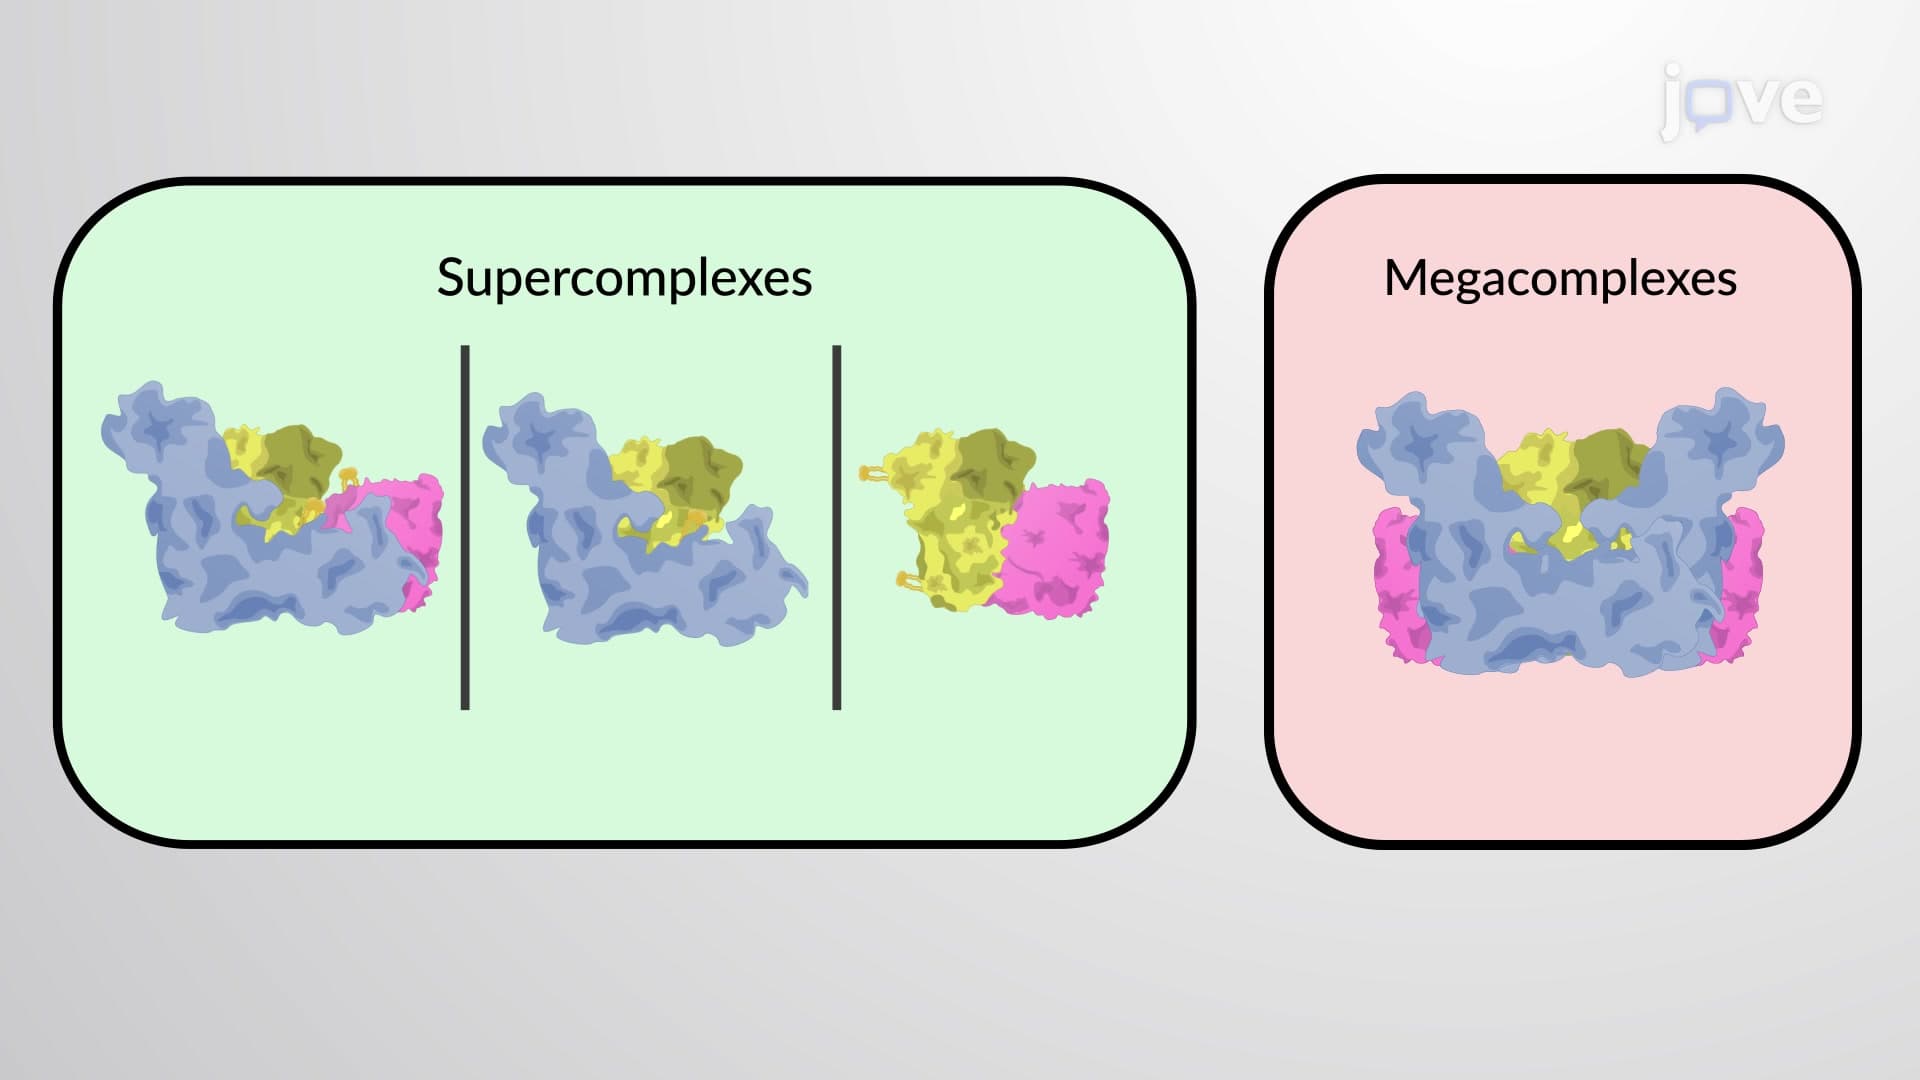
The Supercomplexes in the Crista Membrane
 03:11
Oxidation-Reduction Reactions