Related Experiment Video
Updated: Apr 26, 2026

Purifying the Impure: Sequencing Metagenomes and Metatranscriptomes from Complex Animal-associated Samples
Published on: December 22, 2014
Pyrosequencing reveals transient cystic fibrosis lung microbiome changes with intravenous antibiotics
Daniel J Smith1, Alison C Badrick2, Martha Zakrzewski3
1Dept of Thoracic Medicine, The Prince Charles Hospital, Brisbane, QLD, Australia Iron Metabolism Laboratory, QIMR Berghofer Medical Research Institute, Brisbane, QLD, Australia Lung, Inflammation and Infection Laboratory, QIMR Berghofer Medical Research Institute, Brisbane, QLD, Australia School of Medicine, The University of Queensland, Brisbane, QLD, Australia Both authors contributed equally daniel_j_smith@health.qld.gov.au.

Abstract:
Chronic airway infection in adults with cystic fibrosis (CF) is polymicrobial and the impact of intravenous antibiotics on the bacterial community composition is poorly understood. We employed culture-independent molecular techniques to explore the early effects of i.v. antibiotics on the CF airway microbiome. DNA was extracted from sputum samples collected from adult subjects with CF at three time-points (before starting treatment, and at day 3 and day 8-10 of i.v. antibiotics) during treatment of an infective pulmonary exacerbation. Microbial community profiles were derived through analysis of bacterial-derived 16S ribosomal RNA by pyrosequencing and changes over time were compared. 59 sputum samples were collected during 24 pulmonary exacerbations from 23 subjects. Between treatment onset and day 3 there was a significant reduction in the relative abundance of Pseudomonas and increased microbial diversity. By day 8-10, bacterial community composition was similar to pre-treatment. Changes in community composition did not predict improvements in lung function. The relative abundance of Pseudomonas falls rapidly in subjects with CF receiving i.v. antibiotic treatment for a pulmonary exacerbation and is accompanied by an increase in overall microbial diversity. However, this effect is not maintained beyond the first week of treatment.
Related Concept Videos
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation,...
 01:29
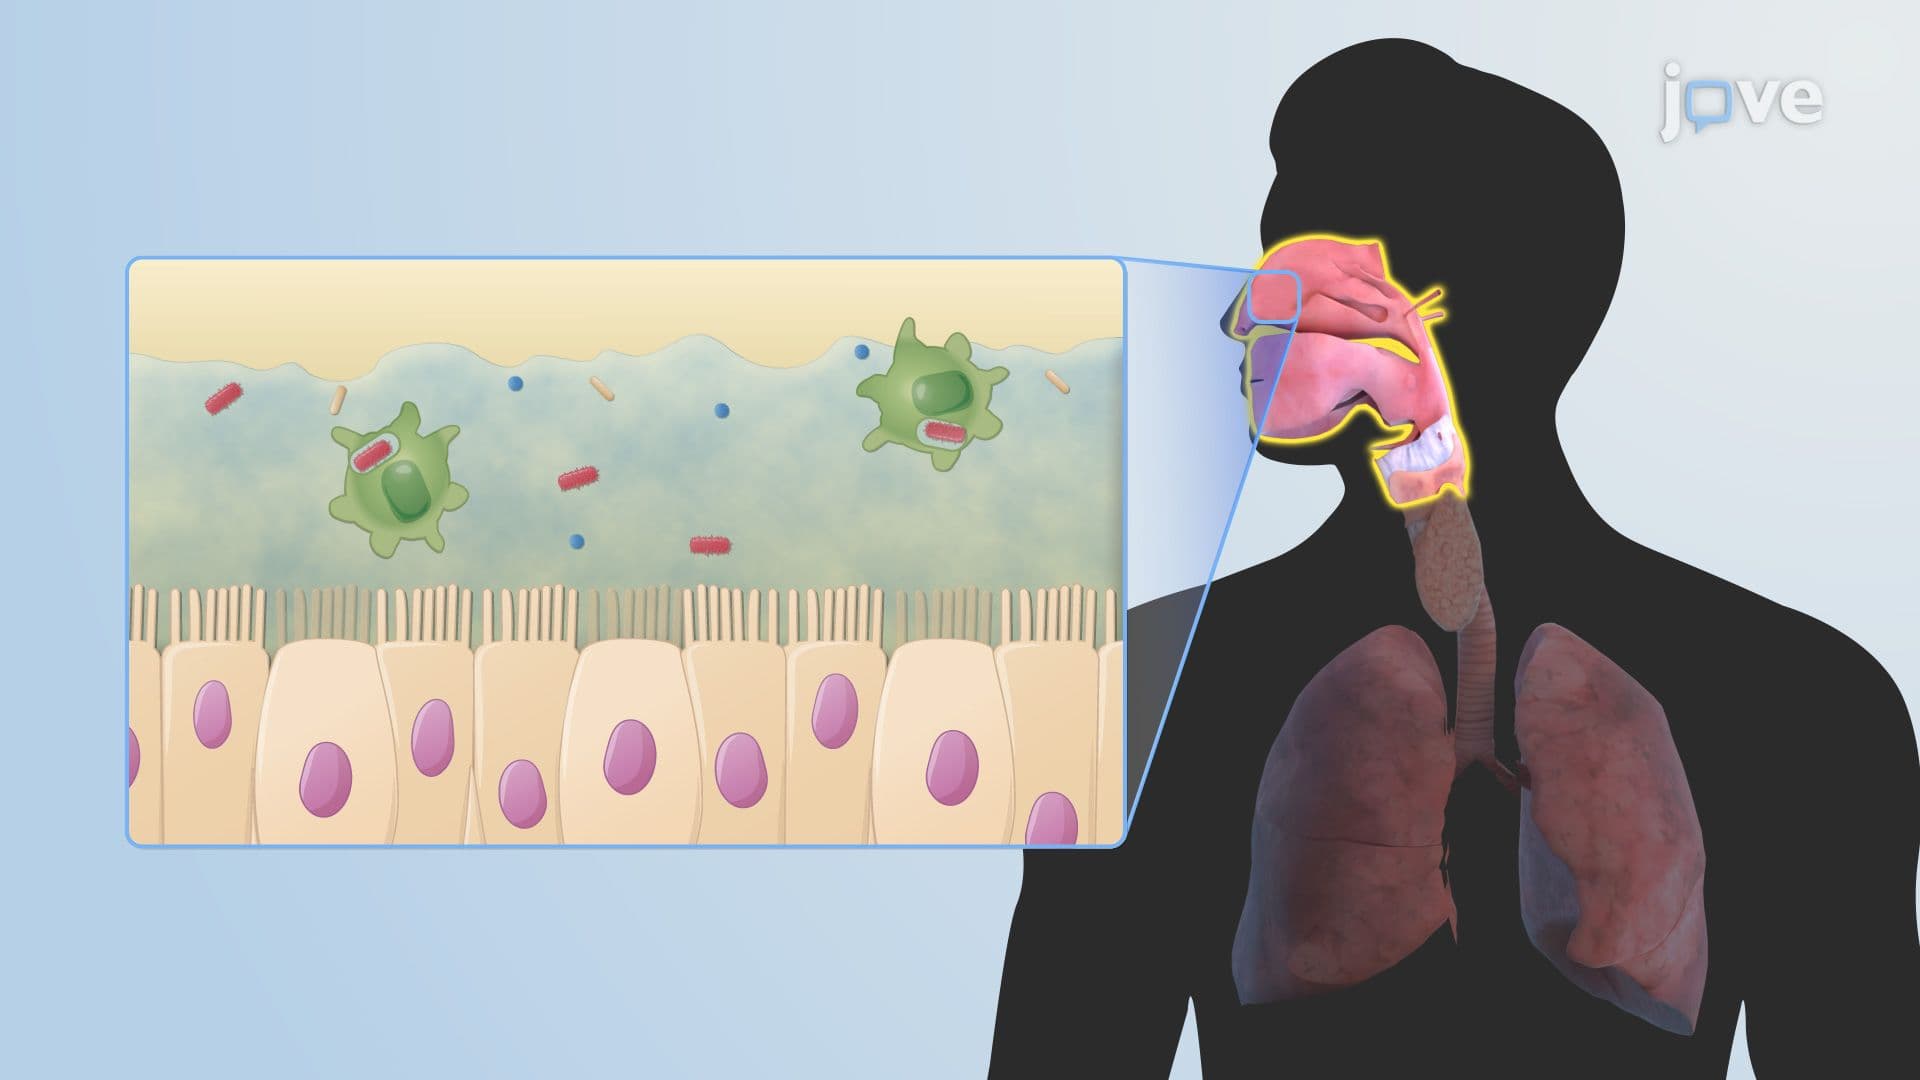
Microbiota of the Respiratory Tract
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic...