
Effects of non-local exchange on core level shifts for gas-phase and adsorbed molecules
M Van den Bossche1, N M Martin2, J Gustafson2
1Department of Applied Physics and Competence Centre for Catalysis, Chalmers University of Technology, Göteborg, Sweden.

Abstract:
Density functional theory calculations are often used to interpret experimental shifts in core level binding energies. Calculations based on gradient-corrected (GC) exchange-correlation functionals are known to reproduce measured core level shifts (CLS) of isolated molecules and metal surfaces with reasonable accuracy. In the present study, we discuss a series of examples where the shifts calculated within a GC-functional significantly deviate from the experimental values, namely the CLS of C 1s in ethyl trifluoroacetate, Pd 3d in PdO and the O 1s shift for CO adsorbed on PdO(101). The deviations are traced to effects of the electronic self-interaction error with GC-functionals and substantially better agreements between calculated and measured CLS are obtained when a fraction of exact exchange is used in the exchange-correlation functional.
Related Concept Videos
 01:27
π Electron Effects on Chemical Shift: Overview
 01:28
Adsorption of Gases on Solids
 01:27
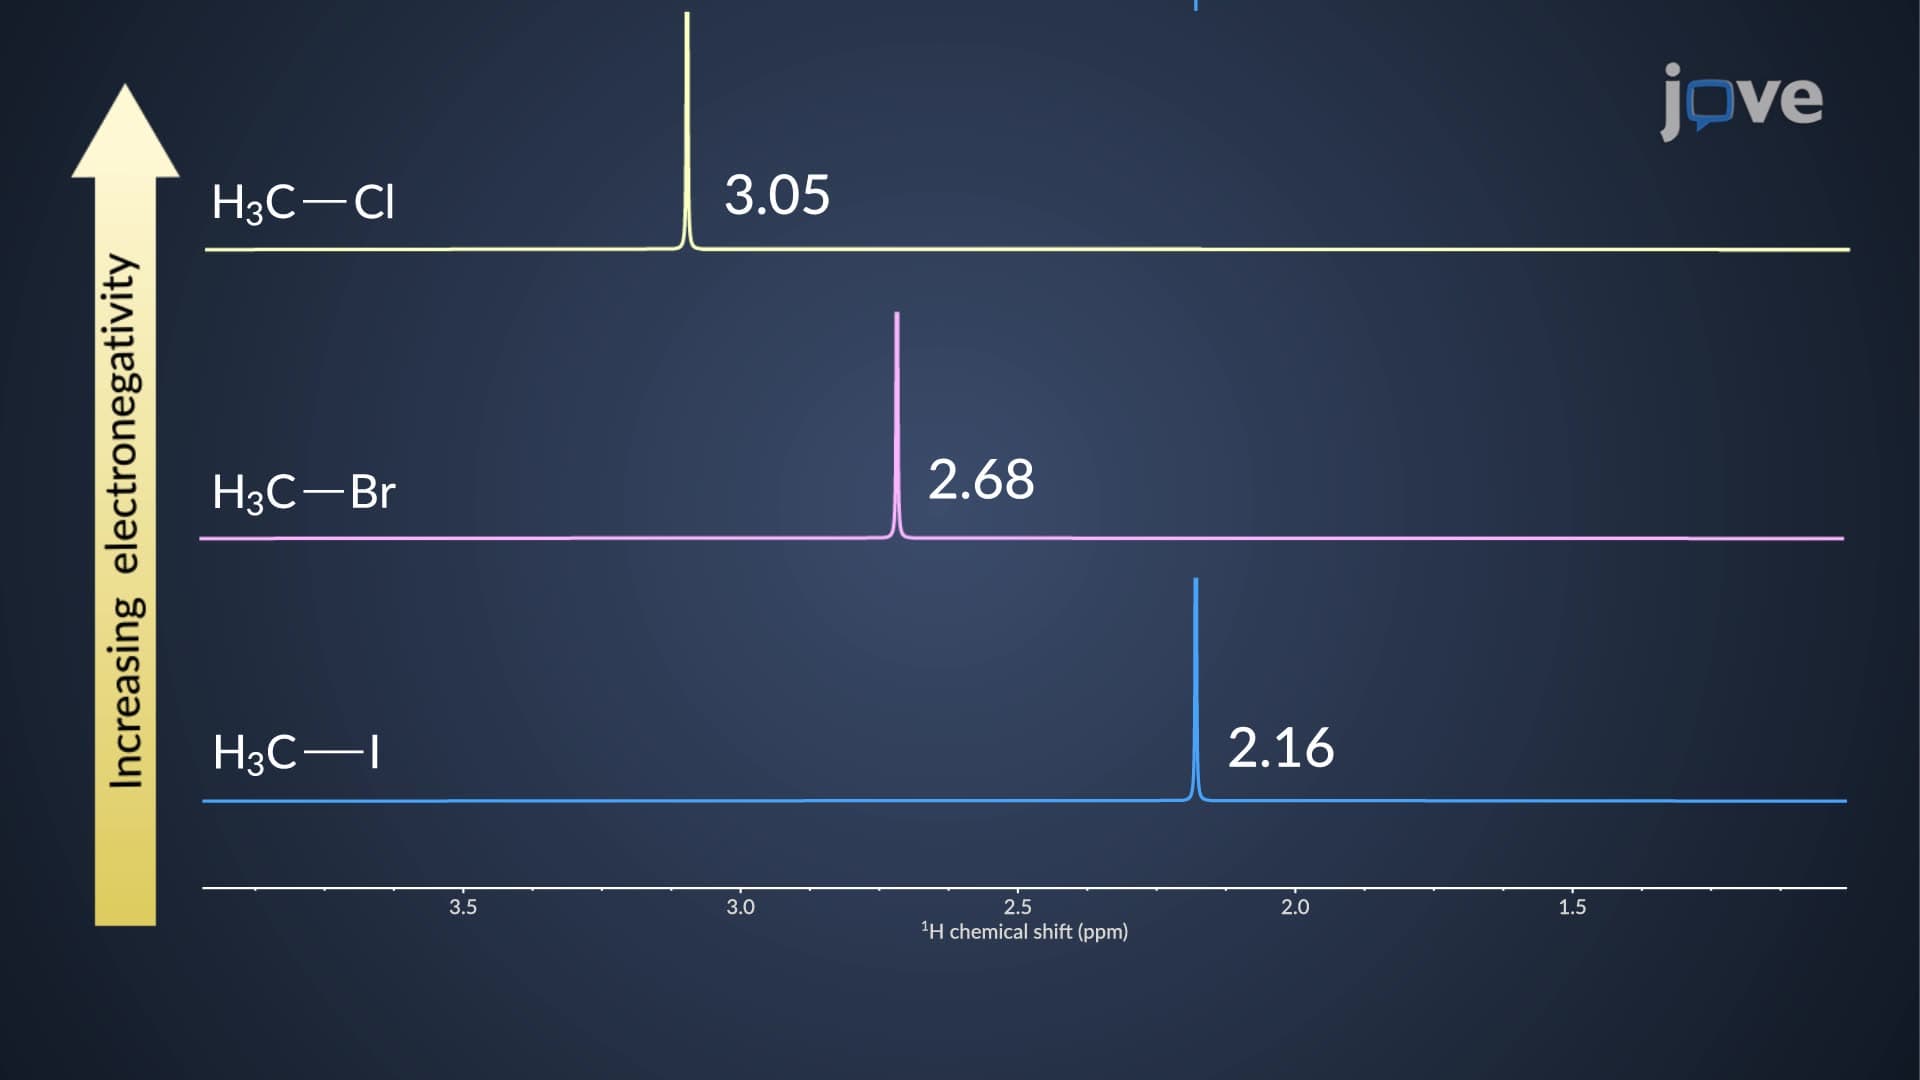
Inductive Effects on Chemical Shift: Overview
 01:14
π Electron Effects on Chemical Shift: Aromatic and Antiaromatic Compounds
 01:17
Chemical Shift: Internal References and Solvent Effects
The internal reference compound generally used in NMR spectroscopy is tetramethylsilane (TMS). TMS is preferred because it is chemically inert, soluble in NMR solvents, and easily removable. Also, the highly shielded methyl protons in TMS yield an intense...
 03:48
Behavior of Gas Molecules: Molecular Diffusion, Mean Free Path, and Effusion