Identification of copy number variants through whole-exome sequencing in autosomal recessive nonsyndromic hearing loss
Related Experiment Video
Updated: Apr 26, 2026
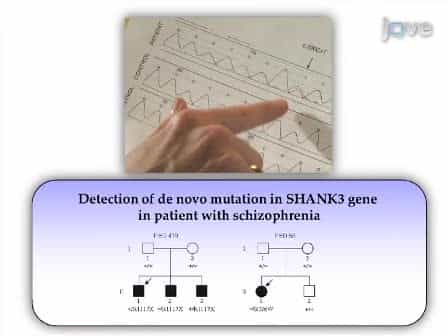
A Strategy to Identify de Novo Mutations in Common Disorders such as Autism and Schizophrenia
Published on: June 15, 2011
Guney Bademci1, Oscar Diaz-Horta, Shengru Guo
11 John P. Hussmann Institute for Human Genomics, Miller School of Medicine, University of Miami , Miami, Florida.

Abstract:
Genetic variants account for more than half of the cases with congenital or prelingual onset hearing loss. Autosomal recessive nonsyndromic hearing loss (ARNSHL) is the most common subgroup. Whole-exome sequencing (WES) has been shown to be effective detecting deafness-causing single-nucleotide variants (SNVs) and insertion/deletions (INDELs). After analyzing the WES data for causative SNVs or INDELs involving previously reported deafness genes in 78 families with ARNSHL, we searched for copy number variants (CNVs) through two different tools in 24 families that remained unresolved. We detected large homozygous deletions in STRC and OTOA in single families. Thus, causative CNVs in known deafness genes explain 2 out of 78 (2.6%) families in our sample set. We conclude that CNVs can be reliably detected through WES and should be the part of pipelines used to clarify genetic basis of hearing loss.
Related Concept Videos
 02:26
Comparing Copy Number Variations and SNPs
Copy number variations or CNVs are the structural variations that cover more than 1kb of DNA sequence. The single nucleotide polymorphism (SNP), on the other hand, is a single nucleotide change or a point mutation that is found in more than 1%...
 01:11
Genome-wide Association Studies-GWAS
GWAS does not require the identification of the target gene involved in...
 01:05
Single Nucleotide Polymorphisms-SNPs
 01:33
Pleiotropy