Related Experiment Video
Updated: Apr 26, 2026
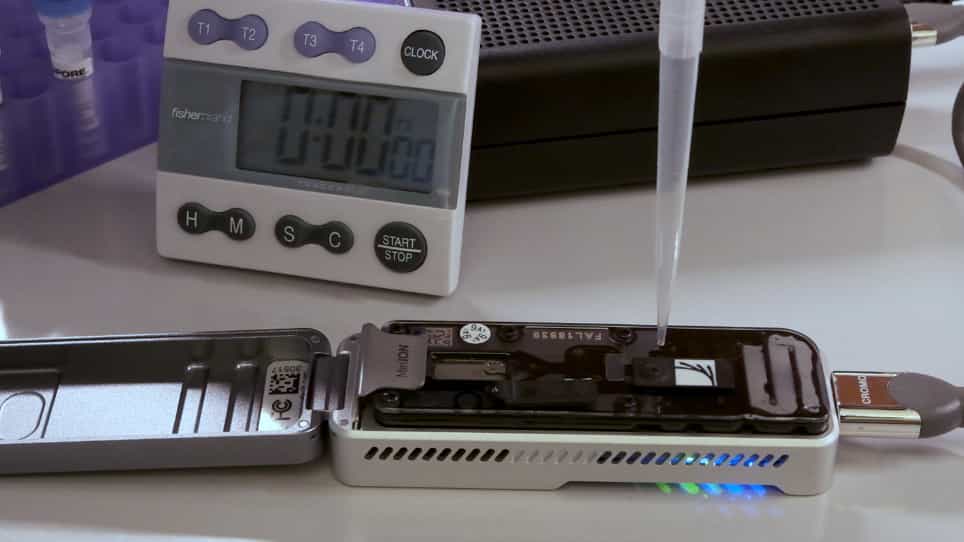

Hybrid De Novo Genome Assembly for the Generation of Complete Genomes of Urinary Bacteria using Short- and Long-read Sequencing Technologies
Published on: August 20, 2021
Employing whole genome mapping for optimal de novo assembly of bacterial genomes
Basil Britto Xavier, Julia Sabirova, Moons Pieter
1Department of Medical Microbiology, Vaccine & Infectious Disease Institute, Universiteit Antwerpen, Antwerp, Belgium. surbhi.malhotra@uantwerpen.be.

Background:
De novo genome assembly can be challenging due to inherent properties of the reads, even when using current state-of-the-art assembly tools based on de Bruijn graphs. Often users are not bio-informaticians and, in a black box approach, utilise assembly parameters such as contig length and N50 to generate whole genome sequences, potentially resulting in mis-assemblies.
Findings:
Utilising several assembly tools based on de Bruijn graphs like Velvet, SPAdes and IDBA, we demonstrate that at the optimal N50, mis-assemblies do occur, even when using the multi-k-mer approaches of SPAdes and IDBA. We demonstrate that whole genome mapping can be used to identify these mis-assemblies and can guide the selection of the best k-mer size which yields the highest N50 without mis-assemblies.
Conclusions:
We demonstrate the utility of whole genome mapping (WGM) as a tool to identify mis-assemblies and to guide k-mer selection and higher quality de novo genome assembly of bacterial genomes.
Related Concept Videos
 03:36
Genome Annotation and Assembly
 01:29
Modern Molecular Taxonomy
 01:57
Sanger Sequencing
 02:02
Genomics
 00:46
Genomic DNA in Prokaryotes
Genomic Diversity in Bacteria
Although bacterial genomes are much...
 03:21
RNA-seq
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...