Related Experiment Video
Updated: Apr 26, 2026
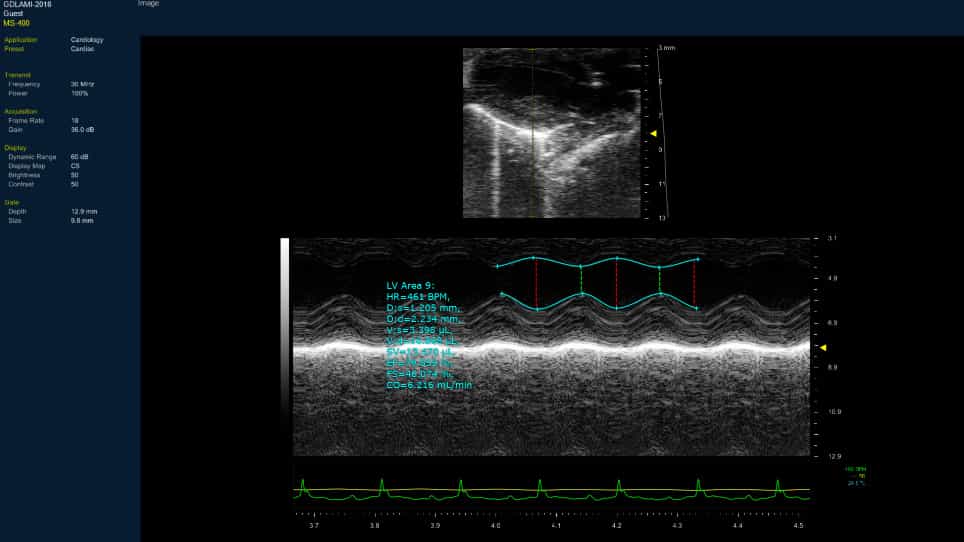
Investigating the Pathogenesis of MYH7 Mutation Gly823Glu in Familial Hypertrophic Cardiomyopathy using a Mouse Model
Published on: August 8, 2022
Detection of mutations in symptomatic patients with hypertrophic cardiomyopathy in Taiwan
Kuan-Rau Chiou1, Chien-Tung Chu2, Min-Ji Charng3
1Division of Cardiology, Department of Medicine, Kaohsiung Veterans General Hospital, Taipei, Taiwan, ROC; School of Medicine, National Yang-Ming University, Taipei, Taiwan, ROC.

Insights
Genetic screening identified mutations in hypertrophic cardiomyopathy (HCM) genes in 34.2% of Taiwanese patients. Researchers discovered five novel mutations, aiding earlier diagnosis and management of this genetic heart condition.
Area of Science:
- Cardiovascular Genetics
- Molecular Cardiology
- Genetic Epidemiology
Background:
- Hypertrophic cardiomyopathy (HCM) is a prevalent genetic heart disorder linked to sudden cardiac death, heart failure, and stroke.
- Understanding the genetic underpinnings of HCM is crucial for risk stratification and management.
Purpose of the Study:
- To determine the prevalence and spectrum of mutations in sarcomeric genes among symptomatic hypertrophic cardiomyopathy patients in Taiwan.
- To identify novel mutations and assess their pathogenicity in the Taiwanese HCM population.
Main Methods:
- Systematic mutation screening of eight key sarcomeric genes (MYH7, MYBPC3, TNNT2, TNNI3, MYL2, MYL3, TPM1, ACTC) in 38 index HCM patients using direct DNA sequencing.
- In silico analysis to predict the pathogenicity of identified mutations.
- Genotype-phenotype correlation studies and family pedigree analysis to assess cosegregation and penetrance.
Main Results:
- Mutations were identified in 13 out of 38 index patients (34.2%), with MYH7, MYBPC3, and TNNT2 being the most frequently affected genes.
- Five novel missense mutations were discovered and predicted as pathological by in silico tools.
- Family studies revealed cosegregation of mutations in affected members, alongside evidence of incomplete penetrance in younger individuals.
Conclusions:
- The study identified 13 hypertrophic cardiomyopathy pedigrees in Taiwan, including five with novel mutations and one with a double mutation.
- MYH7, MYBPC3, and TNNT2 are the most commonly mutated genes in this cohort.
- Findings support the integration of genetic counseling and molecular diagnosis for earlier identification and improved management of at-risk family members.
Background:
Hypertrophic cardiomyopathy (HCM) is a common genetic cardiac disorder associated with sudden death, heart failure, and stroke. The aim of the present study was to evaluate the prevalence and types of mutations in symptomatic patients with HCM in Taiwan.
Methods:
Thirty-eight HCM index patients (mean age 60±16 years) underwent systematic mutation screening of eight sarcomeric genes: β-myosin heavy chain (MYH7), myosin-binding protein C (MYBPC3), troponin T (TNNT2), troponin I (TNNI3), myosin ventricular regulatory light chain 2 (MYL2), myosin ventricular essential light chain 1 (MYL3), α-tropomyosin (TPM1), and cardiac α-actin (ACTC), using direct DNA sequencing. In silico programs predicted damaging amino acids. In the positive families, genotype-phenotype correlation studies were done.
Results:
Overall, 13 mutations were identified in 13 index patients (34.2%). The three most frequently mutated genes were MYH7, MYBPC3, and TNNT2. One patient carried double mutations. Five mutations (MYH7 R147S; MYBPC3 R597Q; MYBPC3 W1007R; TNNI3 E124Q; MYL3 R63C) were novel; all were missense mutations. Analysis using in silico tools showed near consensus to classify these five novel mutations as pathological. Family pedigree analysis showed the presence of cosegregation in at least two affected members in each proband family, but incomplete penetrance in young family members with a positive genotype.
Conclusions:
We identified 13 HCM pedigrees, including 5 carrying novel mutations and 1 with a double mutation. The three most commonly mutated genes were MYH7, MYBPC3, and TNNT2. These results, together with genetic counseling, could lead to earlier diagnosis and better management of family members at risk of HCM.
Related Concept Videos
 01:29
Cardiomyopathy III: Hypertrophic Cardiomyopathy
 01:27
Myocarditis II: Clinical Features and Diagnostic Tests
 01:30
Cardiomyopathy II: Dilated Cardiomyopathy