Related Experiment Video
Updated: Apr 25, 2026

Fingerprinting Cardiolipin in Leukocytes by Mass Spectrometry for a Rapid Diagnosis of Barth Syndrome
Published on: March 23, 2022
ECHS1 mutations in Leigh disease: a new inborn error of metabolism affecting valine metabolism
Heidi Peters1, Nicole Buck1, Ronald Wanders2
11 Murdoch Childrens Research Institute, Victorian Clinical Genetics Services, Royal Children's Hospital, Parkville, Vic 3052, Australia.

Abstract:
Two siblings with fatal Leigh disease had increased excretion of S-(2-carboxypropyl)cysteine and several other metabolites that are features of 3-hydroxyisobutyryl-CoA hydrolase (HIBCH) deficiency, a rare defect in the valine catabolic pathway associated with Leigh-like disease. However, this diagnosis was excluded by HIBCH sequencing and normal enzyme activity. In contrast to HIBCH deficiency, the excretion of 3-hydroxyisobutyryl-carnitine was normal in the children, suggesting deficiency of short-chain enoyl-CoA hydratase (ECHS1 gene). This mitochondrial enzyme is active in several metabolic pathways involving fatty acids and amino acids, including valine, and is immediately upstream of HIBCH in the valine pathway. Both children were compound heterozygous for a c.473C > A (p.A158D) missense mutation and a c.414+3G>C splicing mutation in ECHS1. ECHS1 activity was markedly decreased in cultured fibroblasts from both siblings, ECHS1 protein was undetectable by immunoblot analysis and transfection of patient cells with wild-type ECHS1 rescued ECHS1 activity. The highly reactive metabolites methacrylyl-CoA and acryloyl-CoA accumulate in deficiencies of both ECHS1 and HIBCH and are probably responsible for the brain pathology in both disorders. Deficiency of ECHS1 or HIBCH should be considered in children with Leigh disease. Urine metabolite testing can detect and distinguish between these two disorders.
Related Concept Videos
 01:20
Inborn Errors of Metabolism
 01:18
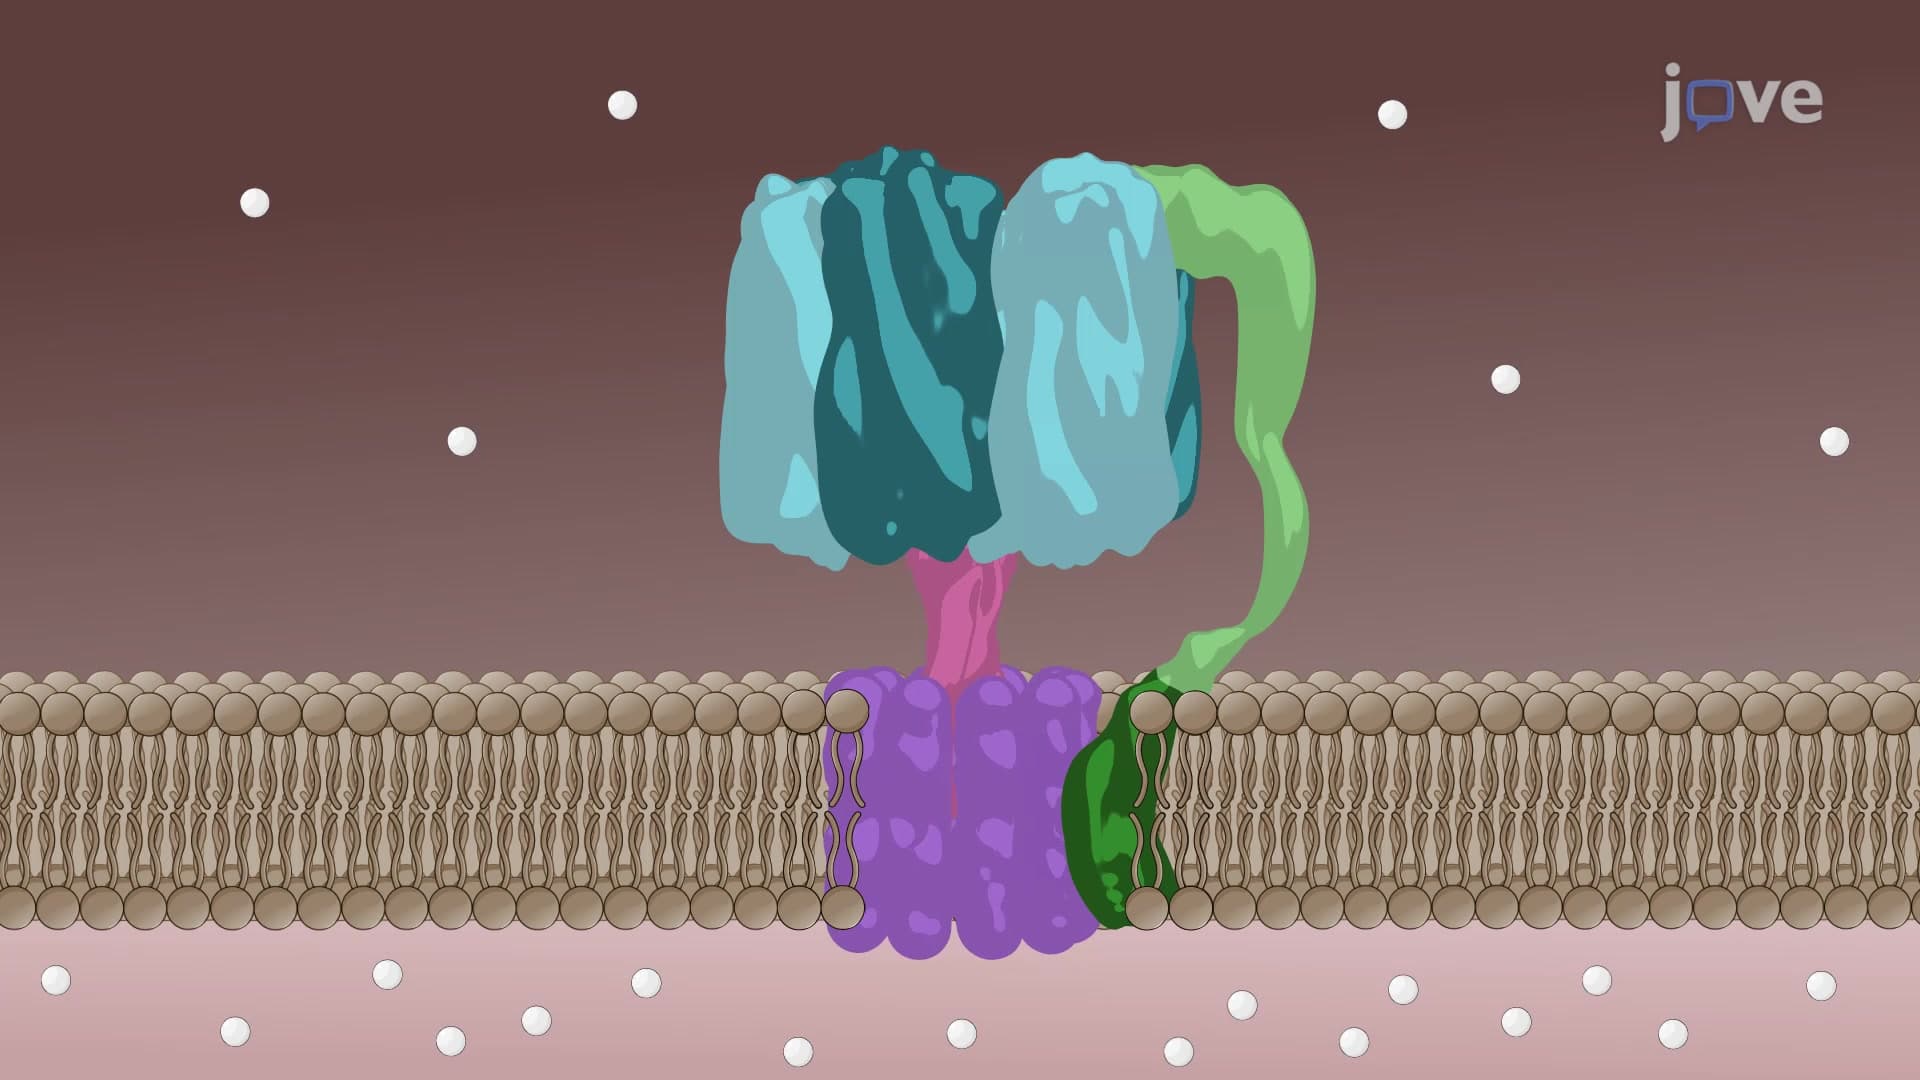
ATP Synthase: Structure
 01:27
Protein Import into the Peroxisomes
Peroxisomal Protein Import:
Peroxisomes lack the genetic machinery required to code for their own proteins. Hence, most peroxisomal membrane, lumenal and transmembrane proteins are synthesized in the cytoplasm or ER and transported to the peroxisome...
 01:21
Smooth Endoplasmic Reticulum
The ER provides optimal conditions for synthesizing steroid hormones and lipids, such as phospholipids and triglycerides. Traditionally, lipid metabolism was considered to be a smooth ER function. However, there is no direct evidence to prove that rough ER is completely excluded from lipid...
 01:22
Lysosomal Hydrolases
 01:31
Translation
Translation is the process of synthesizing proteins from the genetic information carried by messenger RNA (mRNA). Following transcription, it constitutes the final step in the expression of genes. This process is carried out by ribosomes, complexes of protein and specialized RNA molecules. Ribosomes, transfer RNA (tRNA), and other proteins produce a chain of amino acids—the polypeptide—as the end product of translation.
Translation Produces the Building Blocks of...