Related Experiment Video
Updated: Sep 21, 2026
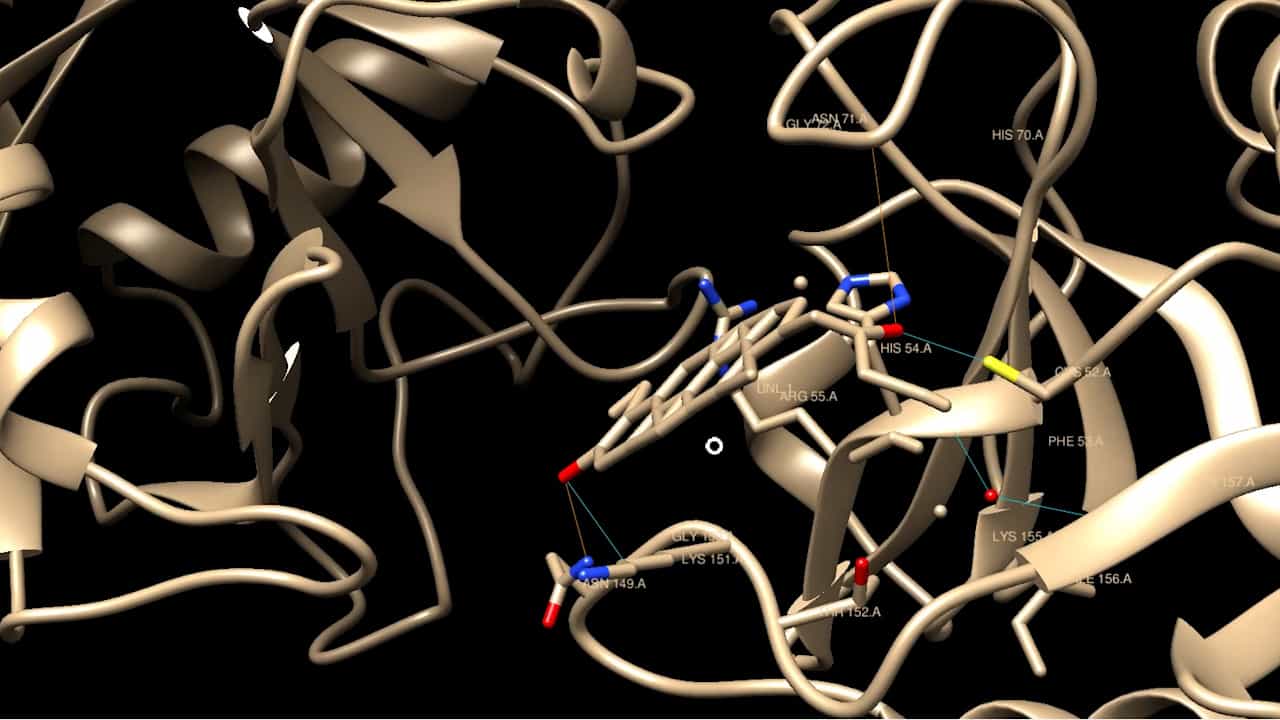

Structure-Guided Design and Development of Novel Cyclophilin A Inhibitors and Ganoderiol-F Derivatives: An In-Silico Approach
Published on: June 23, 2026
Molecular modeling of exquisitely selective c-Met inhibitors through 3D-QSAR and molecular dynamics simulations
Haoliang Yuan1, Jin Zhuang, Shihe Hu
1Key Laboratory of Nuclear Medicine, Ministry of Health, Jiangsu Key Laboratory of Molecular Nuclear Medicine, Jiangsu Institute of Nuclear Medicine , Wuxi, 214063 Jiangsu, P. R. China.

Abstract:
c-Met has been considered as an attractive target for developing antitumor agents. The highly selective c-Met inhibitors provide invaluable opportunities for the combination with other therapies safely to achieve the optimal efficacy. In this work, a series of triazolopyrazine c-Met inhibitors with exquisitely selectivity were investigated using a combination of molecular docking, three-dimensional quantitative structure-activity relationship (3D-QSAR), and molecular dynamics simulation. Comparative molecular field analysis (CoMFA) and comparative molecular similarity index analysis (CoMSIA) models were developed to reveal the structural determinants for c-Met inhibition. Both models were validated to have high reliability and predictability, and contour map analysis suggested feature requirements for different substituents on the scaffold. It is worth noting that an important hydrogen bond rich region was identified in the unique narrow channel, which is distinct from other kinases. Molecular dynamics simulations and binding free energy calculations provided further support that suitable groups in this hydrogen bond rich region made great contributions to the binding of ligands. Moreover, hydrogen bonds with residues of the narrow channel were also indicated to be essential to improve the activity and selectivity. This study will facilitate the discovery and optimization of novel c-Met inhibitors with higher activity and selectivity.
More Related Videos
 10:29
Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
 05:50
Pharmacophore Modeling for Targets with Extensive Ligand Libraries: A Case Study on SARS-CoV-2 Mpro
Published on: September 26, 2025
Related Concept Videos
 02:00
Molecular Models
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...