Related Experiment Video
Updated: Apr 24, 2026


Optimization of Synthetic Proteins: Identification of Interpositional Dependencies Indicating Structurally and/or Functionally Linked Residues
Published on: July 14, 2015
Fast and accurate discovery of degenerate linear motifs in protein sequences
Abdellali Kelil1, Benjamin Dubreuil1, Emmanuel D Levy1
1Département de Biochimie and Centre Robert-Cedergren, Bio-Informatique et Génomique, Université de Montréal, Succursale Centre-Ville, Montreal, Quebec, Canada.

Abstract:
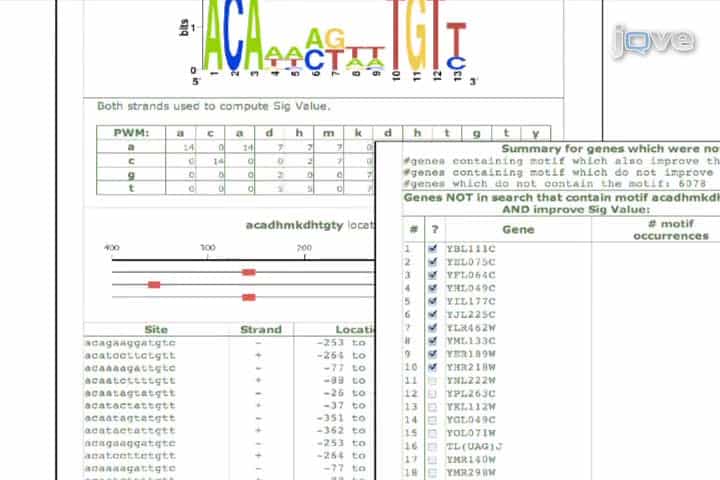
Linear motifs mediate a wide variety of cellular functions, which makes their characterization in protein sequences crucial to understanding cellular systems. However, the short length and degenerate nature of linear motifs make their discovery a difficult problem. Here, we introduce MotifHound, an algorithm particularly suited for the discovery of small and degenerate linear motifs. MotifHound performs an exact and exhaustive enumeration of all motifs present in proteins of interest, including all of their degenerate forms, and scores the overrepresentation of each motif based on its occurrence in proteins of interest relative to a background (e.g., proteome) using the hypergeometric distribution. To assess MotifHound, we benchmarked it together with state-of-the-art algorithms. The benchmark consists of 11,880 sets of proteins from S. cerevisiae; in each set, we artificially spiked-in one motif varying in terms of three key parameters, (i) number of occurrences, (ii) length and (iii) the number of degenerate or "wildcard" positions. The benchmark enabled the evaluation of the impact of these three properties on the performance of the different algorithms. The results showed that MotifHound and SLiMFinder were the most accurate in detecting degenerate linear motifs. Interestingly, MotifHound was 15 to 20 times faster at comparable accuracy and performed best in the discovery of highly degenerate motifs. We complemented the benchmark by an analysis of proteins experimentally shown to bind the FUS1 SH3 domain from S. cerevisiae. Using the full-length protein partners as sole information, MotifHound recapitulated most experimentally determined motifs binding to the FUS1 SH3 domain. Moreover, these motifs exhibited properties typical of SH3 binding peptides, e.g., high intrinsic disorder and evolutionary conservation, despite the fact that none of these properties were used as prior information. MotifHound is available (http://michnick.bcm.umontreal.ca or http://tinyurl.com/motifhound) together with the benchmark that can be used as a reference to assess future developments in motif discovery.
Related Concept Videos
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 02:26
Conservation of Protein Domains Over Different Proteins
A limited set of protein domains often duplicate and recombine during evolution. These domains can be organized in different combinations to...
02:26Conservation of Protein Domains
 02:26
Protein Networks
These interactions can be represented through maps depicting protein-protein interaction networks, represented as nodes and edges. Nodes are circles that are representative of a protein,...
02:26Protein Networks
 02:18
Intrinsically Disordered Proteins