Related Experiment Video
Updated: Apr 23, 2026


Protein WISDOM: A Workbench for In silico De novo Design of BioMolecules
Published on: July 25, 2013
A large-scale test of free-energy simulation estimates of protein-ligand binding affinities
Paulius Mikulskis1, Samuel Genheden, Ulf Ryde
1Department of Theoretical Chemistry, Lund University, Chemical Centre , P.O. Box 124, SE-221 00 Lund, Sweden.

Abstract:
We have performed a large-scale test of alchemical perturbation calculations with the Bennett acceptance-ratio (BAR) approach to estimate relative affinities for the binding of 107 ligands to 10 different proteins. Employing 20-Å truncated spherical systems and only one intermediate state in the perturbations, we obtain an error of less than 4 kJ/mol for 54% of the studied relative affinities and a precision of 0.5 kJ/mol on average. However, only four of the proteins gave acceptable errors, correlations, and rankings. The results could be improved by using nine intermediate states in the simulations or including the entire protein in the simulations using periodic boundary conditions. However, 27 of the calculated affinities still gave errors of more than 4 kJ/mol, and for three of the proteins the results were not satisfactory. This shows that the performance of BAR calculations depends on the target protein and that several transformations gave poor results owing to limitations in the molecular-mechanics force field or the restricted sampling possible within a reasonable simulation time. Still, the BAR results are better than docking calculations for most of the proteins.
More Related Videos
 10:29
Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
 13:26
Determination of Protein-ligand Interactions Using Differential Scanning Fluorimetry
Published on: September 13, 2014
Related Concept Videos
 02:18
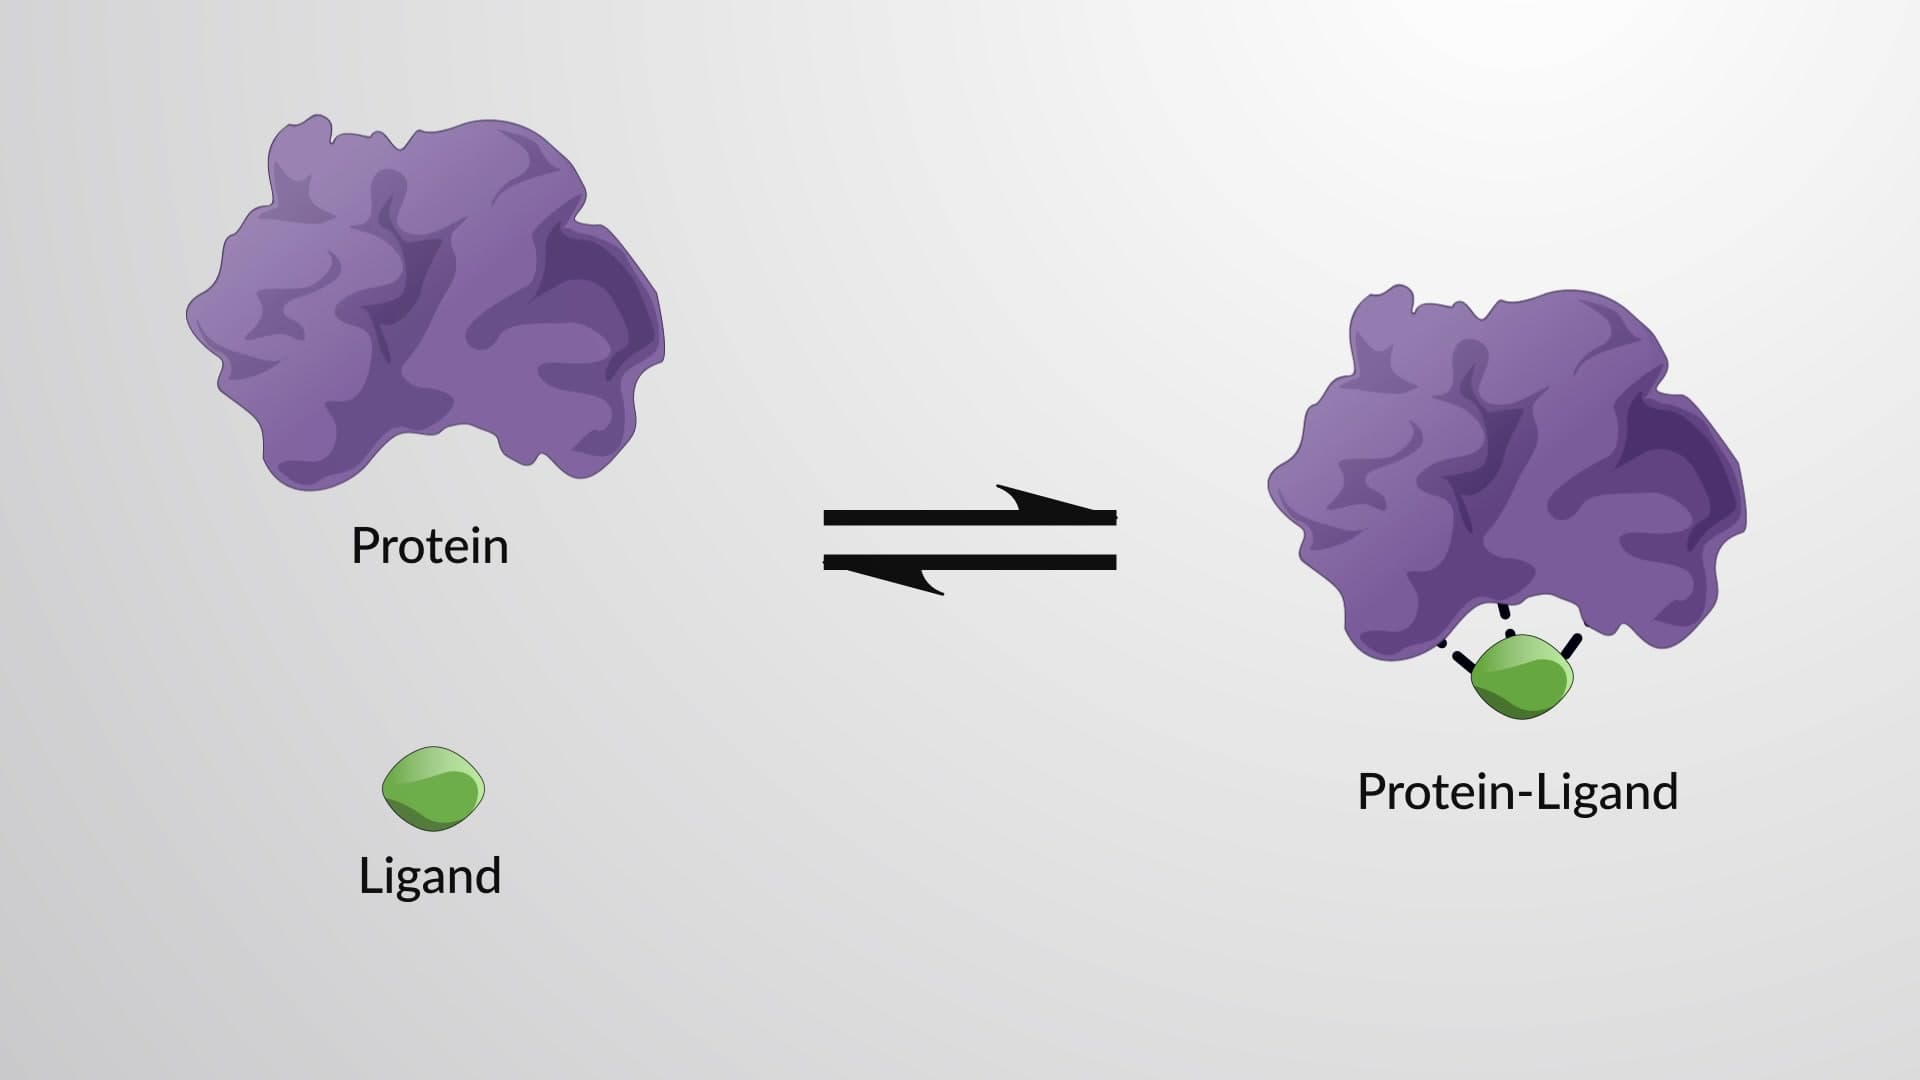
The Equilibrium Binding Constant and Binding Strength
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 01:22
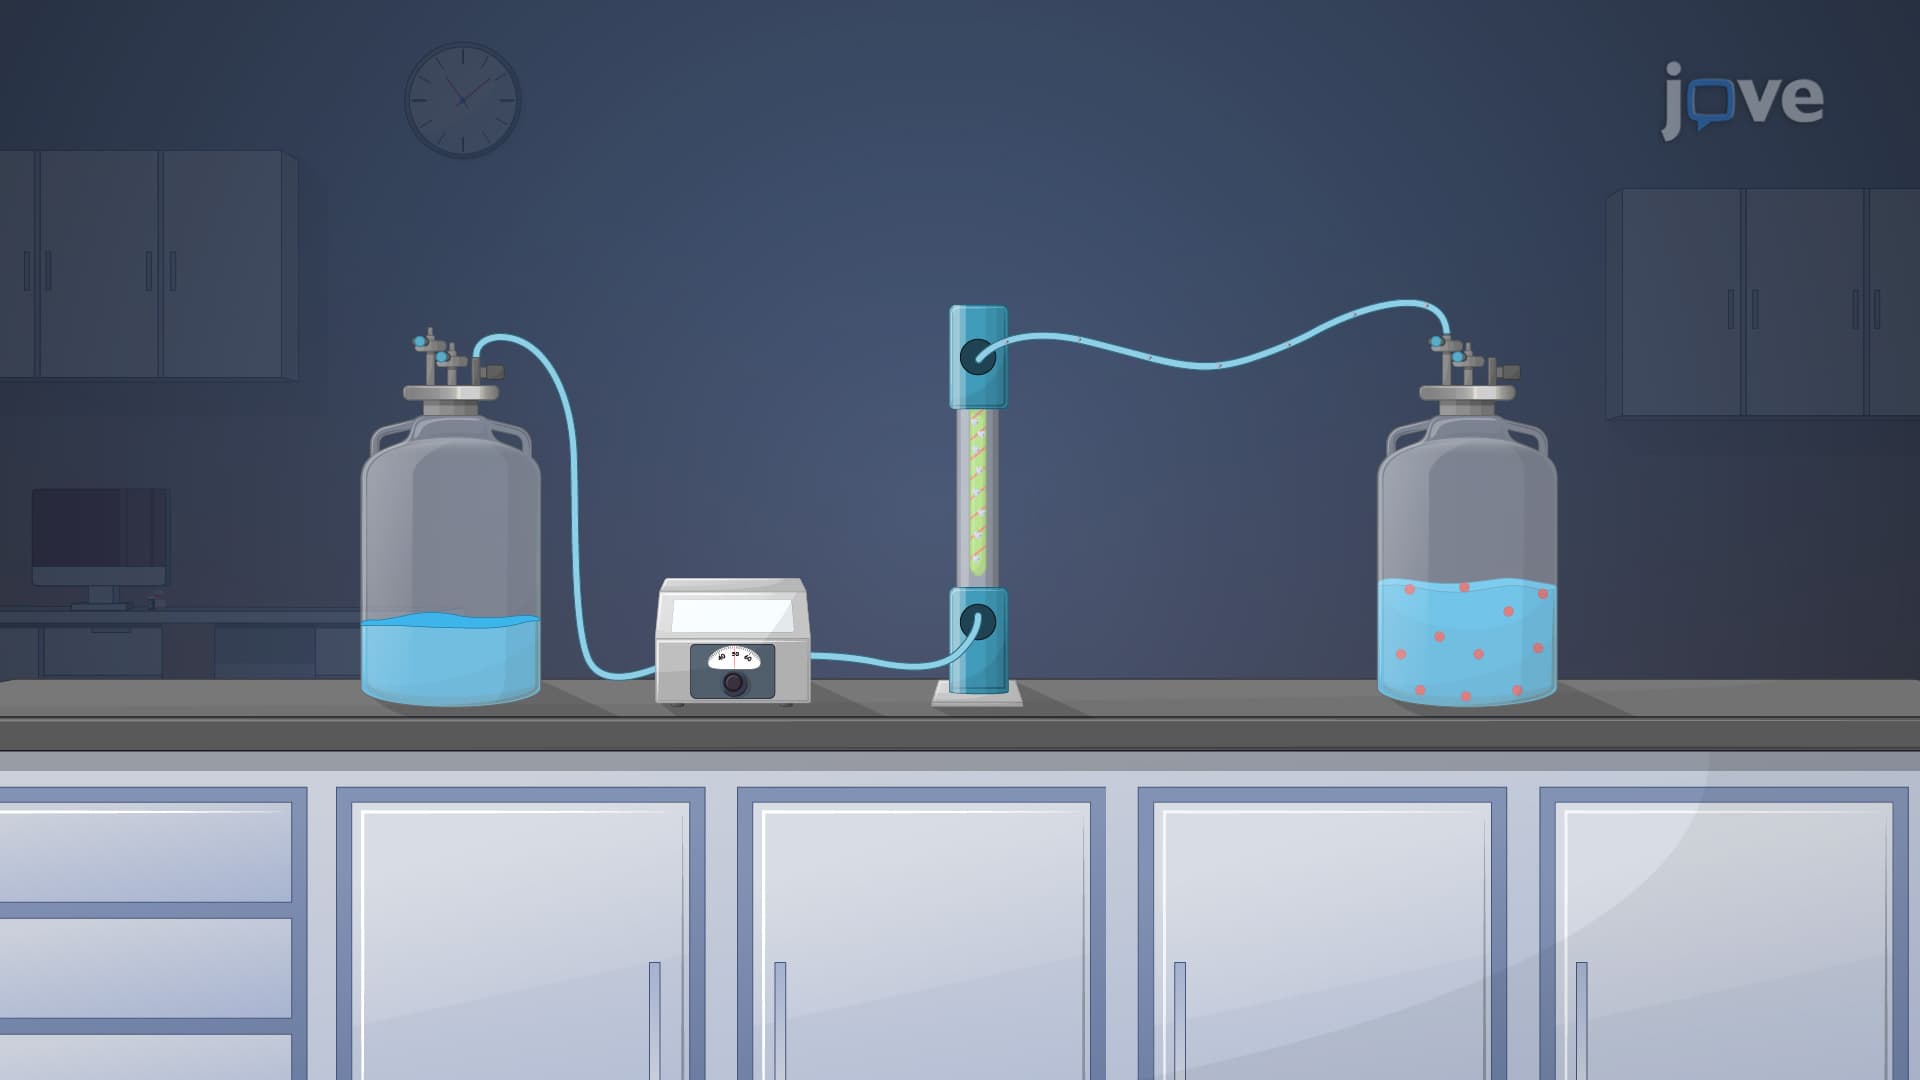
Protein-Drug Binding: Determination Methods
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...