Polyphony: superposition independent methods for ensemble-based drug discovery
Related Experiment Video
Updated: Apr 23, 2026

Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
William R Pitt1, Rinaldo W Montalvão, Tom L Blundell
1Department of Biochemistry, University of Cambridge, 80 Tennis Court Road, Cambridge CB2 1GA, UK. will.pitt@ucb.com.

Background:
Structure-based drug design is an iterative process, following cycles of structural biology, computer-aided design, synthetic chemistry and bioassay. In favorable circumstances, this process can lead to the structures of hundreds of protein-ligand crystal structures. In addition, molecular dynamics simulations are increasingly being used to further explore the conformational landscape of these complexes. Currently, methods capable of the analysis of ensembles of crystal structures and MD trajectories are limited and usually rely upon least squares superposition of coordinates.
Results:
Novel methodologies are described for the analysis of multiple structures of a protein. Statistical approaches that rely upon residue equivalence, but not superposition, are developed. Tasks that can be performed include the identification of hinge regions, allosteric conformational changes and transient binding sites. The approaches are tested on crystal structures of CDK2 and other CMGC protein kinases and a simulation of p38α. Known interaction - conformational change relationships are highlighted but also new ones are revealed. A transient but druggable allosteric pocket in CDK2 is predicted to occur under the CMGC insert. Furthermore, an evolutionarily-conserved conformational link from the location of this pocket, via the αEF-αF loop, to phosphorylation sites on the activation loop is discovered.
Conclusions:
New methodologies are described and validated for the superimposition independent conformational analysis of large collections of structures or simulation snapshots of the same protein. The methodologies are encoded in a Python package called Polyphony, which is released as open source to accompany this paper [http://wrpitt.bitbucket.org/polyphony/].
Related Concept Videos
 01:26
Drug Discovery: Overview
 01:09
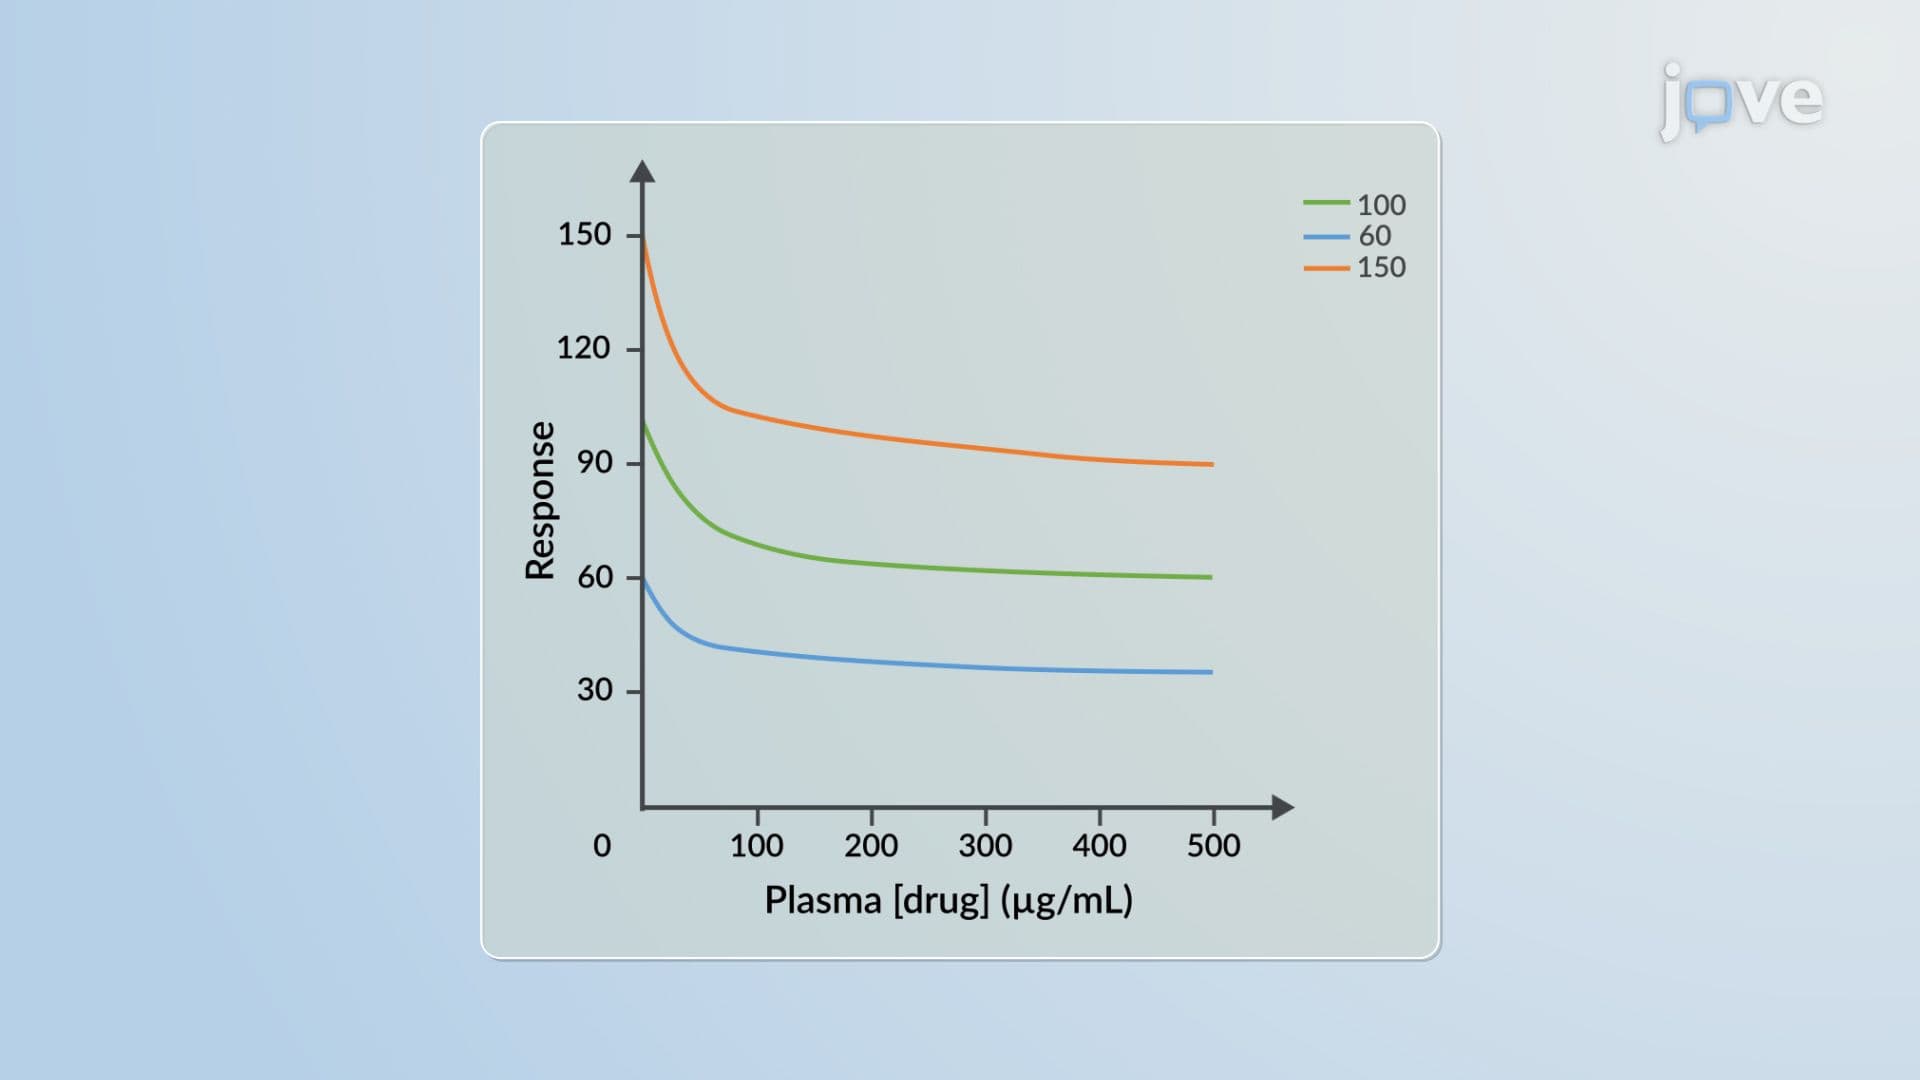
Pharmacodynamic Models: Additive and Proportional Drug Effect Model
 01:27
Combined Effects of Drugs: Synergism
Such synergistic combinations...
 01:29
Pharmacogenomics: Identification of New Drug Targets
 01:22
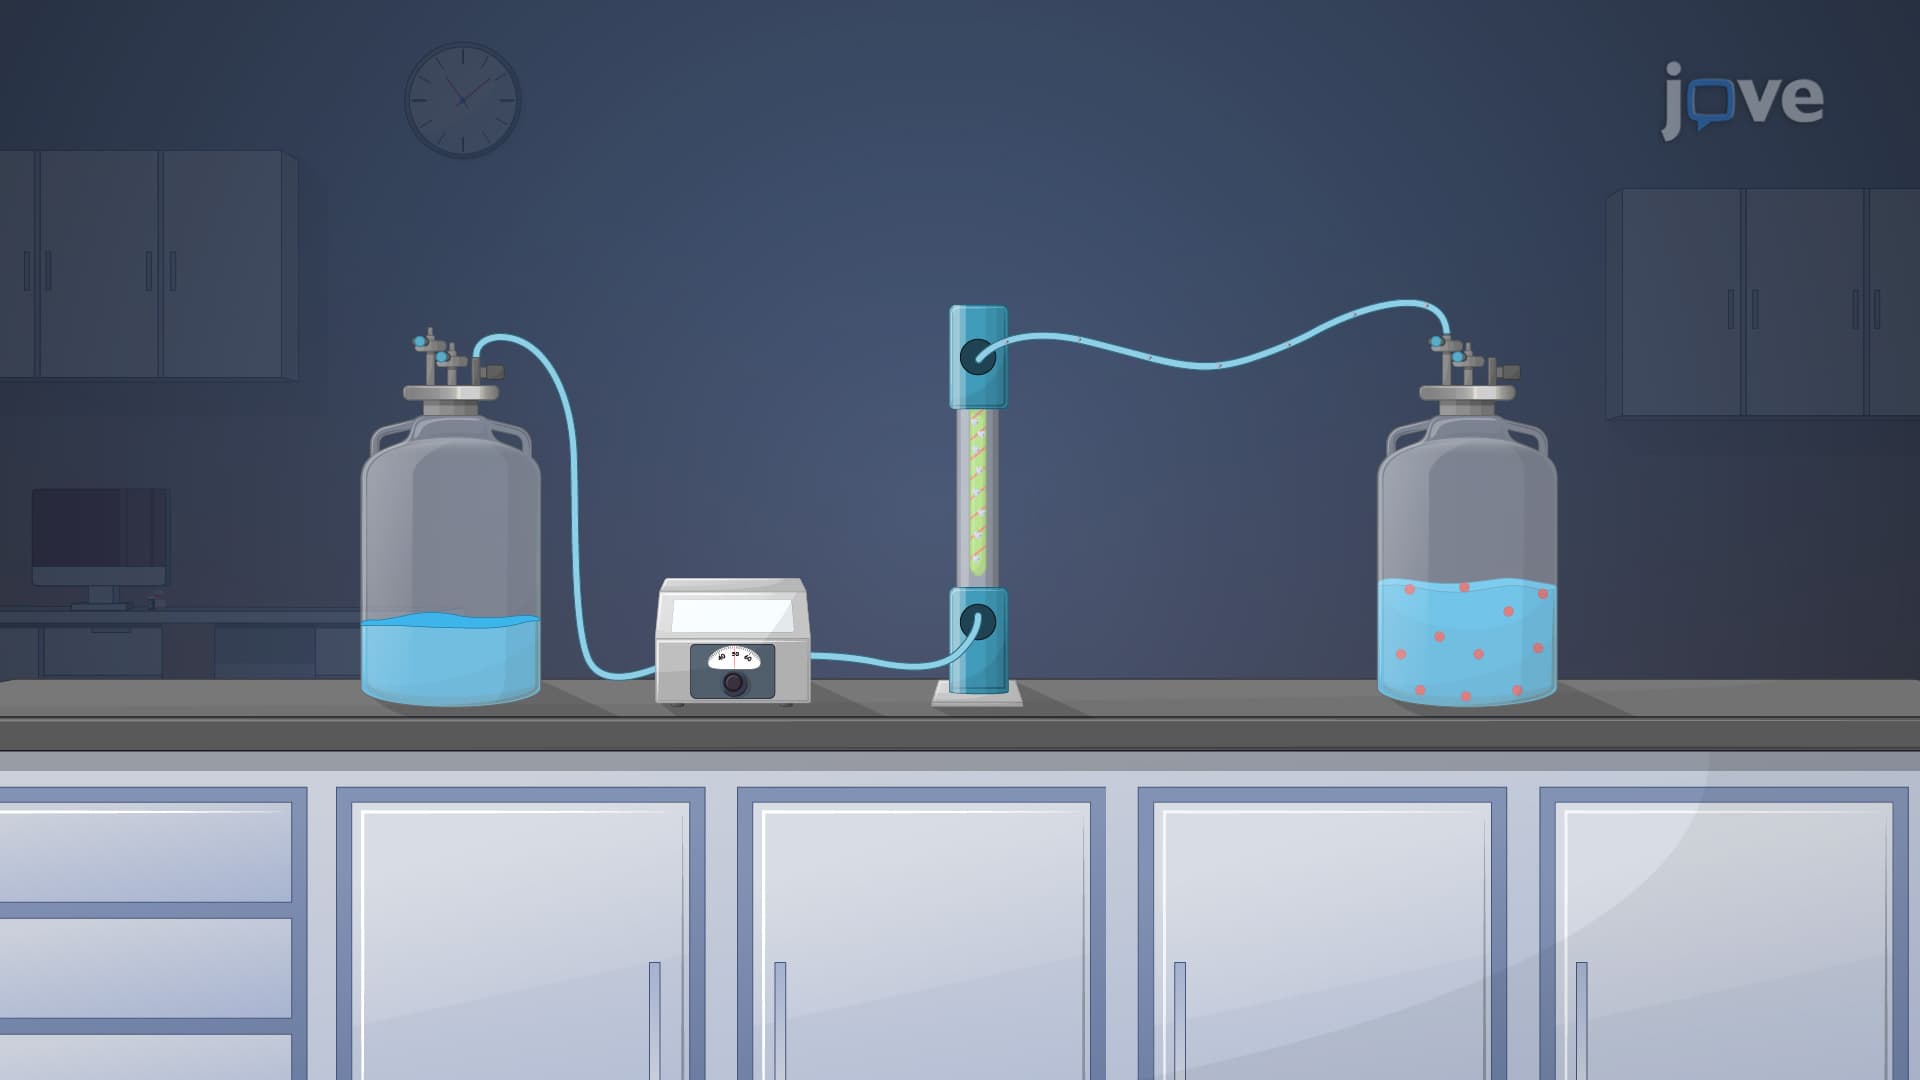
Protein-Drug Binding: Determination Methods
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...
 01:17
In Vitro Drug Dissolution: Alternative Methods