Related Experiment Video
Updated: Apr 23, 2026

Assessing Lysosomal Alkalinization in the Intestine of Live Caenorhabditis elegans
Published on: April 13, 2018
From bedside to cell biology: a century of history on lysosomal dysfunction
Maria Francisca Coutinho1, Liliana Matos1, Sandra Alves1
1Research and Development Unit, Department of Human Genetics, INSA, Portugal.

Abstract:
Lysosomal storage disorders (LSDs) are a group of rare genetic diseases, generally caused by a deficiency of specific lysosomal enzymes, which results in abnormal accumulation of undegraded substrates. The first clinical reports describing what were later shown to be LSDs were published more than a hundred years ago. In general, the history and pathophysiology of LSDs has impacted on our current knowledge of lysosomal biology. Classically, depending on the nature of the substrates, LSDs can be divided into different subgroups. The mucopolysaccharidoses (MPSs) are those caused by impaired degradation of glycosaminoglycans (GAGs). Amongst LSDs, the MPSs are a major group of pathologies with crucial historical relevance, since their study has revealed important biological pathways and highlighted interconnecting pathological cascades which are still being unveiled nowadays. Here we review the major historical discoveries in the field of LSDs and their impact on basic cellular knowledge and practical applications. Attention will be focused on the MPSs, with occasional references to other LSDs. We will show as studies on the metabolic basis of this group of diseases have increased our knowledge of the complex degradative pathways associated with the lysosome and established the basis to the development of specific therapeutic approaches aiming at correcting or, at least ameliorating their associated phenotypes.
Related Concept Videos
 01:22
Lysosomal Hydrolases
 01:36
Delivery Pathways to the Lysosome
Endocytosis
In endocytosis, the cell membrane takes up macromolecules and particles from the surrounding medium. Clathrin-mediated...
 01:31
Lysosomes
 01:27
Protein Import into the Peroxisomes
Peroxisomal Protein Import:
Peroxisomes lack the genetic machinery required to code for their own proteins. Hence, most peroxisomal membrane, lumenal and transmembrane proteins are synthesized in the cytoplasm or ER and transported to the peroxisome...
 01:26
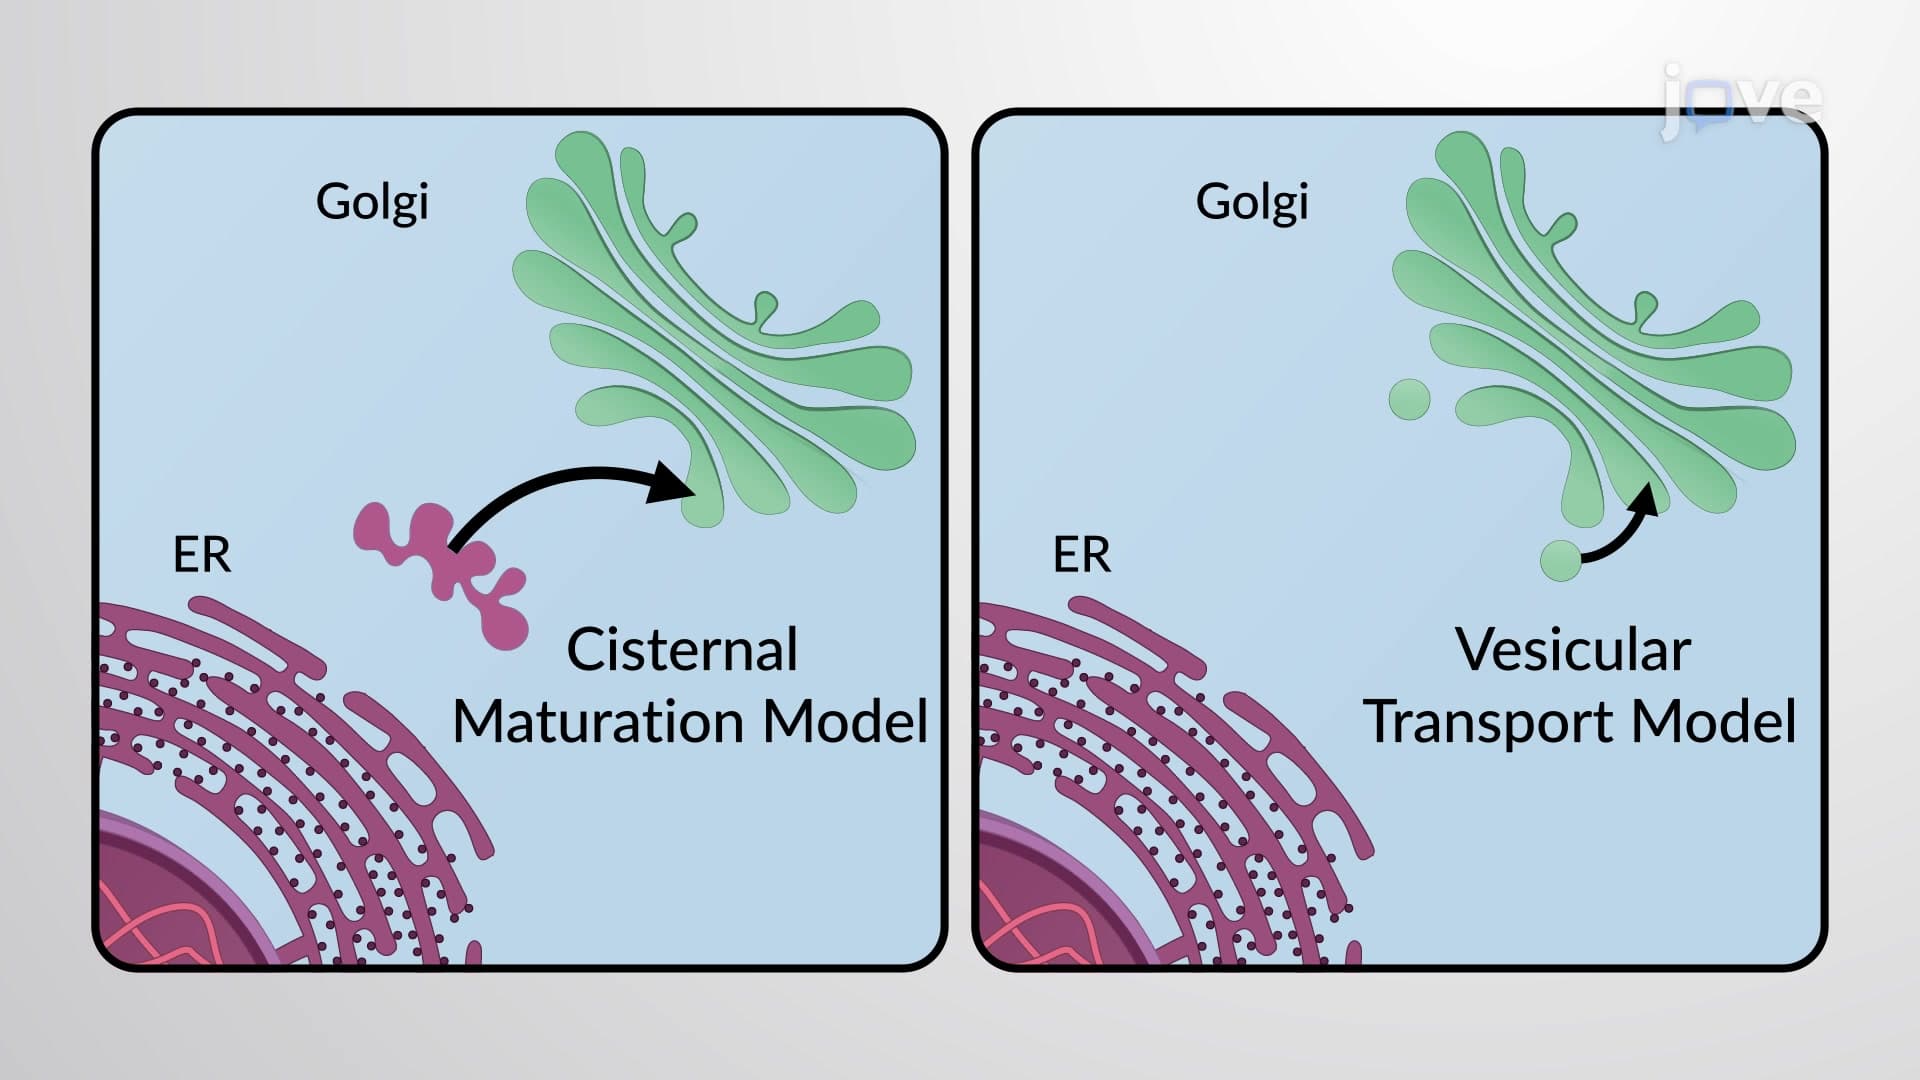
Transport Across the Golgi
 01:38
Intralumenal Vesicles and Multivesicular Bodies