Related Experiment Video
Updated: May 14, 2026
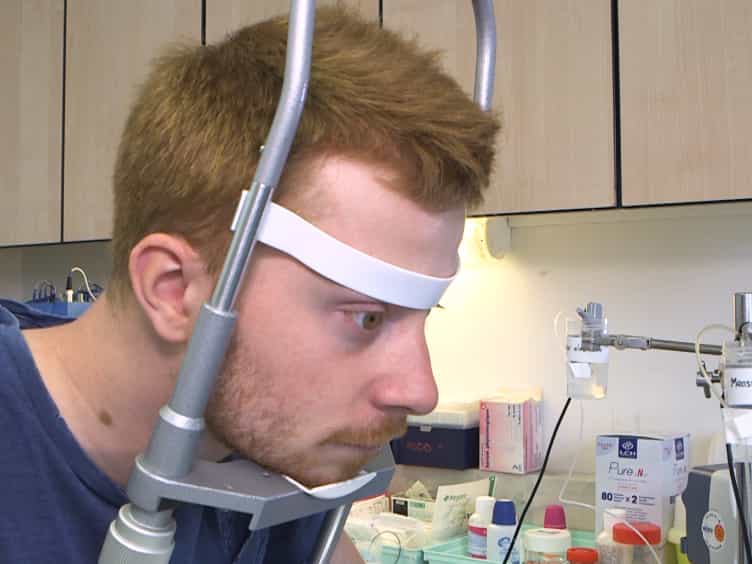
Standardized Measurement of Nasal Membrane Transepithelial Potential Difference NPD
Published on: September 13, 2018
Intestinal current measurement versus nasal potential difference measurements for diagnosis of cystic fibrosis: a
Azadeh Bagheri-Hanson, Sebastian Nedwed, Claudia Rueckes-Nilges
1Department of Pediatrics, Justus-Liebig-University Giessen, Feulgenstrasse 12, 35385 Giessen, Germany. lutz.naehrlich@paediat.med.uni-giessen.de.

Background:
Nasal potential difference (NPD) and intestinal current measurement (ICM) are functional CFTR tests that are used as adjunctive diagnostic tools for cystic fibrosis (CF). Smoking has a systemic negative impact on CFTR function. A diagnostic comparison between NPD and ICM and the impact of smoking on both CFTR tests has not been done.
Methods:
The sweat chloride test, NPD, and ICM were performed in 18 patients with CF (sweat chloride >60 mmol/l), including 6 pancreatic sufficient (PS) patients, and 13 healthy controls, including 8 smokers. The NPD CFTR response to Cl-free and isoproterenol perfusion (Δ0Cl- + Iso) was compared to the ICM CFTR response to forskolin/IBMX, carbachol, and histamine (ΔIsc, forskolin/IBMX+ carbachol+histamine).
Results:
The mean NPD CFTR response and ICM CFTR response between patients with CF and healthy controls was significantly different (p <0.001), but not between patients with CF who were PS and those who were pancreatic insufficient (PI). Smokers have a decreased CFTR response measured by NPD (p = 0.049). For ICM there is a trend towards decreased CFTR response (NS). Three healthy control smokers had NPD responses within the CF-range. In contrast to NPD, there was no overlap of the ICM response between patients with CF and controls.
Conclusions:
ICM is superior to NPD in distinguishing between patients with CF who have a sweat chloride > 60 mmol/l and healthy controls, including smokers. Neither NPD nor ICM differentiated between patients with CF who were PS from those who were PI. Smoking has a negative impact on CFTR function in healthy controls measured by NPD and challenges the diagnostic interpretation of NPD, but not ICM.
Related Concept Videos
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic sinusitis...