Related Experiment Video
Updated: Apr 21, 2026
Atomic Scale Structural Studies of Macromolecular Assemblies by Solid-state Nuclear Magnetic Resonance Spectroscopy
Published on: September 17, 2017
Conformational switching in pyruvic acid isolated in Ar and N₂ matrixes: spectroscopic analysis, anharmonic
Igor Reva1, Cláudio M Nunes1, Malgorzata Biczysko2,3
1†CQC, Department of Chemistry, University of Coimbra, 3004-535 Coimbra, Portugal.

Abstract:
Monomers of pyruvic acid (PA) isolated in cryogenic argon and nitrogen matrixes were characterized by mid- and near-infrared spectroscopy. Interpretation of the experiments was aided by fully anharmonic calculations of the fundamental modes, overtones, and combinations up to two quanta, including their infrared intensities. The initially dominating PA conformer (Tc) has a cis CCOH arrangement and is stabilized by a strong intramolecular H-bond. Selective near-infrared excitation of Tc at the first OH overtone (6630 cm(-1) in Ar, 6643 cm(-1) in N2) induced a large scale conformational conversion to the higher-energy conformer (Tt) with trans CCOH arrangement. Tt was then converted back to Tc by selective NIR irradiation at the first Tt OH overtone (6940 cm(-1) in Ar, 6894 cm(-1) in N2). In N2 matrix, the Tt form was stabilized due to interaction between the OH group and the matrix molecules. This stabilization manifested itself in the absence of Tt → Tc relaxation and in a considerable change of the vibrational Tt signature upon going from argon to nitrogen matrix. In argon, the Tt form spontaneously decayed back to Tc in the dark (characteristic lifetime +16 h). In the presence of broad-band near-infrared light, the Tt → Tc relaxation speed considerably increased. The decay mechanisms are discussed.
More Related Videos
Related Concept Videos
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
 01:15
¹H NMR of Conformationally Flexible Molecules: Variable-Temperature NMR
 01:20
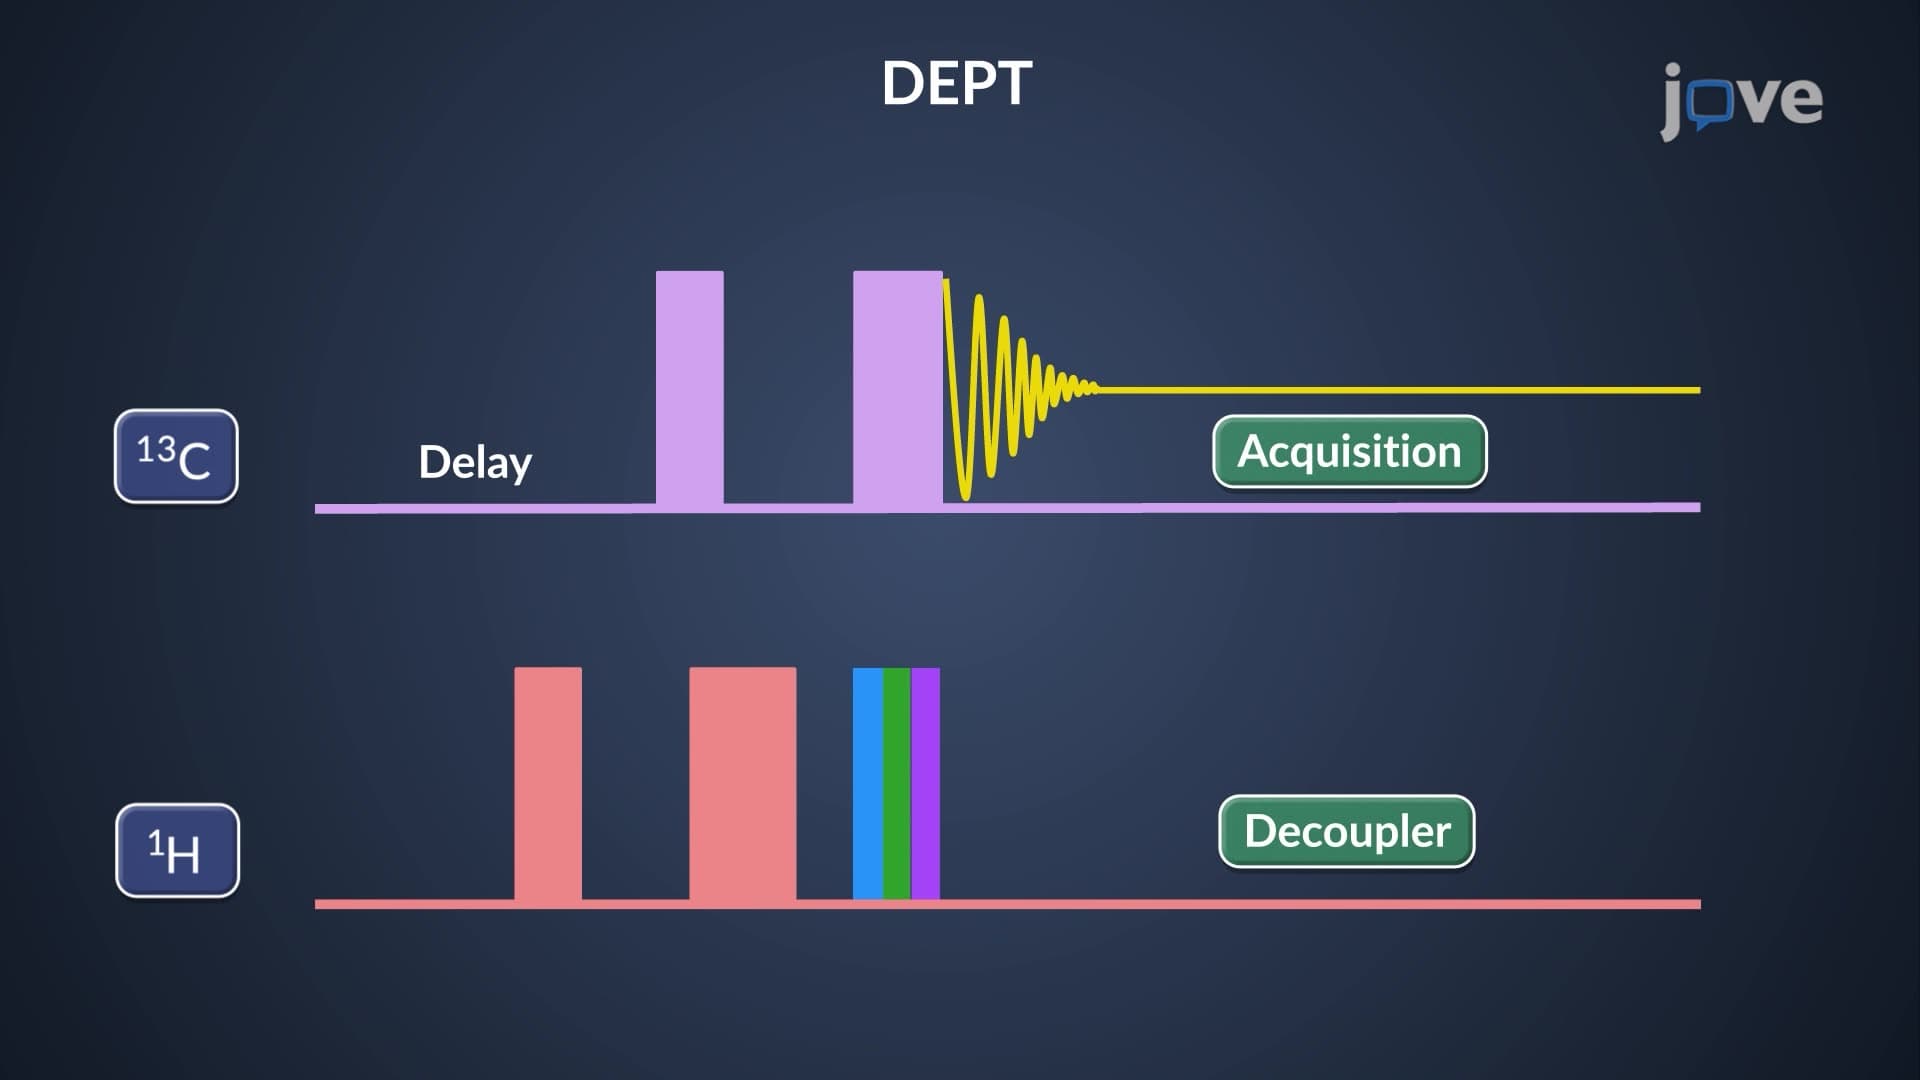
¹³C NMR: Distortionless Enhancement by Polarization Transfer (DEPT)
 01:14
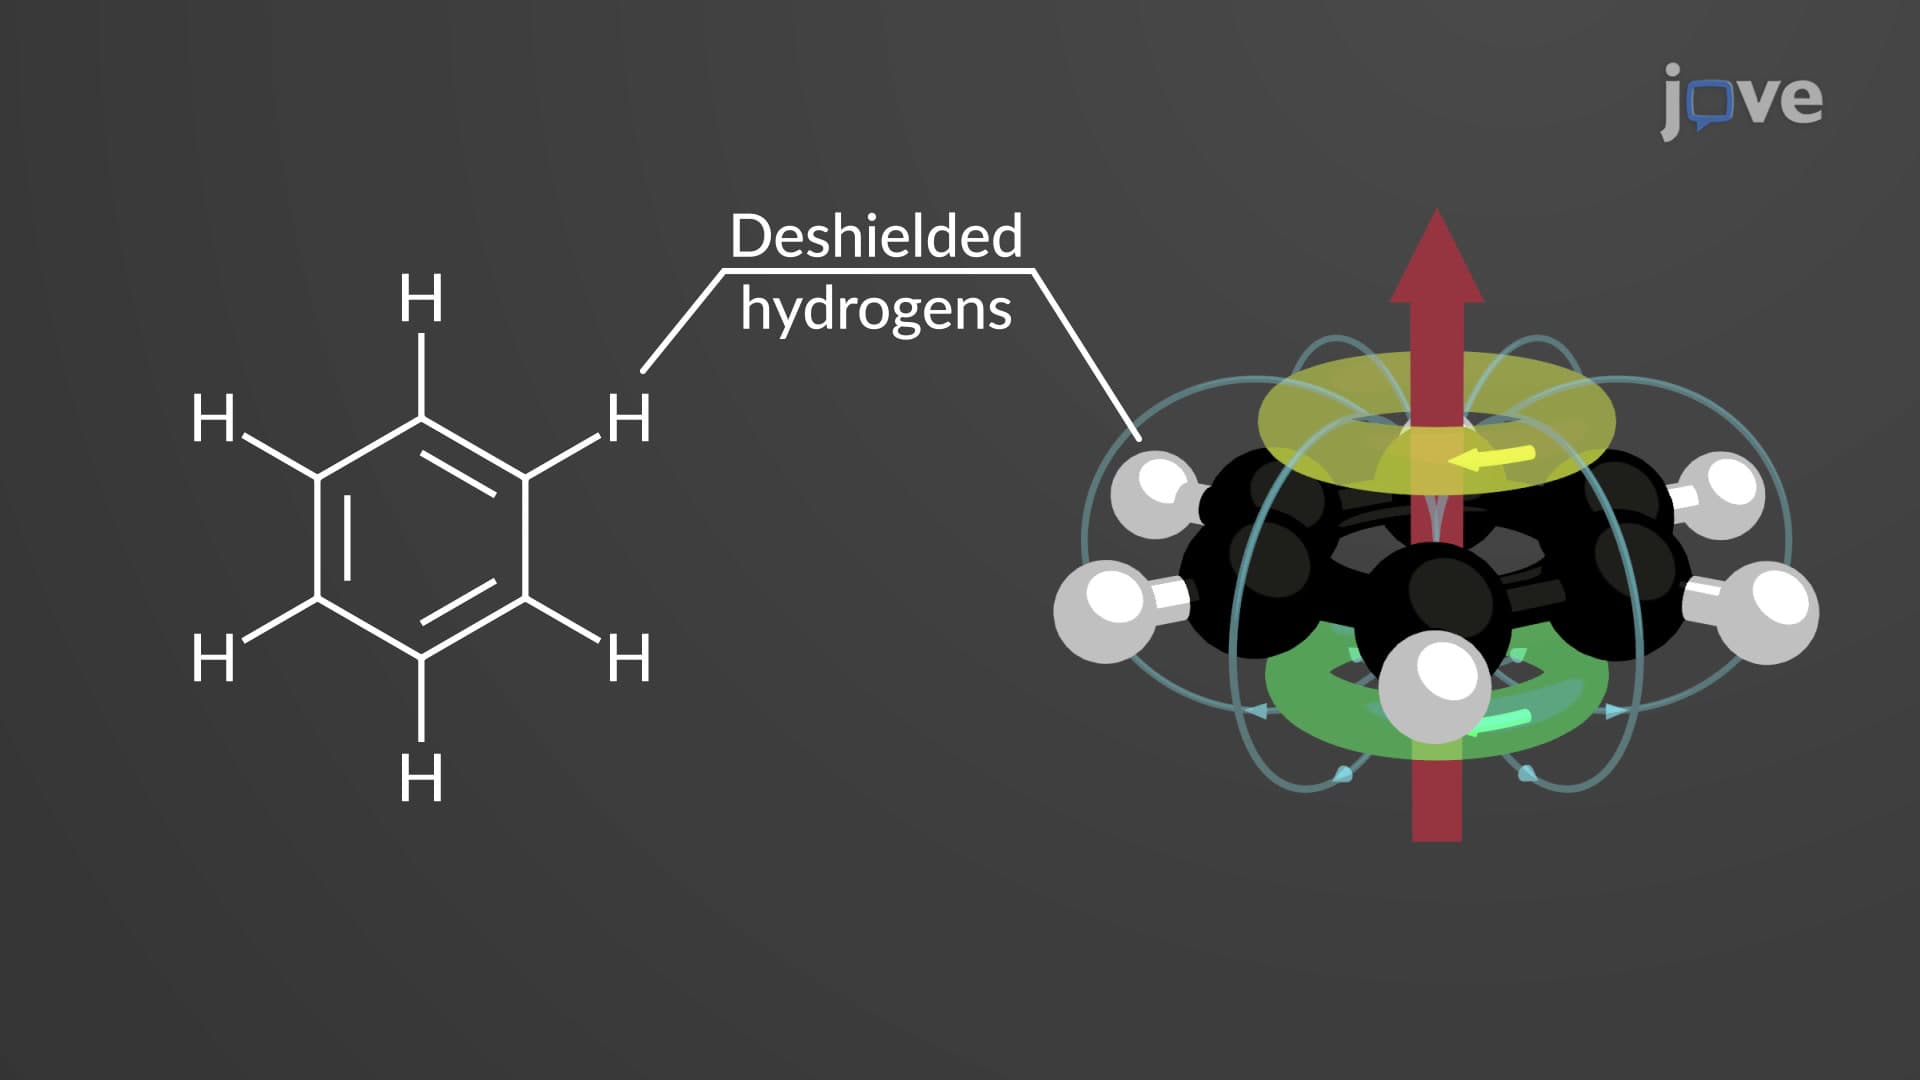
π Electron Effects on Chemical Shift: Aromatic and Antiaromatic Compounds
 01:14
NMR Spectroscopy of Aromatic Compounds
 01:32
UV–Vis Spectroscopy of Conjugated Systems
One of the factors influencing λmax is...