Related Experiment Video
Updated: Apr 21, 2026
Line Shape Analysis of Dynamic NMR Spectra for Characterizing Coordination Sphere Rearrangements at a Chiral Rhenium Polyhydride Complex
Published on: July 27, 2022
Density functional investigation of intermolecular effects on 13C NMR chemical-shielding tensors modeled with
Sean T Holmes1, Robbie J Iuliucci2, Karl T Mueller3
1Department of Chemistry and Biochemistry, University of Delaware, Newark, Delaware 19716, USA.

Abstract:
A quantum-chemical method for modeling solid-state nuclear magnetic resonance chemical-shift tensors by calculations on large symmetry-adapted clusters of molecules is demonstrated. Four hundred sixty five principal components of the (13)C chemical-shielding tensors of 24 organic materials are analyzed. The comparison of calculations on isolated molecules with molecules in clusters demonstrates that intermolecular effects can be successfully modeled using a cluster that represents a local portion of the lattice structure, without the need to use periodic-boundary conditions (PBCs). The accuracy of calculations which model the solid state using a cluster rivals the accuracy of calculations which model the solid state using PBCs, provided the cluster preserves the symmetry properties of the crystalline space group. The size and symmetry conditions that the model cluster must satisfy to obtain significant agreement with experimental chemical-shift values are discussed. The symmetry constraints described in the paper provide a systematic approach for incorporating intermolecular effects into chemical-shielding calculations performed at a level of theory that is more advanced than the generalized gradient approximation. Specifically, NMR parameters are calculated using the hybrid exchange-correlation functional B3PW91, which is not available in periodic codes. Calculations on structures of four molecules refined with density plane waves yield chemical-shielding values that are essentially in agreement with calculations on clusters where only the hydrogen sites are optimized and are used to provide insight into the inherent sensitivity of chemical shielding to lattice structure, including the role of rovibrational effects.
Related Concept Videos
 01:27
π Electron Effects on Chemical Shift: Overview
 01:14
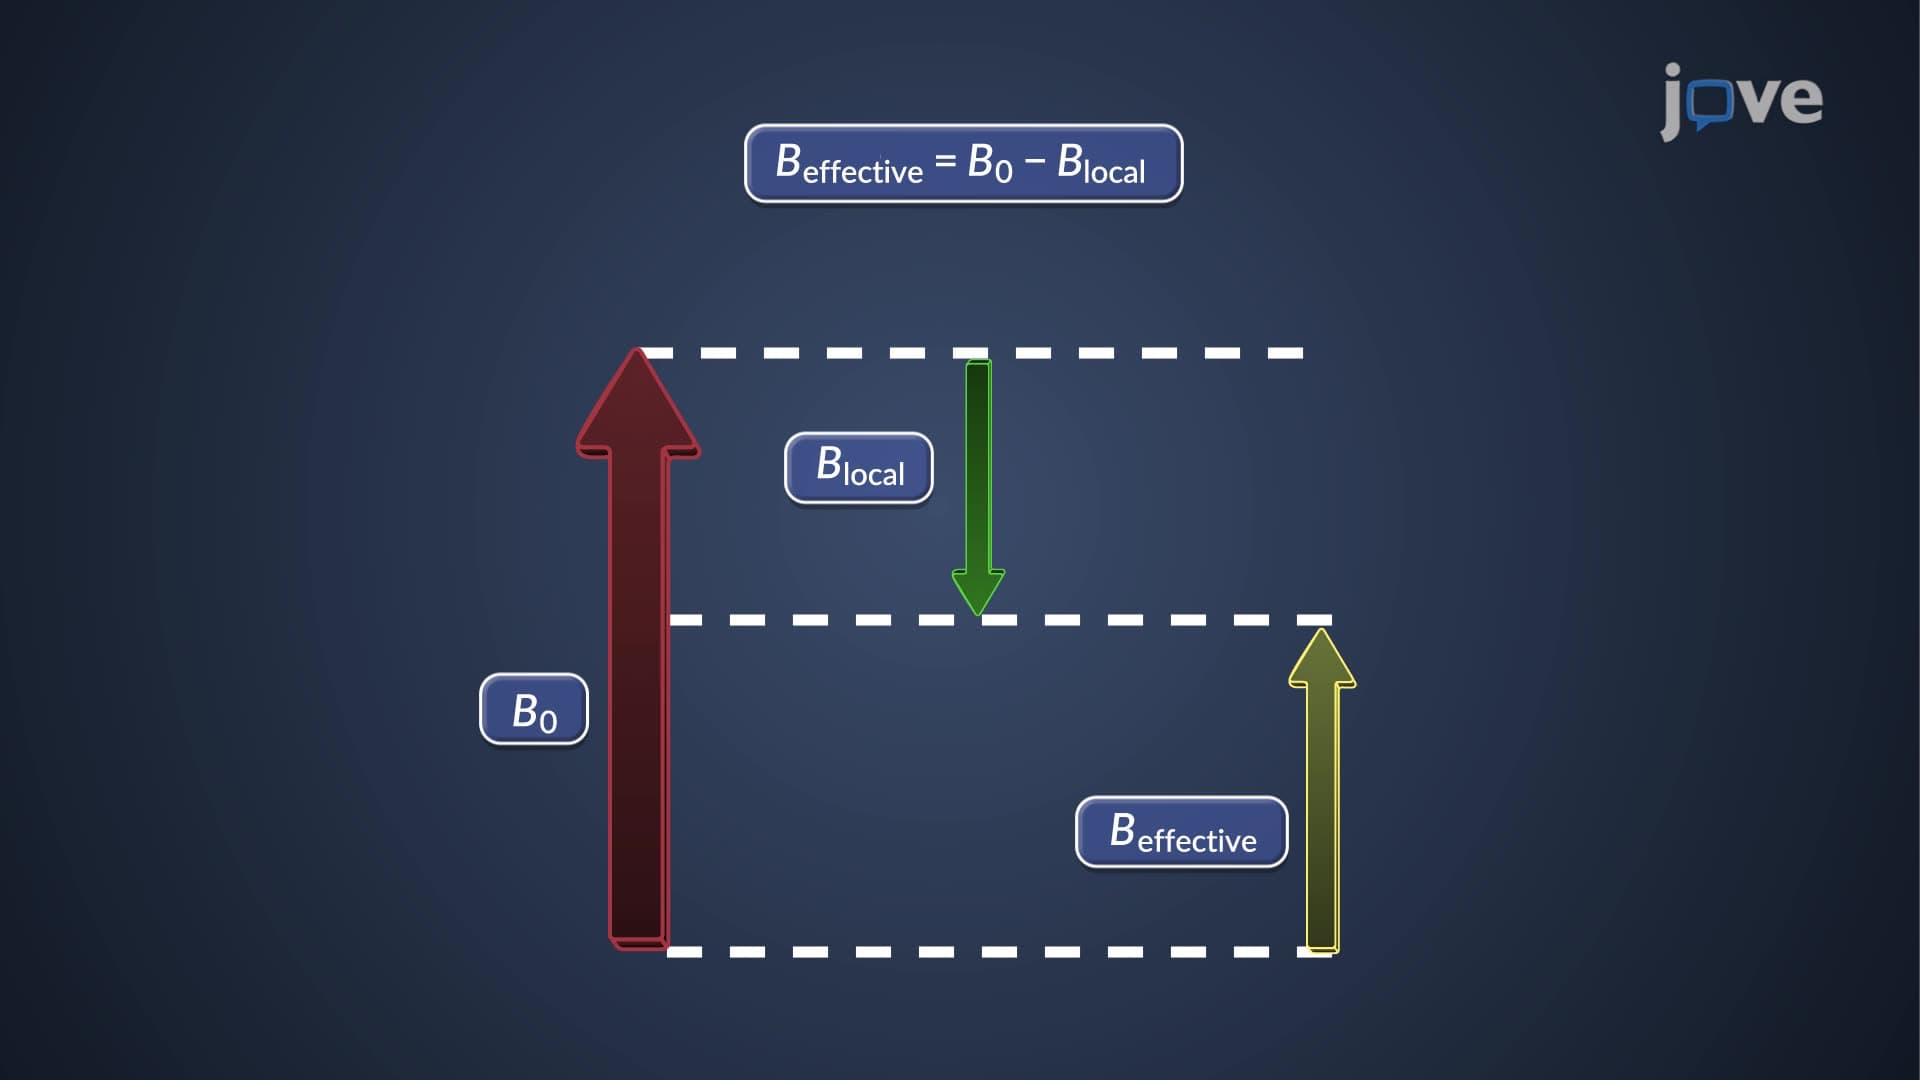
Diamagnetic Shielding of Nuclei: Local Diamagnetic Current
 01:14
π Electron Effects on Chemical Shift: Aromatic and Antiaromatic Compounds
 01:27
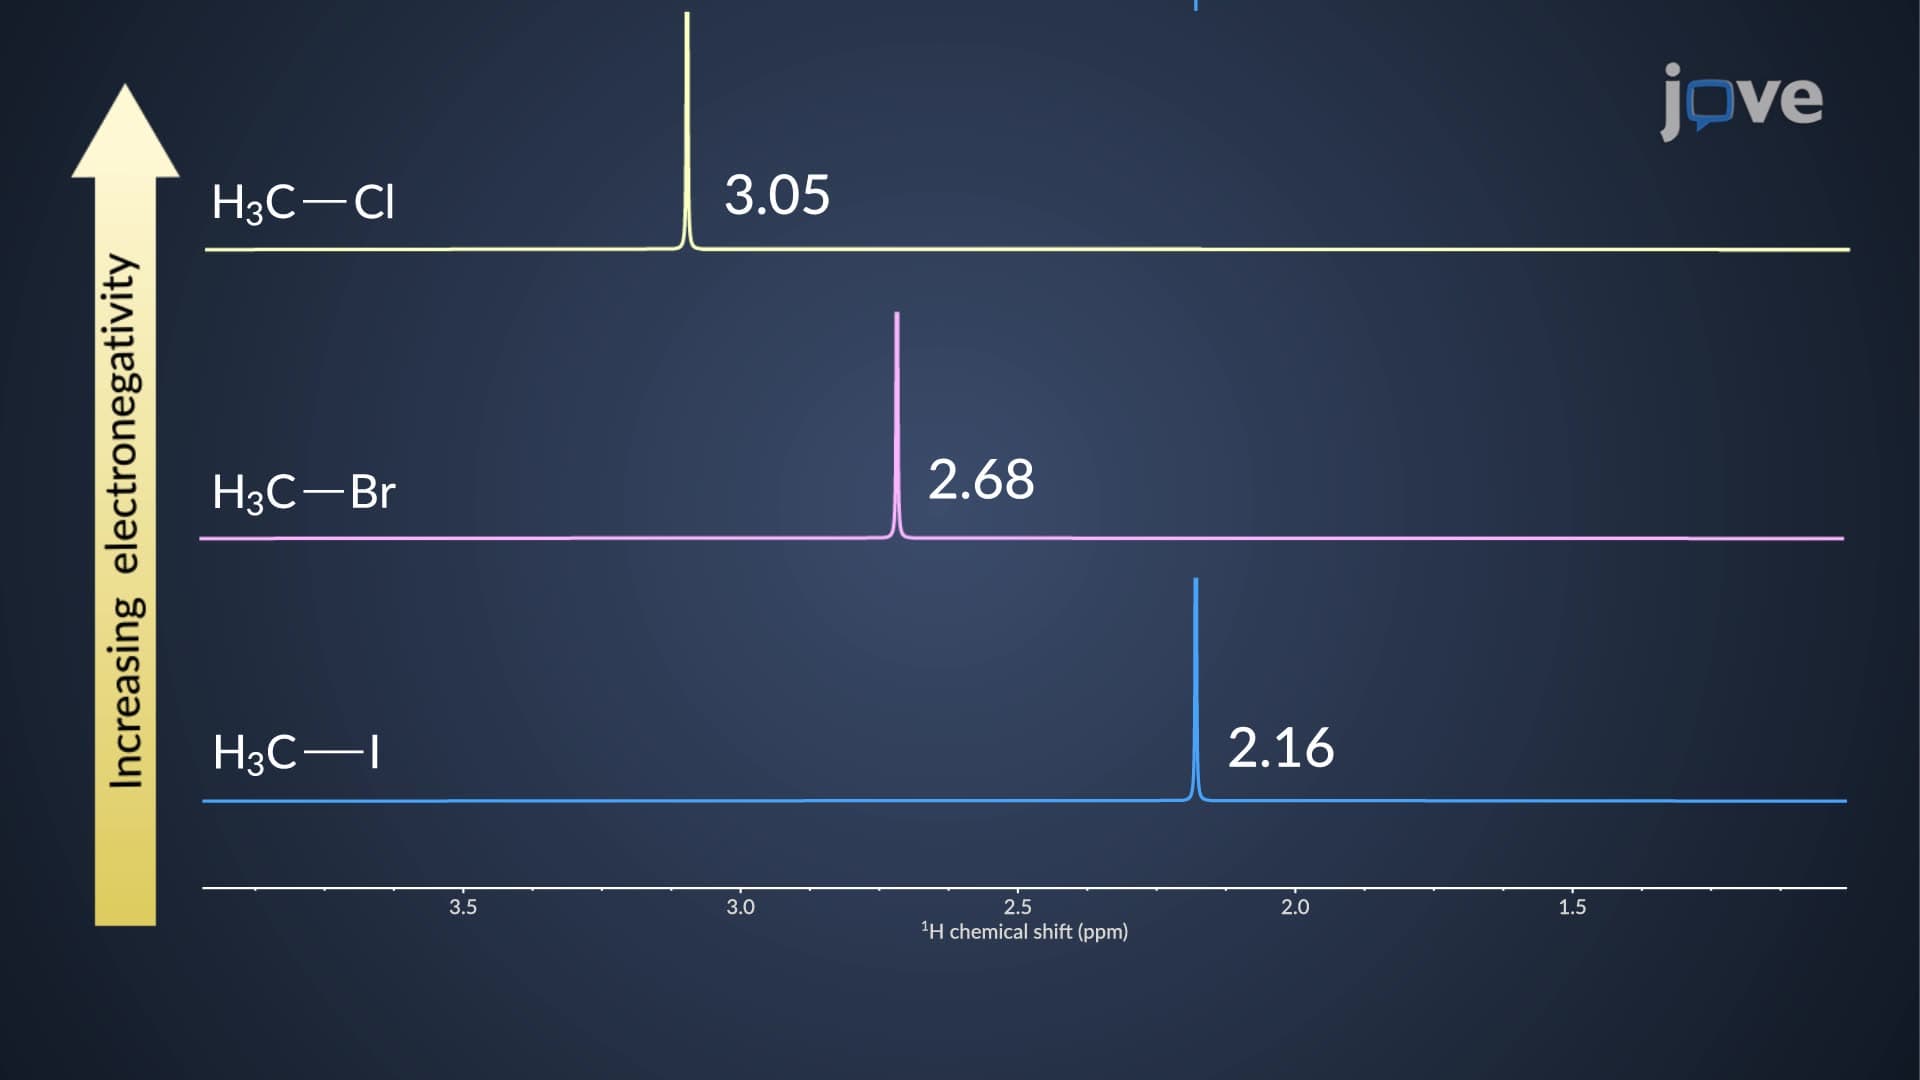
Inductive Effects on Chemical Shift: Overview
 01:17
Chemical Shift: Internal References and Solvent Effects
The internal reference compound generally used in NMR spectroscopy is tetramethylsilane (TMS). TMS is preferred because it is chemically inert, soluble in NMR solvents, and easily removable. Also, the highly shielded methyl protons in TMS yield an intense...
 01:02
¹H NMR: Interpreting Distorted and Overlapping Signals
As Δν decreases and the signals move closer, the doublets appear increasingly distorted. The intensities of the inner lines increase at the cost of those of the outer lines as the signals are...