Related Experiment Video
Updated: Apr 21, 2026
A Fluorescence-based Assay for Characterization and Quantification of Lipid Droplet Formation in Human Intestinal Organoids
Published on: October 13, 2019
Insights from human congenital disorders of intestinal lipid metabolism
1Research Centre, CHU Sainte-Justine and Department of Nutrition, Université de Montréal, Montreal, Quebec H3T 1C5, Canada.

Abstract:
The intestine must challenge the profuse daily flux of dietary fat that serves as a vital source of energy and as an essential component of cell membranes. The fat absorption process takes place in a series of orderly and interrelated steps, including the uptake and translocation of lipolytic products from the brush border membrane to the endoplasmic reticulum, lipid esterification, Apo synthesis, and ultimately the packaging of lipid and Apo components into chylomicrons (CMs). Deciphering inherited disorders of intracellular CM elaboration afforded new insight into the key functions of crucial intracellular proteins, such as Apo B, microsomal TG transfer protein, and Sar1b GTPase, the defects of which lead to hypobetalipoproteinemia, abetalipoproteinemia, and CM retention disease, respectively. These "experiments of nature" are characterized by fat malabsorption, steatorrhea, failure to thrive, low plasma levels of TGs and cholesterol, and deficiency of liposoluble vitamins and essential FAs. After summarizing and discussing the functions and regulation of these proteins for reader's comprehension, the current review focuses on their specific roles in malabsorptions and dyslipidemia-related intestinal fat hyperabsorption while dissecting the spectrum of clinical manifestations and managements. The influence of newly discovered proteins (proprotein convertase subtilisin/kexin type 9 and angiopoietin-like 3 protein) on fat absorption has also been provided. Finally, it is stressed how the overexpression or polymorphism status of the critical intracellular proteins promotes dyslipidemia and cardiometabolic disorders.
Related Concept Videos
 01:27
Protein Import into the Peroxisomes
Peroxisomal Protein Import:
Peroxisomes lack the genetic machinery required to code for their own proteins. Hence, most peroxisomal membrane, lumenal and transmembrane proteins are synthesized in the cytoplasm or ER and transported to the peroxisome...
 01:20
Inborn Errors of Metabolism
 01:24
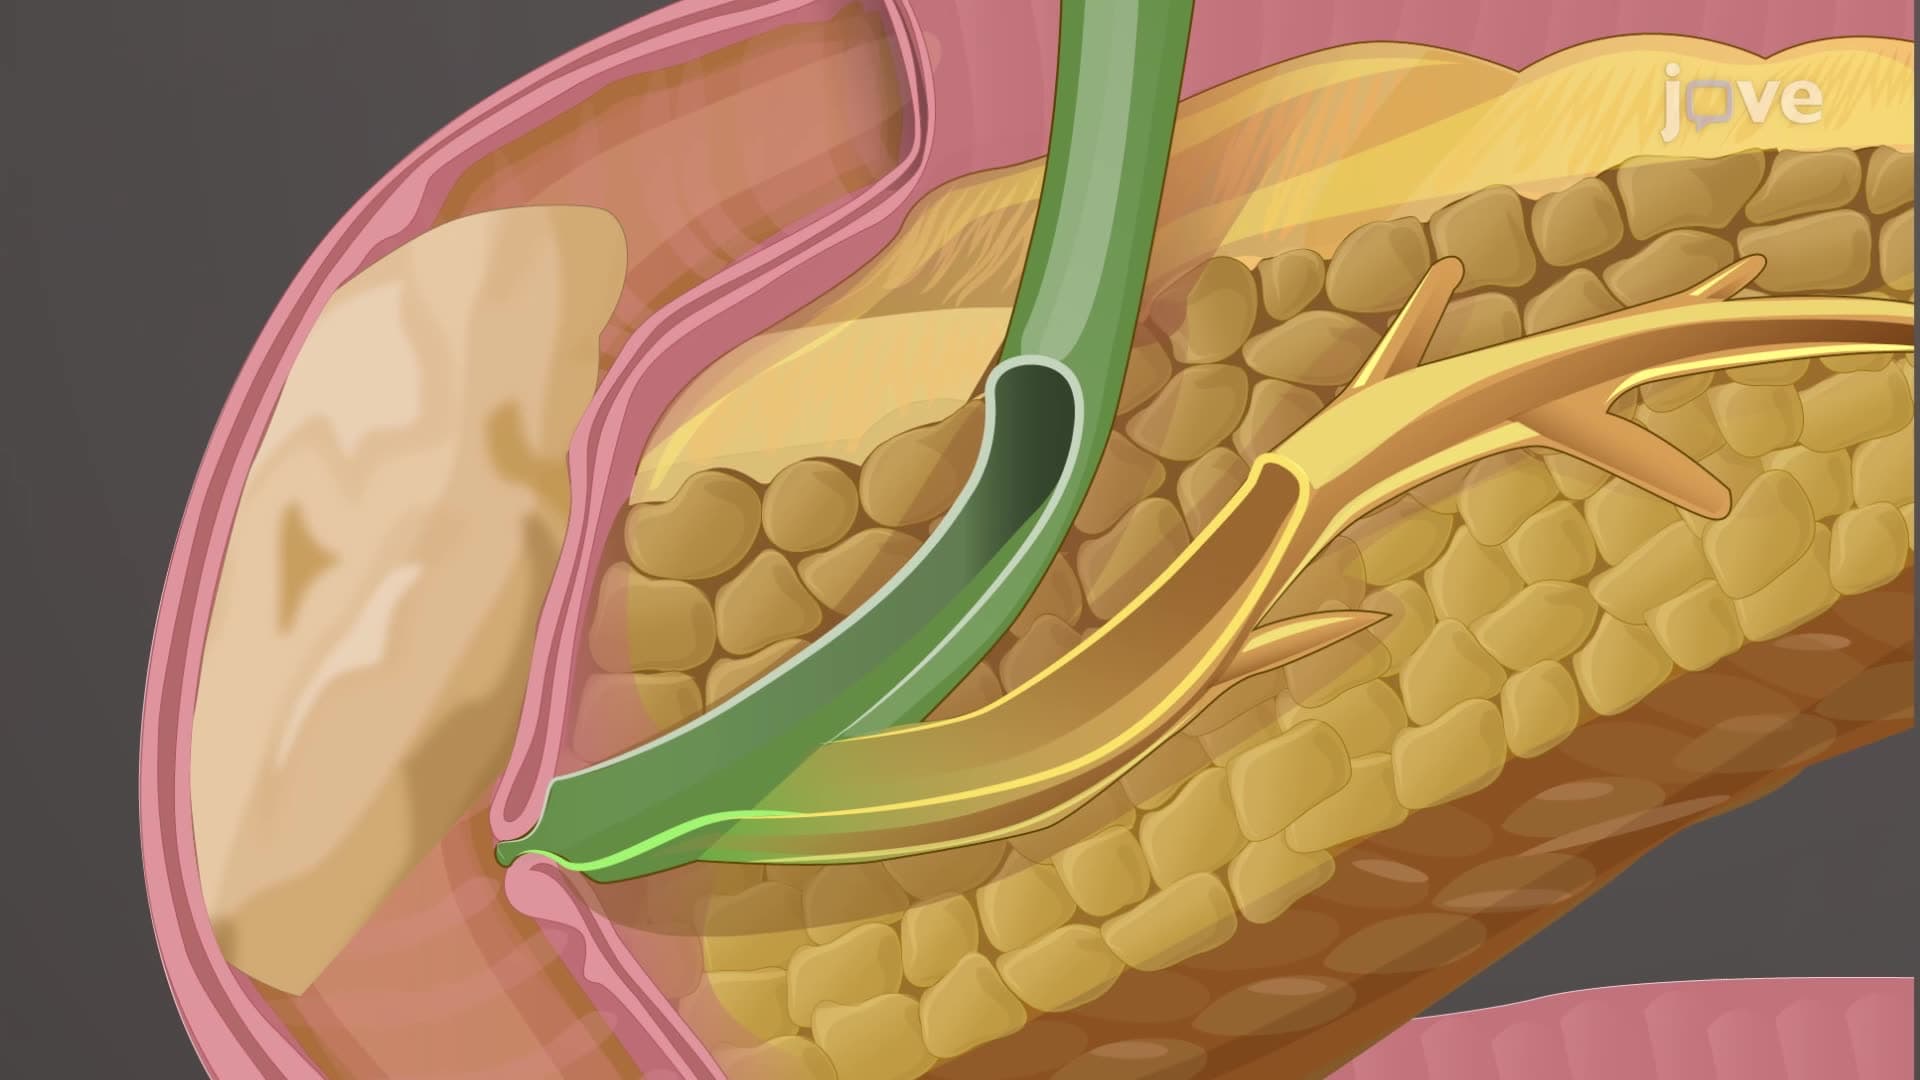
Lipid Absorption
These breakdown products bind with bile salts and lecithin to form micelles, which quickly pass between microvilli to come in close contact with the apical...
 01:24
Overview of Lipid Metabolism
Lipolysis: The Breakdown of Lipids:
Lipolysis is the process of breaking down lipids, particularly triglycerides, into glycerol and fatty acids. This process typically occurs in the adipose tissue and is triggered by various hormones, including glucagon and...
 01:27
Glucose Transporters
Facilitated diffusion-glucose transporters (GLUTs) are encoded by the solute-linked carrier (SLC) family 2, subfamily A gene family, or SLC2A. The 14 GLUT protein members are distributed into three classes:
 01:29
Pharmacogenomics: Identification of New Drug Targets