Related Experiment Video
Updated: Apr 21, 2026
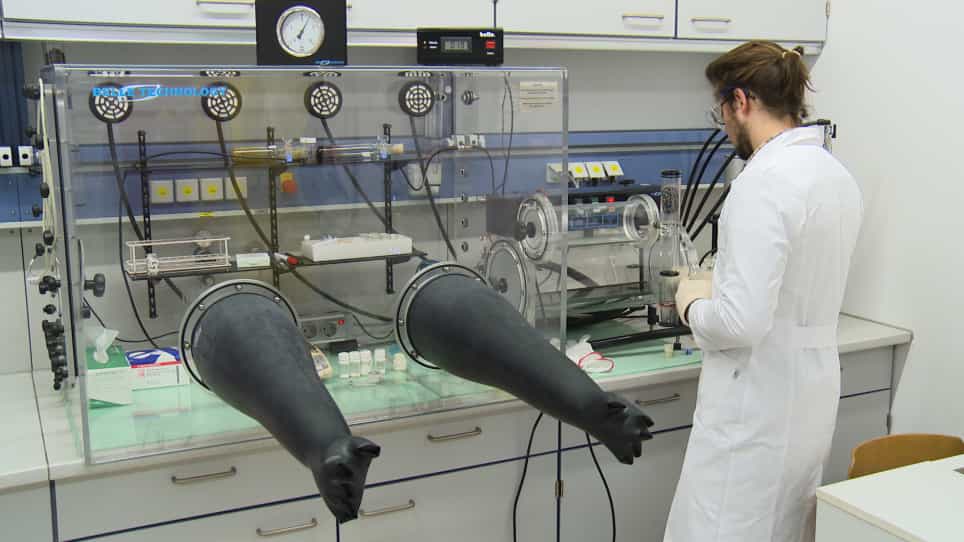

Isolating Free Carbenes, their Mixed Dimers and Organic Radicals
Published on: April 19, 2019
Coinage metals binding as main group elements: structure and bonding of the carbene complexes [TM(cAAC)2] and
Paul Jerabek1, Herbert W Roesky, Guy Bertrand
1Fachbereich Chemie, Philipps-Universität Marburg , Hans-Meerwein-Strasse, D-35032 Marburg, Germany.

Abstract:
Quantum chemical calculations using density functional theory have been carried out for the cyclic (alkyl)(amino)carbene (cAAC) complexes of the group 11 atoms [TM(cAAC)2] (TM = Cu, Ag, Au) and their cations [TM(cAAC)2](+). The nature of the metal-ligand bonding was investigated with the charge and energy decomposition analysis EDA-NOCV. The calculations show that the TM-C bonds in the charged adducts [TM(cAAC)2](+) are significantly longer than in the neutral complexes [TM(cAAC)2], but the cations have much higher bond dissociation energies than the neutral molecules. The intrinsic interaction energies ΔEint in [TM(cAAC)2](+) take place between TM(+) in the (1)S electronic ground state and (cAAC)2. In contrast, the metal-ligand interactions in [TM(cAAC)2] involve the TM atoms in the excited (1)P state yielding strong TM p(π) → (cAAC)2 π backdonation, which is absent in the cations. The calculations suggest that the cAAC ligands in [TM(cAAC)2] are stronger π acceptors than σ donors. The trends of the intrinsic interaction energies and the bond dissociation energies of the metal-ligand bonds in [TM(cAAC)2] and [TM(cAAC)2](+) give the order Au > Cu > Ag. Calculations at the nonrelativistic level give weaker TM-C bonds, particularly for the gold complexes. The trend for the bond strength in the neutral and charged adducts without relativistic effects becomes Cu > Ag > Au. The EDA-NOCV calculations suggest that the weaker bonds at the nonrelativistic level are mainly due to stronger Pauli repulsion and weaker orbital interactions. The NBO picture of the C-TM-C bonding situation does not correctly represent the nature of the metal-ligand interactions in [TM(cAAC)2].
More Related Videos
 09:45
Accessing Valuable Ligand Supports for Transition Metals: A Modified, Intermediate Scale Preparation of 1,2,3,4,5-Pentamethylcyclopentadiene
Published on: March 20, 2017
 19:58
Palladium N-Heterocyclic Carbene Complexes: Synthesis from Benzimidazolium Salts and Catalytic Activity in Carbon-carbon Bond-forming Reactions
Published on: July 30, 2017
Related Concept Videos
 01:18
Aromatic Hydrocarbon Cations: Structural Overview
Removing one hydrogen from the intervening CH2 group...
 02:10
Carbocations
 01:07
Radicals: Electronic Structure and Geometry
Accordingly, the structure of a trivalent radical lies between the geometries of carbocations and carbanions. An sp2-hybridized carbocation is trigonal planar, while an sp3-hybridized carbanion is trigonal pyramidal. Here, the difference in geometry is...
 02:42
Valence Bond Theory
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than...
 01:23
Properties of Organometallic Compounds