
Mito-protective autophagy is impaired in erythroid cells of aged mtDNA-mutator mice
XiuJie Li-Harms1, Sandra Milasta2, John Lynch3
1Department of Pathology.

Abstract:
Somatic mitochondrial DNA (mtDNA) mutations contribute to the pathogenesis of age-related disorders, including myelodysplastic syndromes (MDS). The accumulation of mitochondria harboring mtDNA mutations in patients with these disorders suggests a failure of normal mitochondrial quality-control systems. The mtDNA-mutator mice acquire somatic mtDNA mutations via a targeted defect in the proofreading function of the mtDNA polymerase, PolgA, and develop macrocytic anemia similar to that of patients with MDS. We observed an unexpected defect in clearance of dysfunctional mitochondria at specific stages during erythroid maturation in hematopoietic cells from aged mtDNA-mutator mice. Mechanistically, aberrant activation of mechanistic target of rapamycin signaling and phosphorylation of uncoordinated 51-like kinase (ULK) 1 in mtDNA-mutator mice resulted in proteasome-mediated degradation of ULK1 and inhibition of autophagy in erythroid cells. To directly evaluate the consequence of inhibiting autophagy on mitochondrial function in erythroid cells harboring mtDNA mutations in vivo, we deleted Atg7 from erythroid progenitors of wild-type and mtDNA-mutator mice. Genetic disruption of autophagy did not cause anemia in wild-type mice but accelerated the decline in mitochondrial respiration and development of macrocytic anemia in mtDNA-mutator mice. These findings highlight a pathological feedback loop that explains how dysfunctional mitochondria can escape autophagy-mediated degradation and propagate in cells predisposed to somatic mtDNA mutations, leading to disease.
Insights
Mitochondrial DNA mutations impair autophagy, leading to dysfunctional mitochondria accumulation and anemia in myelodysplastic syndromes (MDS). This study reveals a pathological loop driving disease progression in MDS.
Area of Science:
- Cell Biology
- Genetics
- Hematology
Background:
- Somatic mitochondrial DNA (mtDNA) mutations are implicated in age-related diseases like myelodysplastic syndromes (MDS).
- Accumulation of mutated mitochondria suggests impaired mitochondrial quality control in MDS pathogenesis.
- mtDNA-mutator mice, with defective mtDNA polymerase proofreading, exhibit MDS-like anemia.
Purpose of the Study:
- To investigate the role of mitochondrial quality control failure in MDS.
- To elucidate the mechanism of dysfunctional mitochondria accumulation during erythroid maturation.
- To evaluate the impact of autophagy inhibition on erythroid cells with mtDNA mutations in vivo.
Main Methods:
- Analysis of erythroid maturation in aged mtDNA-mutator mice.
- Investigation of mechanistic target of rapamycin (mTOR) signaling and autophagy pathways.
- Conditional deletion of Atg7 in erythroid progenitors of wild-type and mtDNA-mutator mice.
Main Results:
- Aged mtDNA-mutator mice showed defective clearance of dysfunctional mitochondria during erythroid maturation.
- Aberrant mTOR signaling led to ULK1 degradation and autophagy inhibition in erythroid cells.
- Genetic disruption of autophagy accelerated mitochondrial dysfunction and anemia in mtDNA-mutator mice.
Conclusions:
- Dysfunctional mitochondria escape autophagy-mediated degradation in cells with somatic mtDNA mutations.
- A pathological feedback loop promotes the propagation of mutated mitochondria, contributing to MDS.
- Targeting autophagy may offer therapeutic strategies for MDS associated with mitochondrial dysfunction.
Related Concept Videos
 01:37
Mitochondria
 01:46
Electron Transport Chain: Complex I and II
ROS generation is regulated and maintained at moderate levels necessary...
 01:19
Translocation of Proteins into the Mitochondria
Sorting of outer membrane proteins:
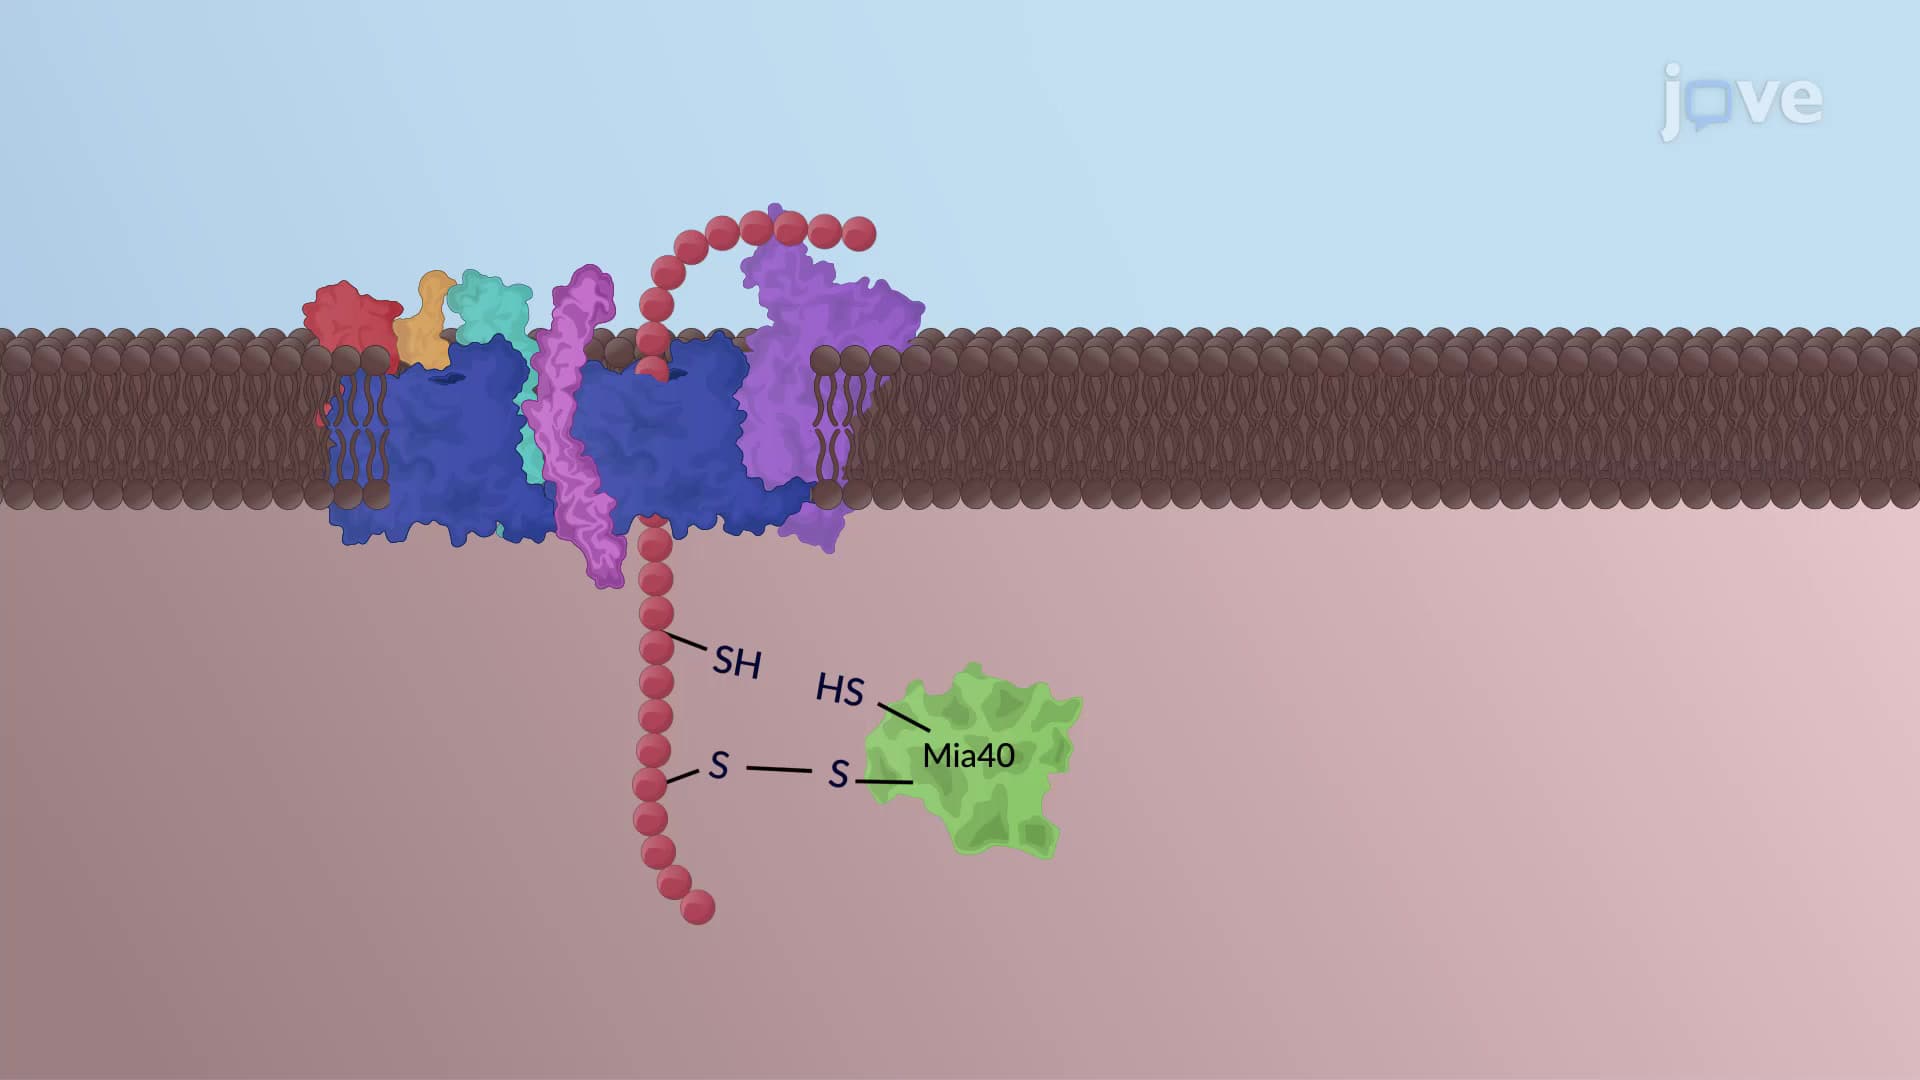
Mitochondrial outer membrane proteins are of two types: the transmembrane, beta-barrel porins, and the membrane-anchored, alpha-helical proteins. Beta-barrel porin precursors are translocated by the TOM complex and inserted into the outer mitochondrial membrane by the SAM complex. In contrast,...