Related Experiment Video
Updated: Apr 19, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Self-interaction corrected density functional calculations of Rydberg states of molecular clusters:
Hildur Gudmundsdóttir1, Yao Zhang2, Peter M Weber2
1Science Institute and Faculty of Physical Sciences VR-III, University of Iceland, 107 Reykjavík, Iceland.

Abstract:
Theoretical calculations of Rydberg excited states of molecular clusters consisting of N,N-dimethylisopropylamine molecules using a Perdew-Zunger self-interaction corrected energy functional are presented and compared with results of resonant multiphoton ionization measurements. The binding energy of the Rydberg electron in the monomer is calculated to be 2.79 eV and 2.27 eV in the 3s and 3p state, respectively, which compares well with measured values of 2.88 eV and 2.21 eV. Three different stable configurations of the dimer in the ground state were found using an energy functional that includes van der Waals interaction. The lowest ground state energy conformation has the two N-atoms widely separated, by 6.2 Å, while the Rydberg state energy is lowest for a configuration where the N-atoms of the two molecules come close together, separated by 3.7 Å. This conformational change is found to lower the Rydberg electron binding energy by 0.2 eV. The self-interaction corrected functional gives a highly localized hole on one of the two molecules, unlike results obtained using the PBE functional or the hybrid B3LYP functional which give a delocalized hole. For the trimer, the self-interaction corrected calculation gives a Rydberg electron binding energy lowered further by 0.13 eV as compared with the dimer. The calculated results compare well with trends observed in experimental measurements. The reduction of the Rydberg electron binding energy with cluster size can be ascribed to an effective delocalization of the positive charge of the hole by the induced and permanent dipole moments of the neighboring molecules. A further decrease observed to occur on a time scale of tens of ps can be ascribed to a structural rearrangement of the clusters in the Rydberg state where molecules rotate to orient their dipoles in response to the formation of the localized hole.

Related Concept Videos
 04:01
Real Gases: Effects of Intermolecular Forces and Molecular Volume Deriving Van der Waals Equation
 01:10
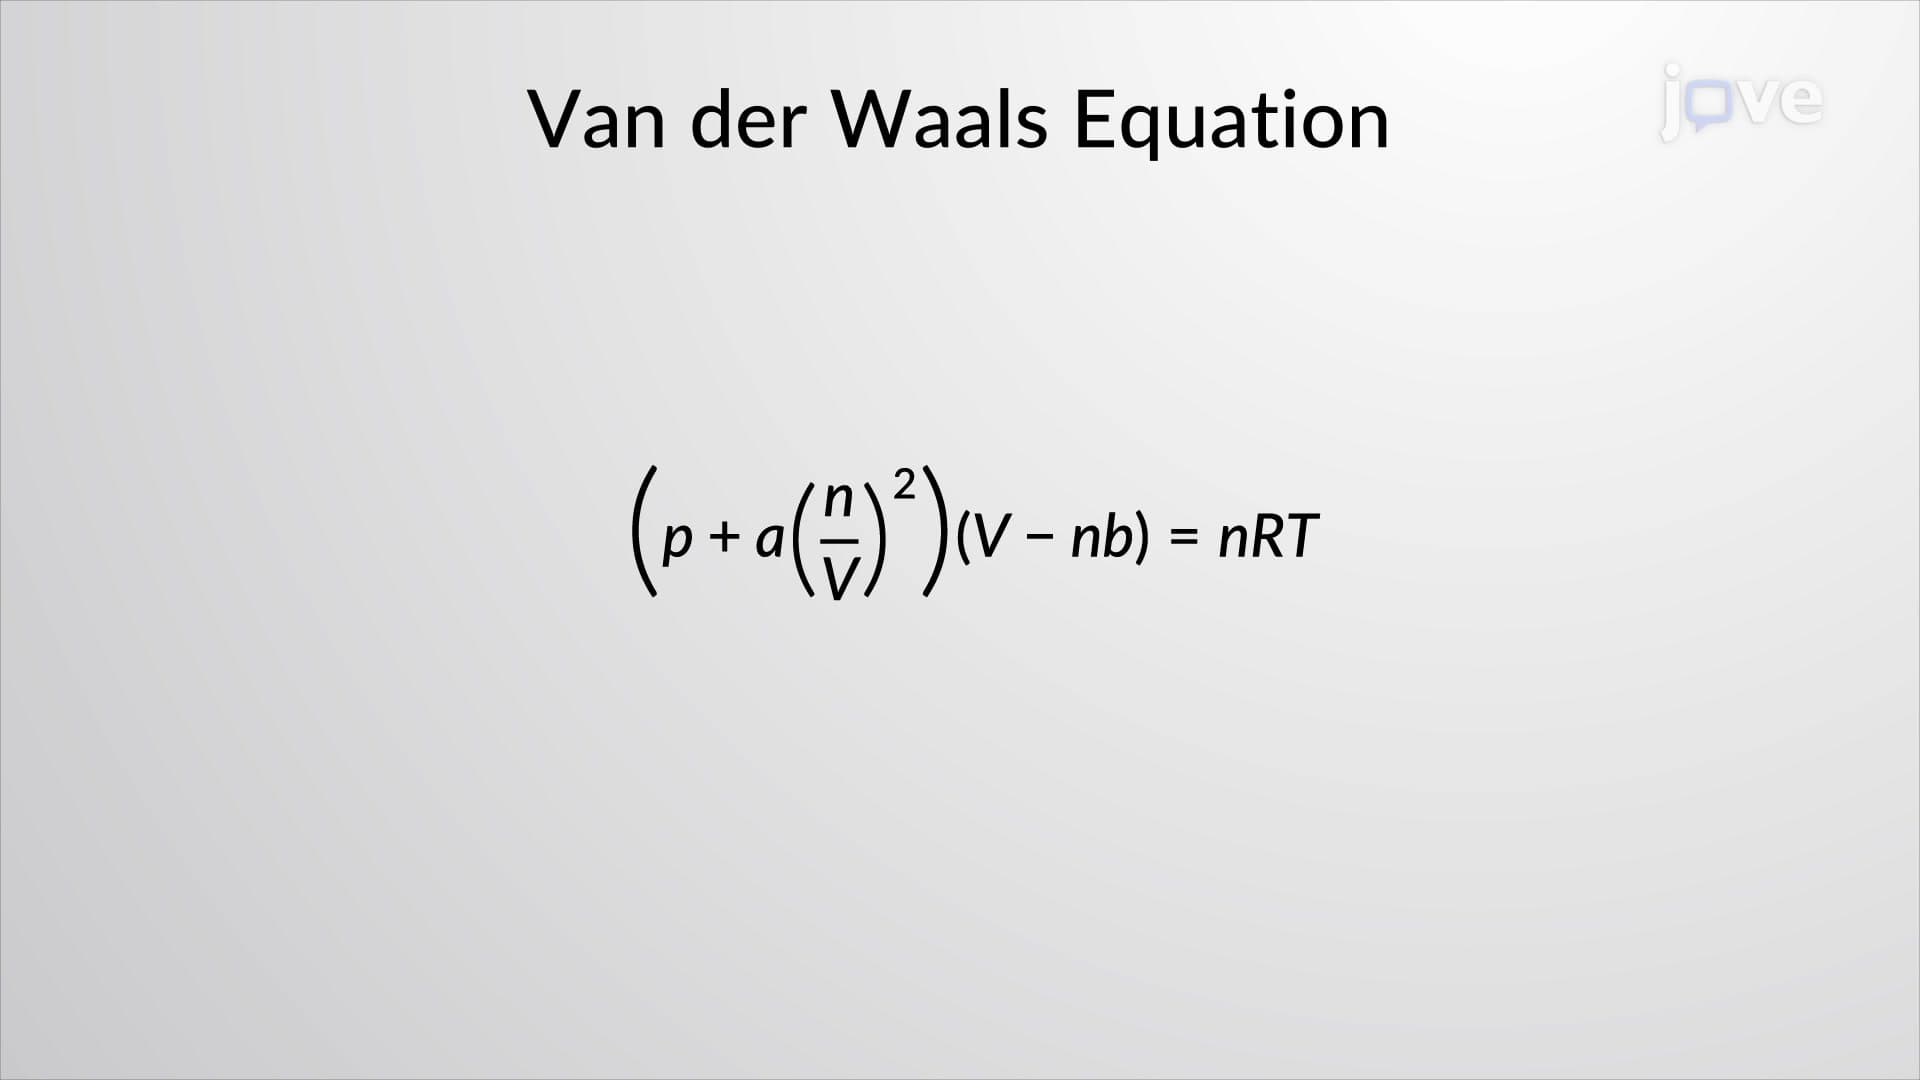
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the...
 01:24
Van der Waals Interactions
 01:26
The Van der Waals Equation
 03:51
Molecular Orbital Theory II
 02:35
Molecular Orbital Theory I