Related Experiment Video
Updated: Apr 19, 2026

X-Ray Crystallography to Study the Oligomeric State Transition of the Thermotoga maritima M42 Aminopeptidase TmPep1050
Published on: May 13, 2020
Prediction of molecular crystal structures by a crystallographic QM/MM model with full space-group symmetry
Philipp Mörschel1, Martin U Schmidt1
1Institut für Anorganische und Analytische Chemie, Goethe-Universität, Max-von-Laue-Strasse 7, D-60438 Frankfurt am Main, Germany.

Abstract:
A crystallographic quantum-mechanical/molecular-mechanical model (c-QM/MM model) with full space-group symmetry has been developed for molecular crystals. The lattice energy was calculated by quantum-mechanical methods for short-range interactions and force-field methods for long-range interactions. The quantum-mechanical calculations covered the interactions within the molecule and the interactions of a reference molecule with each of the surrounding 12-15 molecules. The interactions with all other molecules were treated by force-field methods. In each optimization step the energies in the QM and MM shells were calculated separately as single-point energies; after adding both energy contributions, the crystal structure (including the lattice parameters) was optimized accordingly. The space-group symmetry was maintained throughout. Crystal structures with more than one molecule per asymmetric unit, e.g. structures with Z' = 2, hydrates and solvates, have been optimized as well. Test calculations with different quantum-mechanical methods on nine small organic molecules revealed that the density functional theory methods with dispersion correction using the B97-D functional with 6-31G* basis set in combination with the DREIDING force field reproduced the experimental crystal structures with good accuracy. Subsequently the c-QM/MM method was applied to nine compounds from the CCDC blind tests resulting in good energy rankings and excellent geometric accuracies.
More Related Videos


Related Concept Videos
 02:27
Predicting Molecular Geometry
 01:29
Determination of Crystal Structures
 01:29
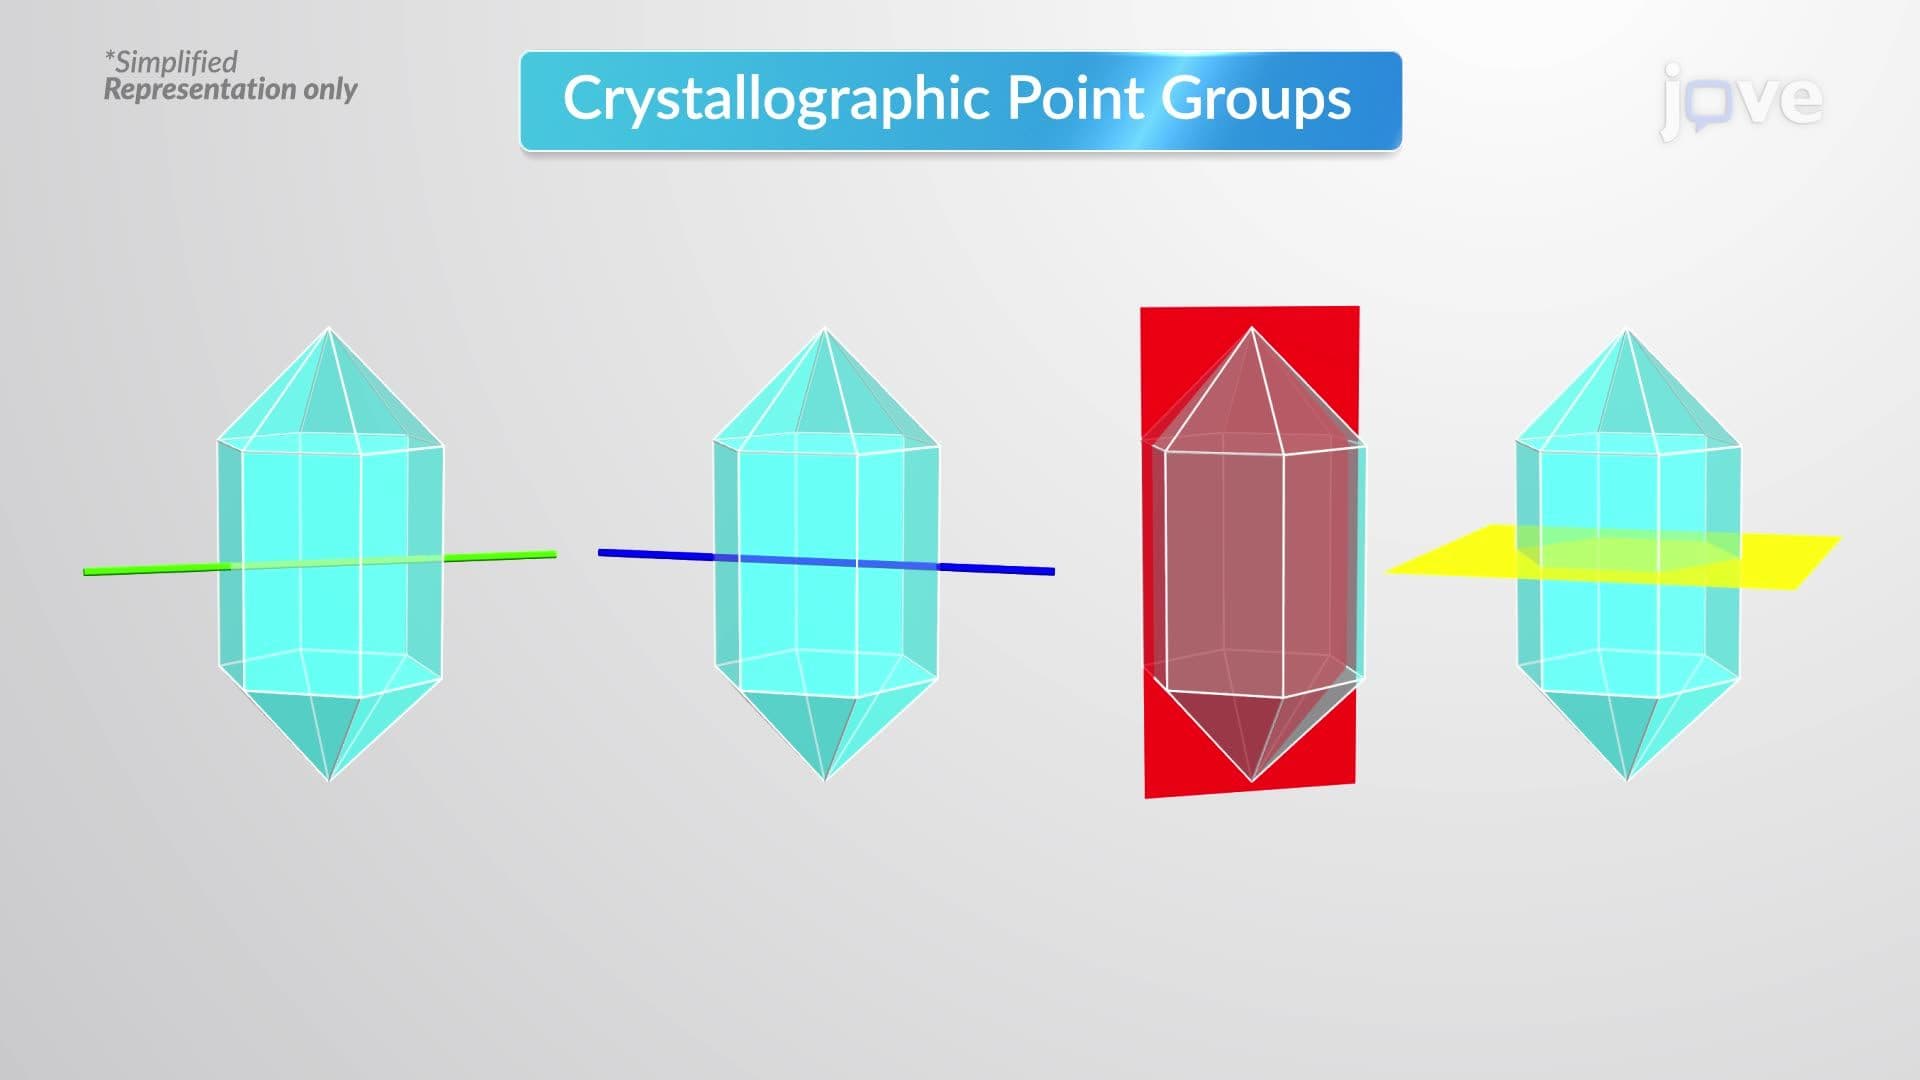
Crystallographic Point Groups
 02:00
Molecular Models
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...