Related Experiment Video
Updated: Apr 19, 2026

Validating Whole Genome Nanopore Sequencing, using Usutu Virus as an Example
Published on: March 11, 2020
Complete genome of a Puumala virus strain from Central Europe
Hanan Sheikh Ali1, Stephan Drewes, Vanessa Weber de Melo
1Friedrich-Loeffler-Institut, Federal Research Institute for Animal Health, OIE Collaborating Centre for Zoonoses in Europe, Institute for Novel and Emerging Infectious Diseases, Südufer 10, 17493, Greifswald - Insel Riems, Germany.

Abstract:
Puumala virus (PUUV) is one of the predominant hantavirus species in Europe causing mild to moderate cases of haemorrhagic fever with renal syndrome. Parts of Lower Saxony in north-western Germany are endemic for PUUV infections. In this study, the complete PUUV genome sequence of a bank vole-derived tissue sample from the 2007 outbreak was determined by a combined primer-walking and RNA ligation strategy. The S, M and L genome segments were 1,828, 3,680 and 6,550 nucleotides in length, respectively. Sliding-window analyses of the nucleotide sequences of all available complete PUUV genomes indicated a non-homogenous distribution of variability with hypervariable regions located at the 3'-ends of the S and M segments. The overall similarity of the coding genome regions to the other PUUV strains ranged between 80.1 and 84.7 % at the level of the nucleotide sequence and between 89.5 and 98.1 % for the deduced amino acid sequences. In comparison to the phylogenetic trees of the complete coding sequences, trees based on partial segments revealed a general drop in phylogenetic support and a lower resolution. The Astrup strain S and M segment sequences showed the highest similarity to sequences of strains from geographically close sites in the Osnabrück Hills region. In conclusion, a primer-walking-mediated strategy resulted in the determination of the first complete nucleotide sequence of a PUUV strain from Central Europe. Different levels of variability along the genome provide the opportunity to choose regions for analyses according to the particular research question, e.g., large-scale phylogenetics or within-host evolution.
More Related Videos
 09:50
Single Cell Micro-aspiration as an Alternative Strategy to Fluorescence-activated Cell Sorting for Giant Virus Mixture Separation
Published on: October 27, 2019
 08:56
Combining Analysis of DNA in a Crude Virion Extraction with the Analysis of RNA from Infected Leaves to Discover New Virus Genomes
Published on: July 27, 2018
Related Concept Videos
 01:29
Viruses of Archaea
 01:29
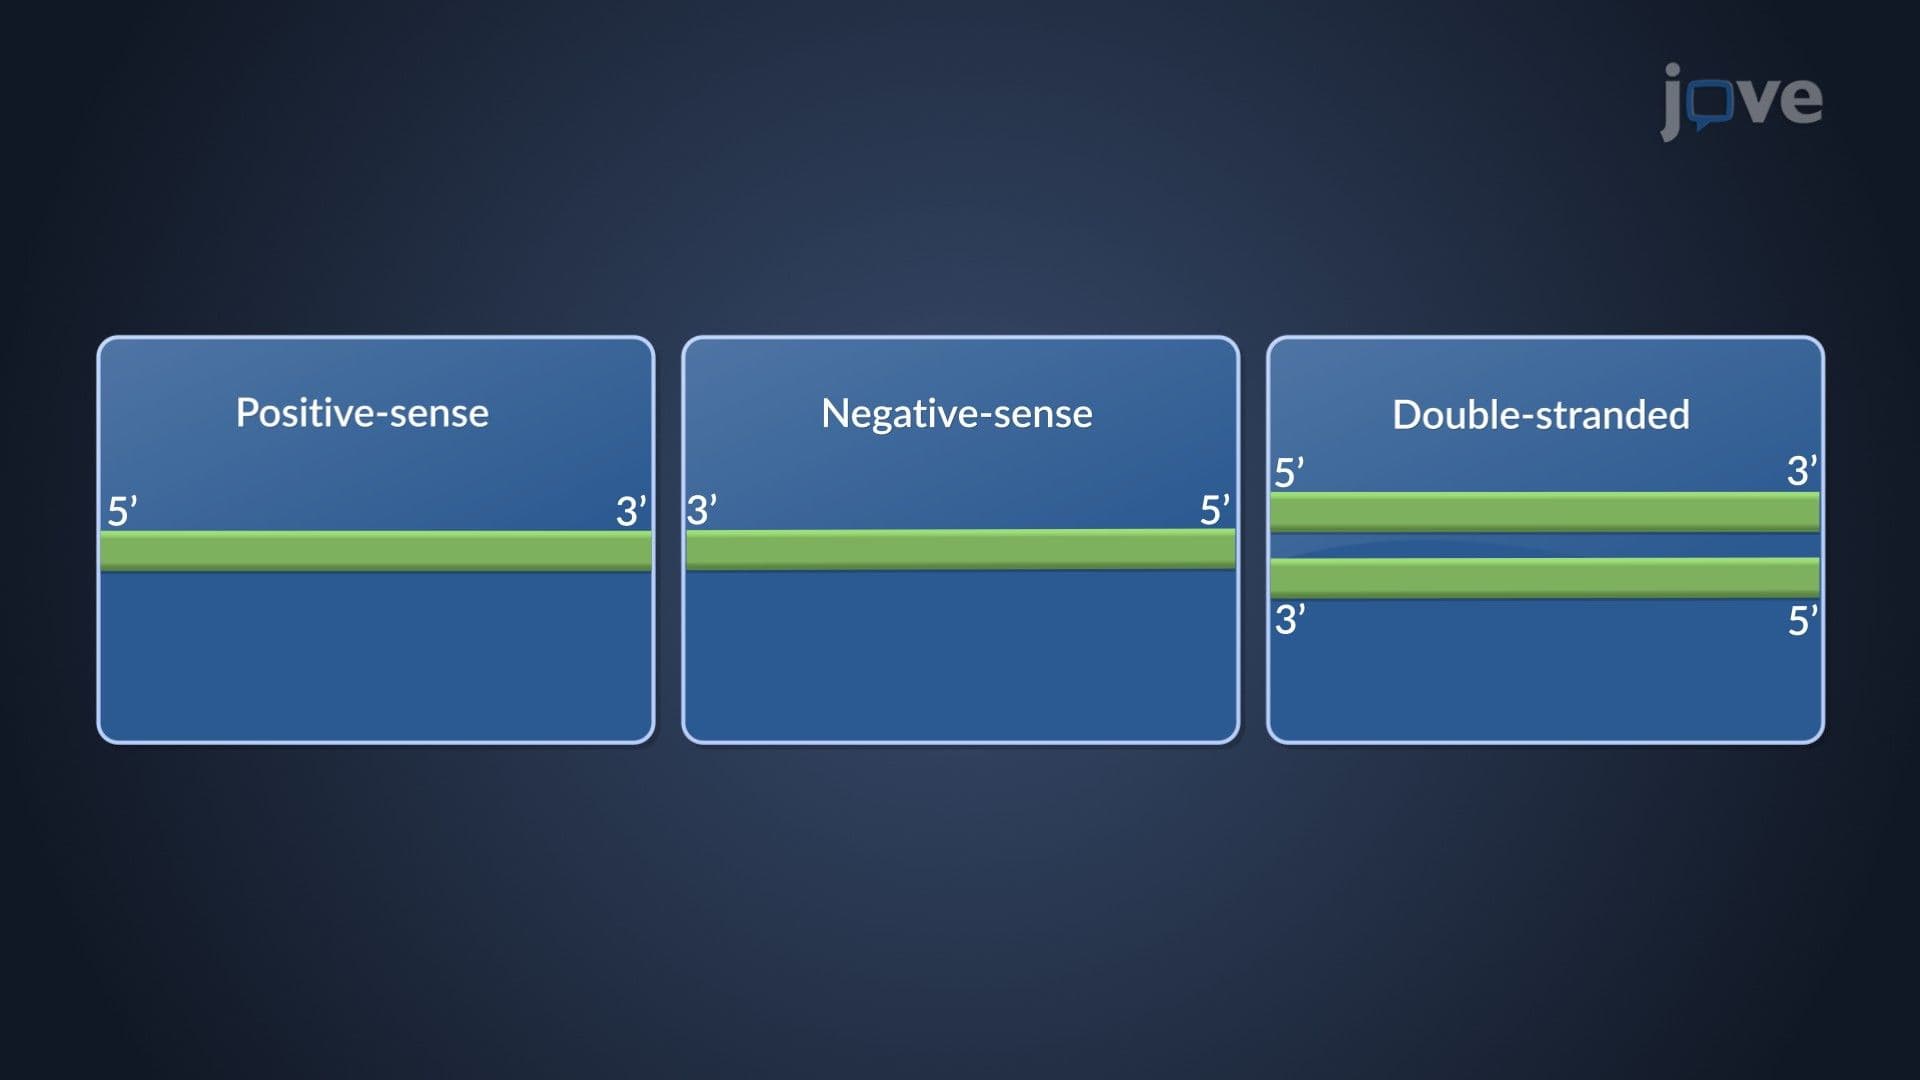
Viruses with RNA Genomes
 01:26
Human Virome