DFT electronic structure calculations, spectroscopic studies, and normal coordinate analysis of
S Muthu1, E Elamuruguporchelvi2, Anitha Varghese3
1Department of Physics, Sri Venkateswara College of Engg, Sriperumbudur 602105, Tamil Nadu, India.

Abstract:
The solid phase FTIR and FT-Raman spectra of 2-[(5-nitro-1,3-thiazol-2-yl)carbamoyl]phenyl acetate (25N2LCPA) have been recorded 450-4000cm(-1) and 100-4000cm(-1) respectively. The normal coordinate analysis was carried out to confirm the precision of the assignments. DFT calculations have been performed giving energies, optimized structures, harmonic vibrational frequencies and IR intensities. The structure of the molecule was optimized and the structural characteristics were determined by density functional theory (DFT) using B3LYP method with 6-31+G(d,p) basis set. The detailed interpretation of the vibrational spectra has been carried out with aid of normal coordinate analysis (NCA) following the scaled quantum mechanical force field methodology. The Vibrational frequencies are calculated in the above method and are compared with experimental frequencies which yield good agreement between observed and calculated frequencies. Stability of the molecule arising from hyper conjugative interactions, charge delocalization has been analyzed using natural bond orbital (NBO) analysis. In addition, Frontiers molecular orbital and molecular electrostatic potential were computed by using Density Functional Theory (DFT) B3LYP/6-31+G(d,p) basis set. The calculated HOMO and LUMO energies show that charge transfer occurs in the molecule.
More Related Videos
 10:42
Preparation of N-2-alkoxyvinylsulfonamides from N-tosyl-1,2,3-triazoles and Subsequent Conversion to Substituted Phthalans and Phenethylamines
Published on: January 3, 2018
 06:46
Facile Preparation of 2Z,4E-Dienamides by the Olefination of Electron-deficient Alkenes with Allyl Acetate
Published on: June 21, 2017
Related Concept Videos
 01:26
Spectroscopy of Carboxylic Acid Derivatives
 01:20
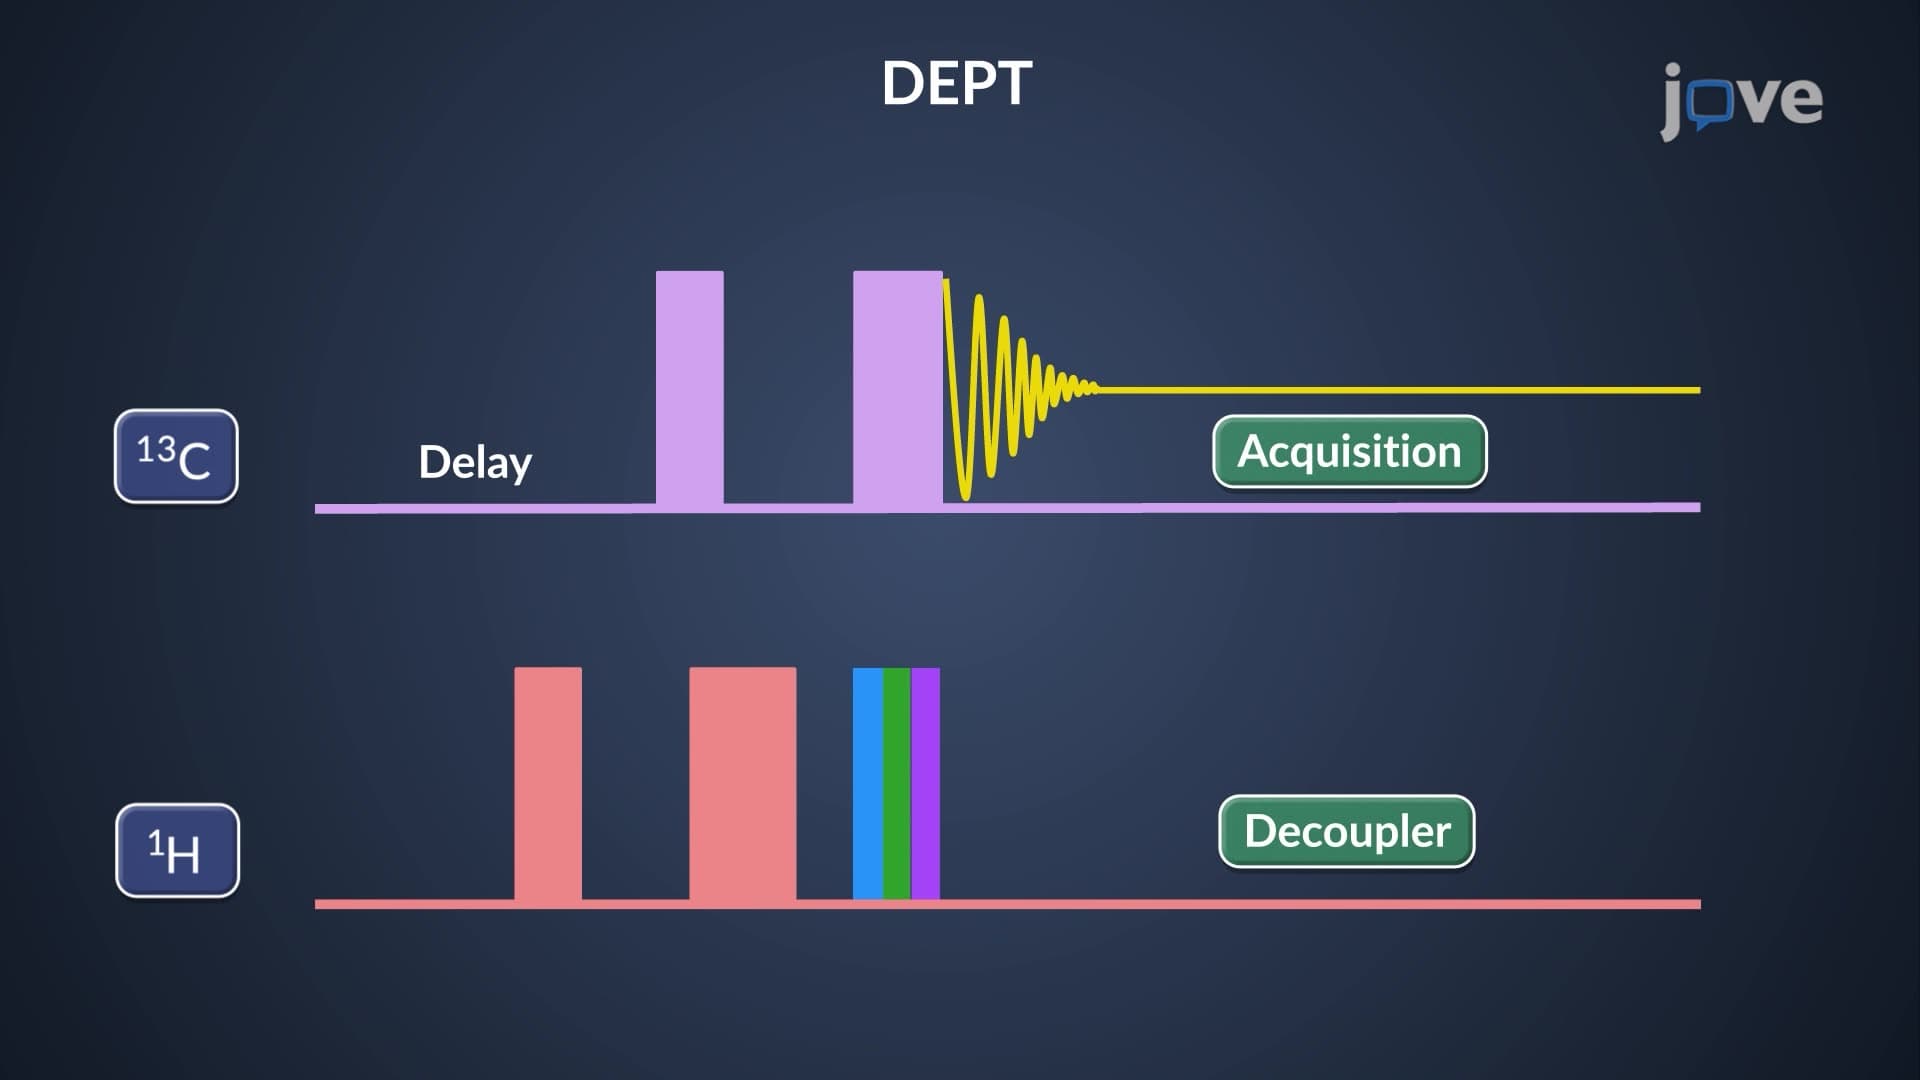
¹³C NMR: Distortionless Enhancement by Polarization Transfer (DEPT)
 01:29
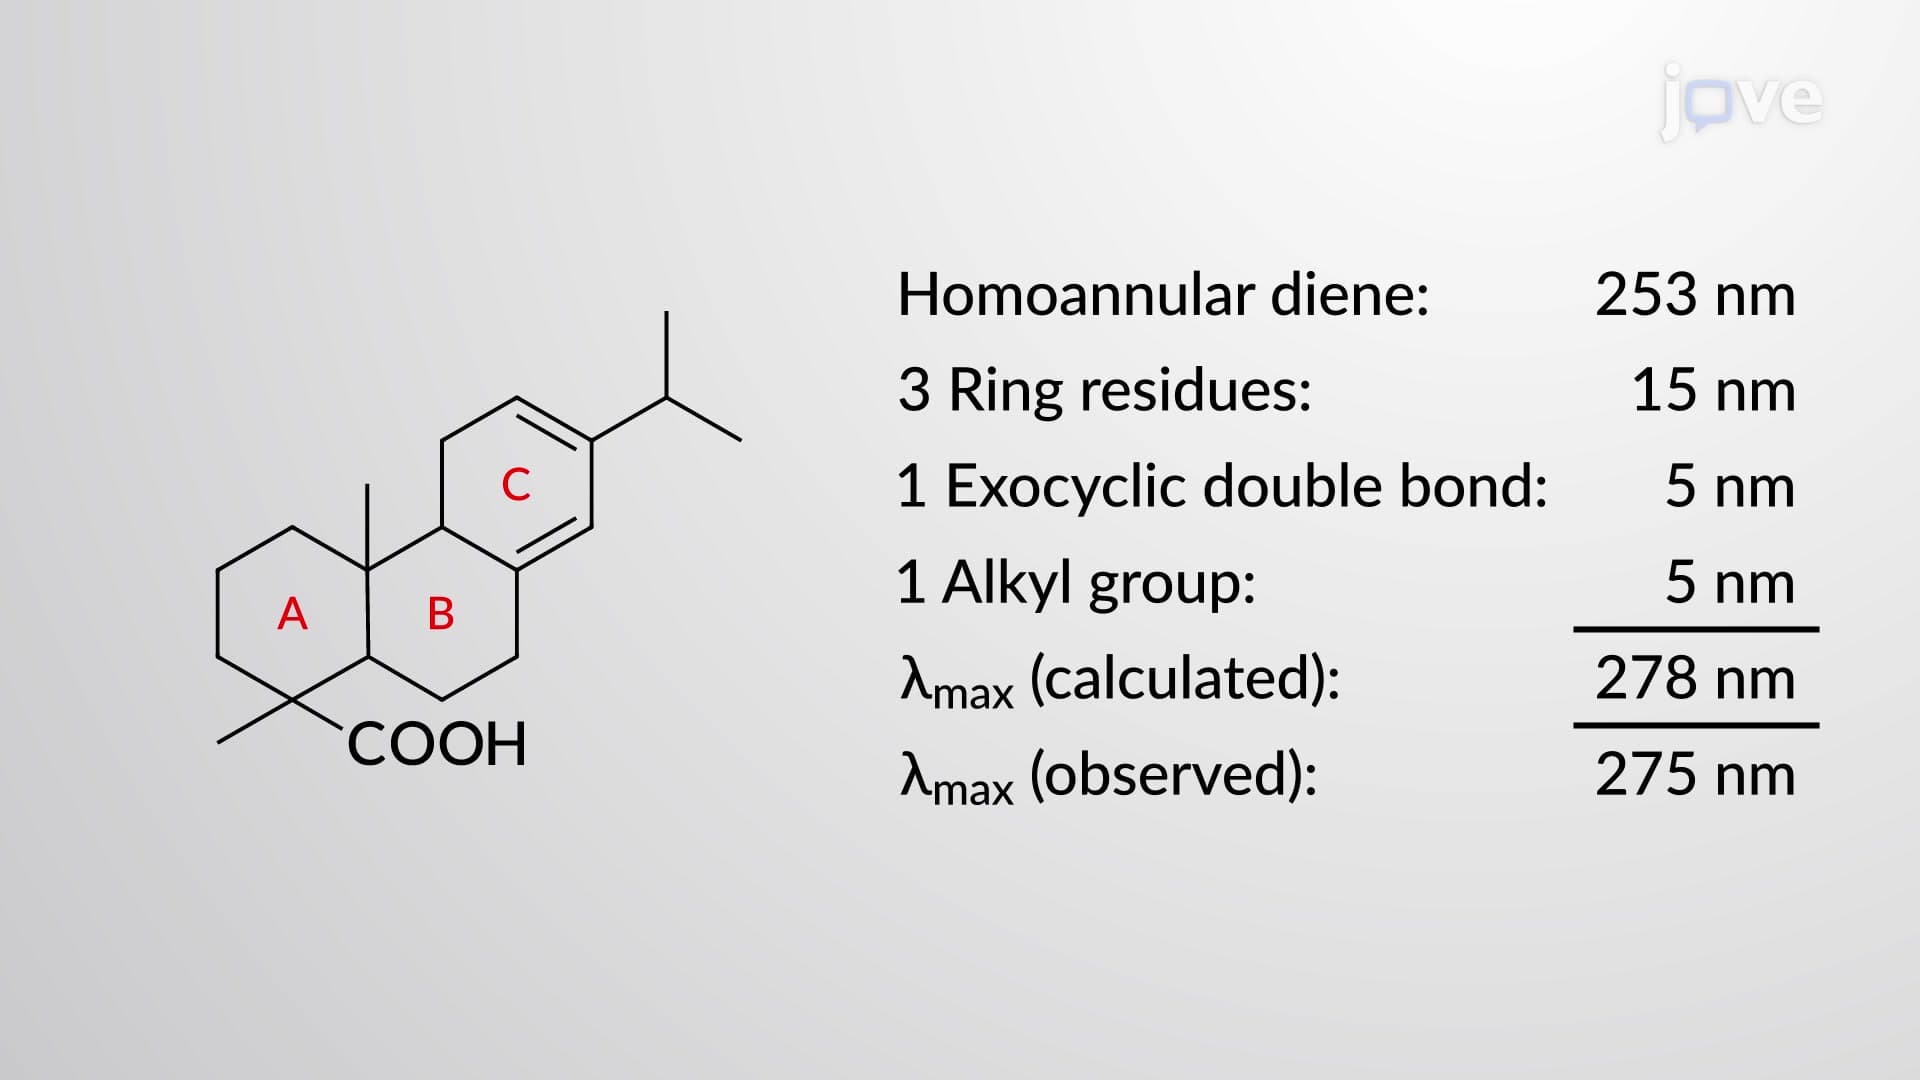
UV–Vis Spectroscopy: Woodward–Fieser Rules
 01:37
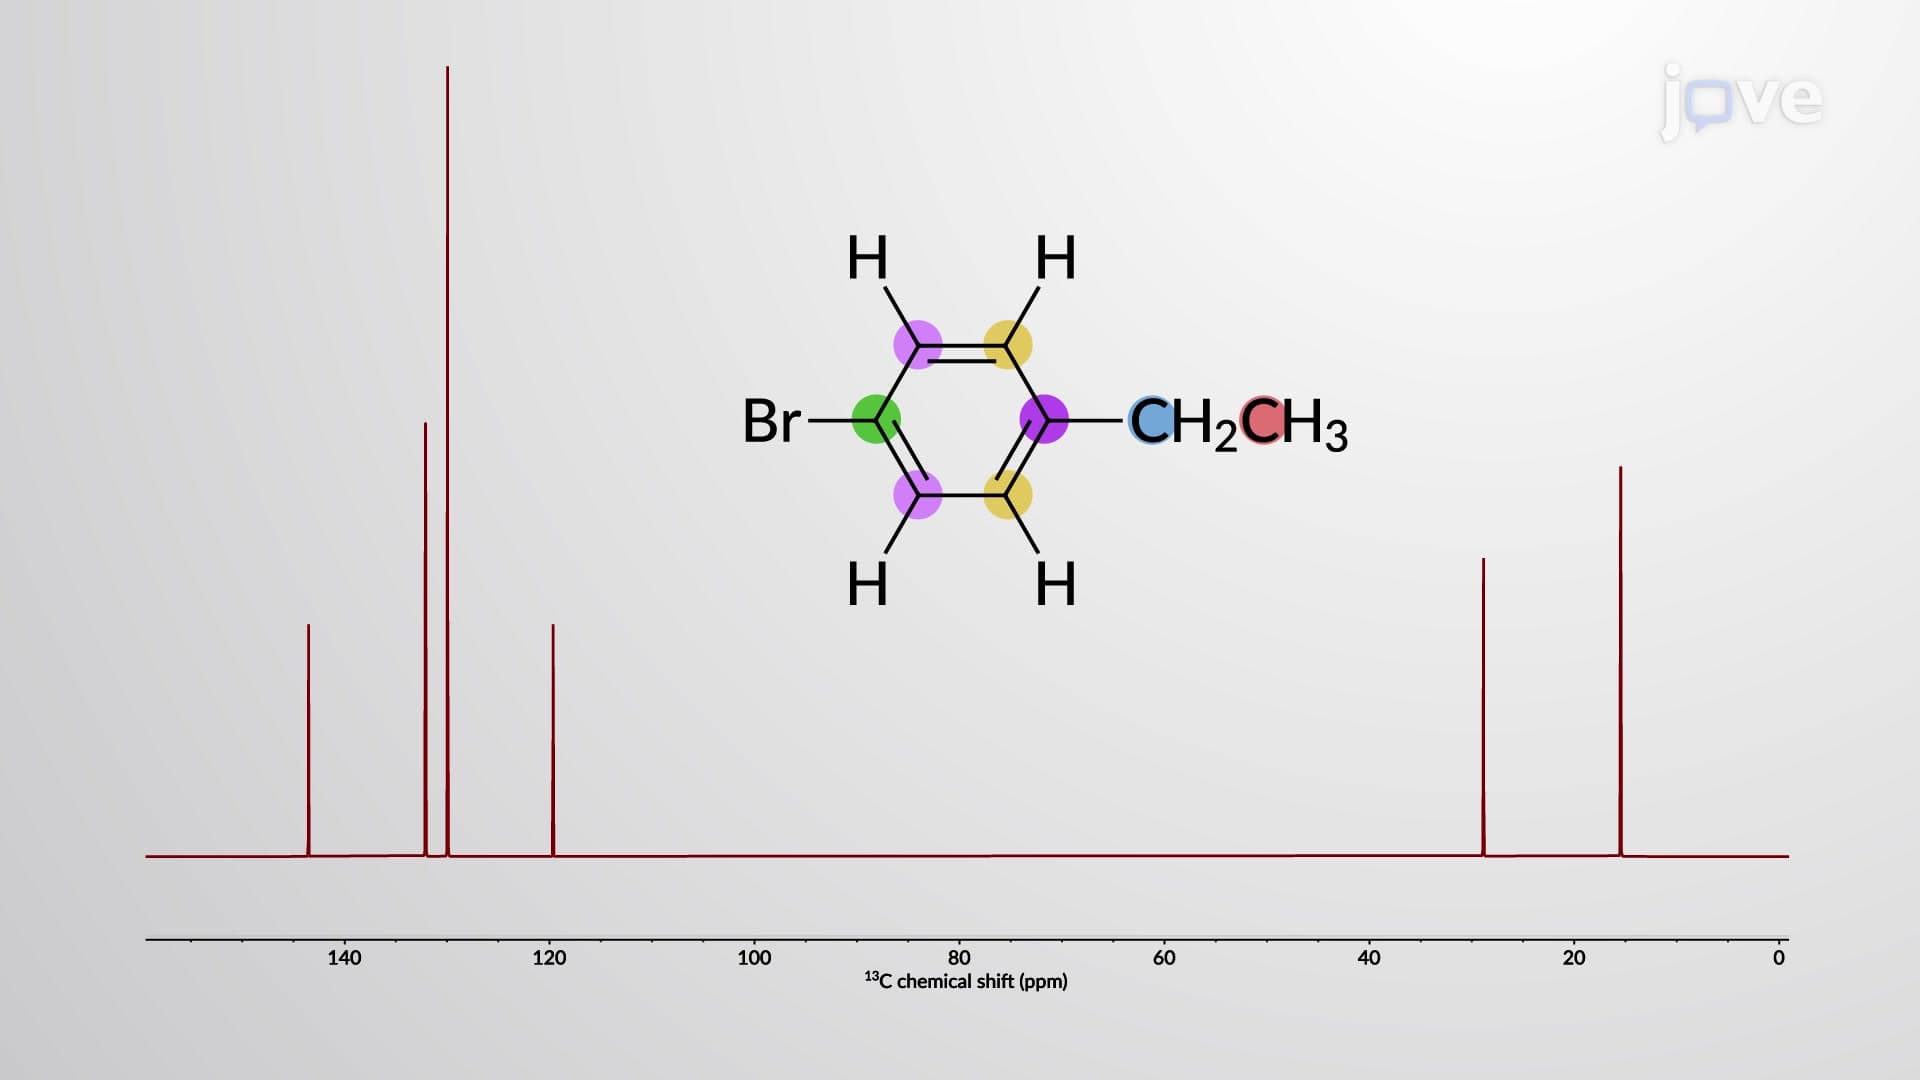
NMR Spectroscopy of Benzene Derivatives
 00:58
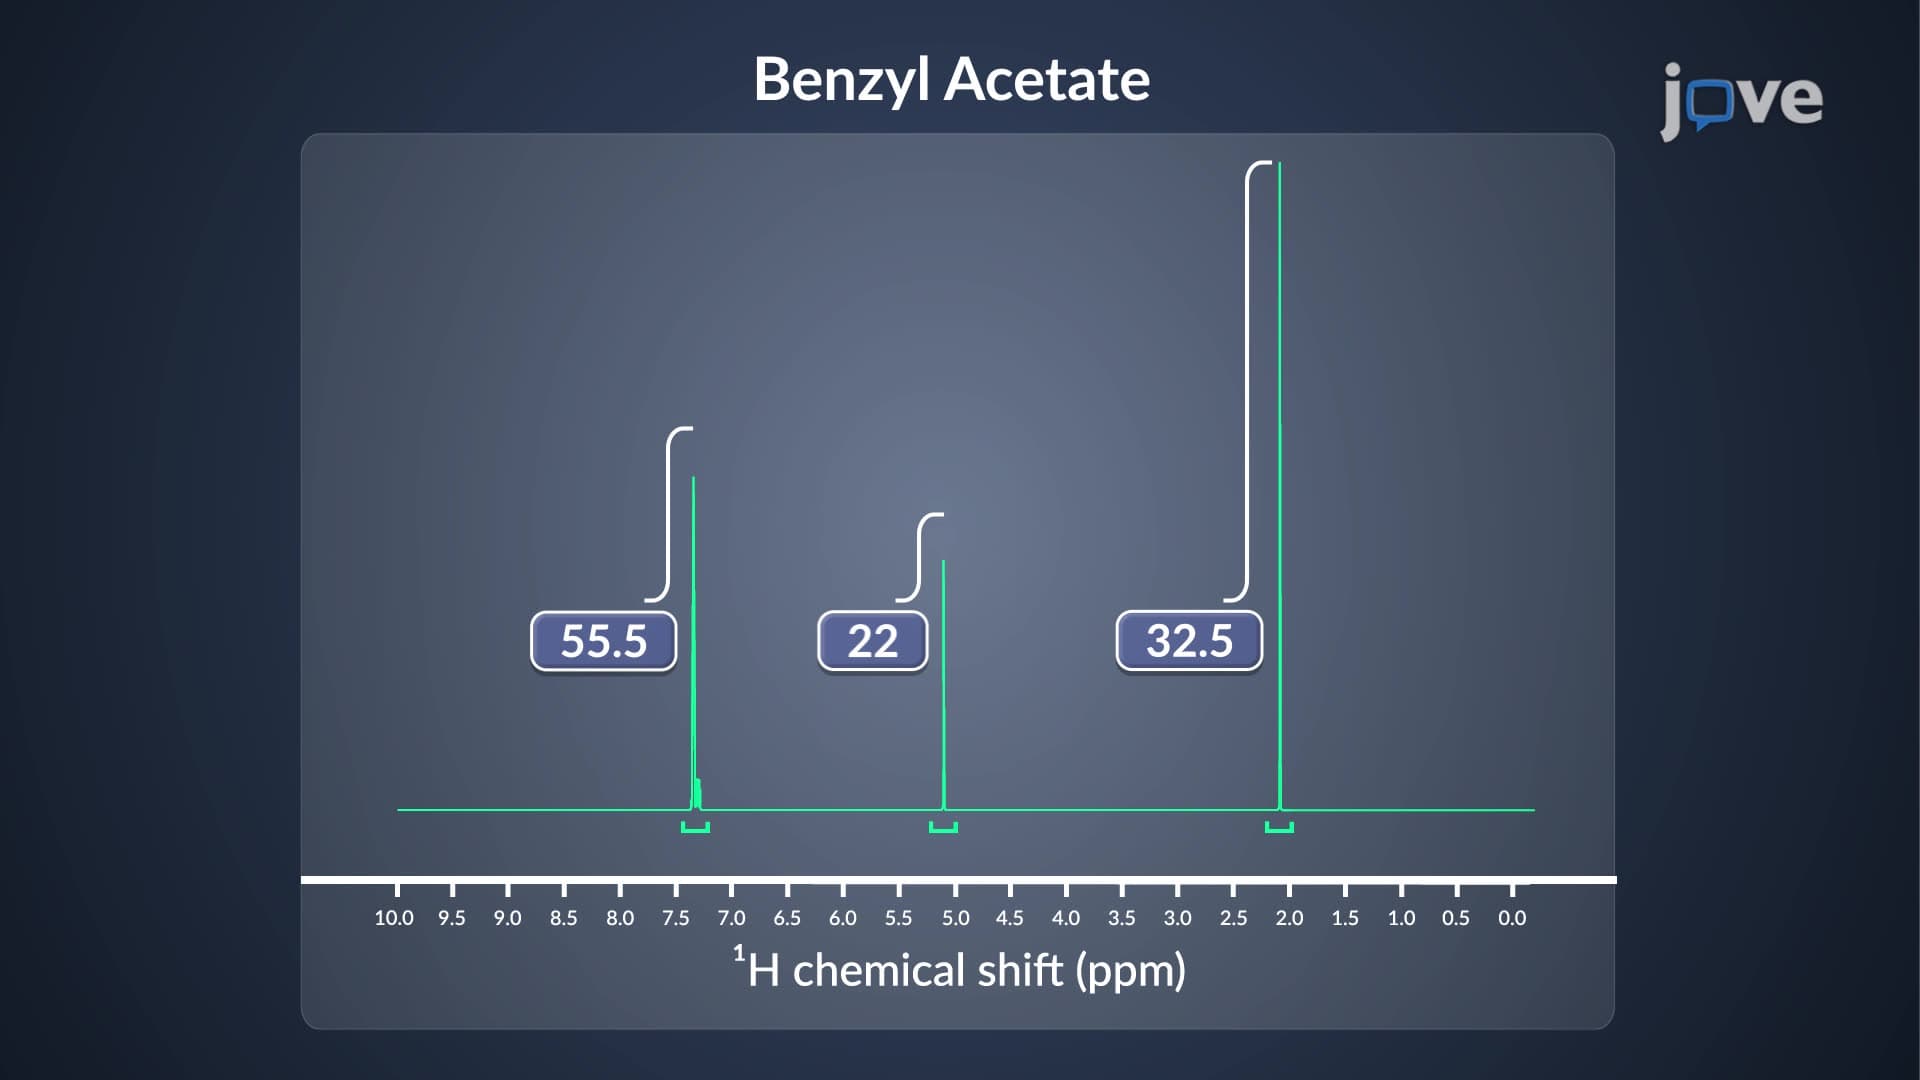
¹H NMR Signal Integration: Overview
 01:14
π Electron Effects on Chemical Shift: Aromatic and Antiaromatic Compounds