Related Experiment Video
Updated: Apr 19, 2026


A Protocol for Computer-Based Protein Structure and Function Prediction
Published on: November 3, 2011
Tri-peptide reference structures for the calculation of relative solvent accessible surface area in protein amino
Christopher M Topham1, Jeremy C Smith2
1Molecular Forces Consulting, 40 Rue Boyssonne, Toulouse 31400, France; Computational Molecular Biophysics, IWR der Universität Heidelberg, Im Neuenheimer Feld 368, Heidelberg D-69120, Germany; University of Tennessee/Oak Ridge National Laboratory, Center for Molecular Biophysics, P.O. Box 2008, Oak Ridge, TN 37831-6309, USA; Department of Biochemistry and Cellular and Molecular Biology, University of Tennessee, M407 Walters Life Sciences, 1414 Cumberland Avenue, Knoxville, TN 37996, USA.

Abstract:
Relative amino acid residue solvent accessibility values allow the quantitative comparison of atomic solvent-accessible surface areas in different residue types and physical environments in proteins and in protein structural alignments. Geometry-optimised tri-peptide structures in extended solvent-exposed reference conformations have been obtained for 43 amino acid residue types at a high level of quantum chemical theory. Significant increases in side-chain solvent accessibility, offset by reductions in main-chain atom solvent exposure, were observed for standard residue types in partially geometry-optimised structures when compared to non-minimised models built from identical sets of proper dihedral angles abstracted from the literature. Optimisation of proper dihedral angles led most notably to marked increases of up to 54% in proline main-chain atom solvent accessibility compared to literature values. Similar effects were observed for fully-optimised tri-peptides in implicit solvent. The relief of internal strain energy was associated with systematic variation in N, C(α) and C(β) atom solvent accessibility across all standard residue types. The results underline the importance of optimisation of 'hard' degrees of freedom (bond lengths and valence bond angles) and improper dihedral angle values from force field or other context-independent reference values, and impact on the use of standardised fixed internal co-ordinate geometry in sampling approaches to the determination of absolute values of protein amino acid residue solvent accessibility. Quantum chemical methods provide a useful and accurate alternative to molecular mechanics methods to perform energy minimisation of peptides containing non-standard (chemically modified) amino acid residues frequently present in experimental protein structure data sets, for which force field parameters may not be available. Reference tri-peptide atomic co-ordinate sets including hydrogen atoms are made freely available.

Related Concept Videos
 01:24
Protein Organization
The primary structure of a protein is its amino acid sequence....
 01:13
Protein Organization
 01:22
Protein Folding
 01:25
Protein Folding
Protein Structure Is Critical to Its Biological Function
Proteins perform a wide range of biological functions such as catalyzing chemical reactions, providing...
 02:15
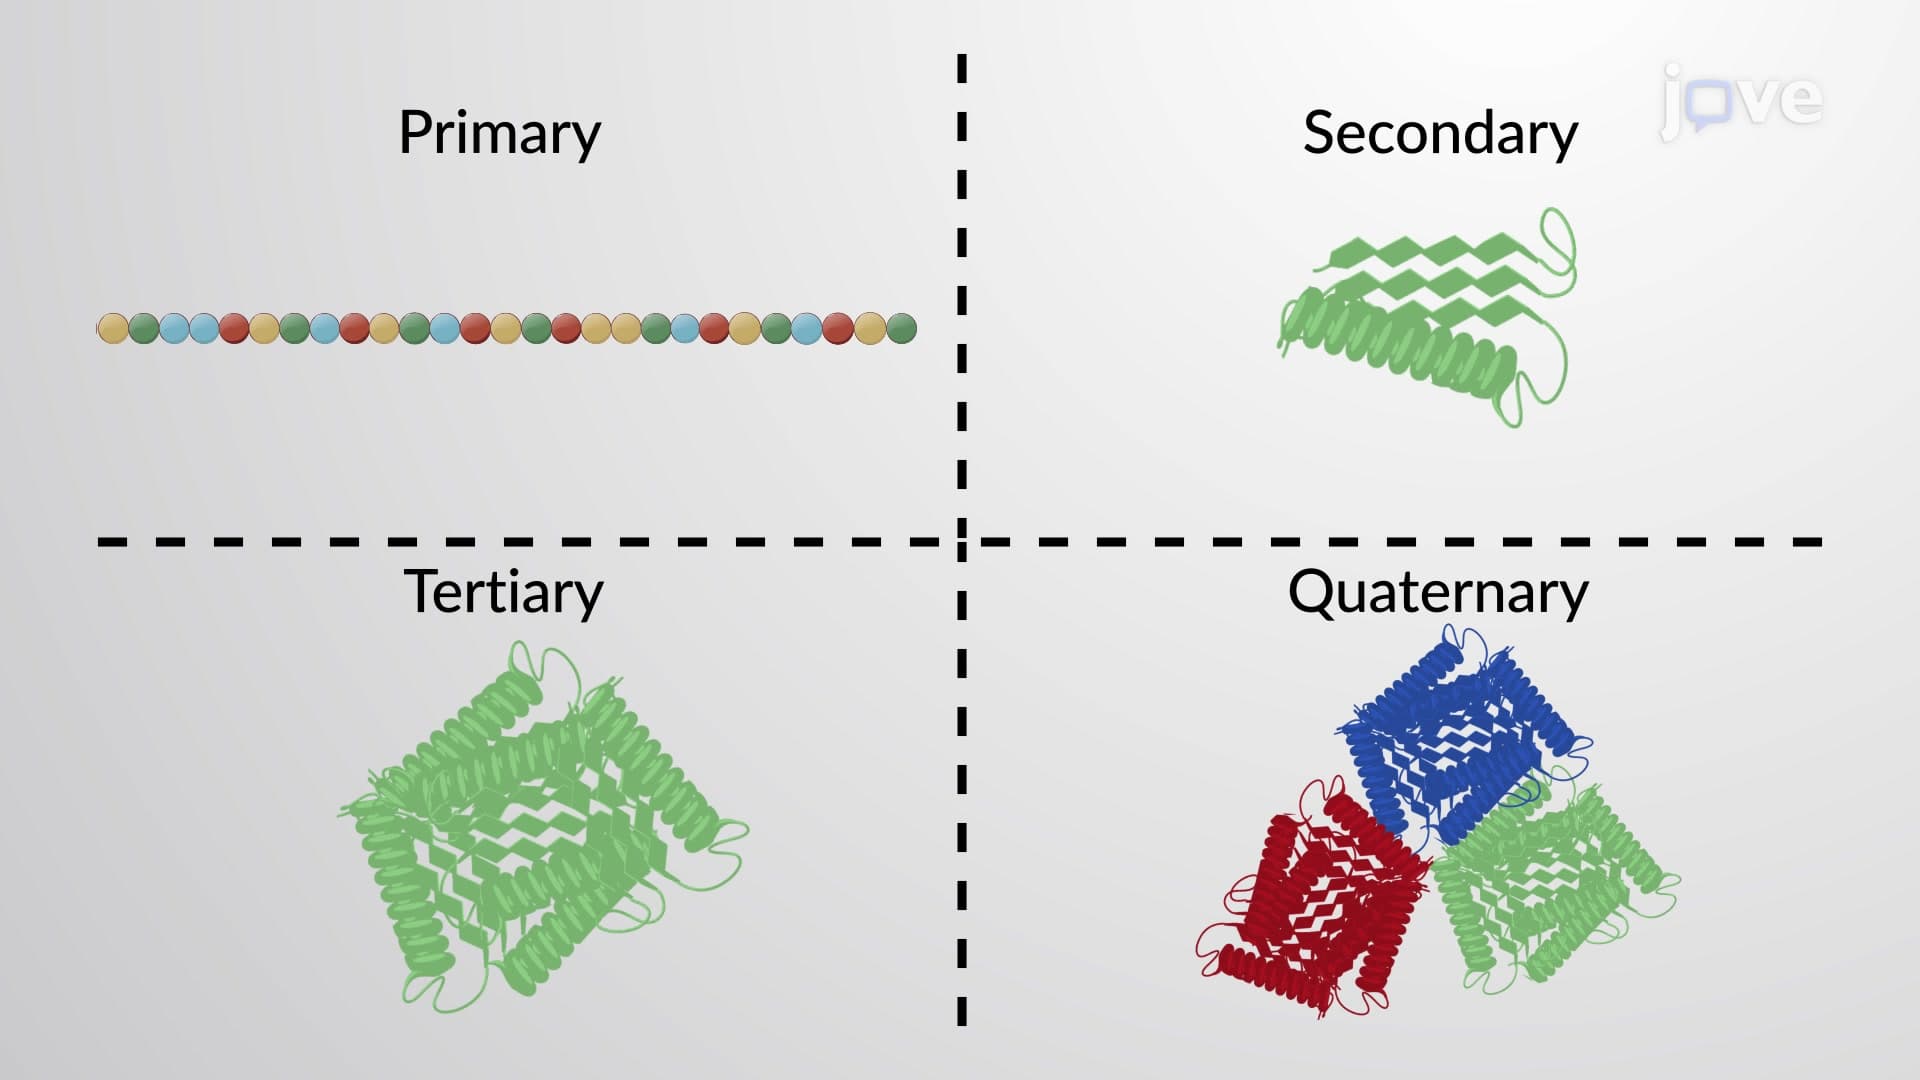
Protein and Protein Structure
A protein's shape is critical to its function. For example, an enzyme...
 03:42
Amino acids