Related Experiment Video
Updated: Apr 18, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Electronic couplings for molecular charge transfer: benchmarking CDFT, FODFT and FODFTB against high-level ab initio
Adam Kubas1, Fruzsina Gajdos, Alexander Heck
1University College London, Department of Physics and Astronomy, Gower Street, London WC1E 6BT, UK. j.blumberger@ucl.ac.uk.

Abstract:
A new database (HAB7-) of electronic coupling matrix elements (Hab) for electron transfer in seven medium-sized negatively charged π-conjugated organic dimers is introduced. Reference data are obtained with spin-component scaled approximate coupled cluster method (SCS-CC2) and large basis sets. Assessed DFT-based approaches include constrained density functional theory (CDFT), fragment-orbital DFT (FODFT), self-consistent charge density functional tight-binding (FODFTB) and the recently described analytic overlap method (AOM). This complements the previously reported HAB11 database where only cationic dimers were considered. The CDFT method in combination with a functional based on PBE and including 50% of exact exchange (HFX) was found to provide best estimates, with a mean relative unsigned error (MRUE) of 8.2%. CDFT couplings systematically increase with decreasing fraction of HFX as a consequence of increasing delocalisation of the SOMO orbital. The FODFT method is found to be very robust underestimating electronic couplings by 28%. The FODFTB and AOM methods, although orders of magnitude more efficient in terms of computational effort than the DFT approaches, perform well with reasonably small errors of 54% and 29%, respectively, translating in errors in the non-adiabatic electron transfer rate of a factor of 2.4 and 1.7, respectively. We discuss carefully various sources of errors and the scope and limitations of all assessed methods taking into account the results obtained for both HAB7- and HAB11 databases.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Related Concept Videos
 01:27
π Electron Effects on Chemical Shift: Overview
 02:28
Electronic Structure of Atoms
An atom comprises protons and neutrons, which are contained inside the dense, central core called the nucleus, with electrons present around the nucleus. Taking into account the wave–particle duality of electrons and the uncertainty in position around the nucleus, quantum mechanics provides a more accurate model for the atomic structure. It describes atomic orbitals as the regions around the nucleus where electrons of discrete energy exist, characterized by four quantum...
 01:22
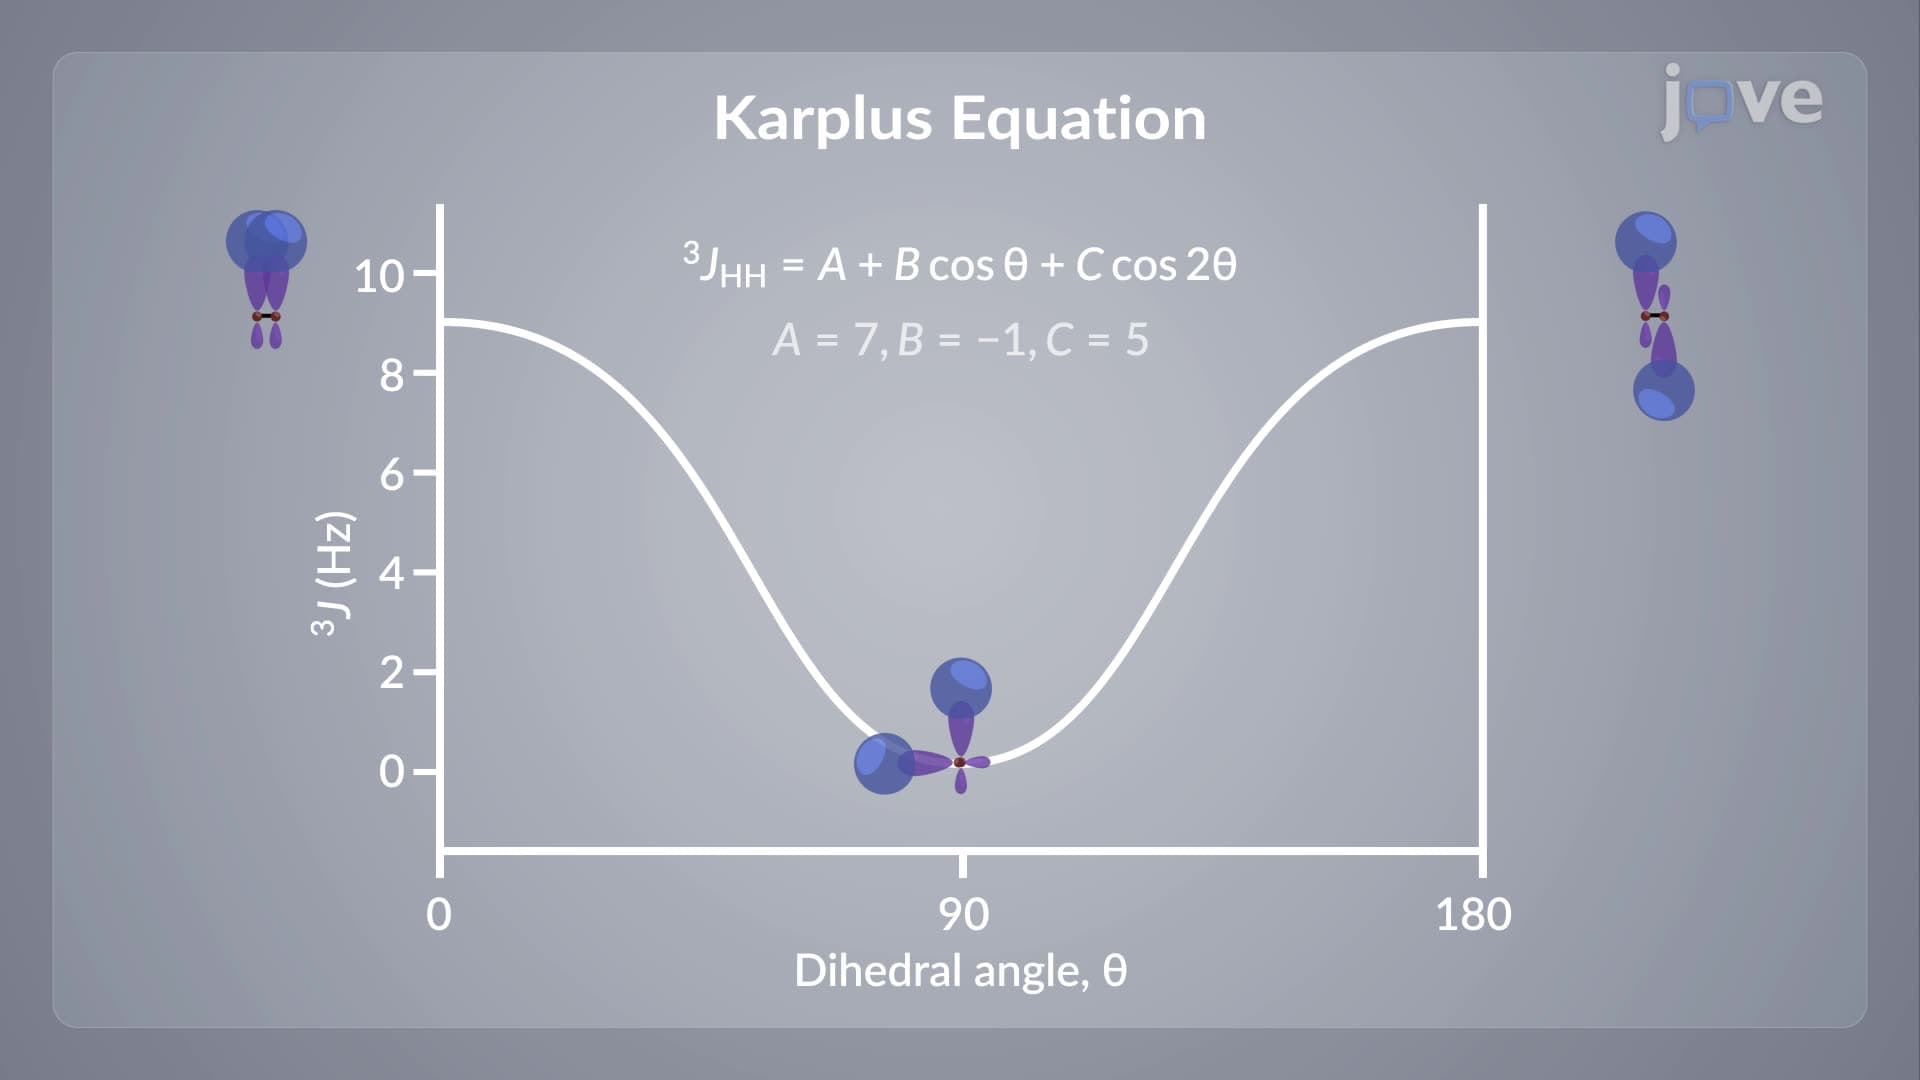
Spin–Spin Coupling: Three-Bond Coupling (Vicinal Coupling)
The extent of coupling depends on the C‑C bond length, the two H‑C‑C angles, any electron-withdrawing substituents, and the dihedral angle between the...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:06
Interfacial Electrochemical Methods: Overview
 01:17
Chemical Shift: Internal References and Solvent Effects
The internal reference compound generally used in NMR spectroscopy is tetramethylsilane (TMS). TMS is preferred because it is chemically inert, soluble in NMR solvents, and easily removable. Also, the highly shielded methyl protons in TMS yield an intense...