Related Experiment Video
Updated: Apr 18, 2026

Evaluation of Integrated Anaerobic Digestion and Hydrothermal Carbonization for Bioenergy Production
Published on: June 15, 2014
Solvation dynamics and energetics of intramolecular hydride transfer reactions in biomass conversion
Samir H Mushrif1, Jithin J Varghese, Chethana B Krishnamurthy
1School of Chemical and Biomedical Engineering, Nanyang Technological University, 62 Nanyang Drive, Singapore 637459. SHMushrif@ntu.edu.sg.

Abstract:
Hydride transfer changes the charge structure of the reactant and thus, may induce reorientation/reorganization of solvent molecules. This solvent reorganization may in turn alter the energetics of the reaction. In the present work, we investigate the intramolecular hydride transfer by taking Lewis acid catalyzed glucose to fructose isomerization as an example. The C2-C1 hydride transfer is the rate limiting step in this reaction. Water and methanol are used as solvents and hydride transfer is simulated in the presence of explicit solvent molecules, treated quantum mechanically and at a finite temperature, using Car-Parrinello molecular dynamics (CPMD) and metadynamics. Activation free energy barrier for hydride transfer in methanol is found to be 50 kJ mol(-1) higher than that in water. In contrast, in density functional theory calculations, using an implicit solvent environment, the barriers are almost identical. Analysis of solvent dynamics and electronic polarization along the molecular dynamics trajectory and the results of CPMD-metadynamics simulation of the hydride transfer process in the absence of any solvent suggest that higher barrier in methanol is a result of non-equilibrium solvation. Methanol undergoes electronic polarization during the hydride transfer step. However, its molecular orientational relaxation is a much slower process that takes place after the hydride transfer, over an extended timescale. This results in non-equilibrium solvation. Water, on the other hand, does not undergo significant electronic polarization and thus, has to undergo minimal molecular reorientation to provide near equilibrium solvation to the transition state and an improved equilibrium solvation to the post hydride shift product state. Hence, the hydride transfer step is also observed to be exergonic in water and endergonic in methanol. The aforementioned explanation is juxtaposed to enzyme catalyzed charge transfer reactions, where the enhanced solvation of the transition and product states by enzymes, due to electrostatic interactions, reduces the activation free energy barrier and the free energy change of the reaction. Similarly, we suggest that, in the intramolecular hydride shift, improved solvation of the transition state and of the product state by water is achieved due to minimal polarization and reorientation, and (near) equilibrium solvation.
More Related Videos
Related Concept Videos
 01:33
Radical Reactivity: Intramolecular vs Intermolecular
 01:26
Radical Substitution: Hydrogenolysis of Alkyl Halides with Tributyltin Hydride
The bonds formed in this reaction are stronger than the bonds broken, making it energetically favorable. The reaction follows a radical chain mechanism similar to radical halogenation...
 02:34
Regioselectivity and Stereochemistry of Acid-Catalyzed Hydration
 01:20
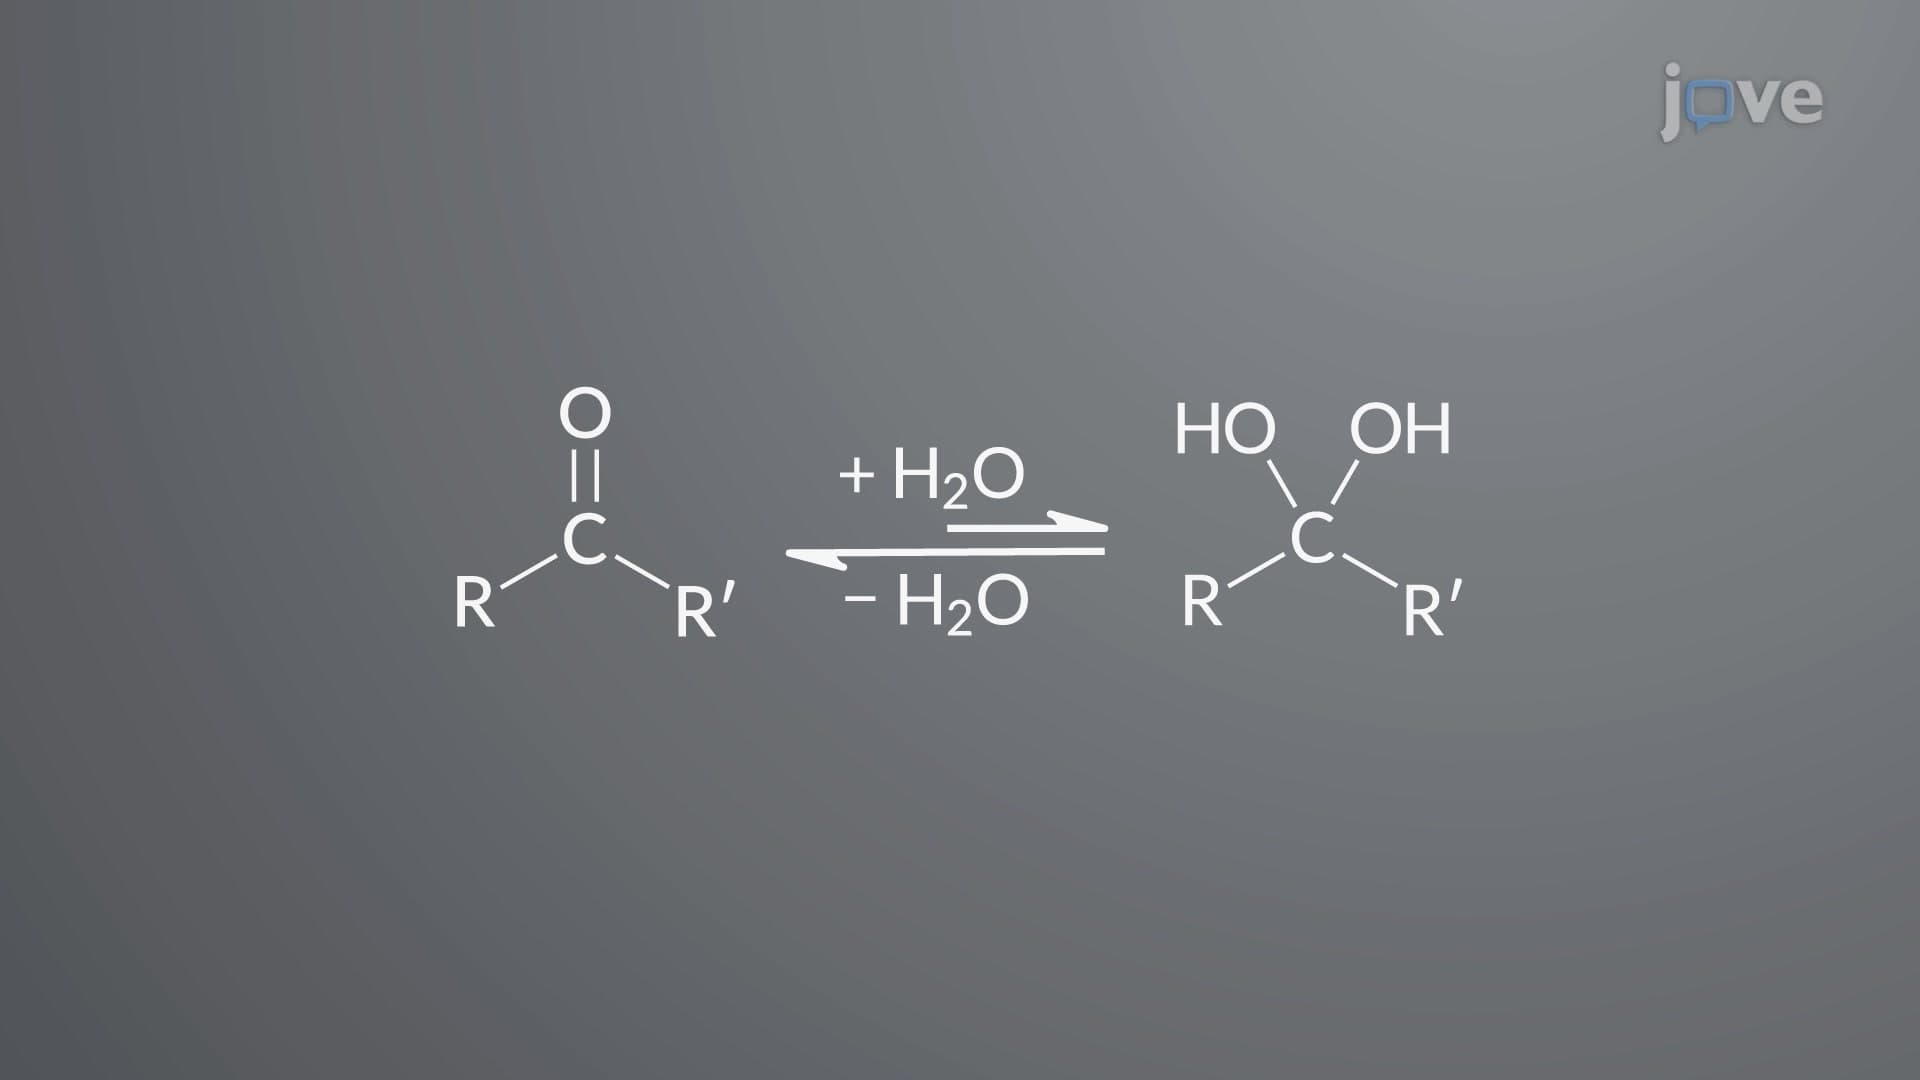
Aldehydes and Ketones with Water: Hydrate Formation
The formation of hydrates is a reversible reaction. Hydrate formation is influenced by steric and electronic factors accompanying the alkyl substituents on the carbonyl group: The rate of hydrate formation increases with a decrease in the number of alkyl groups attached to the carbonyl carbon. Hence,...
 01:19
Acid Halides to Alcohols: LiAlH4 Reduction
The mechanism proceeds in three steps. First, the nucleophilic hydride ion attacks the carbonyl carbon of the acid halide to form a tetrahedral intermediate. Next, the carbonyl group is re-formed, and the halide ion departs as a leaving group, generating an aldehyde. A second nucleophilic attack by the hydride yields an alkoxide ion, which, upon protonation, gives a primary alcohol as...
 02:36
Regioselectivity and Stereochemistry of Hydroboration
Hydroboration proceeds in a concerted fashion with the attack of borane on the π bond, giving a cyclic four-centered transition state. The –BH2 group is bonded to the less substituted carbon and –H to the more substituted carbon. The concerted nature requires the simultaneous addition of –H and –BH2 across the same face of the alkene giving syn...