Related Experiment Video
Updated: Apr 18, 2026

Quantification of Coenzyme A in Cells and Tissues
Published on: September 27, 2019
Mutations in COA3 cause isolated complex IV deficiency associated with neuropathy, exercise intolerance, obesity, and
Elsebet Ostergaard1, Woranontee Weraarpachai2, Kirstine Ravn1
1Department of Clinical Genetics, Copenhagen University Hospital Rigshospitalet, Copenhagen, Denmark.

Background:
We investigated a subject with an isolated cytochrome c oxidase (COX) deficiency presenting with an unusual phenotype characterised by neuropathy, exercise intolerance, obesity, and short stature.
Methods And Results:
Blue-native polyacrylamide gel electrophoresis (BN-PAGE) analysis showed an almost complete lack of COX assembly in subject fibroblasts, consistent with the very low enzymatic activity, and pulse-labelling mitochondrial translation experiments showed a specific decrease in synthesis of the COX1 subunit, the core catalytic subunit that nucleates assembly of the holoenzyme. Whole exome sequencing identified compound heterozygous mutations (c.199dupC, c.215A>G) in COA3, a small inner membrane COX assembly factor, resulting in a pronounced decrease in the steady-state levels of COA3 protein. Retroviral expression of a wild-type COA3 cDNA completely rescued the COX assembly and mitochondrial translation defects, confirming the pathogenicity of the mutations, and resulted in increased steady-state levels of COX1 in control cells, demonstrating a role for COA3 in the stabilisation of this subunit. COA3 exists in an early COX assembly complex that contains COX1 and other COX assembly factors including COX14 (C12orf62), another single pass transmembrane protein that also plays a role in coupling COX1 synthesis with holoenzyme assembly. Immunoblot analysis showed that COX14 was undetectable in COA3 subject fibroblasts, and that COA3 was undetectable in fibroblasts from a COX14 subject, demonstrating the interdependence of these two COX assembly factors.
Conclusions:
The mild clinical course in this patient contrasts with nearly all other cases of severe COX assembly defects that are usually fatal early in life, and underscores the marked tissue-specific involvement in mitochondrial diseases.
Related Concept Videos
 01:43
Electron Transport Chain: Complex III and IV
 01:20
Inborn Errors of Metabolism
 01:41
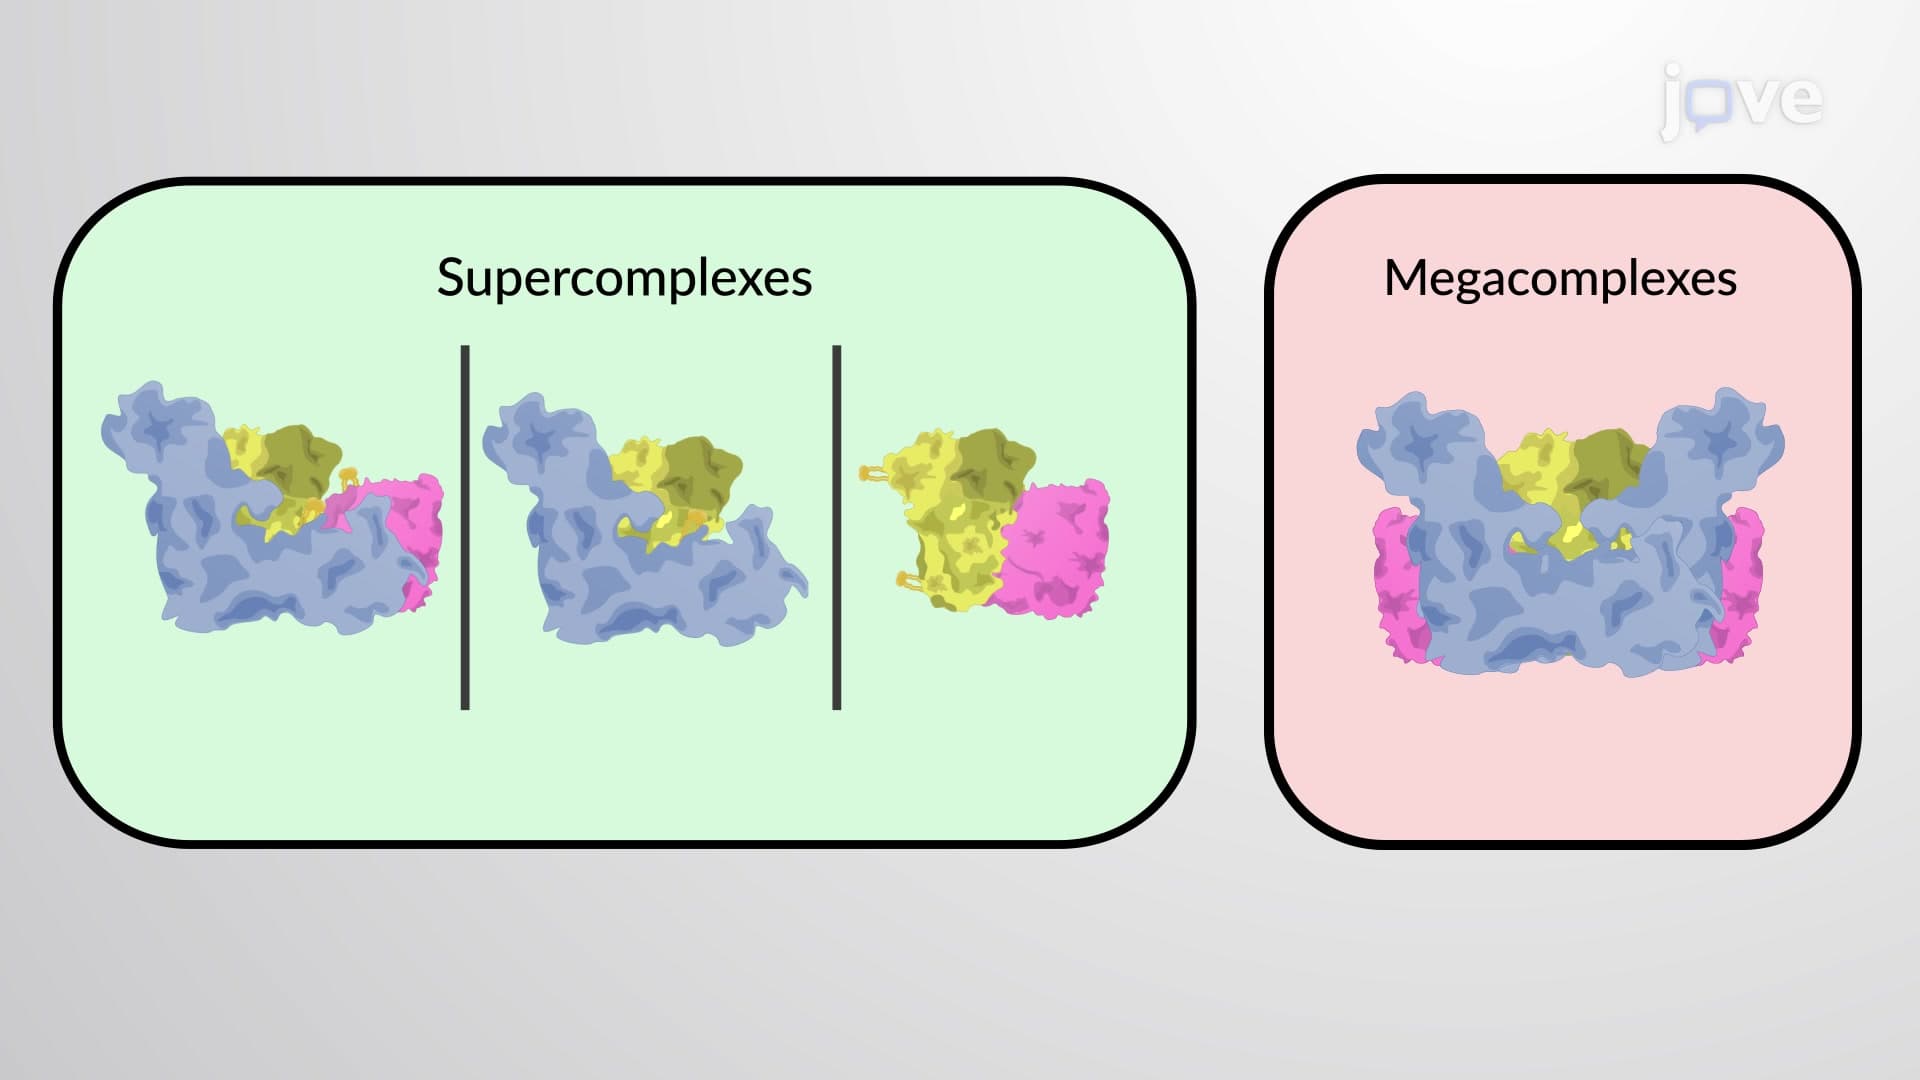
The Supercomplexes in the Crista Membrane
 02:59
Animal Mitochondrial Genetics
 01:48
ATP Synthase: Mechanism
 01:18
Amino Acid Catabolism